Related Experiment Video
Updated: Oct 23, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Pseudospectral implementations of long-range corrected density functional theory
Yixiang Cao1, Mathew D Halls2, Tati Reddy Vadicherla3
1Schrödinger Inc., New York, New York, USA.

Abstract:
We have implemented pseudospectral density-functional theory (DFT) with long-range corrected DFT functionals (PS-LRC) in quantum mechanics package Jaguar, and applied it in the calculations of geometry optimizations, dimmer interaction energies, polarizabilities and first-order hyperpolarizabilities, harmonic vibrational frequencies, S1 and T1 excitation energies, singlet-triplet gaps, charge transfer numbers, oscillator strengths, reaction barrier heights, electron-transfer couplings, and charge-transfer excitation energies. From our accuracy benchmark analysis, PS grids, PS dealiasing functions, PS atomic corrections, PS multigrid strategy, PS length scales, and PS cutoff scheme perform well in PS DFT with LRC density functionals with very small and ignorable deviations when compared to the conventional spectral (CS) method. The timing benchmark study of S1 excitation energy calculations of fullerenes (Cn , n up to 540) demonstrates that PS-LRC achieves 1.4-8.4-fold speedups in SCF, 22-32-fold speedups in Tamm-Dancoff approximation, and 6-15-fold speedups in total wall clock time with an average error 0.004 eV of excitation energies compared to the CS method.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:20
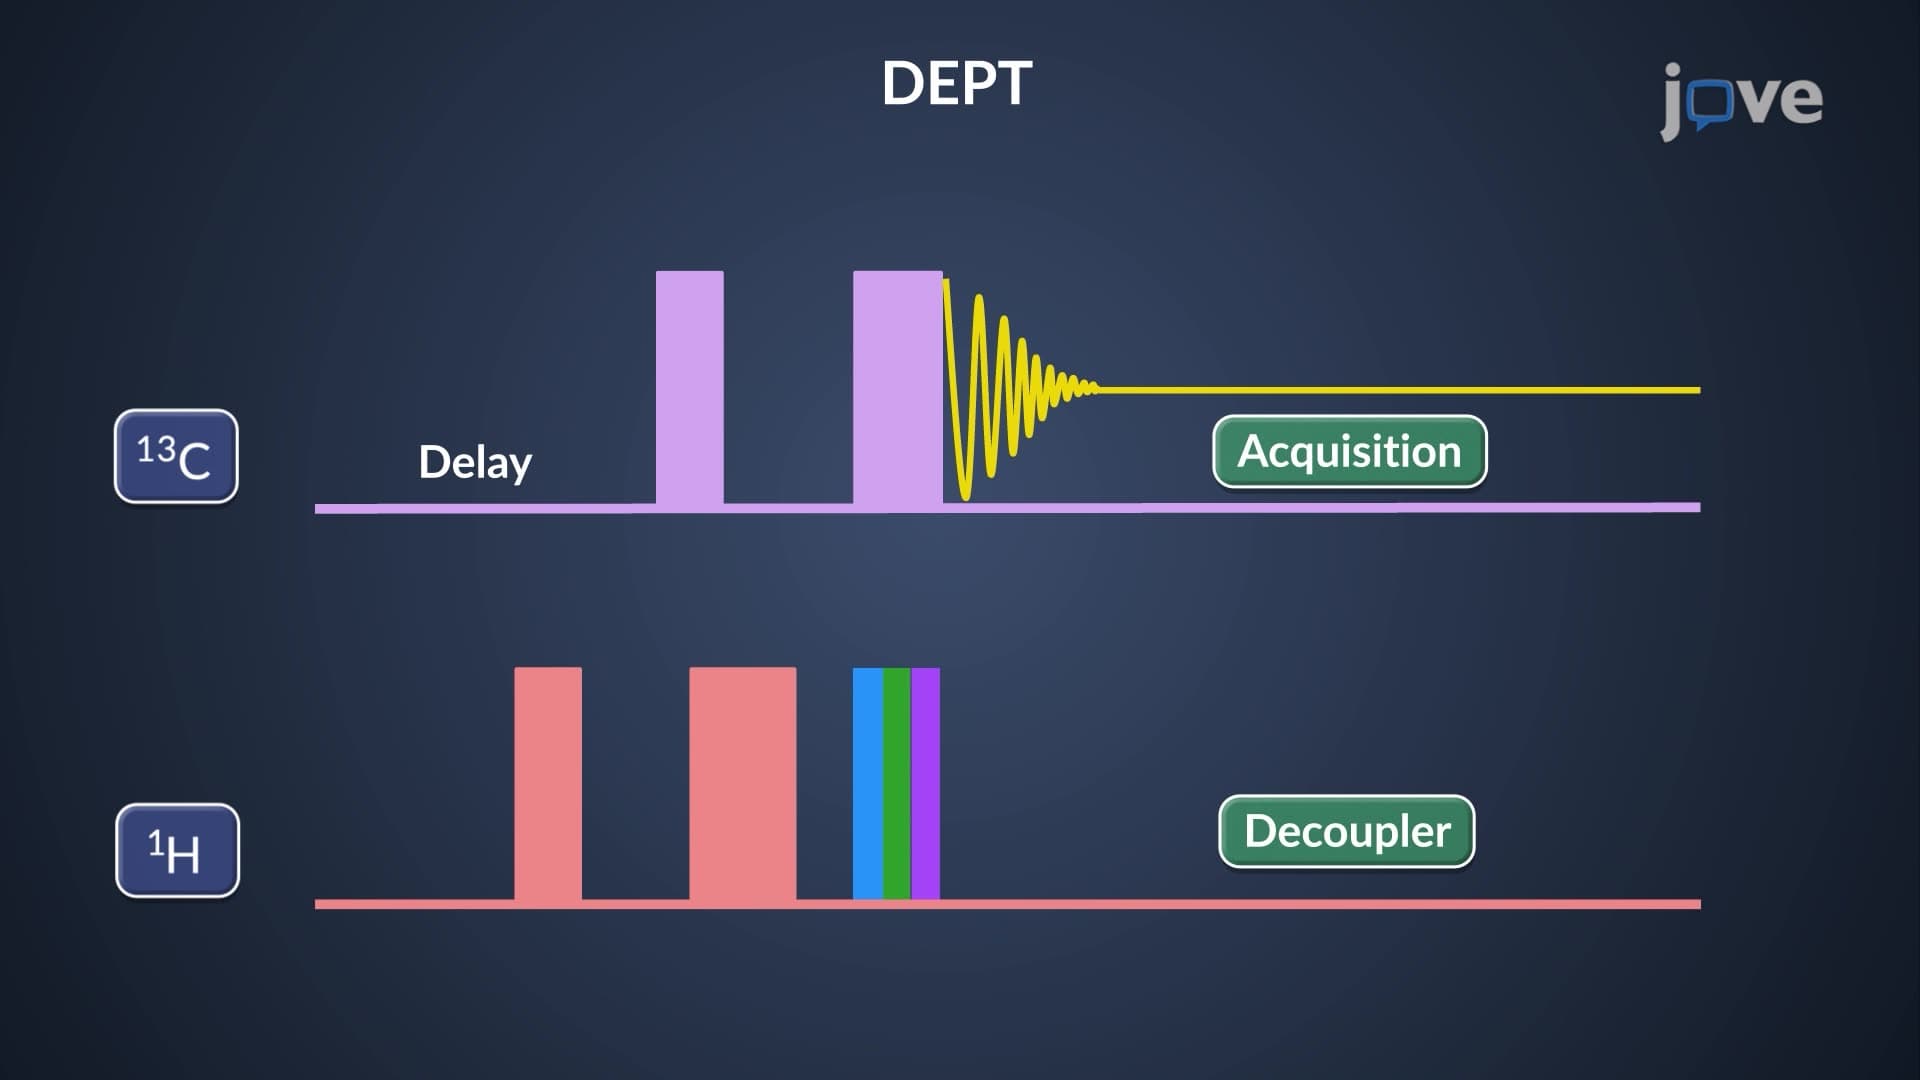
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:10
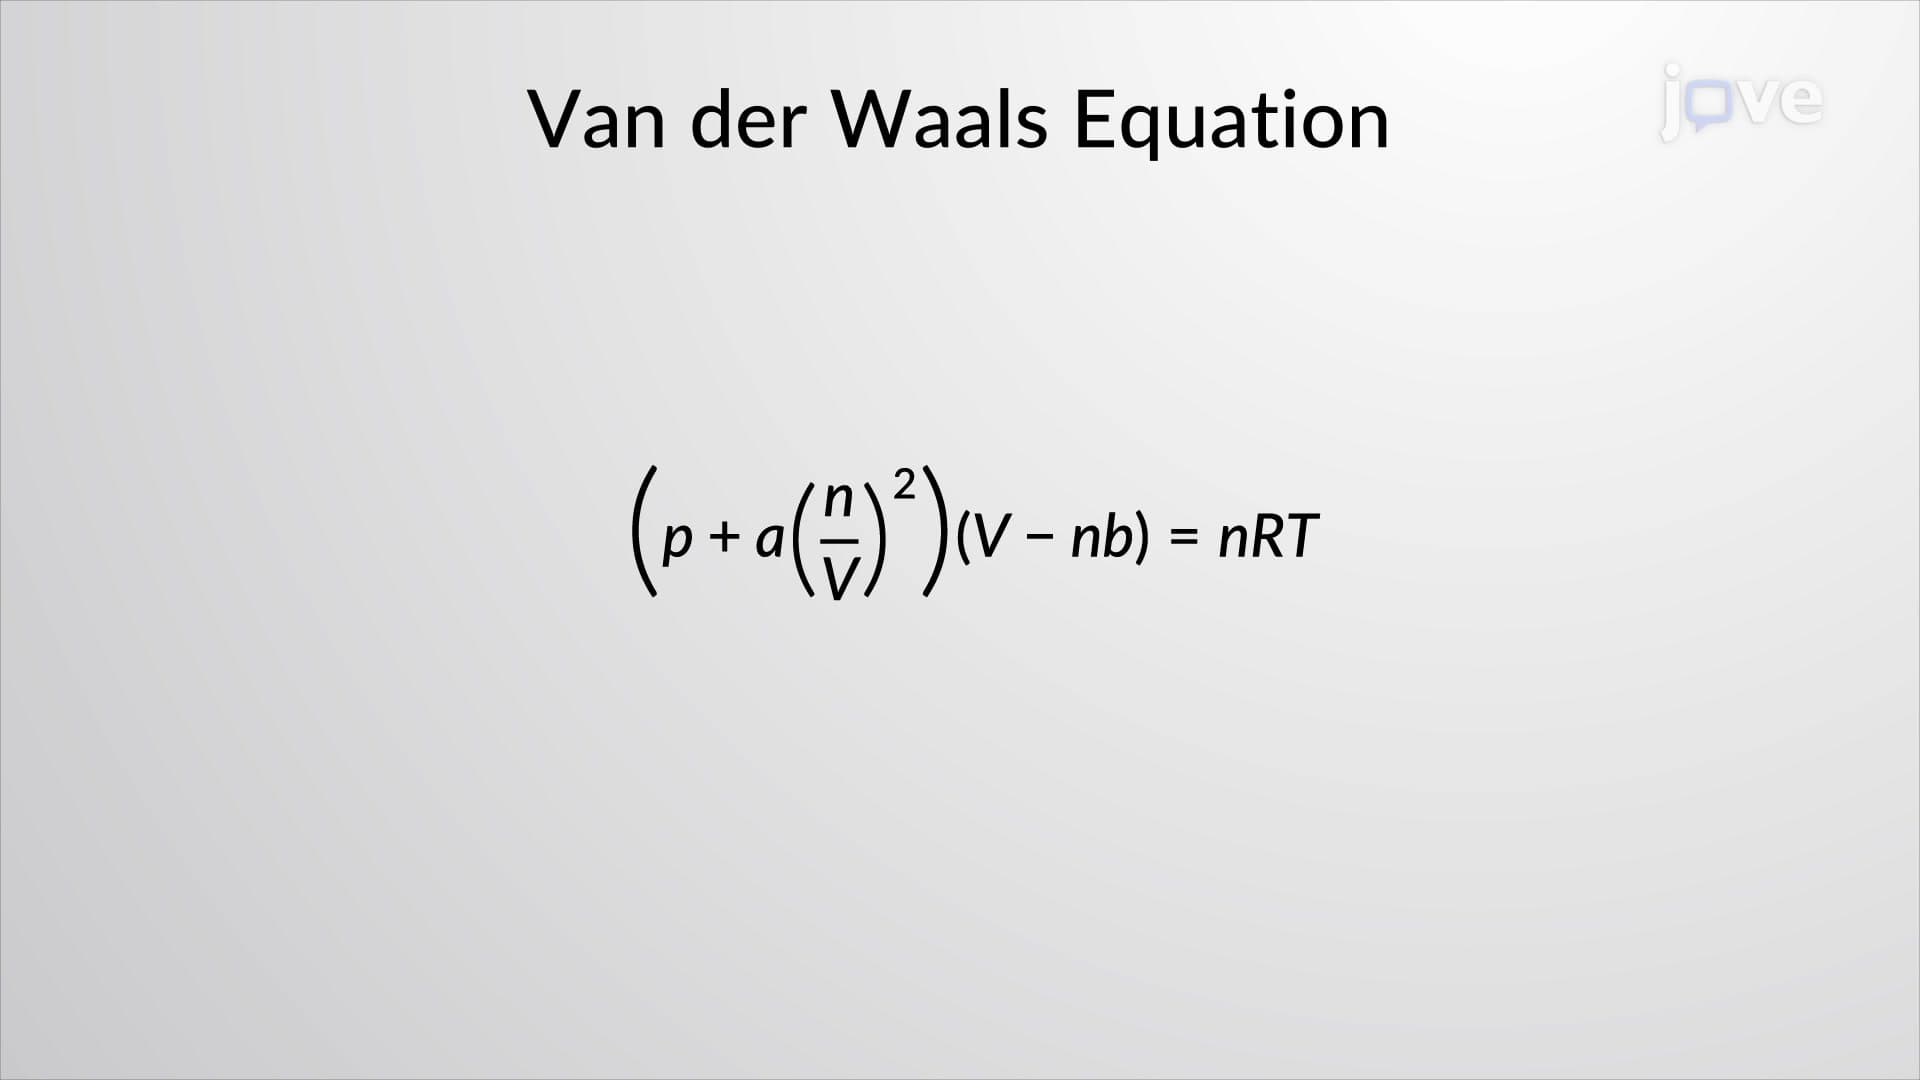
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:26
Linear Approximation in Frequency Domain
In contrast, nonlinear systems do not inherently possess these properties. However, for small deviations around an operating point, a nonlinear system can often be approximated as linear....