Related Experiment Video
Updated: Oct 21, 2025

Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Structure of prion β-oligomers as determined by short-distance crosslinking constraint-guided discrete molecular
Jason J Serpa1, Konstantin I Popov2, Evgeniy V Petrotchenko3,4
1University of Victoria -Genome British Columbia Proteomics Centre, Victoria, British Columbia, Canada.

Abstract:
The conversion of the native monomeric cellular prion protein (PrPC ) into an aggregated pathological β-oligomeric form (PrPβ ) and an infectious form (PrPSc ) is the central element in the development of prion diseases. The structure of the aggregates and the molecular mechanisms of the conformational changes involved in the conversion are still unknown. We applied mass spectrometry combined with chemical crosslinking, hydrogen/deuterium exchange, limited proteolysis, and surface modification for the differential characterization of the native and the urea+acid-converted prion β-oligomer structures to obtain insights into the mechanisms of conversion and aggregation. For the determination of the structure of the monomer and the dimer unit of the β-oligomer, we applied a recently-developed approach for de novo protein structure determination which is based on the incorporation of zero-length and short-distance crosslinking data as intra- and inter-protein constraints in discrete molecular dynamics simulations (CL-DMD). Based on all of the structural-proteomics experimental data and the computationally predicted structures of the monomer units, we propose the potential mode of assembly of the β-oligomer. The proposed β-oligomer assembly provides a clue on the β-sheet nucleation site, and how template-based conversion of the native prion molecule occurs, growth of the prion aggregates, and maturation into fibrils may occur.
Insights
Researchers investigated prion protein (PrP) misfolding in prion diseases. Using advanced structural proteomics, they revealed the assembly mechanism of pathological β-oligomers, offering insights into prion disease progression.
Area of Science:
- Structural Biology
- Neurodegenerative Diseases
- Biochemistry
Background:
- Prion diseases involve the conversion of native cellular prion protein (PrPC) into pathological aggregates (PrPβ, PrPSc).
- The precise structure of these aggregates and the molecular mechanisms driving the conformational changes remain largely unknown.
- Understanding these mechanisms is crucial for deciphering prion disease pathogenesis.
Purpose of the Study:
- To characterize the structural differences between native PrPC and urea+acid-converted PrPβ oligomers.
- To elucidate the molecular mechanisms underlying PrP conversion and aggregation.
- To determine the structure of the monomer and dimer units within the β-oligomer.
Main Methods:
- Applied mass spectrometry combined with chemical crosslinking, hydrogen/deuterium exchange, limited proteolysis, and surface modification.
- Utilized a de novo protein structure determination approach integrating crosslinking data with discrete molecular dynamics (CL-DMD) simulations.
- Determined structures of monomer and dimer units of the β-oligomer.
Main Results:
- Differential structural characterization of native and converted prion β-oligomers was achieved.
- The study determined the structure of the monomer and dimer units of the β-oligomer.
- A potential assembly model for the β-oligomer was proposed based on experimental and computational data.
Conclusions:
- The proposed β-oligomer assembly model offers insights into the β-sheet nucleation site.
- The findings provide clues on how template-based conversion of native prion protein occurs.
- The study sheds light on the growth and maturation of prion aggregates into fibrils.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining,...
 01:22
Protein Folding
 01:24
Protein Organization
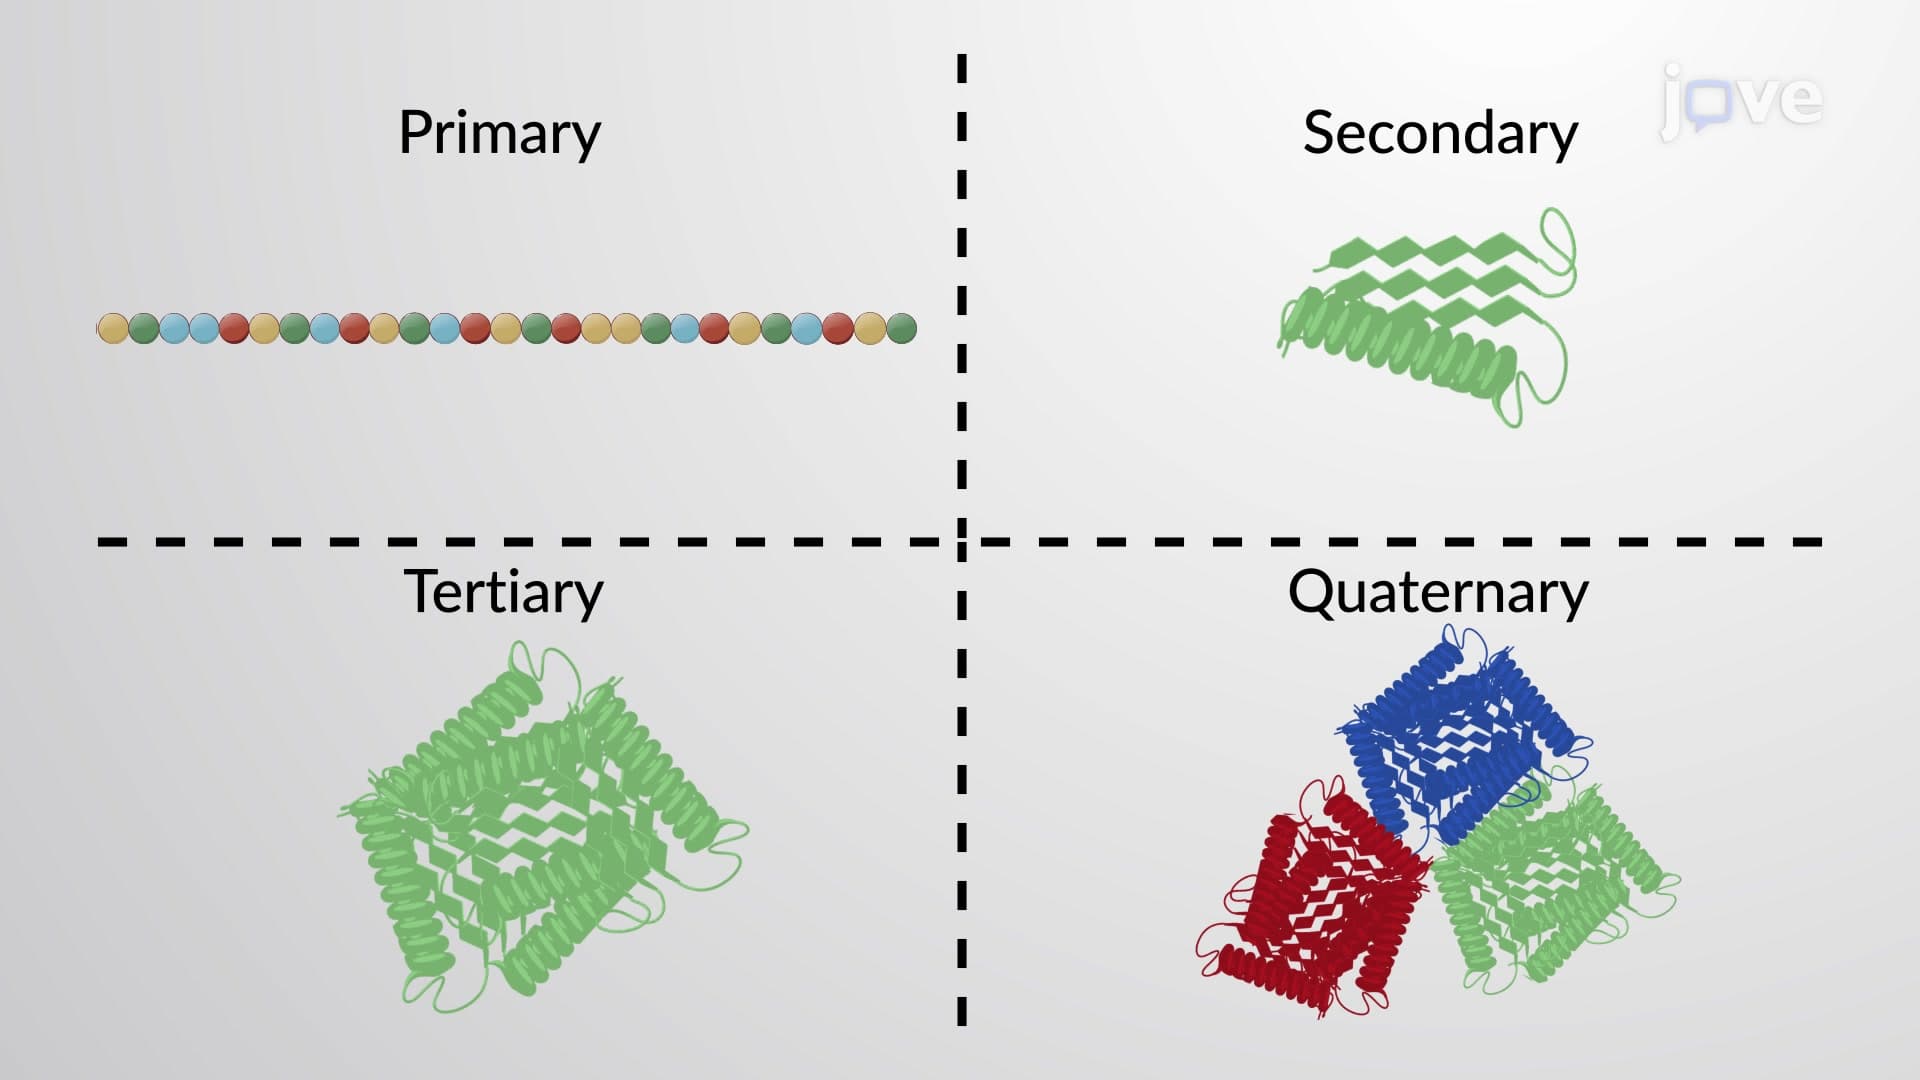
The primary structure of a protein is its amino acid sequence....
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...