
Effect of the Force Field on Molecular Dynamics Simulations of the Multidrug Efflux Protein P-Glycoprotein
1Research School of Chemistry, College of Science, Australian National University, Canberra, ACT 2601, Australia.

Abstract:
Molecular dynamics (MD) simulations have been used extensively to study P-glycoprotein (P-gp), a flexible multidrug transporter that is a key player in the development of multidrug resistance to chemotherapeutics. A substantial body of literature has grown from simulation studies that have employed various simulation conditions and parameters, including AMBER, CHARMM, OPLS, GROMOS, and coarse-grained force fields, drawing conclusions from simulations spanning hundreds of nanoseconds. Each force field is typically parametrized and validated on different data and observables, usually of small molecules and peptides; there have been few comparisons of force field performance on large protein-membrane systems. Here we compare the conformational ensembles of P-gp embedded in a POPC/cholesterol bilayer generated over 500 ns of replicate simulation with five force fields from popular biomolecular families: AMBER 99SB-ILDN, CHARMM 36, OPLS-AA/L, GROMOS 54A7, and MARTINI. We find considerable differences among the ensembles with little conformational overlap, although they correspond to similar extents to structural data obtained from electron paramagnetic resonance and cross-linking studies. Moreover, each trajectory was still sampling new conformations at a high rate after 500 ns of simulation, suggesting the need for more sampling. This work highlights the need to consider known limitations of the force field used (e.g., biases toward certain secondary structures) and the simulation itself (e.g., whether sufficient sampling has been achieved) when interpreting accumulated results of simulation studies of P-gp and other transport proteins.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:16
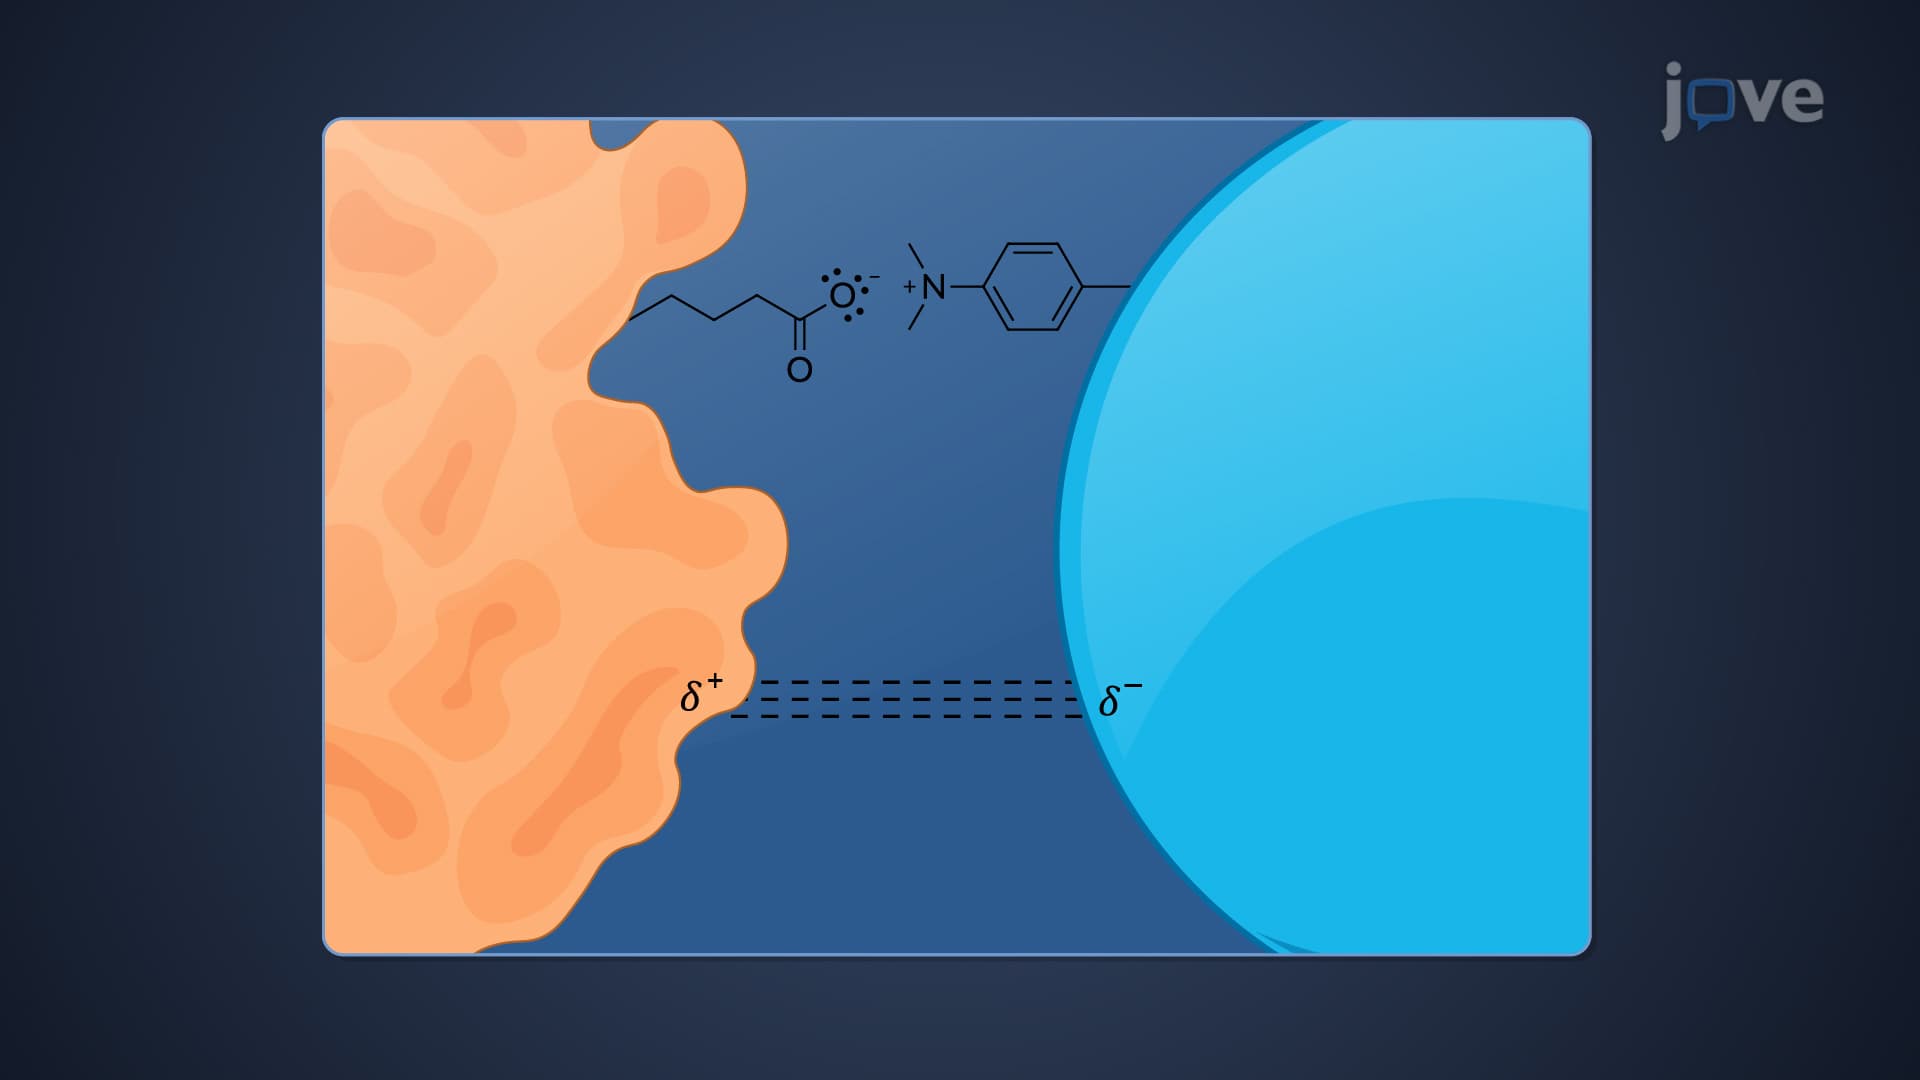
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:06
Carrier-Mediated Transport
Active transport involves two types of membrane-spanning transporters: uptake and efflux. Uptake transporters are expressed in the small...
 01:24
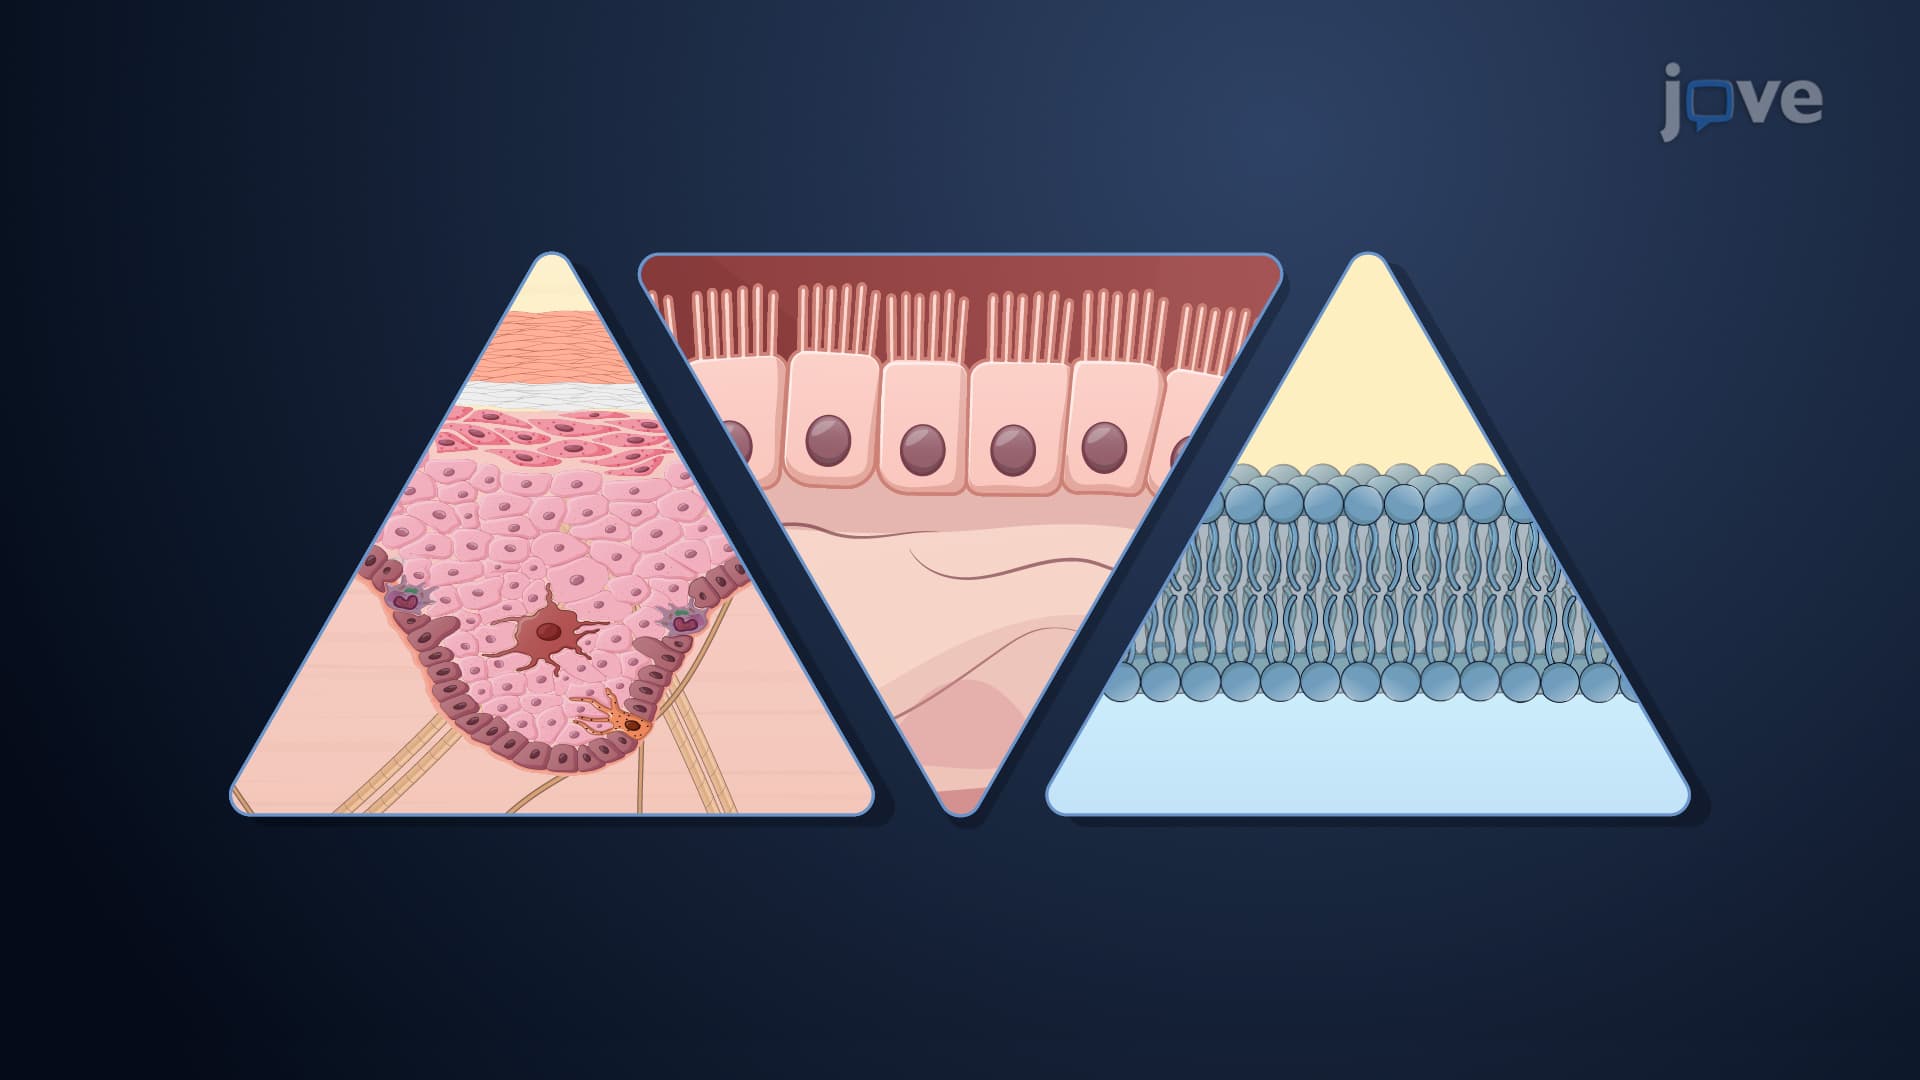
Cellular Membranes and Drug Transport
Phospholipids arrange themselves into a bilayer, with hydrophilic heads oriented outward and hydrophobic tails facing inward.
 01:17
Pore Transport and Ion-Pair Transport
Pore transport, also known as convective transport, is a process where small molecules like urea, water, and sugars rapidly cross cell membranes as though there were channels or pores in the membrane. Although direct microscopic evidence is limited but the concept of pores or channels is widely accepted based on physiological evidence. Despite the lack of direct...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...