Related Experiment Video
Updated: Oct 20, 2025

Author Spotlight: Decoding Mitochondrial Aging
Published on: June 30, 2023
Mitochondrial Abnormalities and Synaptic Damage in Huntington's Disease: a Focus on Defective Mitophagy and
Neha Sawant1, Hallie Morton1, Sudhir Kshirsagar1
1Department of Internal Medicine, Texas Tech University Health Sciences Center, Lubbock, TX, USA.

Insights
Huntington's disease (HD) involves genetic mutations leading to neuron loss. Research shows abnormal mitochondria and impaired removal of damaged mitochondria contribute to HD progression, suggesting new therapeutic targets.
Area of Science:
- Neuroscience
- Genetics
- Cell Biology
Background:
- Huntington's disease (HD) is a fatal genetic disorder characterized by progressive loss of medium spiny neurons (MSN).
- It stems from expanded polyglutamine repeats in the HD gene, causing symptoms like chorea, cognitive decline, and emotional disturbances.
- Cellular dysfunctions implicated in HD include defective axonal transport, altered bioenergetics, and synaptic damage.
Purpose of the Study:
- To review recent research on Huntington's disease progression.
- To discuss advancements in cell and mouse models for HD.
- To explore cellular changes, mitochondrial abnormalities, DNA damage, bioenergetics, oxidative stress, mitophagy, and therapeutic strategies.
Main Methods:
- Review of recent scientific literature on Huntington's disease.
- Analysis of cellular and molecular mechanisms underlying HD.
- Examination of findings from cell and mouse models of HD.
Main Results:
- Mutant huntingtin (mHtt) interacts with dynamin-related protein 1 (DRP1), causing excessive mitochondrial fragmentation and impaired axonal transport.
- Failure to remove damaged mitochondria is an early event in HD progression.
- Abnormal protein interactions and impaired synaptic mitophagy are key pathological features.
Conclusions:
- Mitochondrial dynamics and mitophagy are critical in Huntington's disease pathogenesis.
- Targeting mHtt-DRP1 interactions and enhancing synaptic mitophagy offer promising therapeutic avenues for HD.
- Further research into these cellular mechanisms is crucial for developing effective treatments.
Abstract:
Huntington's disease (HD) is a fatal and pure genetic disease with a progressive loss of medium spiny neurons (MSN). HD is caused by expanded polyglutamine repeats in the exon 1 of HD gene. Clinically, HD is characterized by chorea, seizures, involuntary movements, dystonia, cognitive decline, intellectual impairment, and emotional disturbances. Several years of intense research revealed that multiple cellular changes, including defective axonal transport, protein-protein interactions, defective bioenergetics, calcium dyshomeostasis, NMDAR activation, synaptic damage, mitochondrial abnormalities, and selective loss of medium spiny neurons are implicated in HD. Recent research on mutant huntingtin (mHtt) and mitochondria has found that mHtt interacts with the mitochondrial division protein, dynamin-related protein 1 (DRP1), enhances GTPase DRP1 enzymatic activity, and causes excessive mitochondrial fragmentation and abnormal distribution, leading to defective axonal transport of mitochondria and selective synaptic degeneration. Recent research also revealed that failure to remove dead and/or dying mitochondria is an early event in the disease progression. Currently, efforts are being made to reduce abnormal protein interactions and enhance synaptic mitophagy as therapeutic strategies for HD. The purpose of this article is to discuss recent research in HD progression. This article also discusses recent developments of cell and mouse models, cellular changes, mitochondrial abnormalities, DNA damage, bioenergetics, oxidative stress, mitophagy, and therapeutics strategies in HD.
More Related Videos
Related Concept Videos
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:19
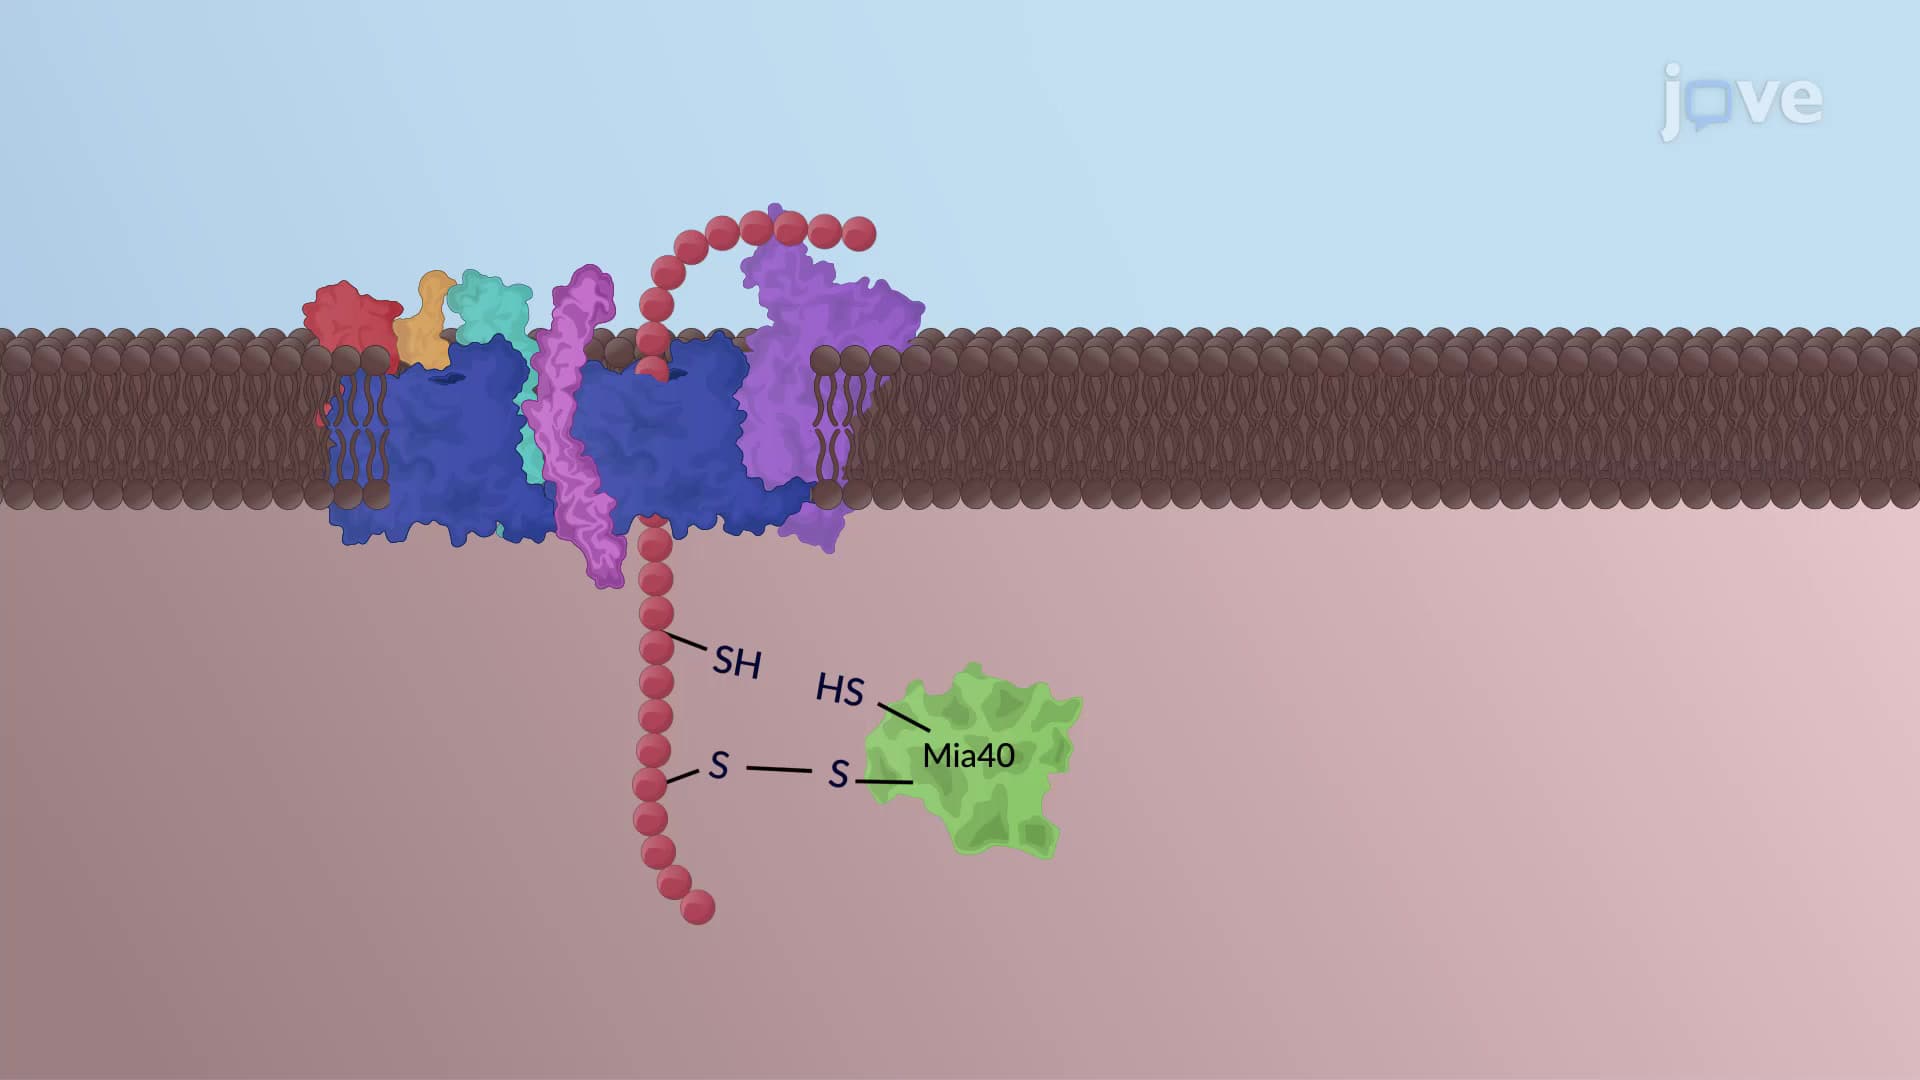
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:22
Lysosomal Hydrolases
 01:37
Mitochondria
 01:48
ATP Synthase: Mechanism
 01:45
Mitochondrial Membranes