Related Experiment Video
Updated: Oct 20, 2025
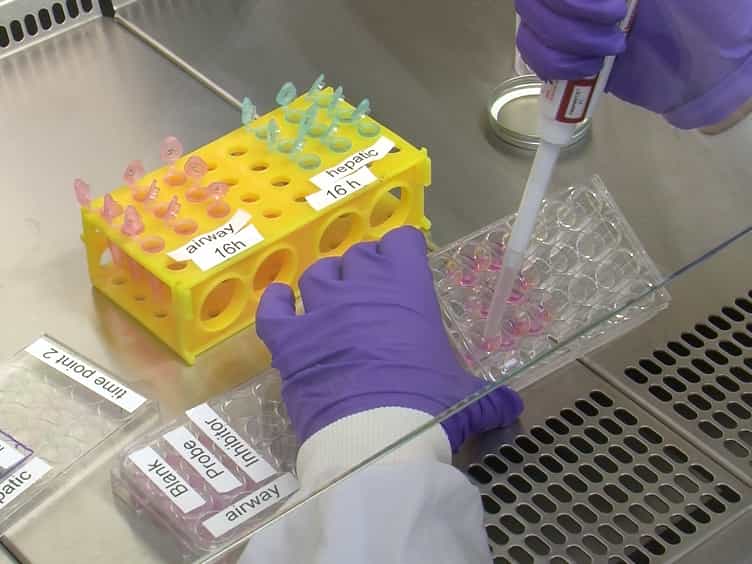

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
The broad phenotypic spectrum of 17α-hydroxylase/17,20-lyase (CYP17A1) deficiency: a case series
Min Sun1,2, Jonathan W Mueller1,2, Lorna C Gilligan1,2
1Institute of Metabolism and Systems Research, College of Medical and Dental Sciences, University of Birmingham, Birmingham, UK.

Context:
17α-Hydroxylase/17,20-lyase deficiency (17OHD) caused by mutations in the CYP17A1 gene is a rare form of congenital adrenal hyperplasia typically characterised by cortisol deficiency, mineralocorticoid excess and sex steroid deficiency.
Objective:
To examine the phenotypic spectrum of 17OHD by clinical and biochemical assessment and corresponding in silico and in vitro functional analysis.
Design:
Case series.
Patients And Results:
We assessed eight patients with 17OHD, including four with extreme 17OHD phenotypes: two siblings presented with failure to thrive in early infancy and two with isolated sex steroid deficiency and normal cortisol reserve. Diagnosis was established by mass spectrometry-based urinary steroid profiling and confirmed by genetic CYP17A1 analysis, revealing homozygous and compound heterozygous sequence variants. We found novel (p.Gly111Val, p.Ala398Glu, p.Ile371Thr) and previously described sequence variants (p.Pro409Leu, p.Arg347His, p.Gly436Arg, p.Phe53/54del, p.Tyr60IlefsLys88X). In vitro functional studies employing an overexpression system in HEK293 cells showed that 17,20-lyase activity was invariably decreased while mutant 17α-hydroxylase activity retained up to 14% of WT activity in the two patients with intact cortisol reserve. A ratio of urinary corticosterone over cortisol metabolites reflective of 17α-hydroxylase activity correlated well with clinical phenotype severity.
Conclusion:
Our findings illustrate the broad phenotypic spectrum of 17OHD. Isolated sex steroid deficiency with normal stimulated cortisol has not been reported before. Attenuation of 17α-hydroxylase activity is readily detected by urinary steroid profiling and predicts phenotype severity.
Significance Statement:
Here we report, supported by careful phenotyping, genotyping and functional analysis, a prismatic case series of patients with congenital adrenal hyperplasia due to 17α-hydroxylase (CYP17A1) deficiency (17OHD). These range in severity from the abolition of function, presenting in early infancy, and unusually mild with isolated sex steroid deficiency but normal ACTH-stimulated cortisol in adult patients. These findings will guide improved diagnostic detection of CYP17A1 deficiency.

Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:35
Pedigree Analysis
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:27
Adrenal Gland Disorders
Adrenal insufficiency, characterized by insufficient cortisol and aldosterone production, leads to conditions like Addison's disease. This disorder, affecting the adrenal cortex, exhibits symptoms such as skin bronzing, dehydration, low blood pressure, fatigue, and weight loss. Congenital adrenal hyperplasia, a genetic ailment causing...
 01:33
Pleiotropy