Related Experiment Video
Updated: Oct 19, 2025

Detection of Glycosaminoglycans by Polyacrylamide Gel Electrophoresis and Silver Staining
Published on: February 25, 2021
Congenital Disorders of Deficiency in Glycosaminoglycan Biosynthesis
Shuji Mizumoto1, Shuhei Yamada1
1Department of Pathobiochemistry, Faculty of Pharmacy, Meijo University, Nagoya, Japan.

Abstract:
Glycosaminoglycans (GAGs) including chondroitin sulfate, dermatan sulfate, and heparan sulfate are covalently attached to specific core proteins to form proteoglycans, which are distributed at the cell surface as well as in the extracellular matrix. Proteoglycans and GAGs have been demonstrated to exhibit a variety of physiological functions such as construction of the extracellular matrix, tissue development, and cell signaling through interactions with extracellular matrix components, morphogens, cytokines, and growth factors. Not only connective tissue disorders including skeletal dysplasia, chondrodysplasia, multiple exostoses, and Ehlers-Danlos syndrome, but also heart and kidney defects, immune deficiencies, and neurological abnormalities have been shown to be caused by defects in GAGs as well as core proteins of proteoglycans. These findings indicate that GAGs and proteoglycans are essential for human development in major organs. The glycobiological aspects of congenital disorders caused by defects in GAG-biosynthetic enzymes including specific glysocyltransferases, epimerases, and sulfotransferases, in addition to core proteins of proteoglycans will be comprehensively discussed based on the literature to date.
Related Concept Videos
 01:05
Proteoglycans
 01:23
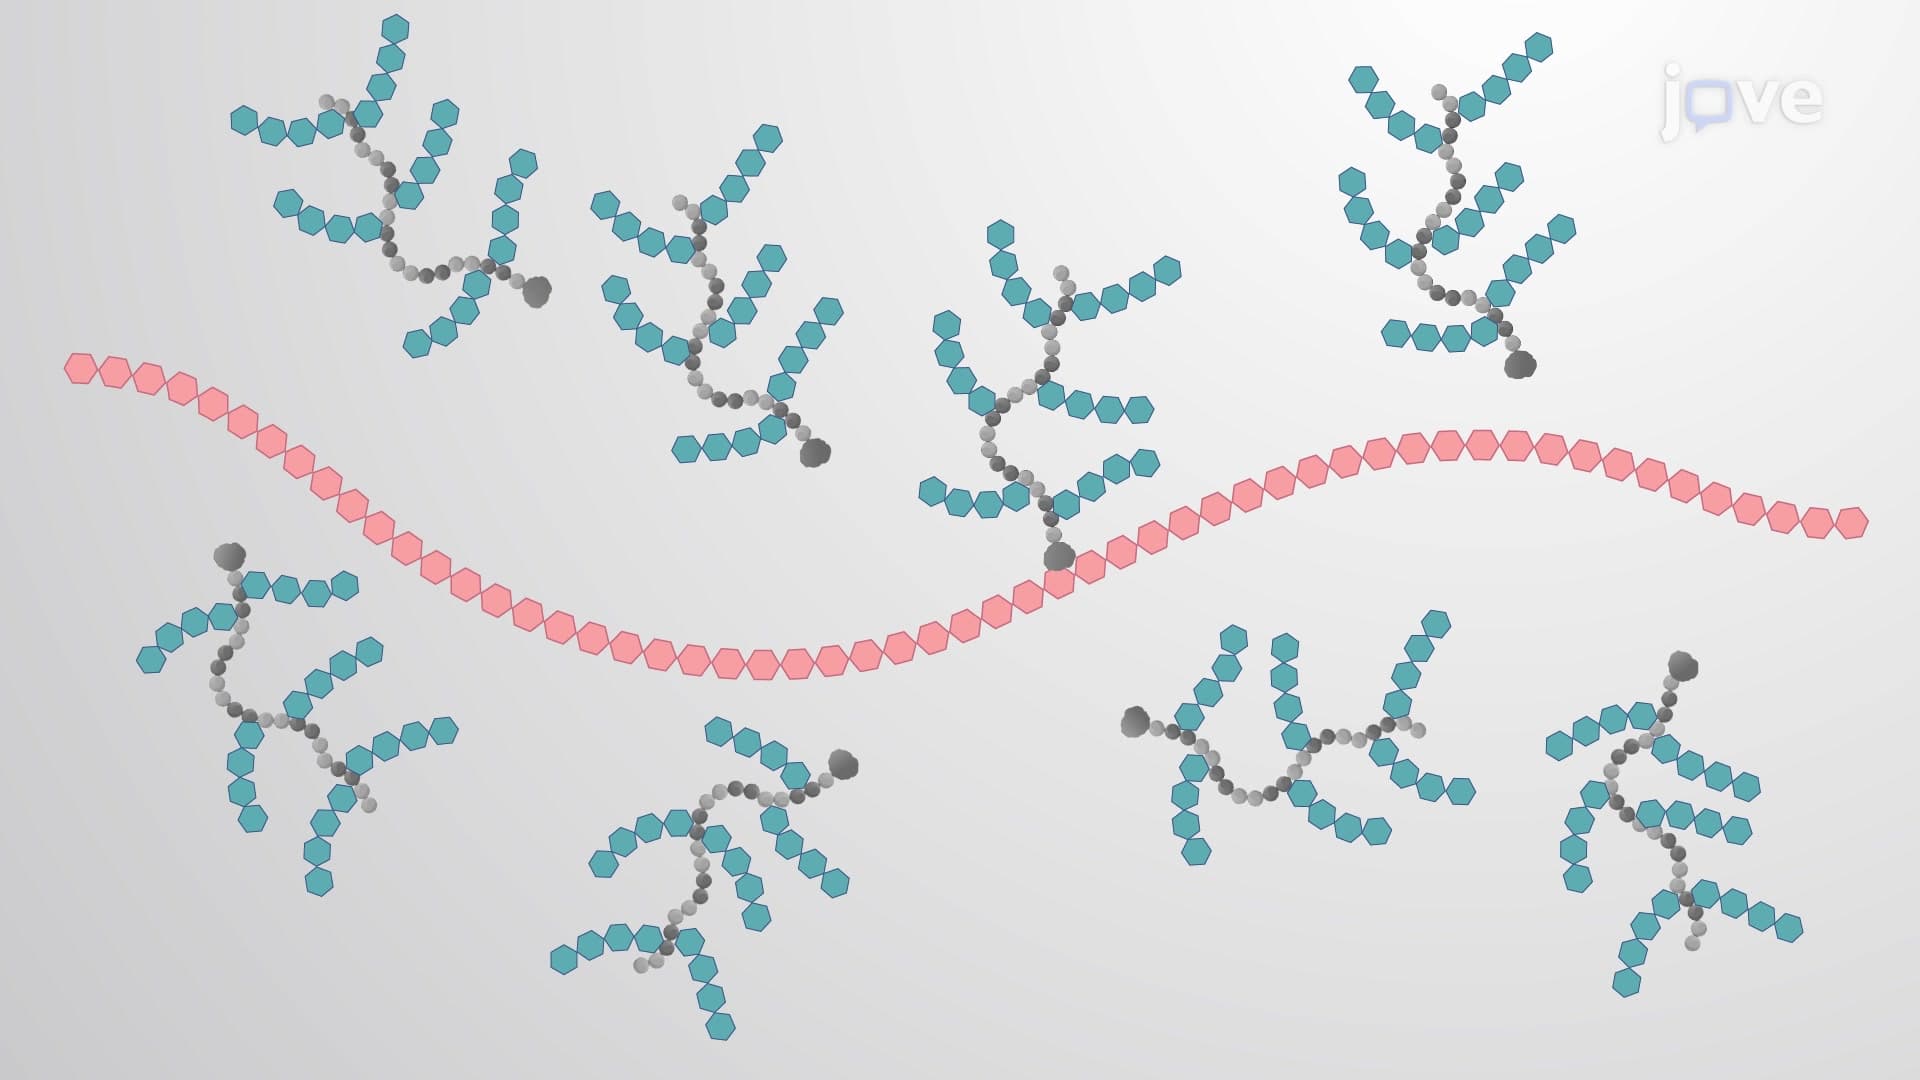
Glycosaminoglycans
GAGS are found in the extracellular matrix of vertebrates, invertebrates, and bacteria. Due to their polar nature they attract water, and serve as excellent lubricants or shock absorbers in an animal body.
Hyaluronic...
 01:20
Inborn Errors of Metabolism
 01:22
Lysosomal Hydrolases
 01:24
Oligosaccharide Assembly
Multiple sugar molecules that may or may...
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...