A phosphine-free Mn(I)-NNS catalyst for asymmetric transfer hydrogenation of acetophenone: a theoretical prediction
Yaqi Zhao1, Lin Zhang1, Min Pu1
1State Key Laboratory of Chemical Resource Engineering, Institute of Computational Chemistry, College of Chemistry, Beijing University of Chemical Technology, Beijing, 100029, China. leim@mail.buct.edu.cn.

Abstract:
The density functional theory (DFT) method was employed to investigate the reaction mechanism of the hydrogen activation and asymmetric transfer hydrogenation (ATH) of acetophenone catalyzed by a well-defined phosphine-free Mn(I)-NNS complex. The calculation results indicate that the Mn-NNS complex has potential high catalytic hydrogenation activity. Meanwhile, the hydrogen transfer step of this reaction is proposed to be a concerted but asynchronous process, and the hydride transfer precedes proton transfer. This work also pointed out that the stereoselectivity of ATH catalyzed by the Mn(I)-NNS complex mainly originates from the noncovalent interaction between the substrate and the catalyst. Additionally, the catalytic activities of Mn-NNS complexes with different X ligands (X = CO, Cl, H, OMe, NCMe, CCMe, and CHCHMe) were compared, and the calculated total reaction energy barriers were all viable, which indicates that these Mn-NNS complexes show higher CO bond hydrogenation activity under mild conditions. This theoretical study predicts that the reactions catalyzed by complexes with H and NCMe ligands exhibit high stereoselectivity with enantiomeric excess (ee) values of 97% and 93%, respectively.
Related Concept Videos
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
 01:28
Preparation of 1° Amines: Gabriel Synthesis
Strong bases like NaOH or KOH deprotonate the phthalimide to form the corresponding anion, which acts as a nucleophile. Further, the anion attacks an...
 02:34
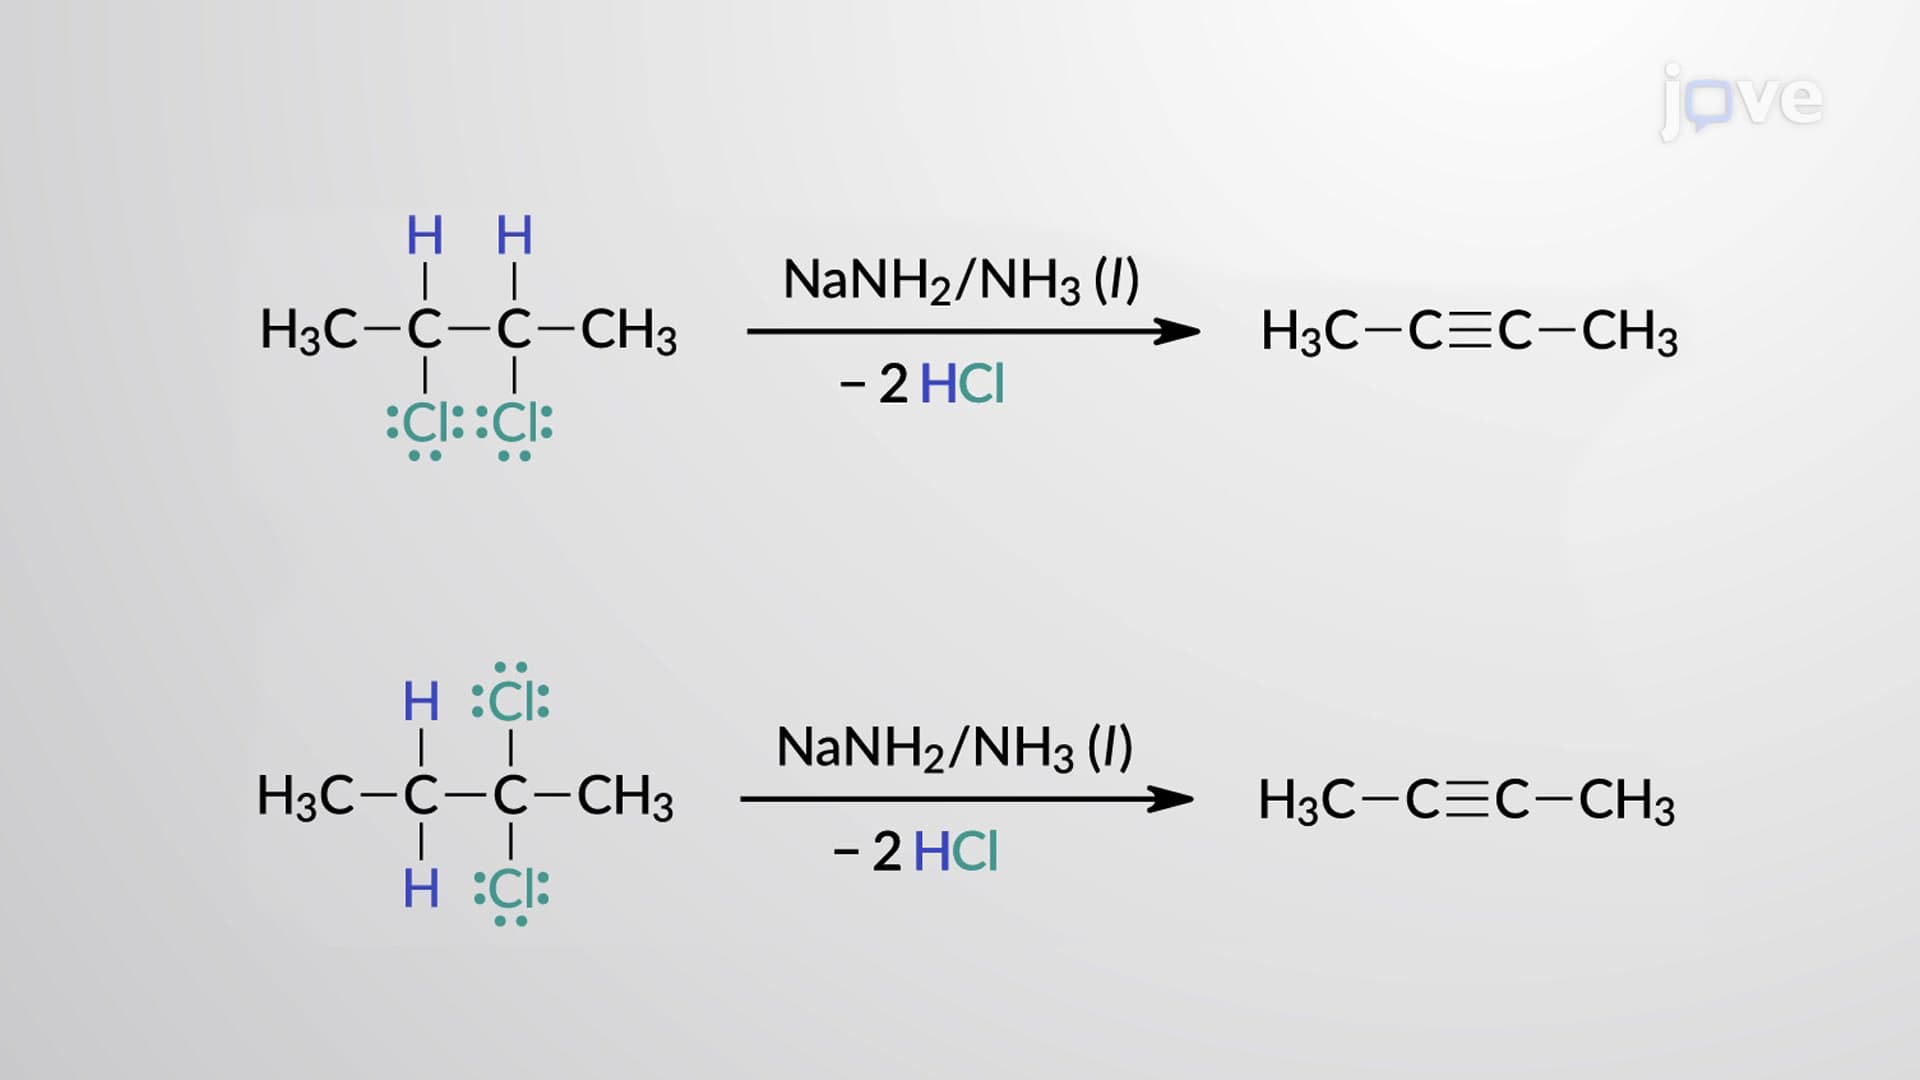
Preparation of Alkynes: Dehydrohalogenation
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.