Related Experiment Video
Updated: Oct 17, 2025
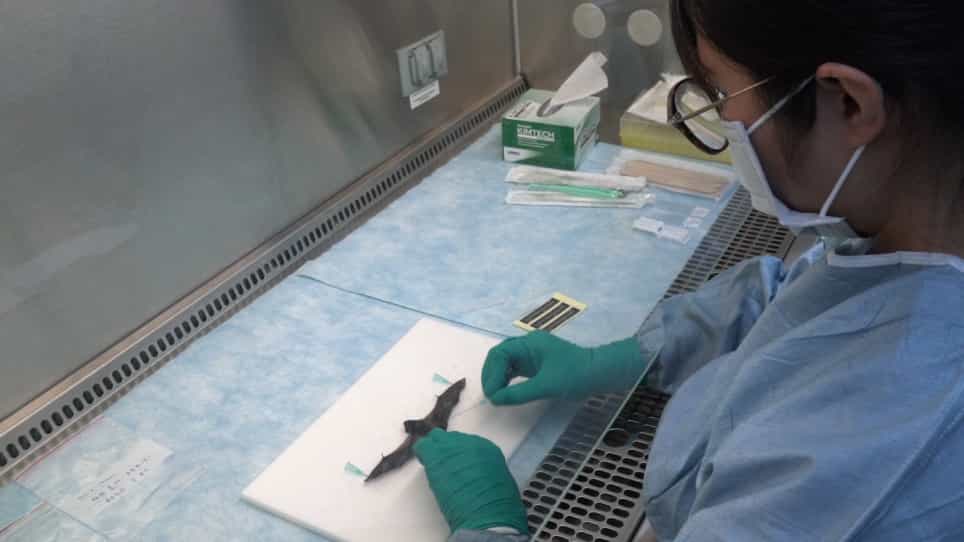

Standard Operating Procedure for Lyssavirus Surveillance of the Bat Population in Taiwan
Published on: August 27, 2019
Host genetic factors associated with the range limit of a European hantavirus
Moritz Saxenhofer1,2, Anton Labutin1, Thomas A White1
1Institute of Ecology and Evolution, University of Bern, Bern, Switzerland.

Abstract:
The natural host ranges of many viruses are restricted to very specific taxa. Little is known about the molecular barriers between species that lead to the establishment of this restriction or generally prevent virus emergence in new hosts. Here, we identify genomic polymorphisms in a natural rodent host associated with a strong genetic barrier to the transmission of European Tula orthohantavirus (TULV). We analysed the very abrupt spatial transition between two major phylogenetic clades in TULV across the comparatively much wider natural hybrid zone between evolutionary lineages of their reservoir host, the common vole (Microtus arvalis). Genomic scans of 79,225 single nucleotide polymorphisms (SNPs) in 323 TULV-infected host individuals detected 30 SNPs that were consistently associated with the TULV clades CEN.S or EST.S in two replicate sampling transects. Focusing the analysis on 199 voles with evidence of genomic admixture at the individual level (0.1-0.9) supported statistical significance for all 30 loci. Host genomic variation at these SNPs explained up to 37.6% of clade-specific TULV infections. Genes in the vicinity of associated SNPs include SAHH, ITCH and two members of the Syngr gene family, which are involved in functions related to immune response or membrane transport. This study demonstrates the relevance of natural hybrid zones as systems not only for studying processes of evolutionary divergence and speciation, but also for the detection of evolving genetic barriers for specialized parasites.
More Related Videos
 10:33
Multi-locus Variable-number Tandem-repeat Analysis of the Fish-pathogenic Bacterium Yersinia ruckeri by Multiplex PCR and Capillary Electrophoresis
Published on: June 17, 2019
 07:15
Determining the Likelihood of Variant Pathogenicity Using Amino Acid-level Signal-to-Noise Analysis of Genetic Variation
Published on: January 16, 2019
Related Concept Videos
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:43
Incomplete Dominance
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:11
Genetic Lingo
 01:09
Mutation, Gene Flow, and Genetic Drift