Related Experiment Video
Updated: Oct 16, 2025
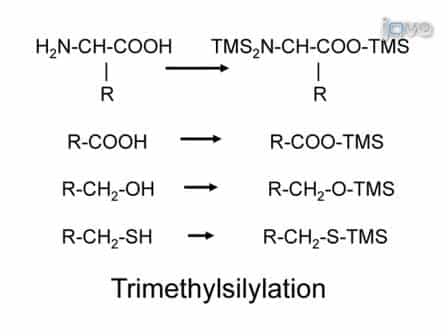

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Hyperammonemia in Inherited Metabolic Diseases
Graziela Schmitt Ribas1,2, Franciele Fátima Lopes3, Marion Deon3
1Departamento de Análises Clínicas, Faculdade de Farmácia, Universidade Federal do Rio Grande do Sul, Porto Alegre, RS, Brazil. grazielaribas@yahoo.com.br.

Abstract:
Ammonia is a neurotoxic compound which is detoxified through liver enzymes from urea cycle. Several inherited or acquired conditions can elevate ammonia concentrations in blood, causing severe damage to the central nervous system due to the toxic effects exerted by ammonia on the astrocytes. Therefore, hyperammonemic patients present potentially life-threatening neuropsychiatric symptoms, whose severity is related with the hyperammonemia magnitude and duration, as well as the brain maturation stage. Inherited metabolic diseases caused by enzymatic defects that compromise directly or indirectly the urea cycle activity are the main cause of hyperammonemia in the neonatal period. These diseases are mainly represented by the congenital defects of urea cycle, classical organic acidurias, and the defects of mitochondrial fatty acids oxidation, with hyperammonemia being more severe and frequent in the first two groups mentioned. An effective and rapid treatment of hyperammonemia is crucial to prevent irreversible neurological damage and it depends on the understanding of the pathophysiology of the diseases, as well as of the available therapeutic approaches. In this review, the mechanisms underlying the hyperammonemia and neurological dysfunction in urea cycle disorders, organic acidurias, and fatty acids oxidation defects, as well as the therapeutic strategies for the ammonia control will be discussed.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:23
Urea Cycle
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:22
Lysosomal Hydrolases
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:18
Amino Acid Catabolism