Related Experiment Video
Updated: Oct 13, 2025

In Situ SIMS and IR Spectroscopy of Well-defined Surfaces Prepared by Soft Landing of Mass-selected Ions
Published on: June 16, 2014
The size-dependent influence of palladium doping on the structures of cationic gold clusters
Piero Ferrari1, Laia Delgado-Callico2, Olga V Lushchikova3
1Quantum Solid-State Physics, Department of Physics and Astronomy, KU Leuven Leuven Belgium piero.ferrari@kuleuven.be.

Abstract:
The physicochemical properties of small metal clusters strongly depend on their precise geometry. Determining such geometries, however, is challenging, particularly for clusters formed by multiple elements. In this work, we combine infrared multiple photon dissociation spectroscopy and density functional theory calculations to investigate the lowest-energy structures of Pd doped gold clusters, PdAu + (n ≤ 10). The high-quality experimental spectra allow for an unambiguous determination of the structures adopted by the clusters. Our results show that the Pd-Au interaction is so large that the structures of PdAu + and Au + are very different. Pd doping induces a 2D to 3D transition at much smaller cluster sizes than for pure Au + clusters. PdAu + clusters are three-dimensional from n = 4, whereas for Au + this transition only takes place at n = 7. Despite the strong Au-Pd interaction, the Au + cluster geometries remain recognizable in PdAu + up to n = 7. This is particularly clear for PdAu6 +. In PdAu8 + and PdAu9 +, Pd triggers major rearrangements of the Au clusters, which adopt pyramidal shapes. For PdAu4 + we find a geometry that was not considered in previous studies, and the geometry found for PdAu8 + does not correspond to the lowest-energy structure predicted by DFT, suggesting kinetic trapping during formation. This work demonstrates that even with the continuous improvement of computational methods, unambiguous assignment of cluster geometries still requires a synergistic approach, combining experiment and computational modelling.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:57
Coordination Number and Geometry
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:09
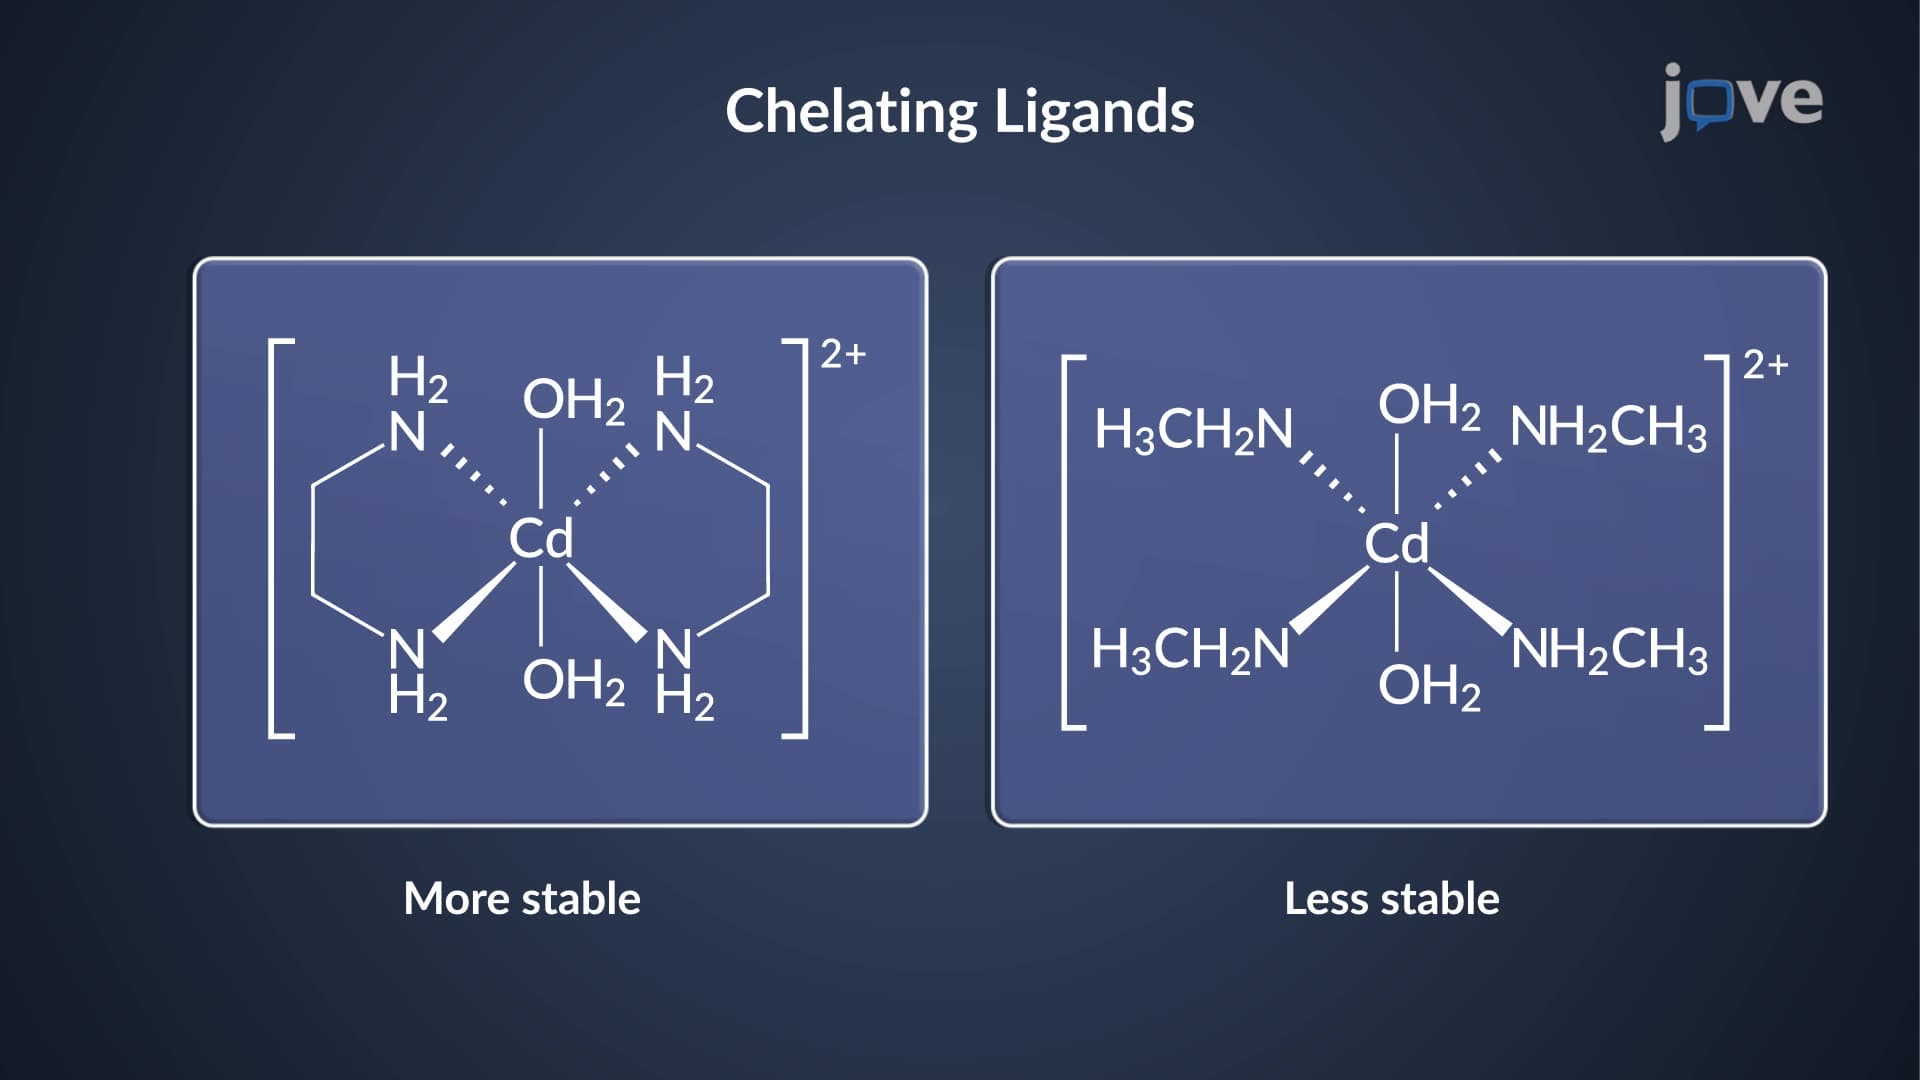
Complexation Equilibria: Factors Influencing Stability of Complexes