Related Experiment Video
Updated: Oct 13, 2025
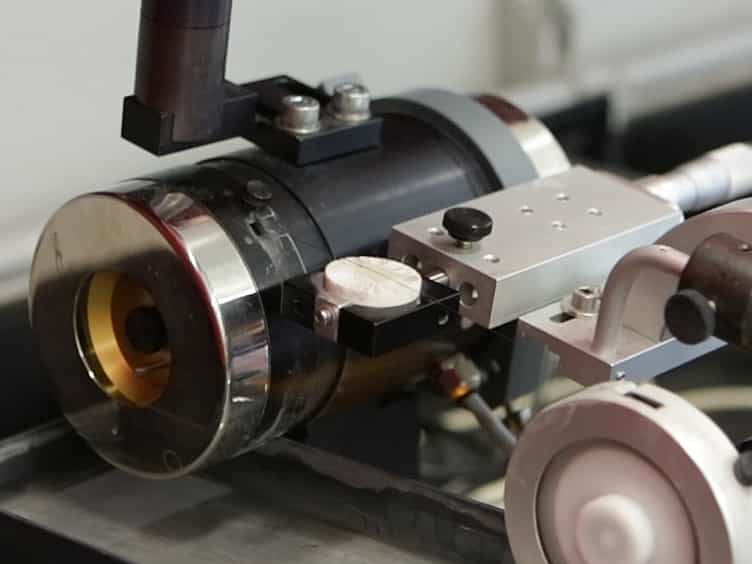
Spark Plasma Sintering Apparatus Used for the Formation of Strontium Titanate Bicrystals
Published on: February 9, 2017
Band Structure Engineering and Optical Properties of Pristine and Doped Monoclinic Zirconia (m-ZrO2): Density
El-Sayed R Khattab1, Sayed S Abd El Rehim1, Walid M I Hassan2
1Department of Chemistry, Faculty of Science, Ain-Shams University, Cairo 11566, Egypt.

Abstract:
Recently, monoclinic ZrO2 has received great technological importance because of its remarkable dielectric properties, high chemical stability, and high melting point. Herein, we introduce first-principles calculations using the Hubbard approach (DFT + U) to study the effects of doping with Nb and W on the electronic and optical properties of pristine ZrO2. The introduction of dopant atoms into the pristine crystal structure led to the displacement of the bandgap edges and reallocation of the Fermi level. The valence band maximum (VBM) shifted upward, resulting in band gap tightening from 5.79 to 0.89 for ZrO2: Nb and to 1.33 eV for ZrO2: W. The optical absorption of doped crystals extended into the visible and near-infrared regions. Partial density of states (PDOS) calculations showed valence band dependency on the O 2p orbital energy, with the conduction band predominantly composed of Nb 4d and W 5d. For pristine ZrO2, the results obtained for the imaginary and real parts of the dielectric function, the refractive index, and the reflectivity show good agreement with the available experimental and theoretical results. For ZrO2:W, we checked the dopant location effect, and the obtained results showed no significant effect on the calculated values of the band gap with a maximum difference of 0.17 eV. Significant band gap tightening and optical properties of our systems indicate that these systems could be promising candidates for photoelectrochemical energy conversion (PEC) applications.
More Related Videos
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:22
Structures of Solids
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:33
Lattice Centering and Coordination Number
Types of Unit Cells
Imagine taking a large number of identical...