Related Experiment Video
Updated: Oct 13, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Dipole Moment Calculations Using Multiconfiguration Pair-Density Functional Theory and Hybrid Multiconfiguration
Aleksandr O Lykhin1, Donald G Truhlar2, Laura Gagliardi1,3
1Department of Chemistry, Pritzker School of Molecular Engineering, The James Franck Institute and Chicago Center for Theoretical Chemistry, The University of Chicago, Chicago, Illinois 60637, United States.

Abstract:
The dipole moment is the molecular property that most directly indicates molecular polarity. The accuracy of computed dipole moments depends strongly on the quality of the calculated electron density, and the breakdown of single-reference methods for strongly correlated systems can lead to poor predictions of the dipole moments in those cases. Here, we derive the analytical expression for obtaining the electric dipole moment by multiconfiguration pair-density functional theory (MC-PDFT), and we assess the accuracy of MC-PDFT for predicting dipole moments at equilibrium and nonequilibrium geometries. We show that MC-PDFT dipole moment curves have reasonable behavior even for stretched geometries, and they significantly improve upon the CASSCF results by capturing more electron correlation. The analysis of a dataset consisting of 18 first-row transition-metal diatomics and 6 main-group polyatomic molecules with a multireference character suggests that MC-PDFT and its hybrid extension (HMC-PDFT) perform comparably to CASPT2 and MRCISD+Q methods and have a mean unsigned deviation of 0.2-0.3 D with respect to the best available dipole moment reference values. We explored the dependence of the predicted dipole moments upon the choice of the on-top density functional and active space, and we recommend the tPBE and hybrid tPBE0 on-top choices for the functionals combined with the moderate correlated-participating-orbitals scheme for selecting the active space. With these choices, the mean unsigned deviations (in debyes) of the calculated equilibrium dipole moments from the best estimates are 0.77 for CASSCF, 0.29 for MC-PDFT, 0.24 for HMC-PDFT, 0.28 for CASPT2, and 0.25 for MRCISD+Q. These results are encouraging because the computational cost of MC-PDFT or HMC-PDFT is largely reduced compared to the CASPT2 and MRCISD+Q methods.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 01:30
Electric Dipoles and Dipole Moment
Theoretically, studying electric dipoles leads to understanding why the resultant electric forces around us are weak. Since electric forces are strong, remnant net charges are rare. Hence, the interaction between dipoles helps us understand electrical interactions in...
 03:37
Molecular Shape and Polarity
 02:48
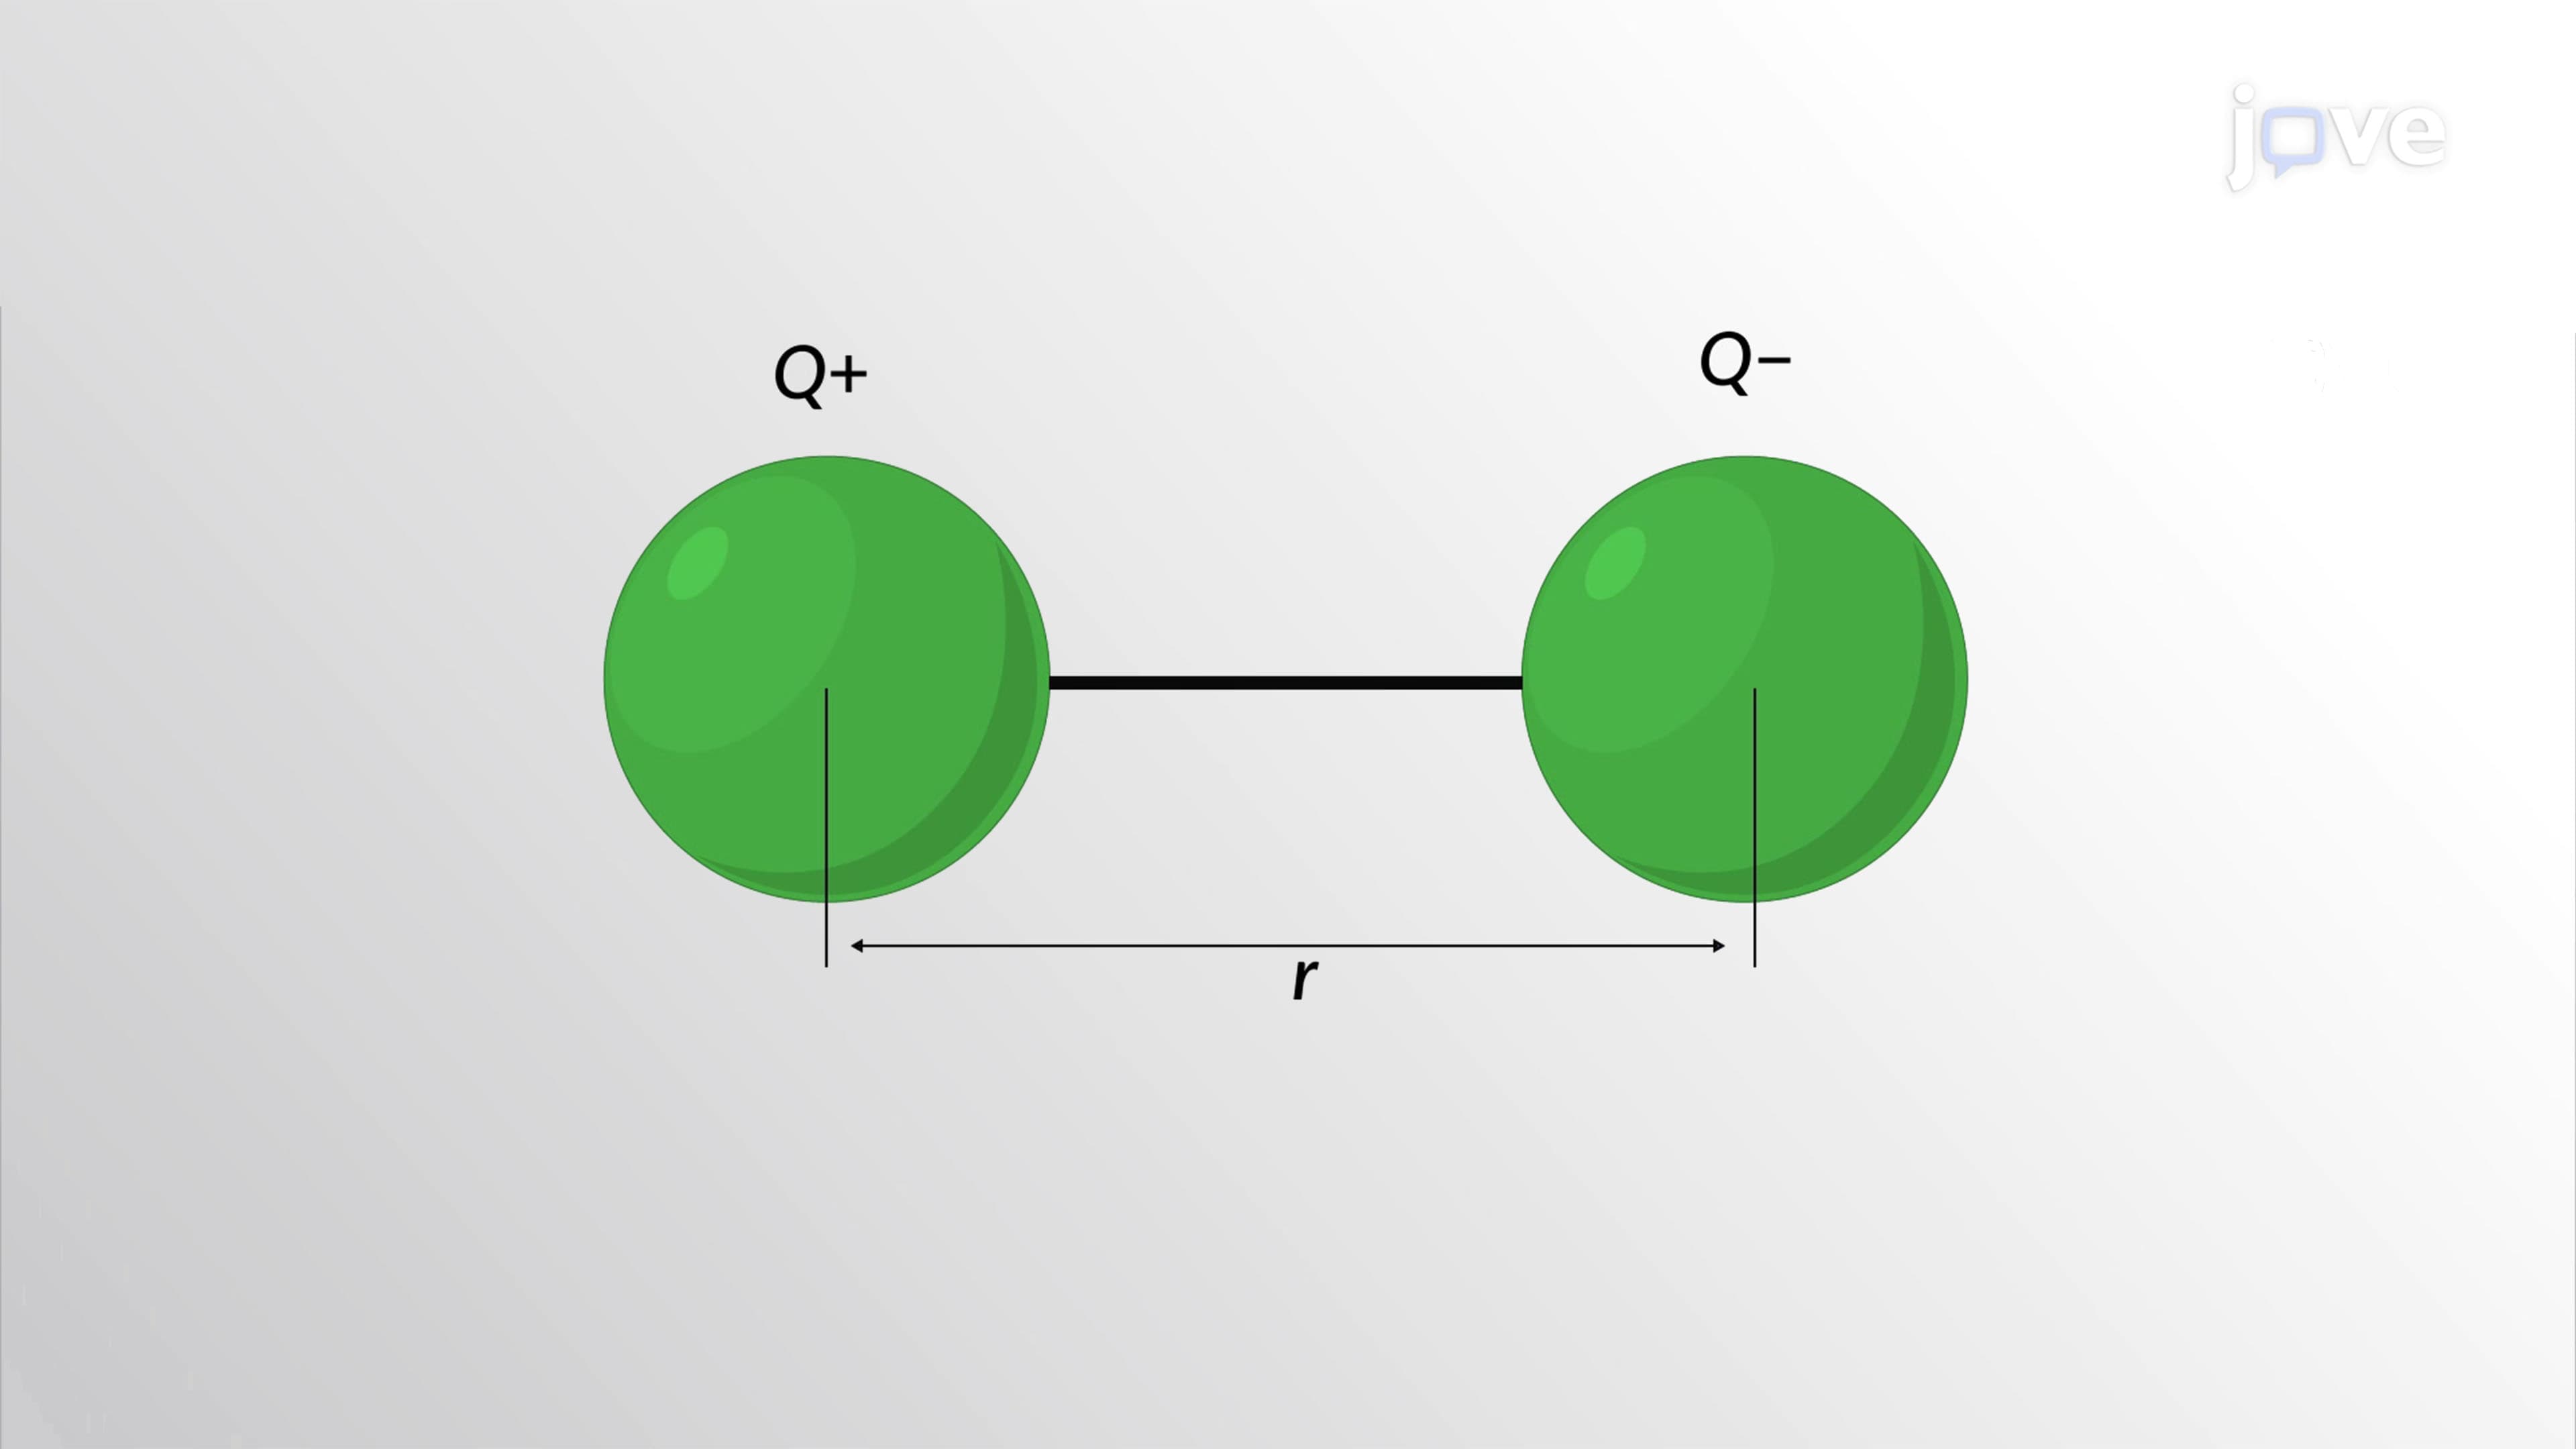
Bond Polarity, Dipole Moment, and Percent Ionic Character
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding