Related Experiment Video
Updated: Oct 12, 2025

Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
DFT calculation, molecular docking, and molecular dynamics simulation study on substituted phenylacetamide and
Vidyasrilekha Yele1, Dilep Kumar Sigalapalli2, Srikanth Jupudi1
1Department of Pharmaceutical Chemistry, JSS College of Pharmacy, JSS Academy of Higher Education & Research, Ooty, Tamil Nadu, India.

Abstract:
The atomic and molecular properties of the title compounds were calculated by Jaguar using a basis set B3LYP/6-31G**++ with hybrid DFT in the gas phase, to determine the chemical reactivity. Analysis of quantum chemical features such as HOMO and LUMO explained that the electronic charge transfer occurred within the system through conjugated paths of the selected compounds. The nucleophilic and electrophilic reactive sites are recognized from the molecular electrostatic potential plot. Electrophilic and nucleophilic attack-prone molecular sites were predicted by mapping ALIE value to the molecular surface. The bond dissociation energy of the high active compound 15 (2-chloro-N-(2-(2-(2-(2-chlorobenzoyl)hydrazineyl)-2-oxoethoxy)phenyl)acetamide) was calculated to assess the probability of compound autoxidation or degradation. Further, molecular docking, binding free energy calculations, and ADMET profile of the degradation products (DPs) of compound 15 was carried out to determine the binding affinity and toxicity profile of the formed DPs compared with the parent compound. A 150-ns molecular dynamics (MD) simulation was performed to evaluate the binding stability of the compound 15/4URL complex using Desmond. Binding free energy and binding affinity of the complex were computed for 100 trajectory frames using the MM-GBSA approach.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:19
Diazonium Group Substitution: –OH and –H
 01:34
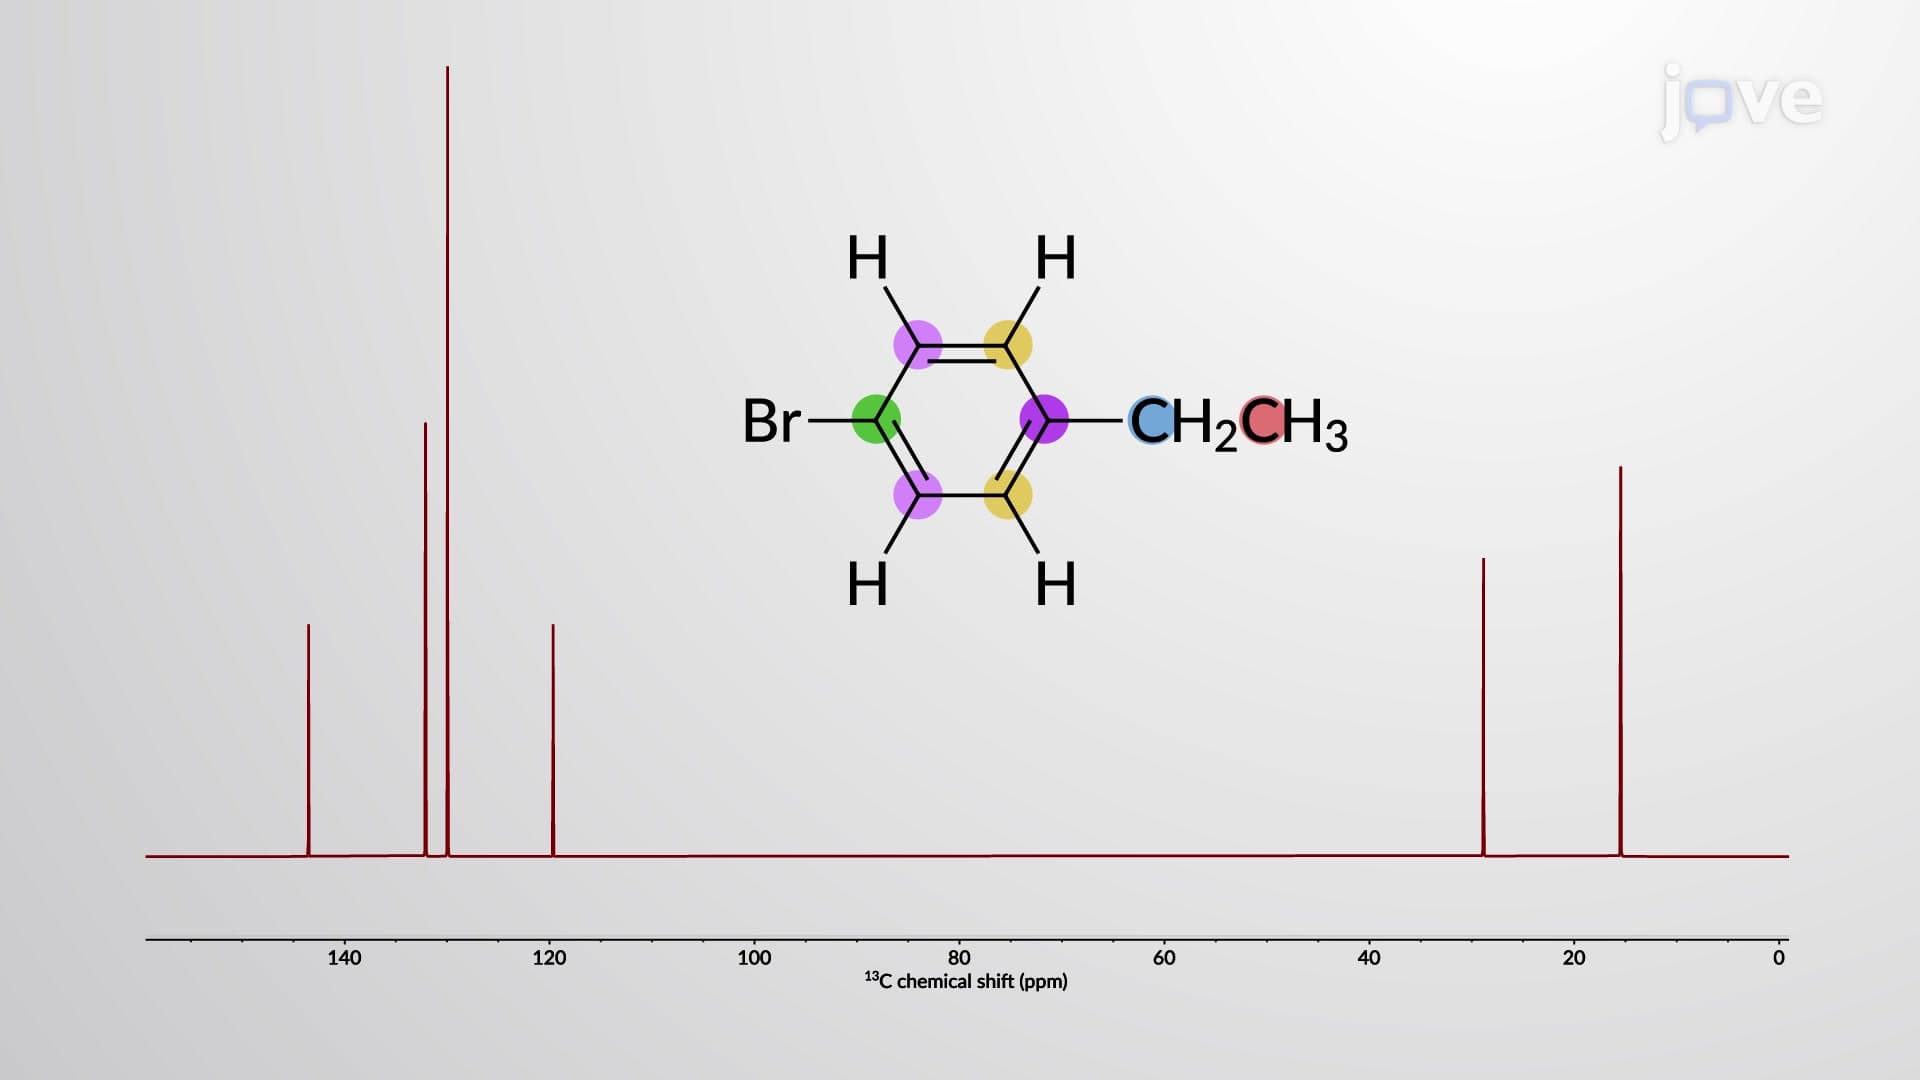
NMR Spectroscopy of Benzene Derivatives
 01:18
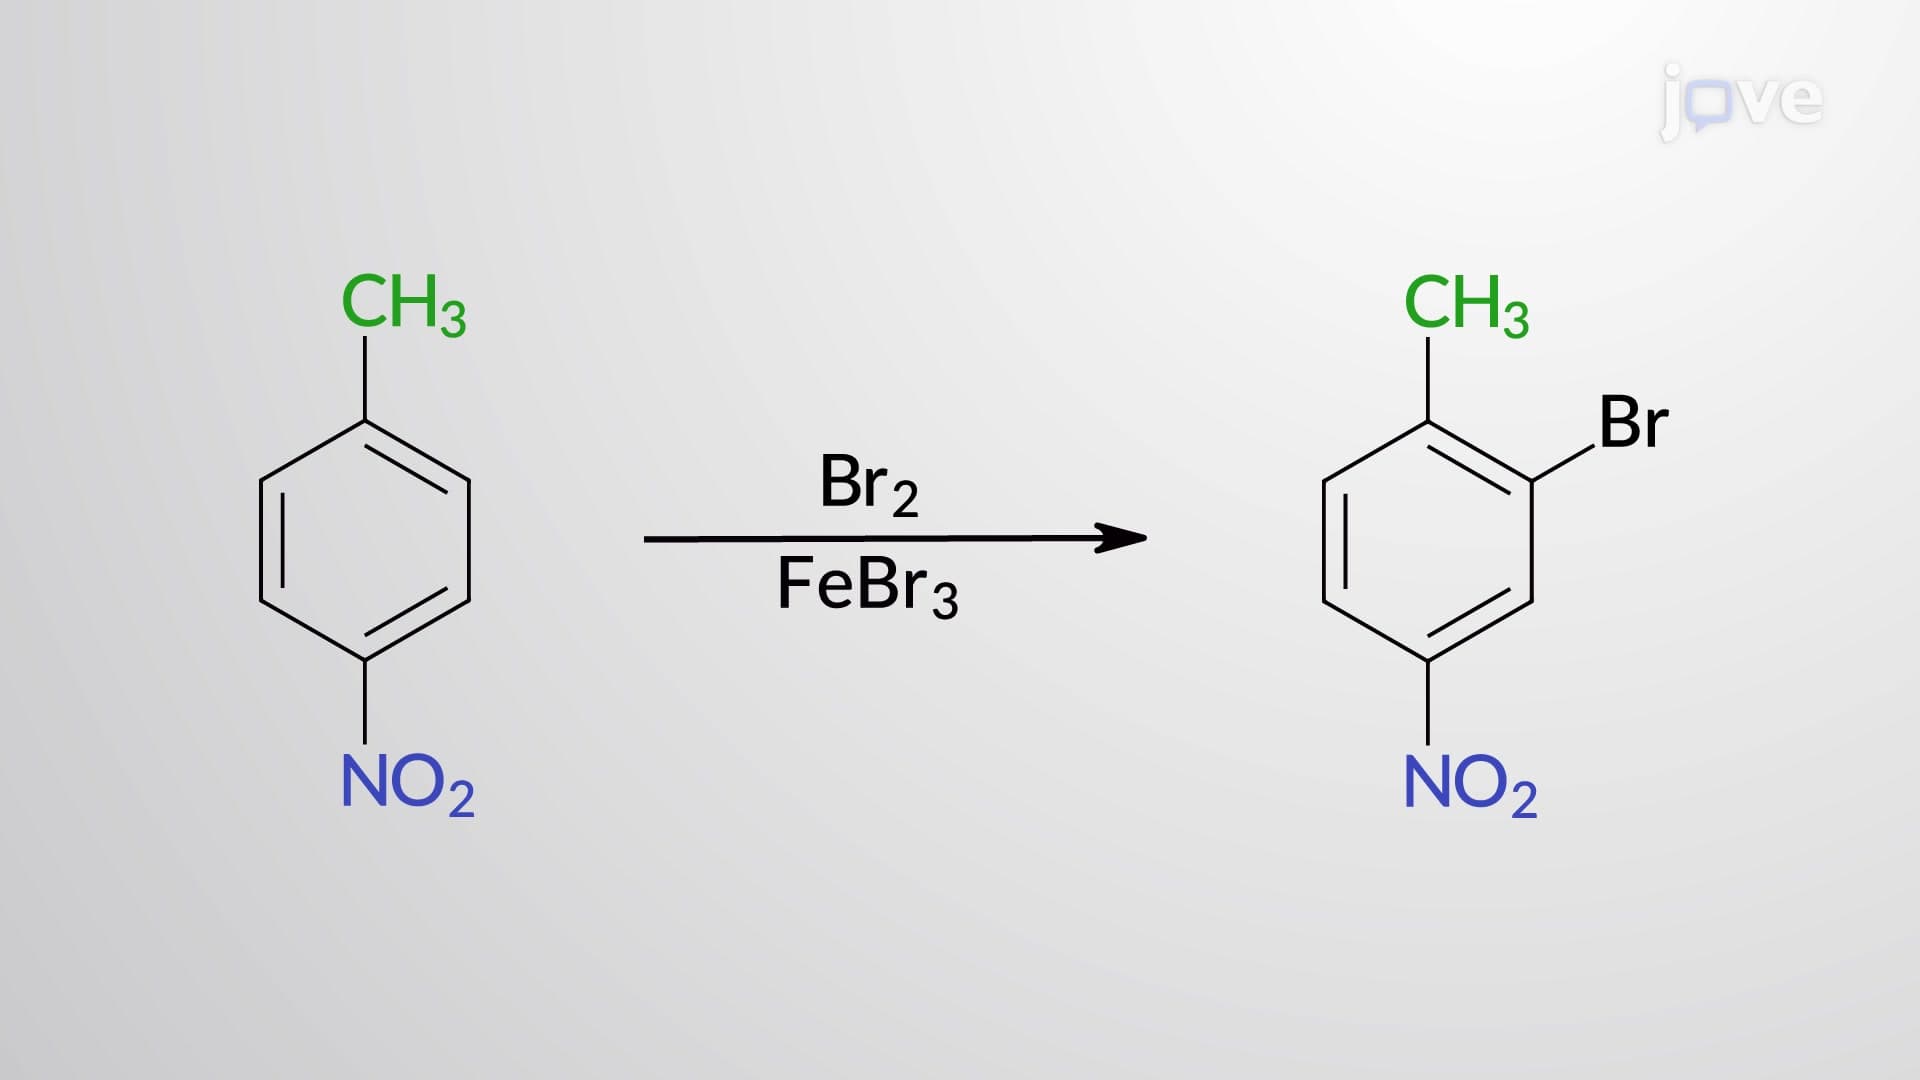
Directing and Steric Effects in Disubstituted Benzene Derivatives
 01:23
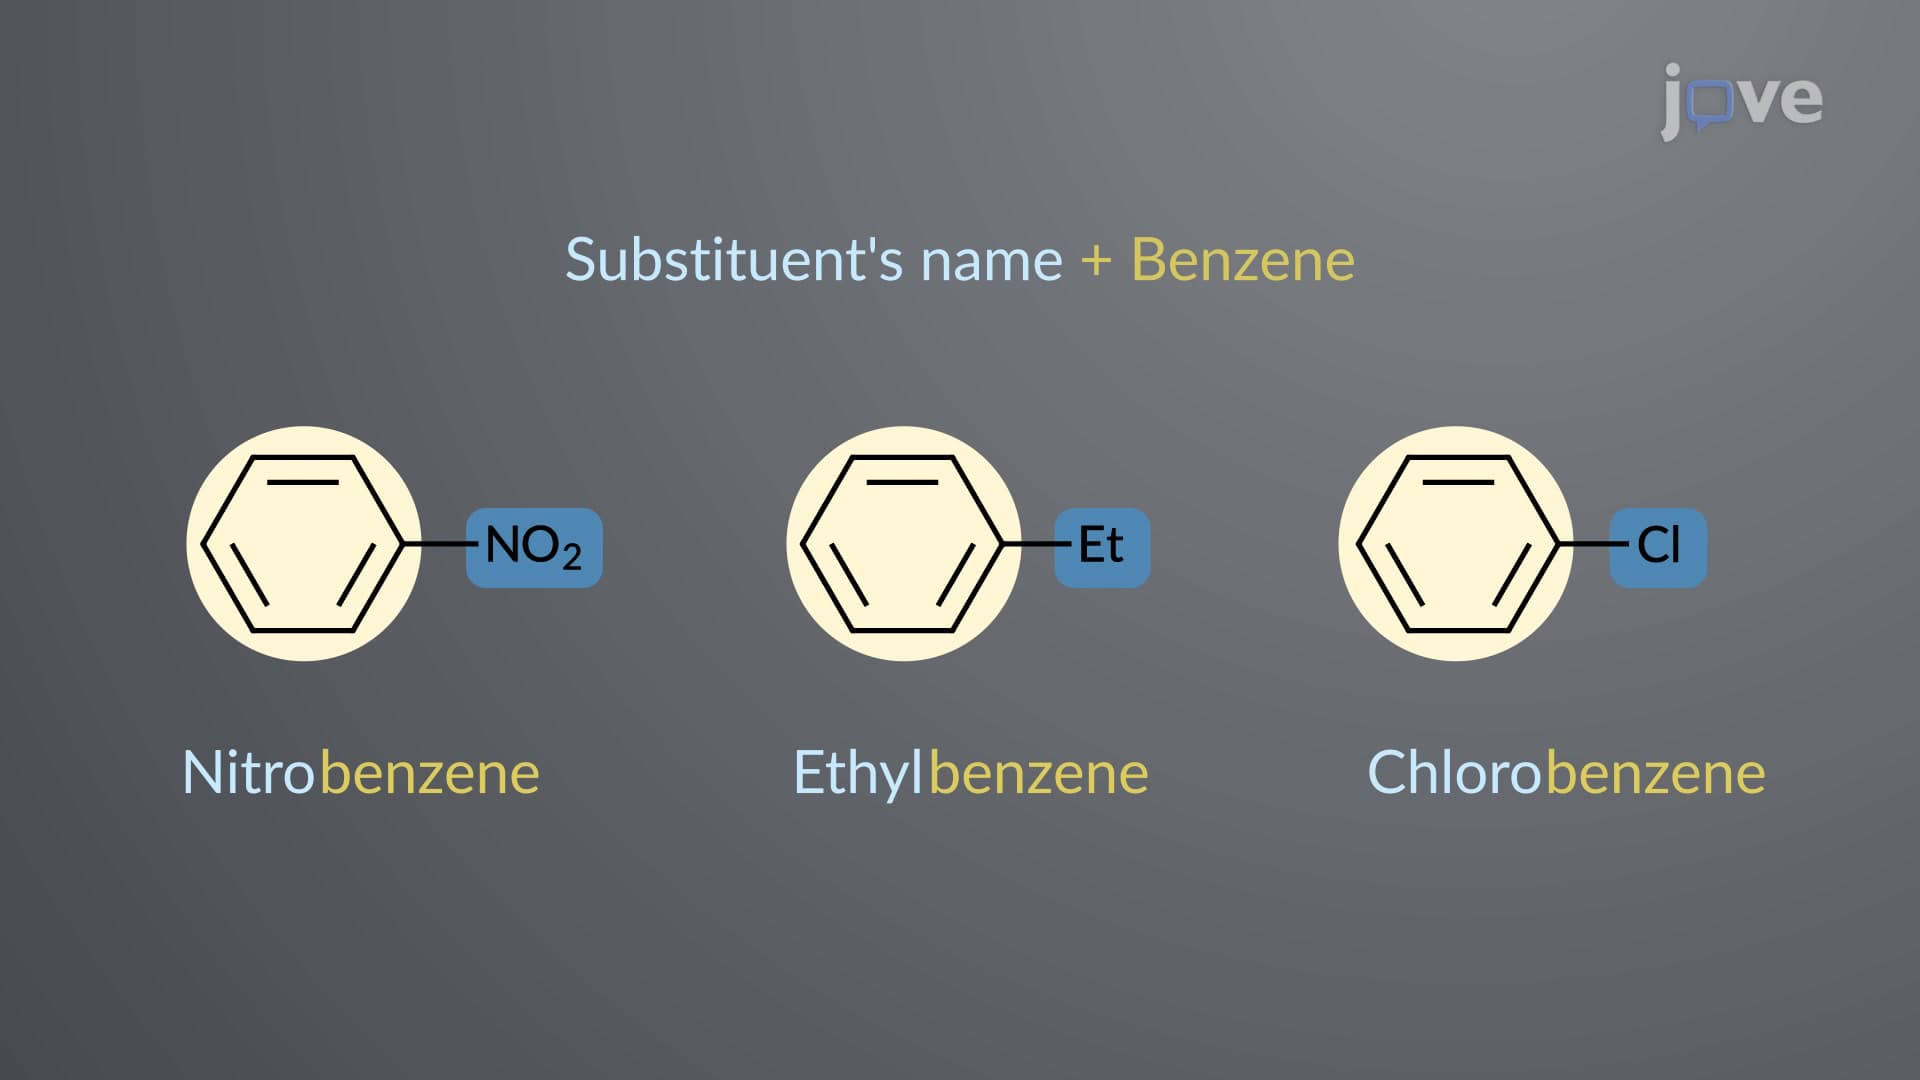
Nomenclature of Aromatic Compounds with a Single Substituent
 01:20
Diazonium Group Substitution with Halogens and Cyanide: Sandmeyer and Schiemann Reactions
 01:09
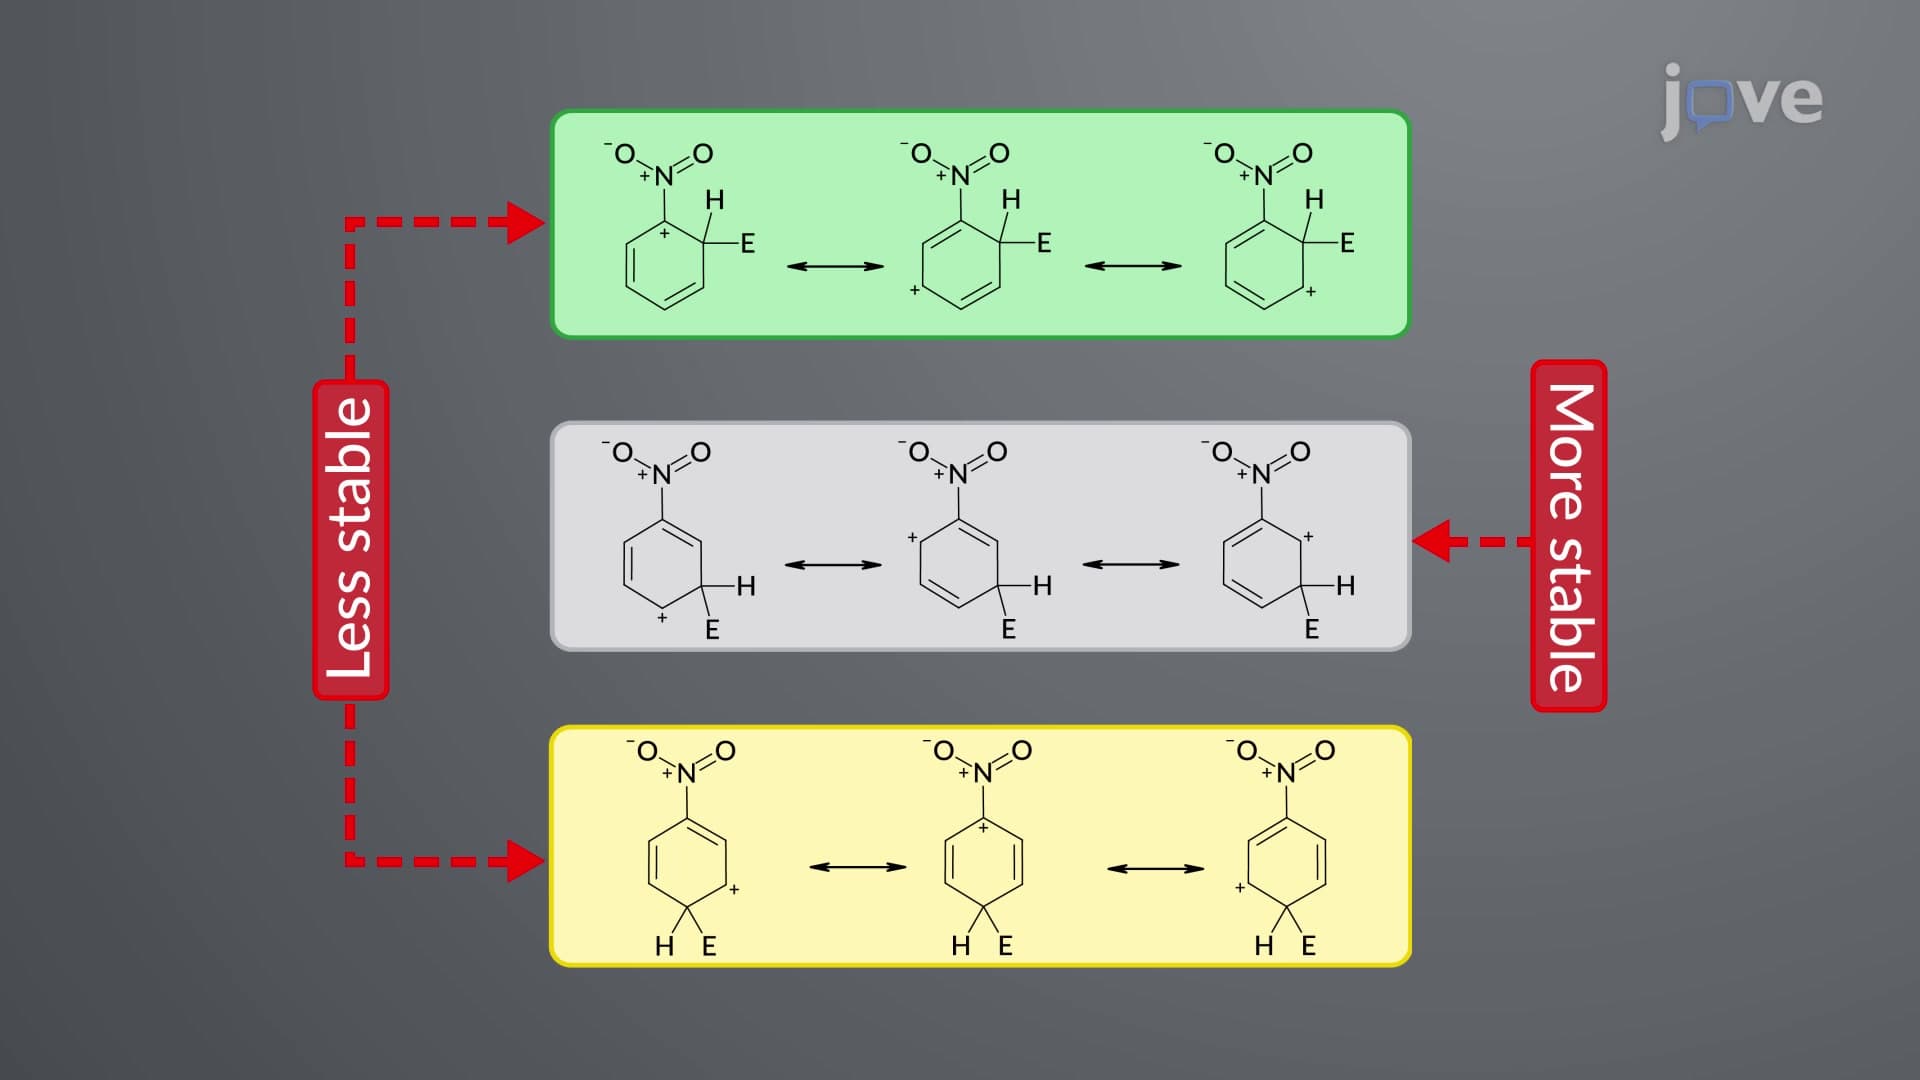
Directing Effect of Substituents: meta-Directing Groups