Related Experiment Video
Updated: Oct 11, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Near-Exact CCSDT Energetics from Rank-Reduced Formalism Supplemented by Non-iterative Corrections
1Faculty of Chemistry, University of Warsaw, Pasteura 1, 02-093 Warsaw, Poland.

Abstract:
We introduce a non-iterative energy correction, added on top of the rank-reduced coupled-cluster method with single, double, and triple substitutions, that accounts for excitations excluded from the parent triple excitation subspace. The formula for the correction is derived by employing the coupled-cluster Lagrangian formalism, with an additional assumption that the parent excitation subspace is closed under the action of the Fock operator. Owing to the rank-reduced form of the triple excitation amplitudes tensor, the computational cost of evaluating the correction scales as N7, where N is the system size. The accuracy and computational efficiency of the proposed method is assessed for both total and relative correlation energies. We show that the non-iterative correction can fulfill two separate roles. If the accuracy level of a fraction of kJ/mol is sufficient for a given system, the correction significantly reduces the dimension of the parent triple excitation subspace needed in the iterative part of the calculations. Simultaneously, it enables reproducing the exact CCSDT results to an accuracy level below 0.1 kJ/mol, with a larger, yet still reasonable, dimension of the parent excitation subspace. This typically can be achieved at a computational cost only several times larger than required for the CCSD(T) method. The proposed method retains the black-box features of the single-reference coupled-cluster theory; the dimension of the parent excitation subspace remains the only additional parameter that has to be specified.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 03:25
Free Energy Changes for Nonstandard States
where R is the gas constant (8.314 J/K·mol), T is the absolute temperature in kelvin, and Q is the reaction quotient. This equation may be used to predict the spontaneity of a process under any given set of conditions.
Reaction Quotient...
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 02:42
Formal Charges
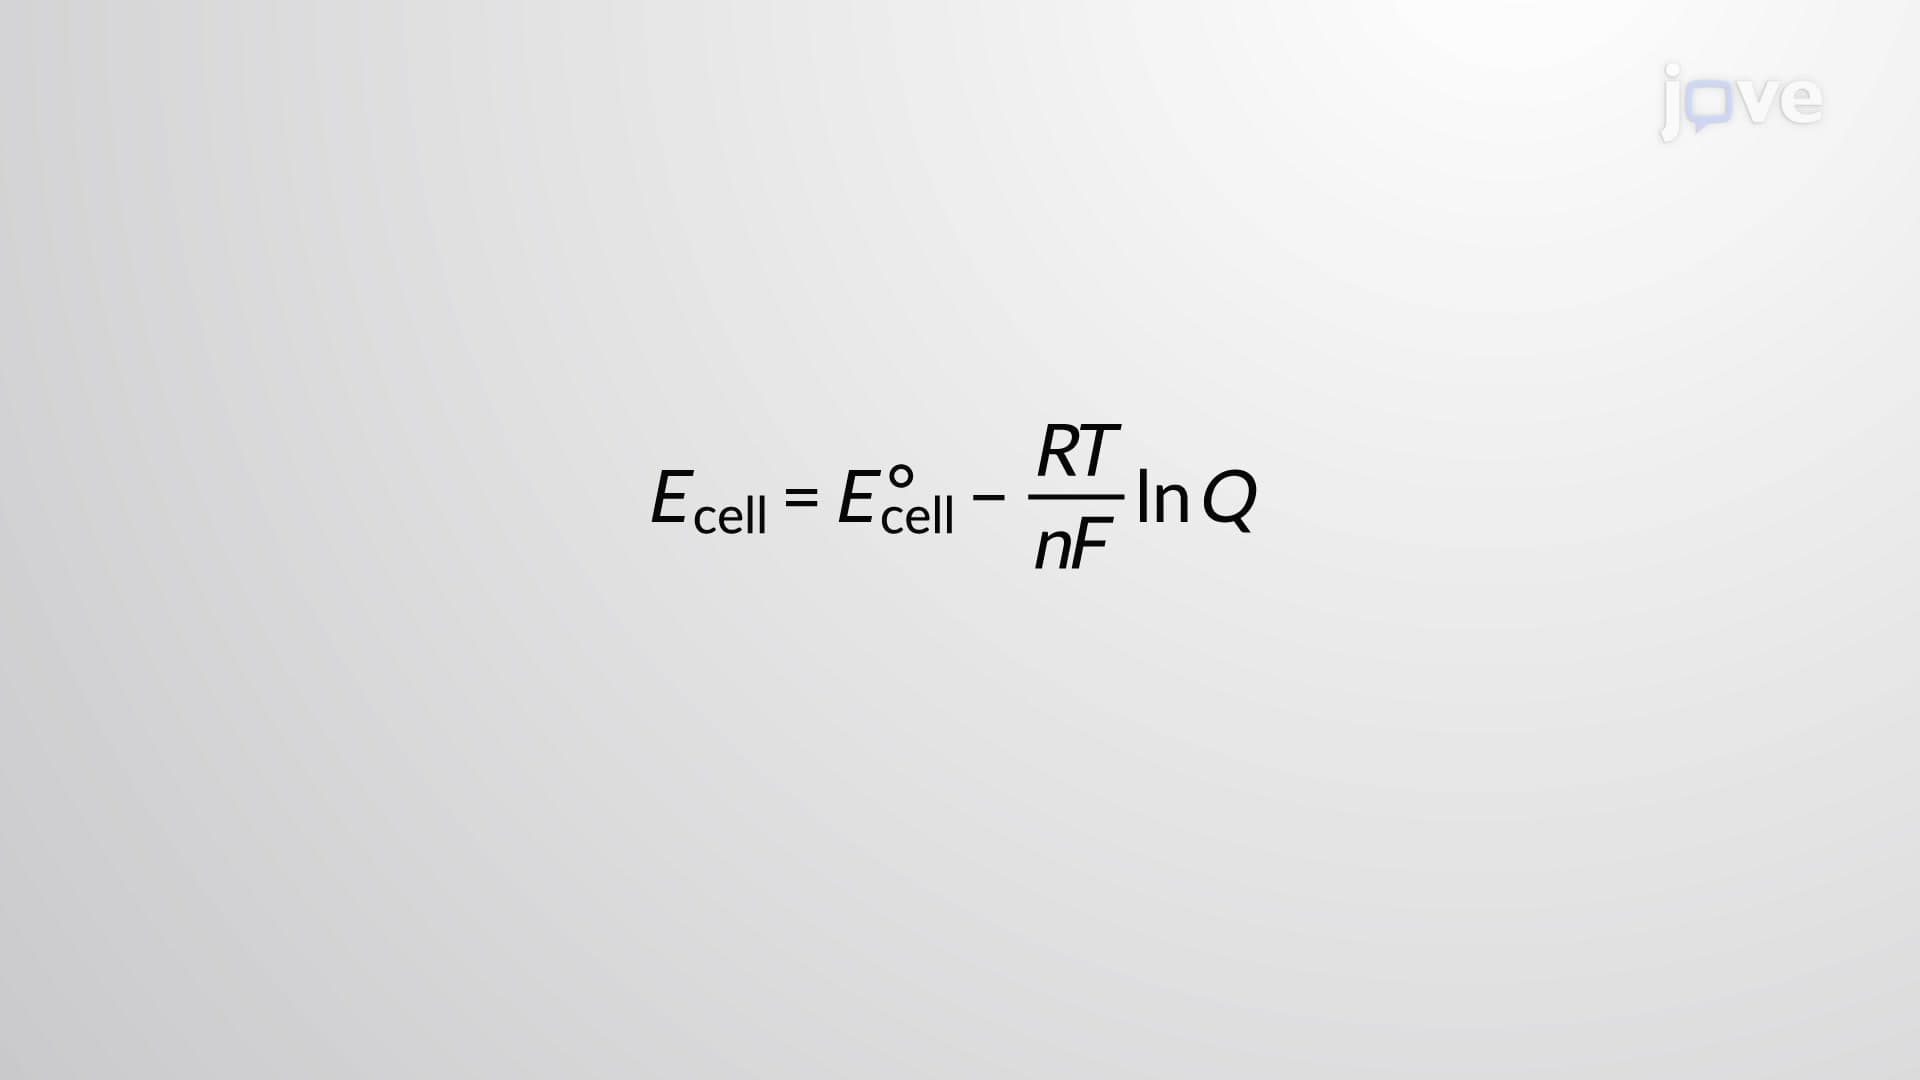 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.
 01:16
Potential-Energy Criterion for Equilibrium
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...