Related Experiment Video
Updated: May 5, 2026
A Quick Phenotypic Neurological Scoring System for Evaluating Disease Progression in the SOD1-G93A Mouse Model of ALS
Published on: October 6, 2015
JAB1 deletion in oligodendrocytes causes senescence-induced inflammation and neurodegeneration in mice
Cristina Rivellini1, Emanuela Porrello1, Giorgia Dina1
1Institute of Experimental Neurology (INSPE), Division of Neuroscience, IRCCS San Raffaele Scientific Institute, Milan, Italy.

Abstract:
Oligodendrocytes are the primary target of demyelinating disorders, and progressive neurodegenerative changes may evolve in the CNS. DNA damage and oxidative stress are considered key pathogenic events, but the underlying molecular mechanisms remain unclear. Moreover, animal models do not fully recapitulate human diseases, complicating the path to effective treatments. Here we report that mice with cell-autonomous deletion of the nuclear COP9 signalosome component CSN5 (JAB1) in oligodendrocytes develop DNA damage and defective DNA repair in myelinating glial cells. Interestingly, oligodendrocytes lacking JAB1 expression underwent a senescence-like phenotype that fostered chronic inflammation and oxidative stress. These mutants developed progressive CNS demyelination, microglia inflammation, and neurodegeneration, with severe motor deficits and premature death. Notably, blocking microglia inflammation did not prevent neurodegeneration, whereas the deletion of p21CIP1 but not p16INK4a pathway ameliorated the disease. We suggest that senescence is key to sustaining neurodegeneration in demyelinating disorders and may be considered a potential therapeutic target.
Insights
Cell-specific deletion of CSN5 (JAB1) in oligodendrocytes causes DNA damage and senescence, leading to neurodegeneration in demyelinating disorders. Targeting senescence may offer a novel therapeutic strategy for these conditions.
Area of Science:
- Neuroscience
- Cell Biology
- Genetics
Background:
- Demyelinating disorders primarily affect oligodendrocytes, leading to central nervous system (CNS) neurodegeneration.
- DNA damage and oxidative stress are implicated in pathogenesis, but molecular drivers remain elusive.
- Existing animal models inadequately represent human demyelinating diseases, hindering therapeutic development.
Purpose of the Study:
- To investigate the role of the COP9 signalosome component CSN5 (JAB1) in oligodendrocyte function and its contribution to demyelinating disorders.
- To elucidate the molecular mechanisms linking oligodendrocyte dysfunction to neurodegeneration and CNS inflammation.
Main Methods:
- Cell-autonomous deletion of JAB1 in oligodendrocytes of mice.
- Assessment of DNA damage, DNA repair, and senescence markers in glial cells.
- Analysis of CNS pathology, including demyelination, neuroinflammation (microglia activation), and neurodegeneration.
- Evaluation of motor function and survival rates in mutant mice.
- Genetic manipulation to assess the role of specific pathways (p21CIP1, p16INK4a) and inflammation in disease progression.
Main Results:
- Oligodendrocytes lacking JAB1 exhibited DNA damage and impaired DNA repair.
- JAB1-deficient oligodendrocytes displayed a senescence-like phenotype, promoting chronic inflammation and oxidative stress.
- Mice with JAB1-deleted oligodendrocytes developed progressive demyelination, neuroinflammation, neurodegeneration, motor deficits, and premature death.
- Blocking microglia inflammation did not halt neurodegeneration, but deleting p21CIP1 ameliorated disease severity.
- The p16INK4a pathway did not appear to be critical in this model.
Conclusions:
- Oligodendrocyte senescence, driven by JAB1 deficiency, is a critical factor in sustaining neurodegeneration in demyelinating disorders.
- Targeting cellular senescence represents a potential therapeutic avenue for treating demyelinating diseases.
Related Concept Videos
 01:23
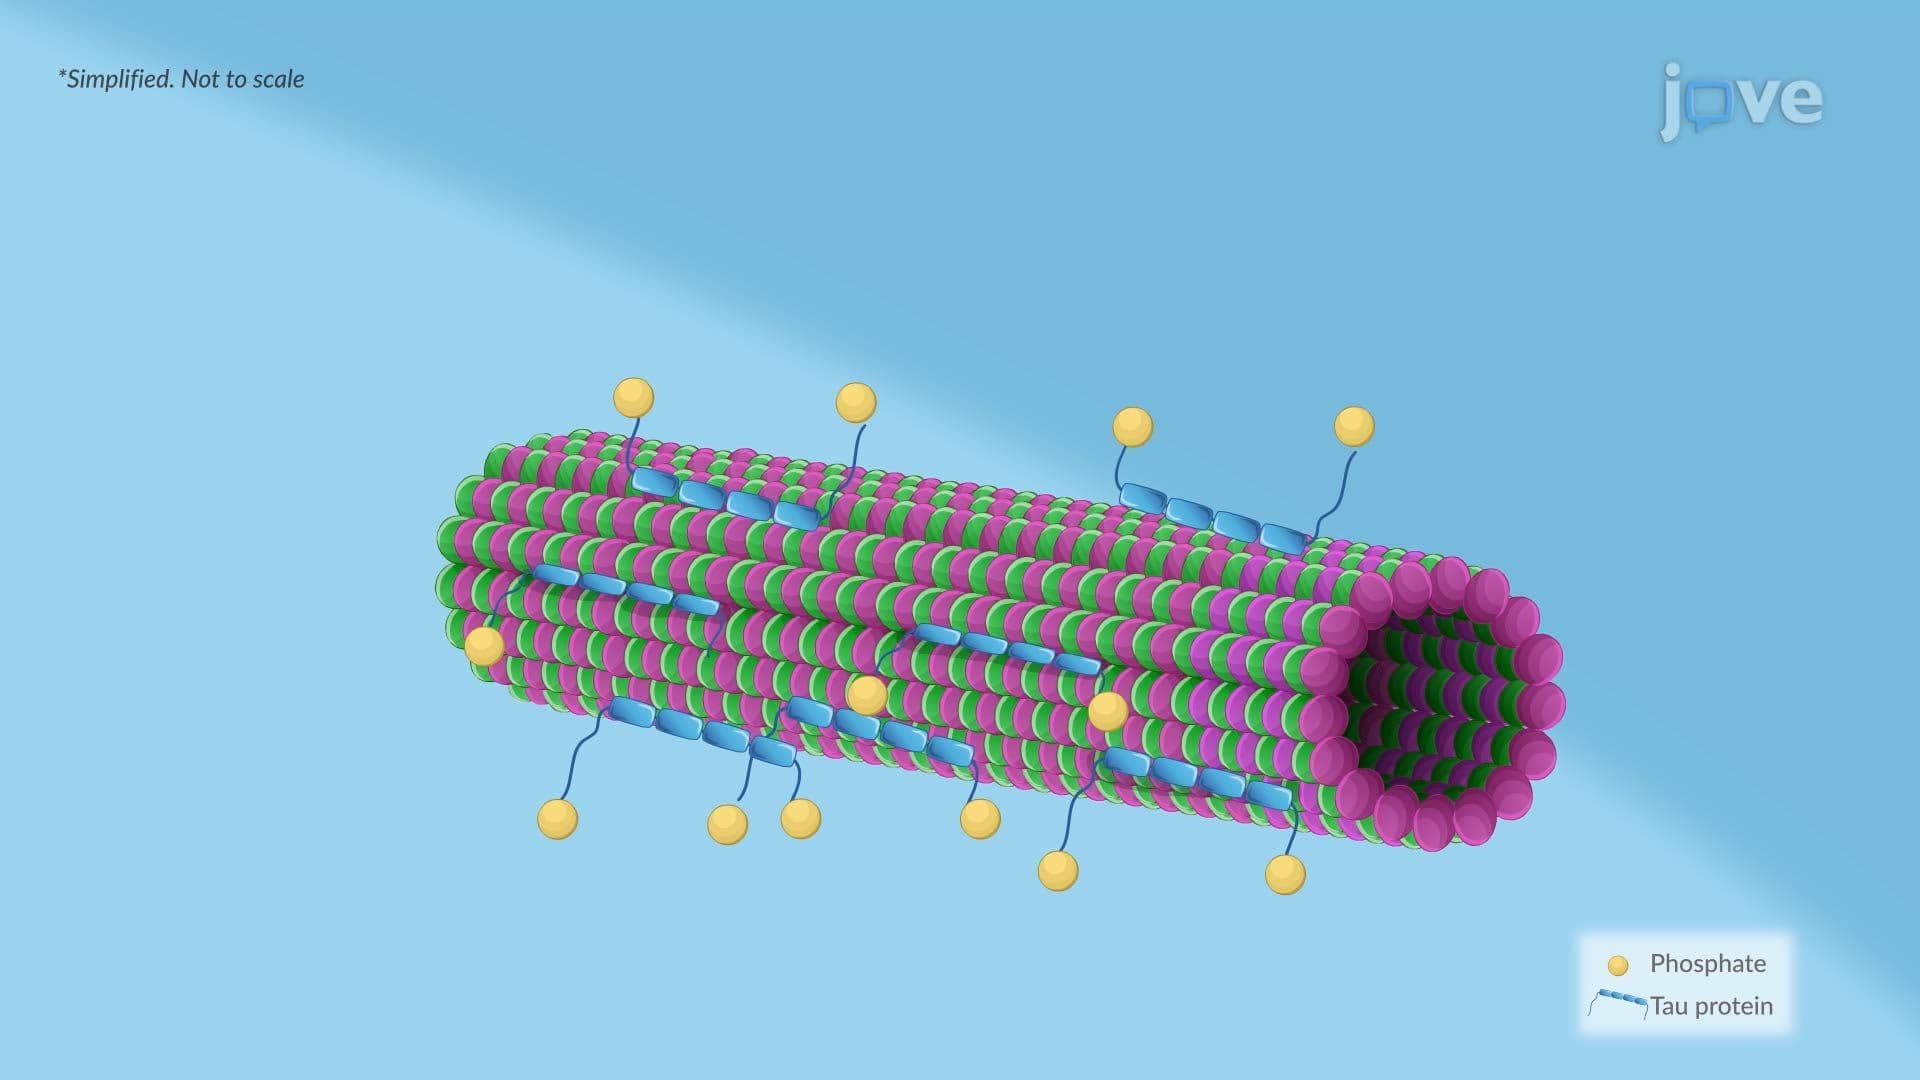
Alzheimer Disease ll: Pathophysiology
 01:24
Parkinson Disease ll: Pathophysiology