Related Experiment Video
Updated: Oct 10, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Response Surface Study on Molecular Docking Simulations of Citalopram and Donepezil as Potent CNS Drugs
Radin Alikhani1, Ahmad Ebadi2,3, Pari Karami4
1Students Research Committee, School of Pharmacy, Ardabil University of Medical Sciences, Ardabil, Iran.

Abstract:
Computer-aided drug design provides broad structural modifications to evolving bioactive molecules without an immediate requirement to observe synthetic restraints or tedious protocols. Subsequently, the most promising guidelines with regard to synthetic and biological resources may be focused on upcoming steps. Molecular docking is common in-silico drug design techniques since it predicts ligand-receptor interaction modes and associated binding affinities. Current docking simulations suffer serious constraints in estimating accurate ligand-receptor binding affinities despite several advantages and historical results. Response surface method (RSM) is an efficient statistical approach for modeling and optimization of various pharmaceutical systems. With the aim of unveiling the full potential of RSM in optimizing molecular docking simulations, this study particularly focused on binding affinity prediction of citalopram-serotonin transporter (SERT) and donepezil-acetyl cholinesterase (AChE) complexes. For this purpose, Box-Behnken design of experiments (DOE) was used to develop a trial matrix for simultaneous variations of AutoDock4.2 driven binding affinity data with selected factor levels. Responses of all docking trials were considered as estimated protein inhibition constants with regard to validated data for each drug. The output matrix was subjected to statistical analysis and constructing polynomial quadratic models. Numerical optimization steps to attain ideal docking accuracies revealed that more accurate results might be envisaged through the best combination of factor levels and considering factor interactions. Results of the current study indicated that the application of RSM in molecular docking simulations might lead to optimized docking protocols with more stable estimates of ligand-target interactions and hence better correlation of in-silico in-vitro data.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:21
Drugs Affecting Neurotransmitter Release or Uptake
 01:16
Protein-Drug Binding: Mechanism and Kinetics
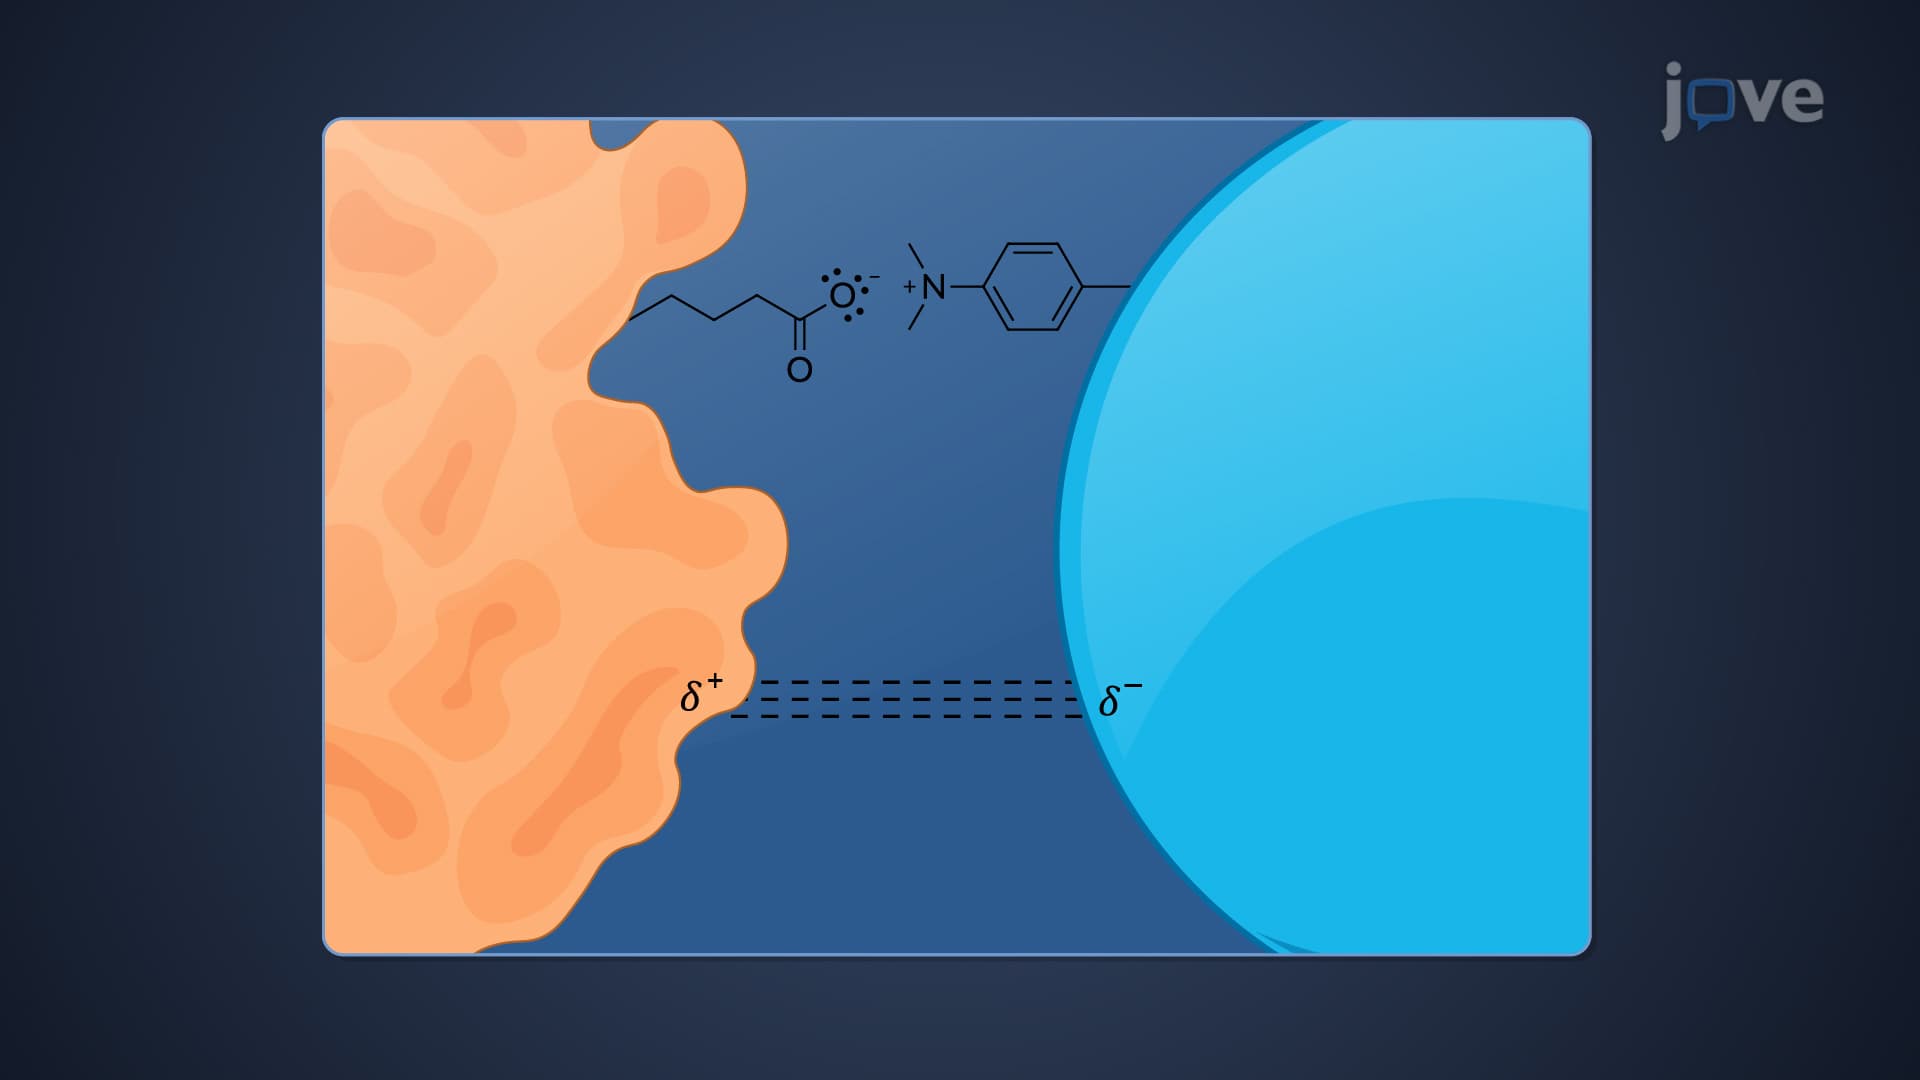
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...