Related Experiment Video
Updated: Oct 9, 2025
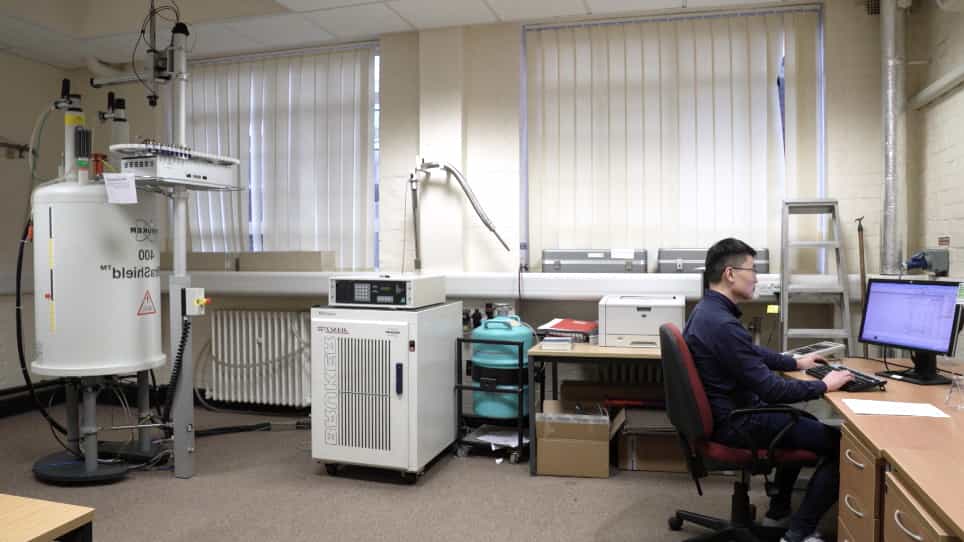

A New Straightforward Method for Lipophilicity logP Measurement using 19F NMR Spectroscopy
Published on: January 30, 2019
Relating Conformational Equilibria to Conformer-Specific Lipophilicities: New Opportunities in Drug Discovery
Bruno Linclau1,2, Zhong Wang1, Benjamin Jeffries1
1School of Chemistry, University of Southampton, Highfield, Southampton, SO17 1BJ, UK.

Abstract:
Efficient drug discovery is based on a concerted effort in optimizing bioactivity and compound properties such as lipophilicity, and is guided by efficiency metrics that reflect both aspects. While conformation-activity relationships and ligand conformational control are known strategies to improve bioactivity, the use of conformer-specific lipophilicities (logp) is much less explored. Here we show how conformer-specific logp values can be obtained from knowledge of the macroscopic logP value, and of the equilibrium constants between the individual species in water and in octanol. This is illustrated with fluorinated amide rotamers, with integration of rotamer 19 F NMR signals as a facile, direct method to obtain logp values. The difference between logp and logP optimization is highlighted, giving rise to a novel avenue for lipophilicity control in drug discovery.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 08:53
Using Scaffold Liposomes to Reconstitute Lipid-proximal Protein-protein Interactions In Vitro
Published on: January 11, 2017
Related Concept Videos
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles...
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:14
Drug Dissolution: Requirements and Profile Comparison
 01:27
Pharmaceutical Alternatives: Polymorphic Form-Related and Particle Size-Related Therapeutic Nonequivalence