Related Experiment Video
Updated: Oct 8, 2025

Author Spotlight: Enhanced Histone PTM Isomer Identification Through LC-TIMS-ToF MS/MS and PASEF
Published on: January 12, 2024
Characterization of Structural Hemoglobin Variants by Top-Down Mass Spectrometry and R Programming Tools for Rapid
Yuan Lin1, Archana M Agarwal2,3, Alan G Marshall1,4
1Department of Chemistry and Biochemistry, Florida State University, Tallahassee, Florida 32308, United States.

Hemoglobinopathies are one of the most prevalent genetic disorders, affecting millions throughout the world. These are caused by pathogenic variants in genes that control the production of hemoglobin (Hb) subunits. As the number of known Hb variants has increased, it has become more challenging to obtain unambiguous results from routine chromatographic assays employed in the clinical laboratory. Top-down proteomic analysis of Hb by mass spectrometry is a definitive method to directly characterize the sequences of intact subunits. Here, we apply "chimeric ion loading" to characterize Hb β subunit variants. In this technique, product ions derived from complementary dissociation techniques are accumulated in a multipole storage device before delivery to a 21 T Fourier-transform ion cyclotron resonance mass spectrometer for simultaneous detection. To further improve the efficiency of identification of Hb variants and localization of the mutation site(s), we developed an R programming script, "Variants Identifier", to search top-down data against a database containing accurate intact mass differences and diagnostic ions from investigated Hb variants. A second R script, "PredictDiag", was developed and employed to determine relevant diagnostic ions for additional Hb variants with known sequences. These two R scripts were successfully applied to the identification of a Hb δ-β fusion protein and other Hb variants. The combination of chimeric ion loading and the above R scripts enables rapid and reliable interpretation of top-down mass spectrometry data, regardless of activation type, for Hb variant identification.
Hemoglobinopathies are one of the most prevalent genetic disorders, affecting millions throughout the world. These are caused by pathogenic variants in genes that control the production of hemoglobin (Hb) subunits. As the number of known Hb variants has increased, it has become more challenging to obtain unambiguous results from routine chromatographic assays employed in the clinical laboratory. Top-down proteomic analysis of Hb by mass spectrometry is a definitive method to directly characterize the sequences of intact subunits. Here, we apply "chimeric ion loading" to characterize Hb β subunit variants. In this technique, product ions derived from complementary dissociation techniques are accumulated in a multipole storage device before delivery to a 21 T Fourier-transform ion cyclotron resonance mass spectrometer for simultaneous detection. To further improve the efficiency of identification of Hb variants and localization of the mutation site(s), we developed an R programming script, "Variants Identifier", to search top-down data against a database containing accurate intact mass differences and diagnostic ions from investigated Hb variants. A second R script, "PredictDiag", was developed and employed to determine relevant diagnostic ions for additional Hb variants with known sequences. These two R scripts were successfully applied to the identification of a Hb δ-β fusion protein and other Hb variants. The combination of chimeric ion loading and the above R scripts enables rapid and reliable interpretation of top-down mass spectrometry data, regardless of activation type, for Hb variant identification.
More Related Videos
Related Concept Videos
 01:57
Gene Families
Occasionally these regions can be adapted to take on new roles within the organism, becoming novel genes...
 01:15
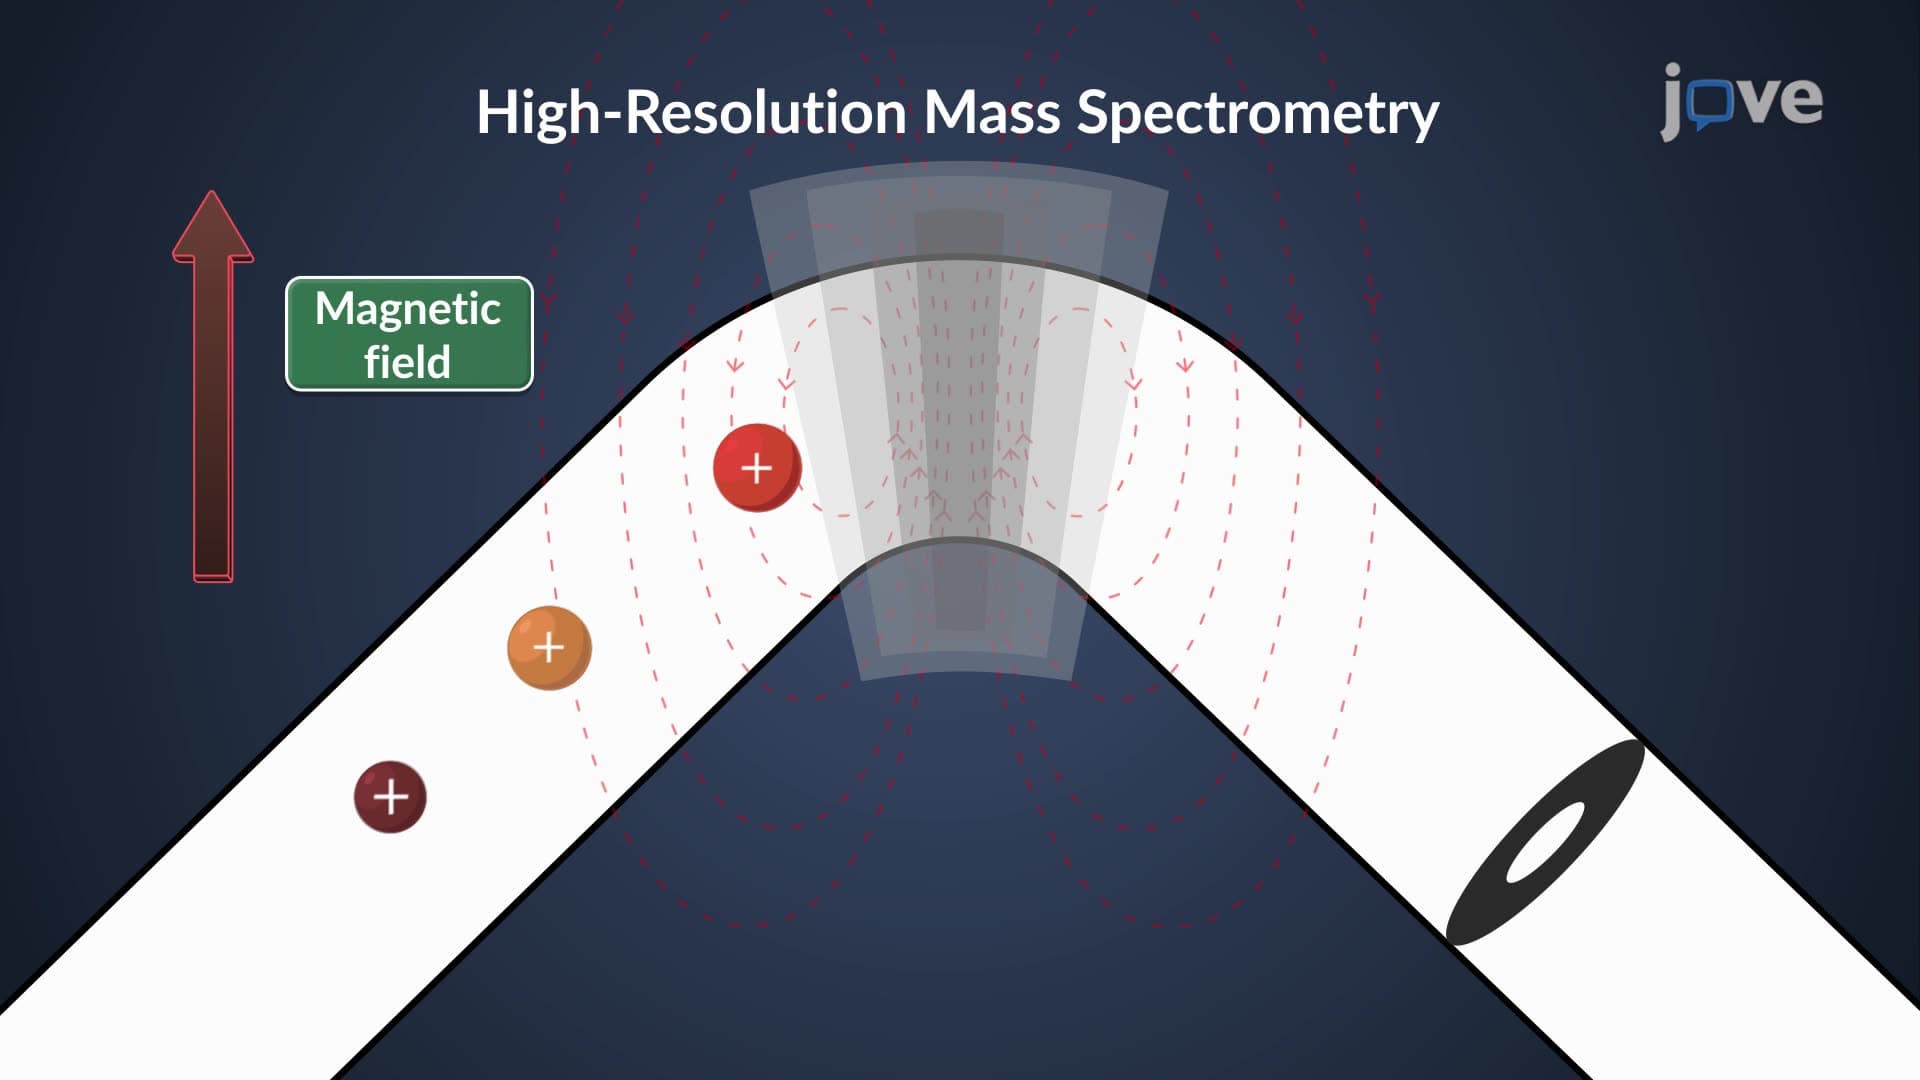
High-Resolution Mass Spectrometry (HRMS)
 02:21
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...