Related Experiment Video
Updated: Oct 8, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Theoretical Calculation of Core-Excited States along Dissociative Pathways beyond Second-Order Perturbation Theory
Meng Huang1, Chenyang Li1,2, Francesco A Evangelista1
1Department of Chemistry and Cherry Emerson Center for Scientific Computation, Emory University, Atlanta, Georgia 30322, United States.

Abstract:
We extend the multireference driven similarity renormalization (MR-DSRG) method to compute core-excited states by combining it with a GASSCF treatment of orbital relaxation and static electron correlation effects. We consider MR-DSRG treatments of dynamical correlation truncated at the level of perturbation theory (DSRG-MRPT2/3) and iterative linearized approximations with one- and two-body operators [MR-LDSRG(2)] in combination with a spin-free exact-two-component (X2C) one-electron treatment of scalar relativistic effects. This approach is calibrated and tested on a series of 16 core-excited states of five closed- and open-shell diatomic molecules containing first-row elements (C, N, and O). All GASSCF-MR-DSRG theories show excellent agreement with experimental adiabatic transitions energies, with mean absolute errors ranging between 0.17 and 0.35 eV, even for the challenging partially doubly excited states of the N2+ molecule. The vibrational structure of all these transitions, obtained from using a full potential energy scan, shows a mean absolute error as low as 25 meV for DSRG-MRPT2 and 12/13 meV for DSRG-MRPT3 and MR-LDSRG(2). We generally find that a treatment of dynamical correlation that goes beyond the second-order level in perturbation theory improves the accuracy of the potential energy surface, especially in the bond-dissociation region.
More Related Videos

Related Concept Videos
 02:13
Energy Diagrams, Transition States, and Intermediates
 03:51
Molecular Orbital Theory II
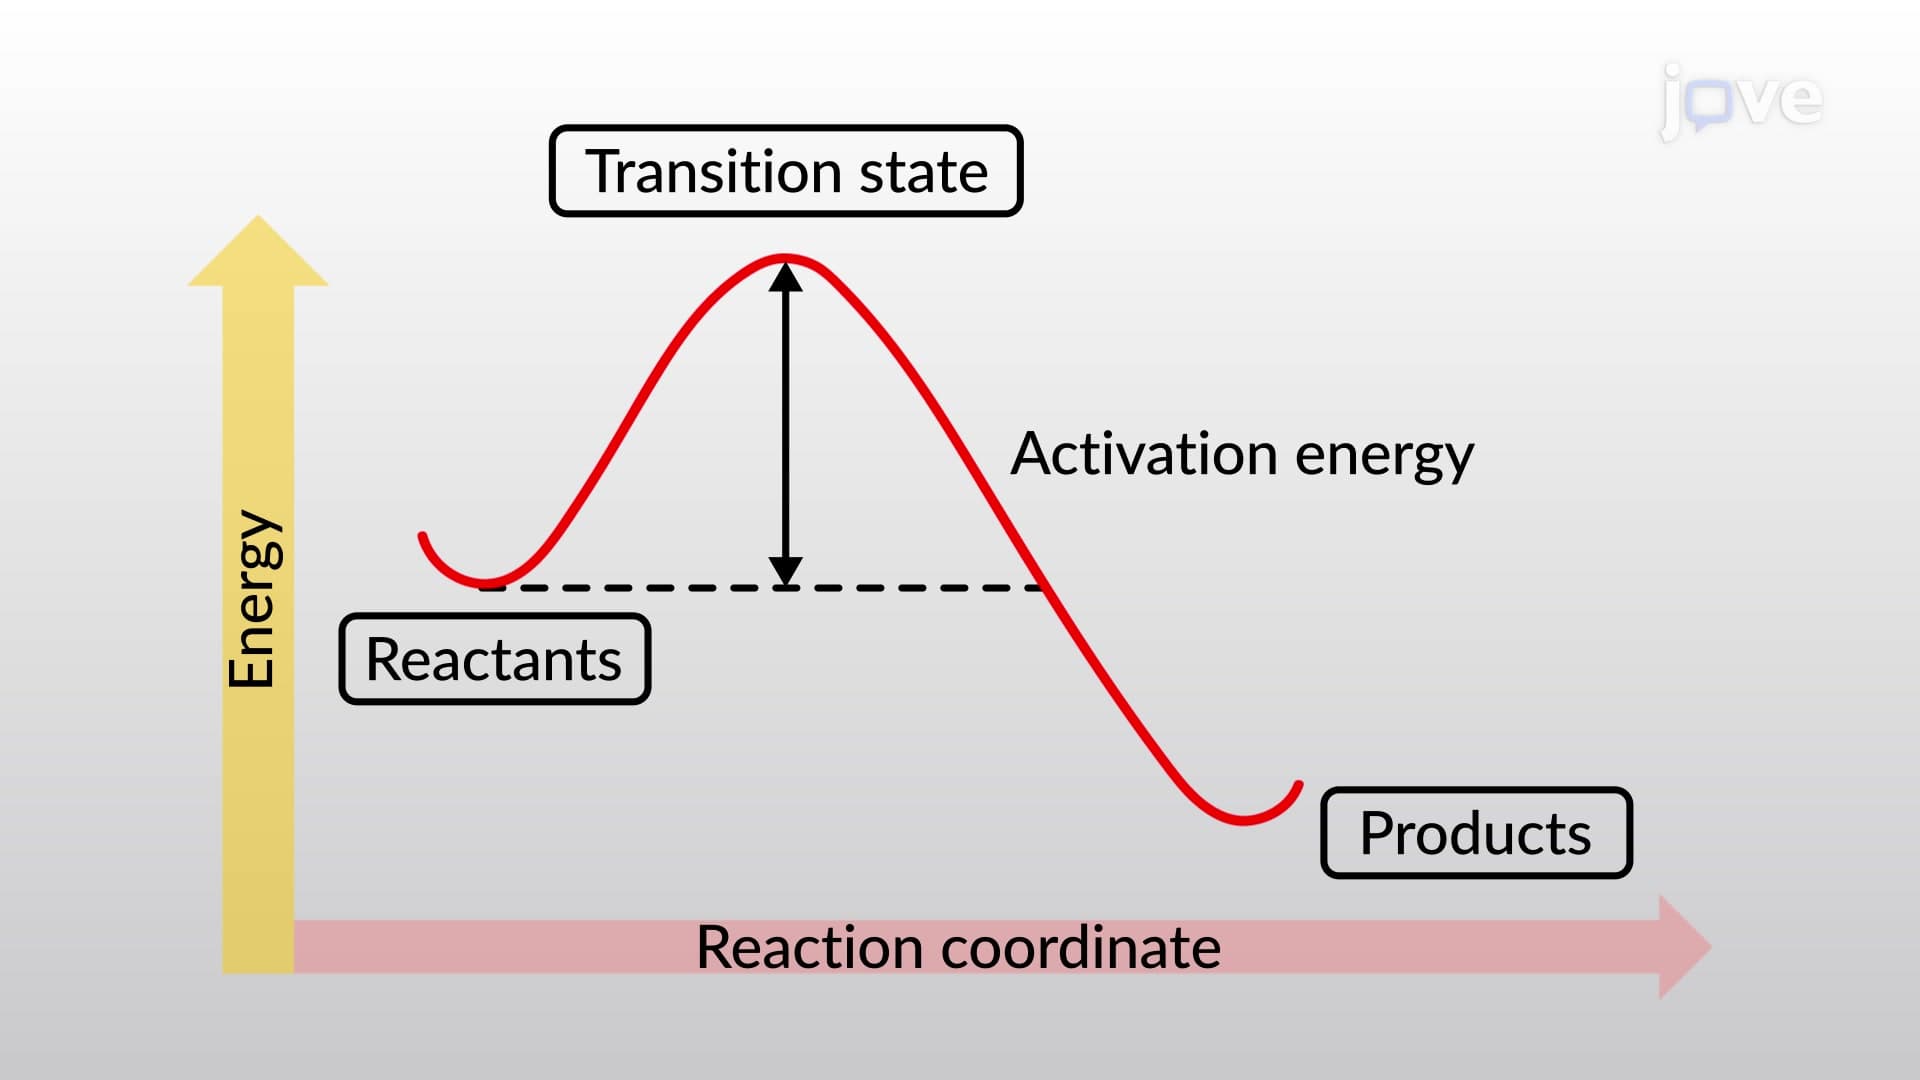 02:13
Bond Dissociation Energy and Activation Energy
 02:35
Molecular Orbital Theory I
 01:27
π Electron Effects on Chemical Shift: Overview
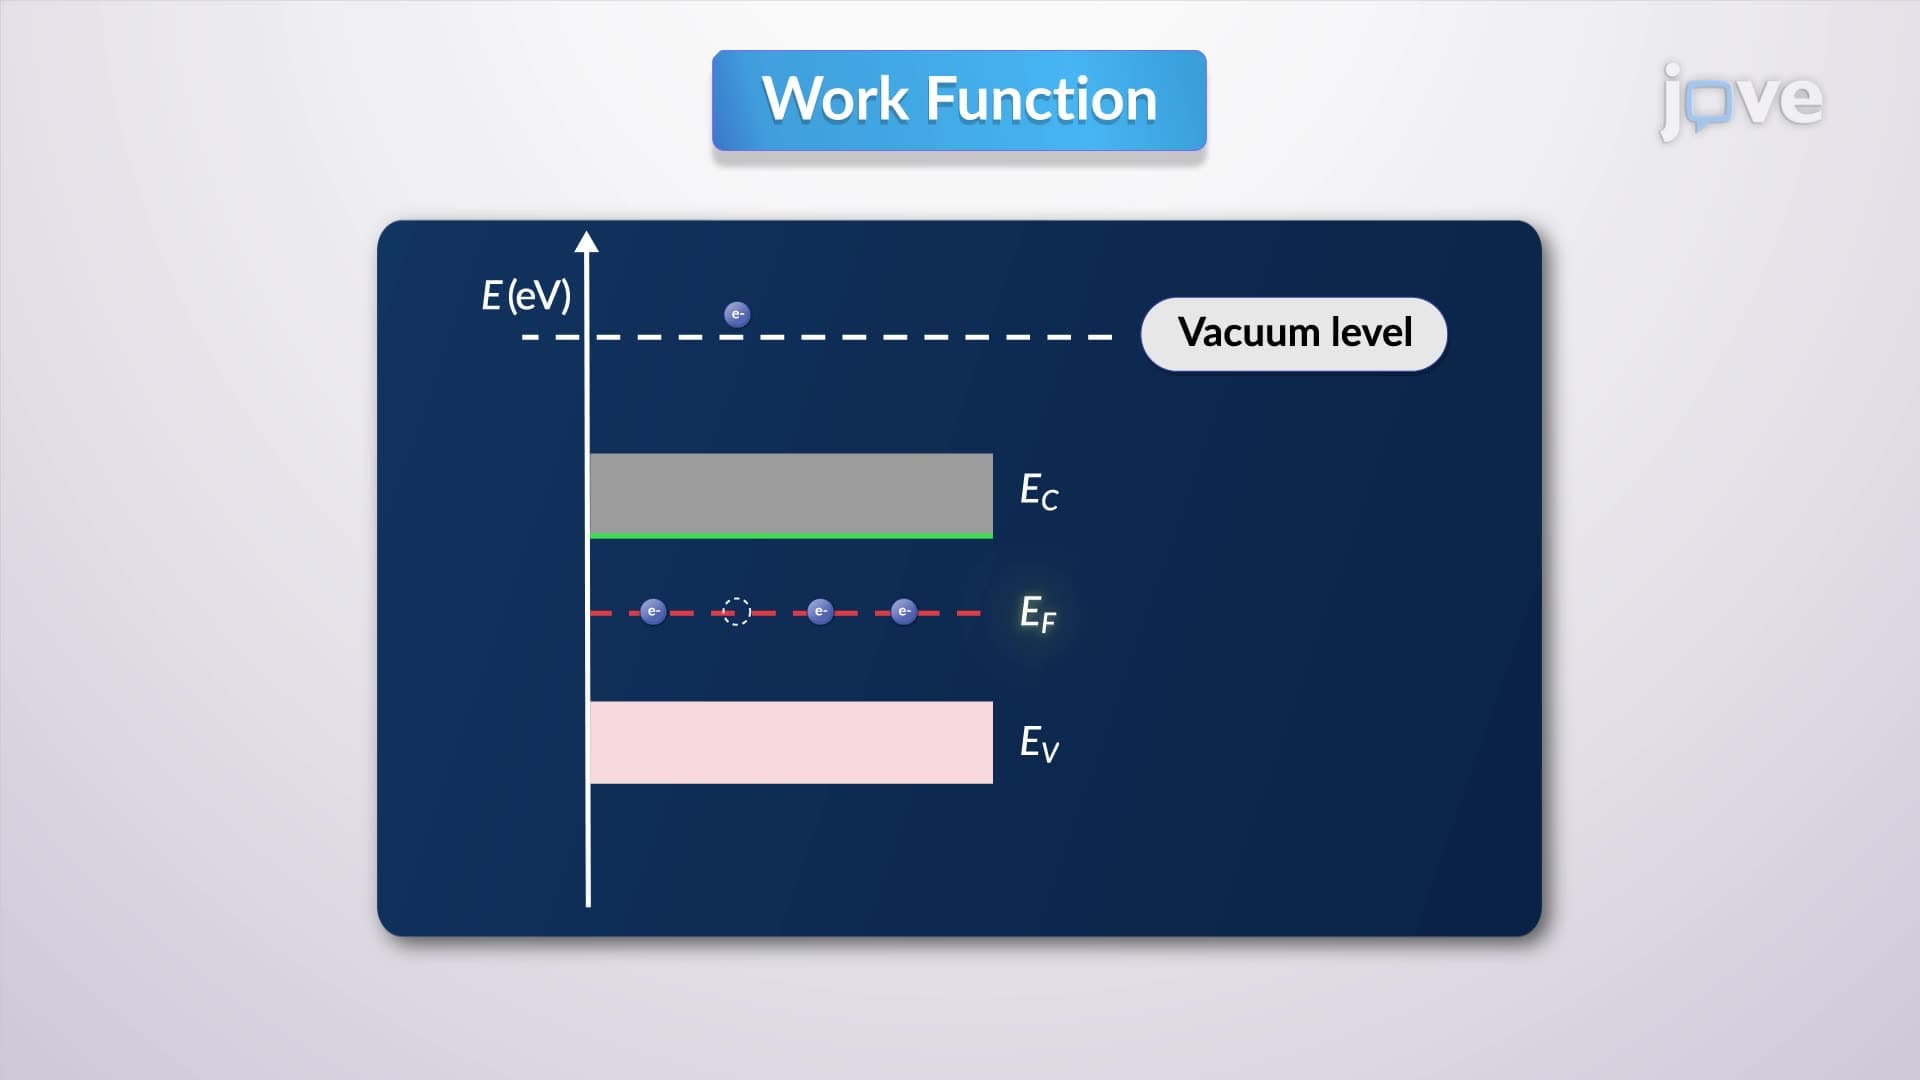 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...