A DMRG/CASPT2 Investigation of Metallocorroles: Quantifying Ligand Noninnocence in Archetypal 3d and 4d Element
Quan Manh Phung1,2, Yasin Muchammad1, Takeshi Yanai1,2
1Department of Chemistry, Graduate School of Science, Nagoya University, Furo-cho, Chikusa-ku, Nagoya, Aichi 464-8602, Japan.

Abstract:
Hybrid density functional theory (B3LYP) and density matrix renormalization group (DMRG) theory have been used to quantitatively compare the degree of ligand noninnocence (corrole radical character) in seven archetypal metallocorroles. The seven complexes, in decreasing order of corrole noninnocent character, are Mn[Cor]Cl > Fe[Cor]Cl > Fe[Cor](NO) > Mo[Cor]Cl2 > Ru[Cor](NO) ≈ Mn[Cor]Ph ≈ Fe[Cor]Ph ≈ 0, where [Cor] refers to the unsubstituted corrolato ligand. DMRG-based second-order perturbation theory calculations have also yielded detailed excited-state energetics data on the compounds, shedding light on periodic trends involving middle transition elements. Thus, whereas the ground state of Fe[Cor](NO) (S = 0) is best described as a locally S = 1/2 {FeNO}7 unit antiferromagnetically coupled to a corrole A' radical, the calculations confirm that Ru[Cor](NO) may be described as simply {RuNO}6-Cor3-, that is, having an innocent corrole macrocycle. Furthermore, whereas the ferromagnetically coupled S = 1{FeNO}7-Cor•2- state of Fe[Cor](NO) is only ∼17.5 kcal/mol higher than the S = 0 ground state, the analogous triplet state of Ru[Cor](NO) is higher by a far larger margin (37.4 kcal/mol) relative to the ground state. In the same vein, Mo[Cor]Cl2 exhibits an adiabatic doublet-quartet gap of 36.1 kcal/mol. The large energy gaps associated with metal-ligand spin coupling in Ru[Cor](NO) and Mo[Cor]Cl2 reflect the much greater covalent character of 4d-π interactions relative to analogous interactions involving 3d orbitals. As far as excited-state energetics is concerned, DMRG-CASPT2 calculations provide moderate validation for hybrid density functional theory (B3LYP) for qualitative purposes, but underscore the possibility of large errors (>10 kcal/mol) in interstate energy differences.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 00:43
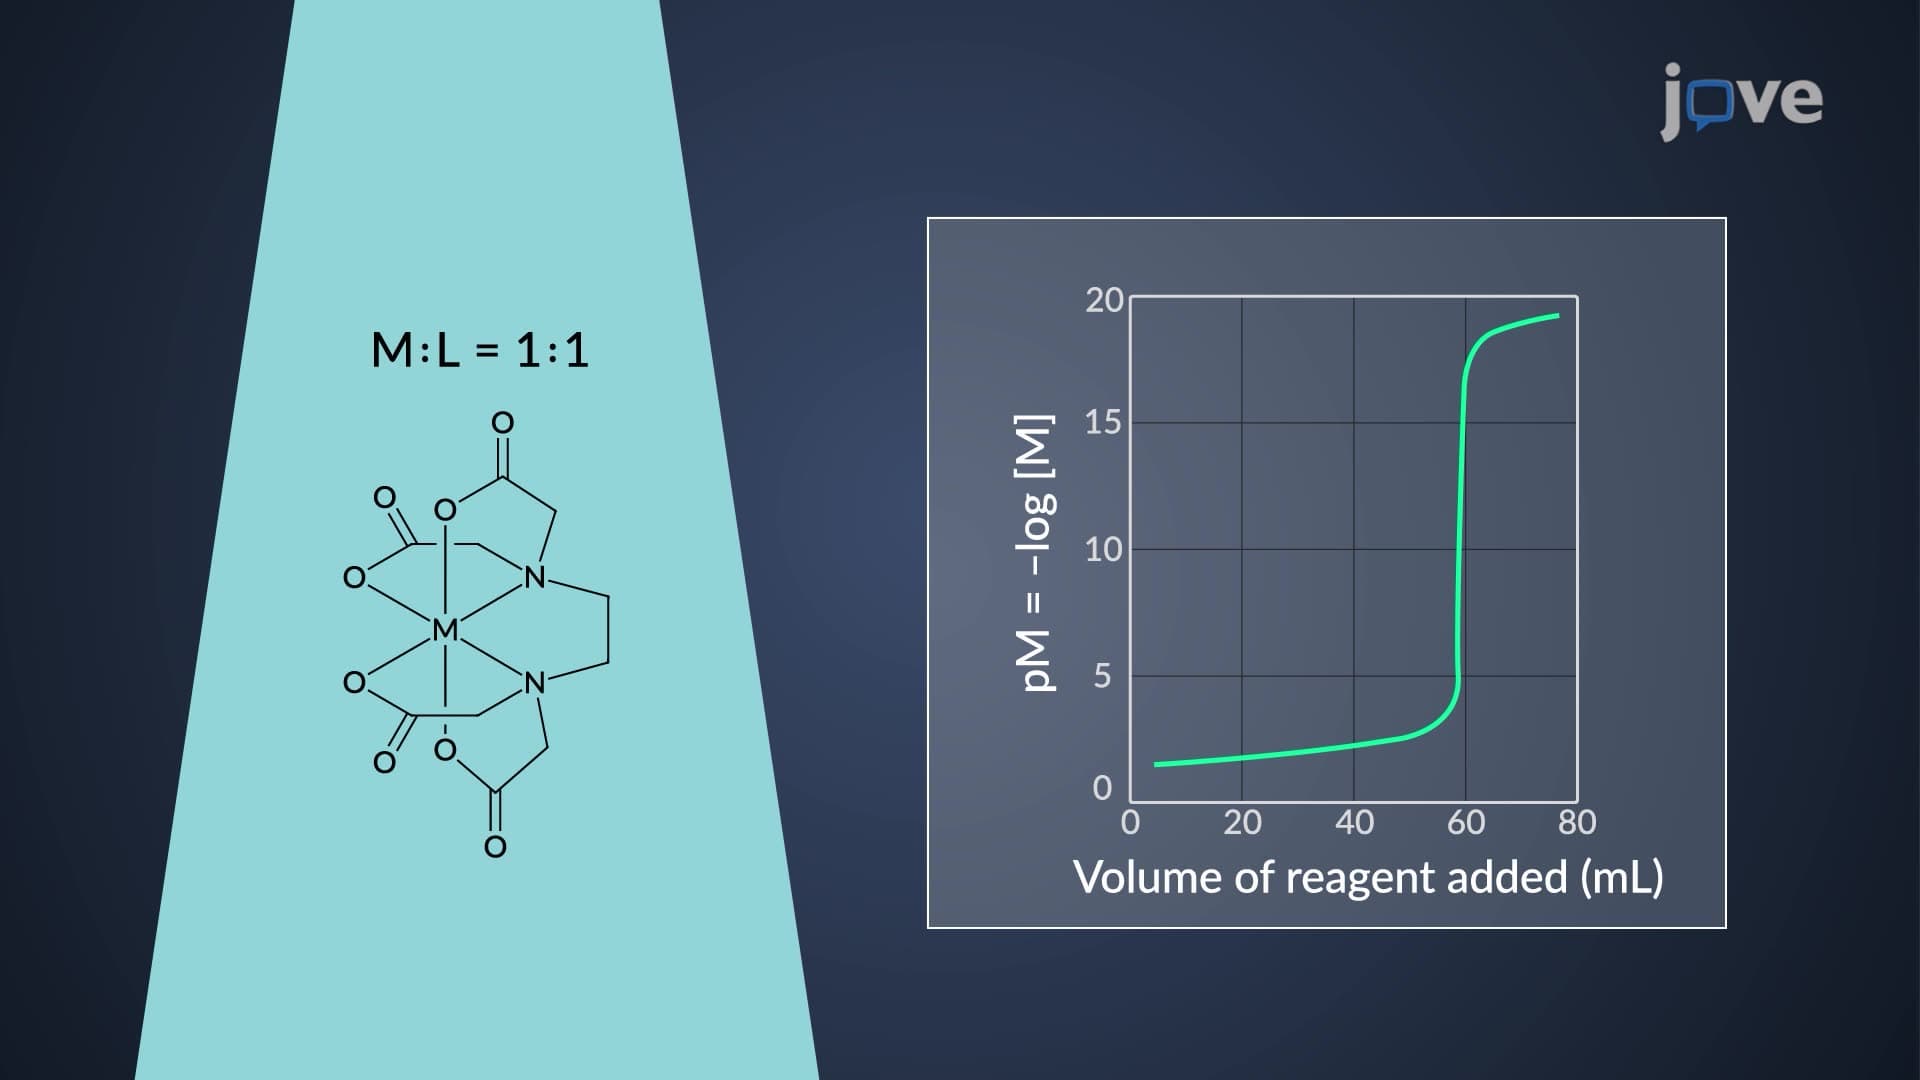
Complexometric Titration: Ligands
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...