
Clinical and genetic analysis of combined oxidative phosphorylation defificiency-10 caused by MTO1 mutation
Chaofan Zhou1, Jin Wang2, Qinle Zhang1
1Guangxi Key Laboratory of Reproductive Health and Birth Defect Prevention, Maternal and Child Health Hospital of Guangxi Zhuang Autonomous Region, Nanning, Guangxi 530003, China; Guangxi Key Laboratory of Precision Medicine for Genetic Diseases, Maternal and Child Health Hospital of Guangxi Zhuang Autonomous Region, Nanning, Guangxi 530003, China; Guangxi Key Laboratory of Birth Defects Research and Prevention, Maternal and Child Health Hospital of Guangxi Zhuang Autonomous Region, Nanning, Guangxi 530003, China; Guangxi Key Laboratory of Birth Defects and Stem Cell Biobank, Maternal and Child Health Hospital of Guangxi Zhuang Autonomous Region, Nanning, Guangxi 530003, China; Genetic and Metabolic Central Laboratory, Birth Defects Prevention and Control Institute of Guangxi Zhuang Autonomous Region, Nanning, Guangxi 530003, China; Guangxi Clinical Research Center for Pediatric Diseases, China.

Abstract:
The mitochondrial translation optimization factor 1(MTO1) gene mutations had been reported to be linked to combined oxidative phosphorylation defificiency-10 (COXPD10). In this study, we presented the detailed clinical features and genetic analysis of the patient with two variants in MTO1, and reviewed 42 different cases available in publications. Whole exome sequencing and bioinformatics analysis were employed to detect the genetic variants of a 6-month-old boy with metabolic disorder and multiple organ failure; Sanger sequencing was performed to confirm the origin of variants; and clinical data of the patients was retrospectively collected and analyzed. Variant classification was followed to ACMG guidline. The proband was diagnosed with multiple organ failure, severe pneumonia, sepsis, hyperlactatemia, metabolic acidosis, and moderate anemia. Compound heterozygous mutations in the coding region of MTO1 gene (c.1291C > T/p.Arg431Trp and c.1390C > T/p.Arg464Cys) were identified, and the results of family verification experiment showed that the mutations were inherited from the parents, respectively. Combined with clinical symptoms, the patient was diagnosed as COXPD10. In summary, hallmark features of MTO1 mutations were lactic acidosis and hypertrophic cardiomyopathy. Of note, patients with the same genetic mutation may not have the same clinical presentation. Additional MTO1 defificiency cases will help to make genotype-phenotype correlations clearer.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:19
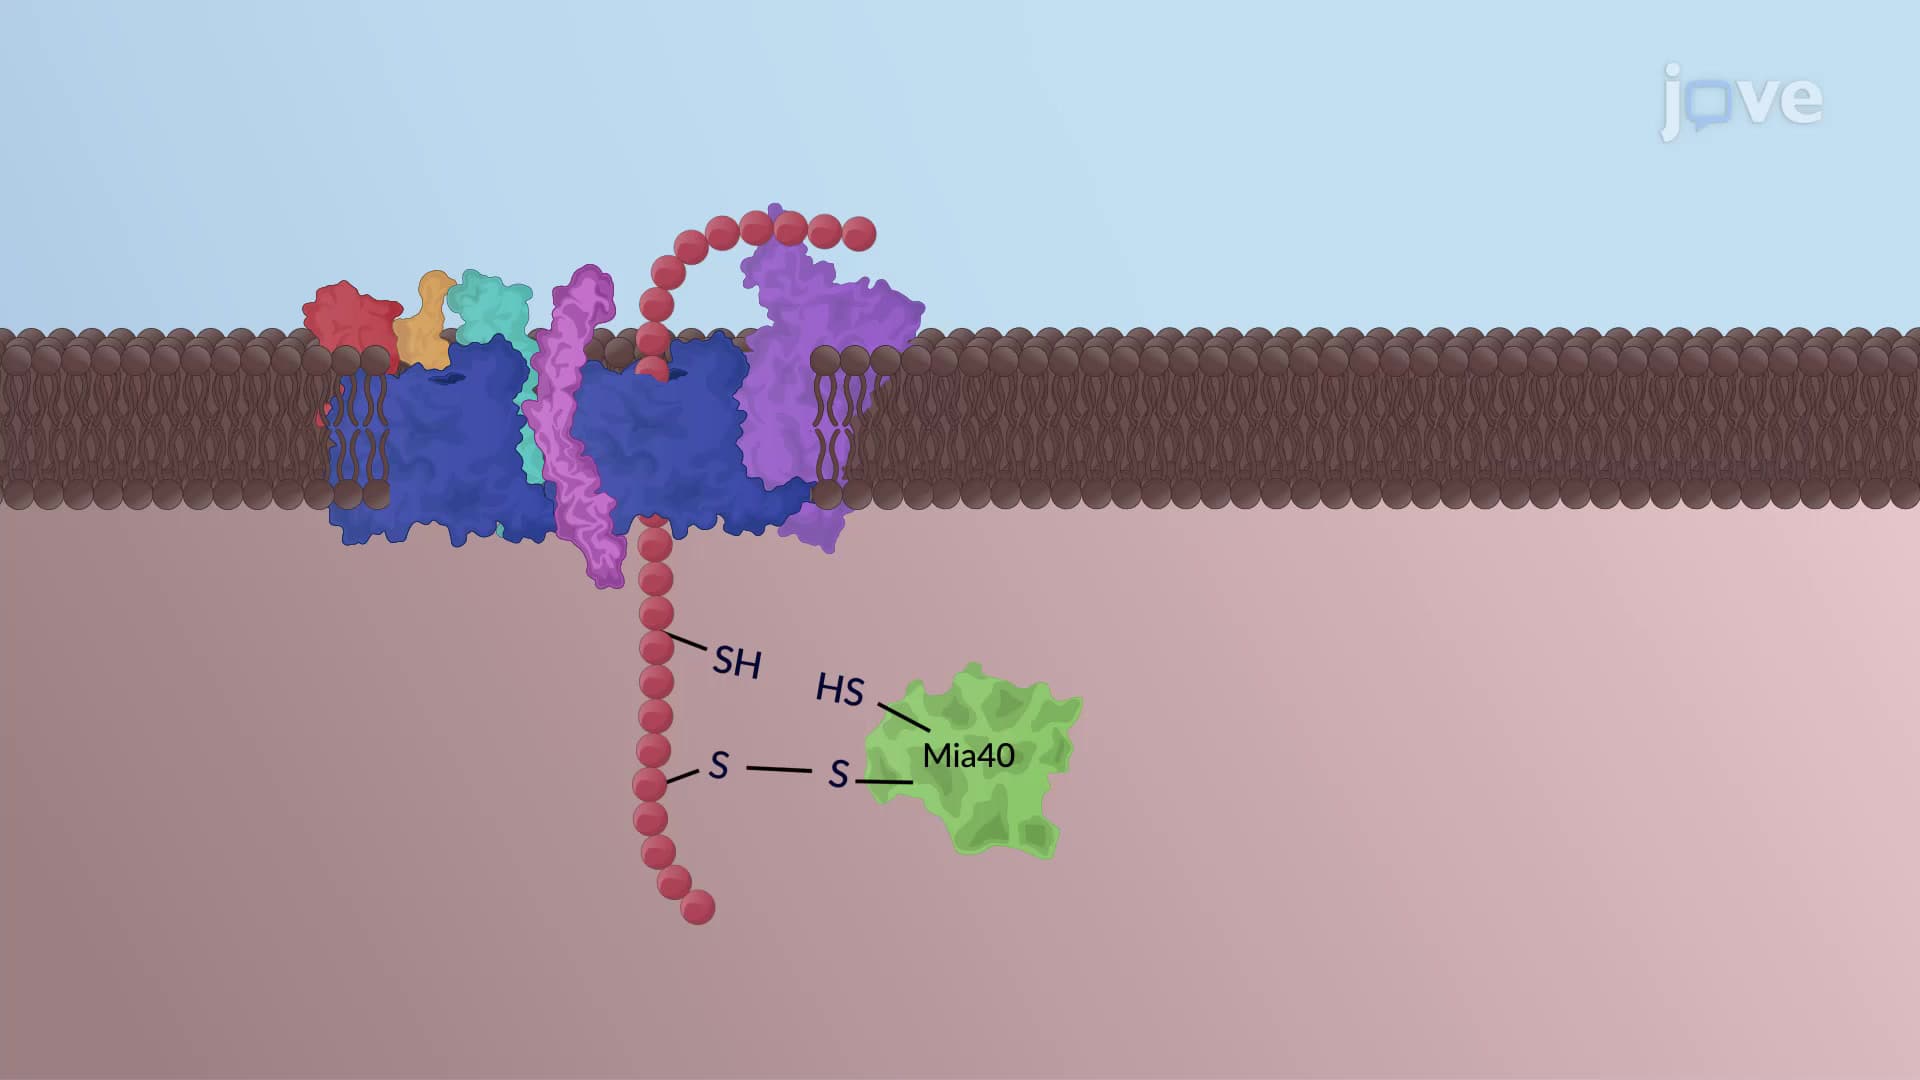
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...