Related Experiment Video
Updated: Oct 6, 2025

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Recent Advances in First-Principles Based Molecular Dynamics
François Mouvet1, Justin Villard1, Viacheslav Bolnykh1
1Laboratory of Computational Chemistry and Biochemistry, Institute of Chemical Sciences and Engineering, École Polytechnique Fédérale de Lausanne, CH-1015 Lausanne, Switzerland.

Abstract:
First-principles molecular dynamics (FPMD) and its quantum mechanical-molecular mechanical (QM/MM) extensions are powerful tools to follow the real-time dynamics of a broad variety of systems in their ground as well as electronically excited states. The continued advances in computational power have enabled simulations of QM regions of larger sizes for more extended time scales. In addition, development of the parallel algorithms has boosted the performance of QM/MM methods even on existing computer architectures. In the case of density functional-based FPMD, systems of several hundreds to thousands of atoms can now be customarily simulated for tens to hundreds of picoseconds. In spite of this progress, the time scale limitations remain severe, especially when high-rung exchange-correlation functionals or high-level wave function based quantum mechanical methods are used. To ameliorate this, a large number of enhanced sampling methods have been introduced but most of the approaches that have been developed to increase the efficiency of FPMD based simulations sacrifice the real-time dynamics in favor of enhancing sampling. Here, we present some recent advances in boosting the efficiency of FPMD based simulations while keeping the full dynamic information. These include a highly efficient recent implementation of FPMD-based QM/MM simulations that not only enables fully flexible combinations of different electronic structure methods and force fields via a highly efficient communication library, it also fully exploits parallelism for both quantum and classical descriptions. The second type of acceleration methods we discuss is a large family of specially devised multiple-time-step algorithms that make use of suitable breakups of the total nuclear forces into fast components that can be calculated via lower level methods and slowly varying correction forces evaluated with a high-level method at long time intervals. The computational gain of this scheme mostly depends on the cost difference between the two methods and advantageous combinations can yield large speedups without compromising the accuracy of the high-level method. And finally, the third class of FPMD acceleration methods presented here are machine learning models to accelerated FPMD and their powerful combinations with multiple-time-step techniques. The combination of all the approaches enables substantial speedups of FPMD simulations of several orders of magnitude while fully preserving the real-time dynamics and accuracy.

Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:00
Molecular Models
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 01:12
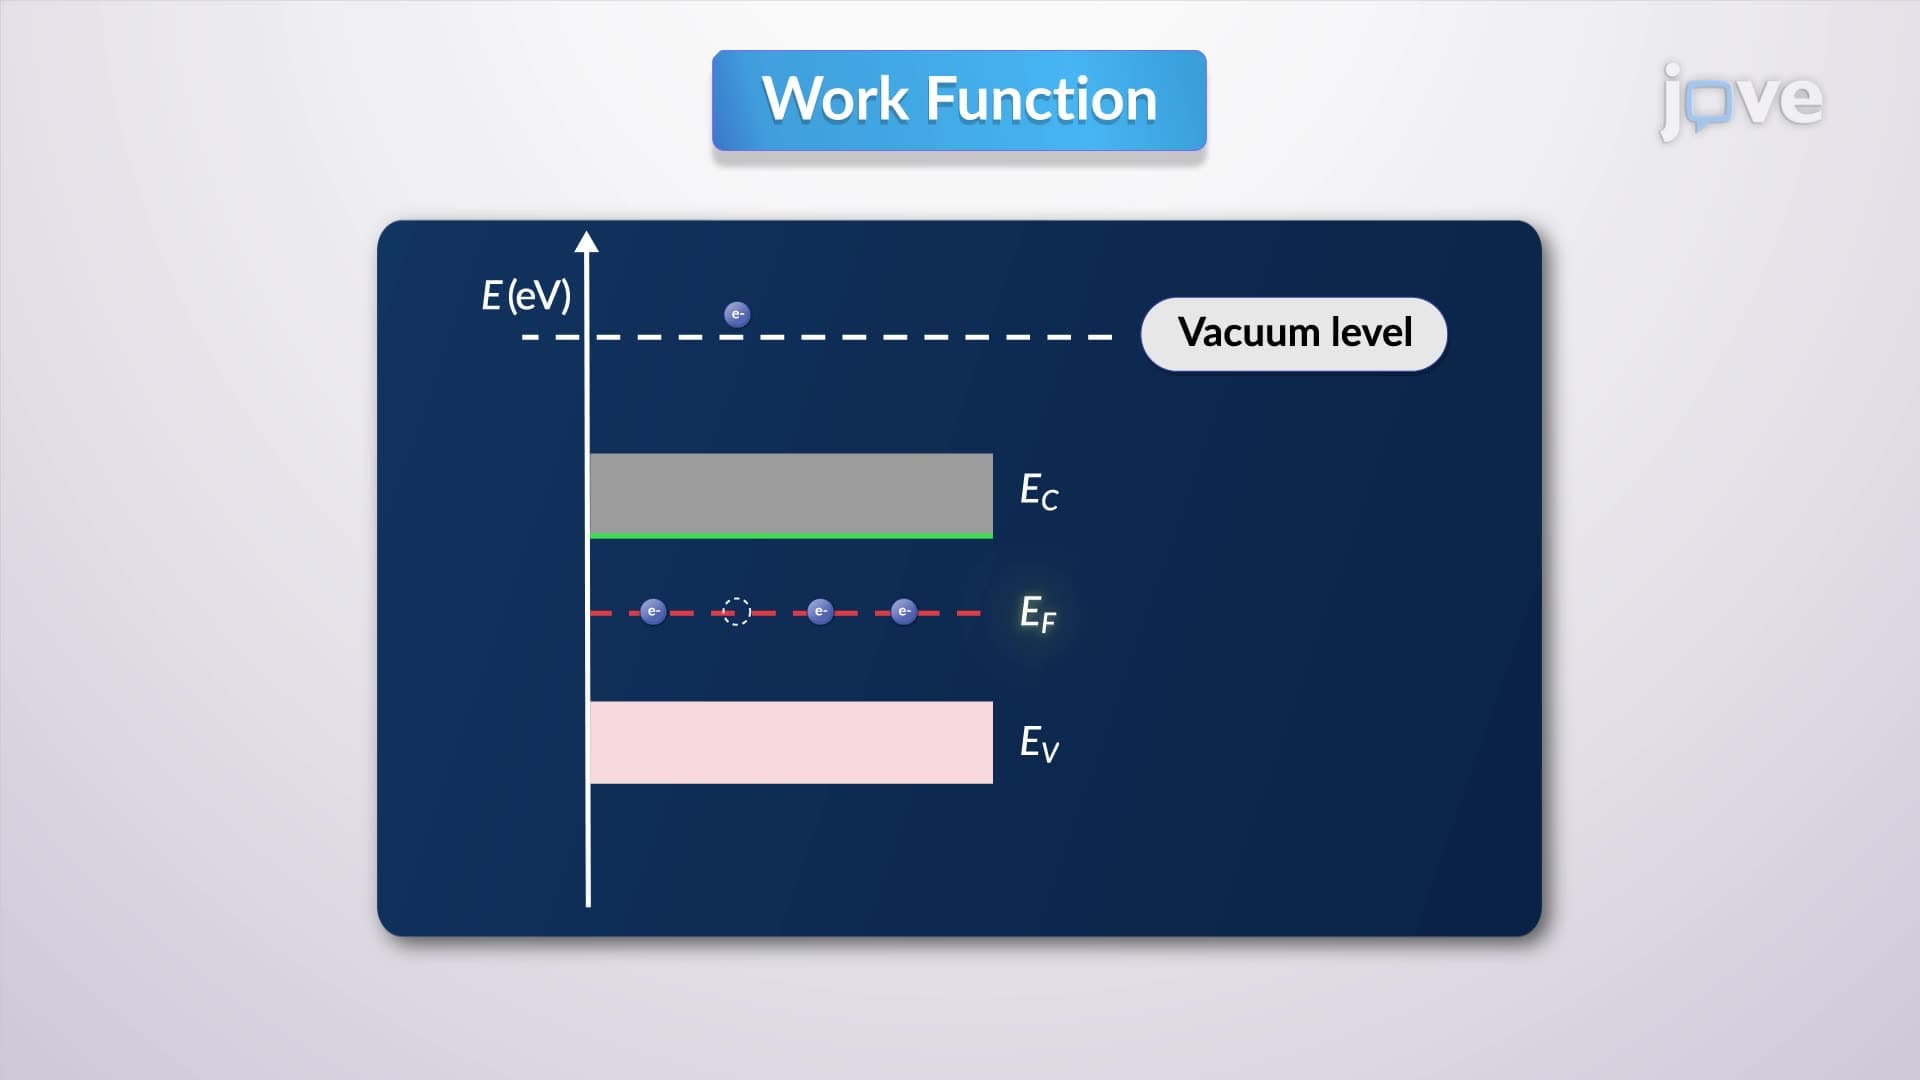
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...