
The Effect of Geometry, Spin, and Orbital Optimization in Achieving Accurate, Correlated Results for Iron-Sulfur
Carlos Mejuto-Zaera1,2, Demeter Tzeli3,4, David Williams-Young2
1University of California, Berkeley, California 94720, United States.

Abstract:
Iron-sulfur clusters comprise an important functional motif in the catalytic centers of biological systems, capable of enabling important chemical transformations at ambient conditions. This remarkable capability derives from a notoriously complex electronic structure that is characterized by a high density of states that is sensitive to geometric changes. The spectral sensitivity to subtle geometric changes has received little attention from correlated, large active space calculations, owing partly to the exceptional computational complexity for treating these large and correlated systems accurately. To provide insight into this aspect, we report the first Complete Active Space Self Consistent Field (CASSCF) calculations for different geometries of the [Fe(II/III)4S4(SMe)4]-2 clusters using two complementary, correlated solvers: spin-pure Adaptive Sampling Configuration Interaction (ASCI) and Density Matrix Renormalization Group (DMRG). We find that the previously established picture of a double-exchange driven magnetic structure, with minute energy gaps (<1 mHa) between consecutive spin states, has a weak dependence on the underlying geometry. However, the spin gap between the singlet and the spin state 2S + 1 = 19, corresponding to a maximal number of Fe-d electrons being unpaired and of parallel spin, is strongly geometry dependent, changing by a factor of 3 upon slight deformations that are still within biologically relevant parameters. The CASSCF orbital optimization procedure, using active spaces as large as 86 electrons in 52 orbitals, was found to reduce this gap compared to typical mean-field orbital approaches. Our results show the need for performing large active space calculations to unveil the challenging electronic structure of these complex catalytic centers and should serve as accurate starting points for fully correlated treatments upon inclusion of dynamical correlation outside the active space.
More Related Videos
 04:09
Demonstrating the Simplicity and In Situ Temperature Monitoring of the Mechanochemical Synthesis of Metal Chalcogenides Suitable for Thermoelectrics
Published on: August 30, 2024
 10:57
Synthesis and Performance Characterizations of Transition Metal Single Atom Catalyst for Electrochemical CO2 Reduction
Published on: April 10, 2018
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:22
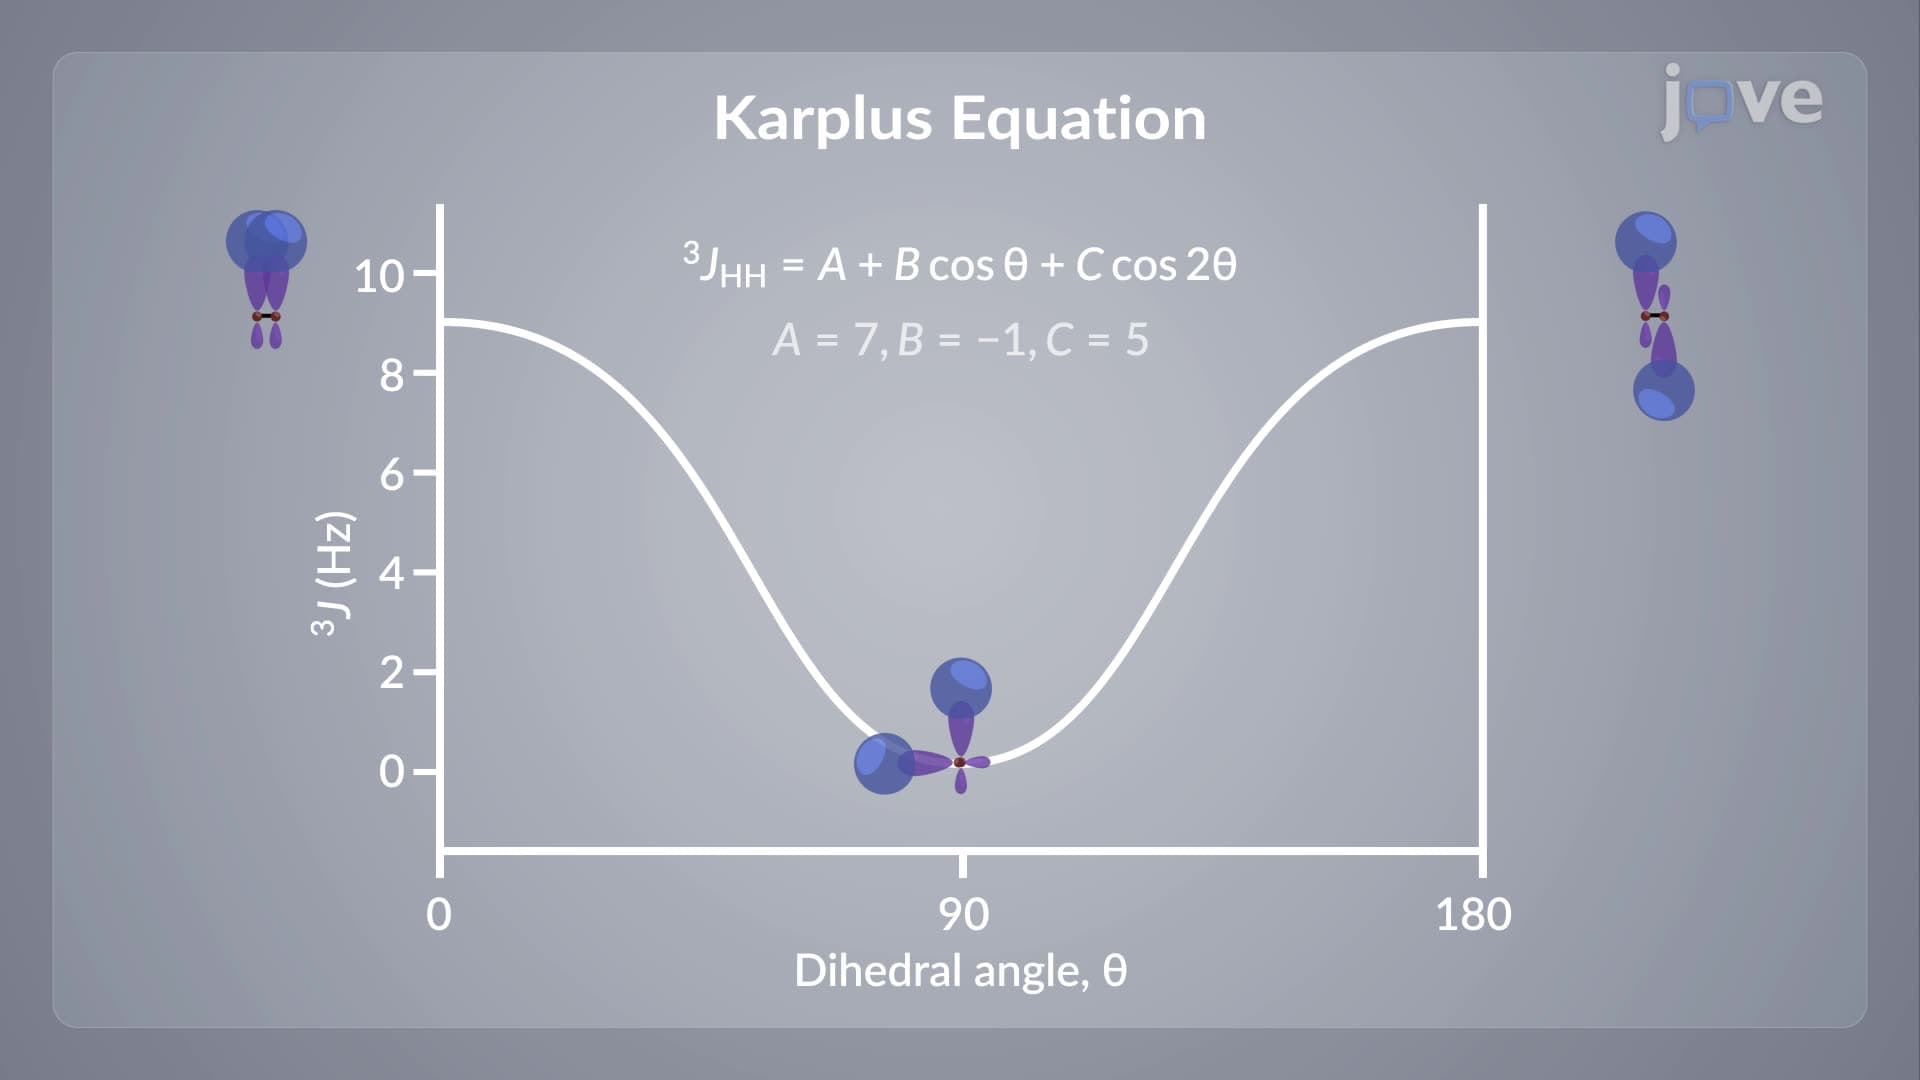
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:08
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must...
 02:42
Valence Bond Theory
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...