Related Experiment Video
Updated: Oct 6, 2025


Selecting Multiple Biomarker Subsets with Similarly Effective Binary Classification Performances
Published on: October 11, 2018
Finding Maximal Exact Matches Using the r-Index
Massimiliano Rossi1, Marco Oliva1, Paola Bonizzoni2
1Department of Computer and Information Science and Engineering, University of Florida, Gainesville, Florida, USA.

Abstract:
Efficiently finding maximal exact matches (MEMs) between a sequence read and a database of genomes is a key first step in read alignment. But until recently, it was unknown how to build a data structure in [Formula: see text] space that supports efficient MEM finding, where r is the number of runs in the Burrows-Wheeler Transform. In 2021, Rossi et al. showed how to build a small auxiliary data structure called thresholds in addition to the r-index in [Formula: see text] space. This addition enables efficient MEM finding using the r-index. In this article, we present the tool that implements this solution, which we call MONI. Namely, we give a high-level view of the main components of the data structure and show how the source code can be downloaded, compiled, and used to find MEMs between a set of sequence reads and a set of genomes.

Related Concept Videos
 01:09
Wilcoxon Signed-Ranks Test for Matched Pairs
 01:21
Friedman Two-way Analysis of Variance by Ranks
 01:02
Ranks
 01:02
Quantifying and Rejecting Outliers: The Grubbs Test
 01:11
Introduction to R
 01:19
Routh-Hurwitz Criterion II
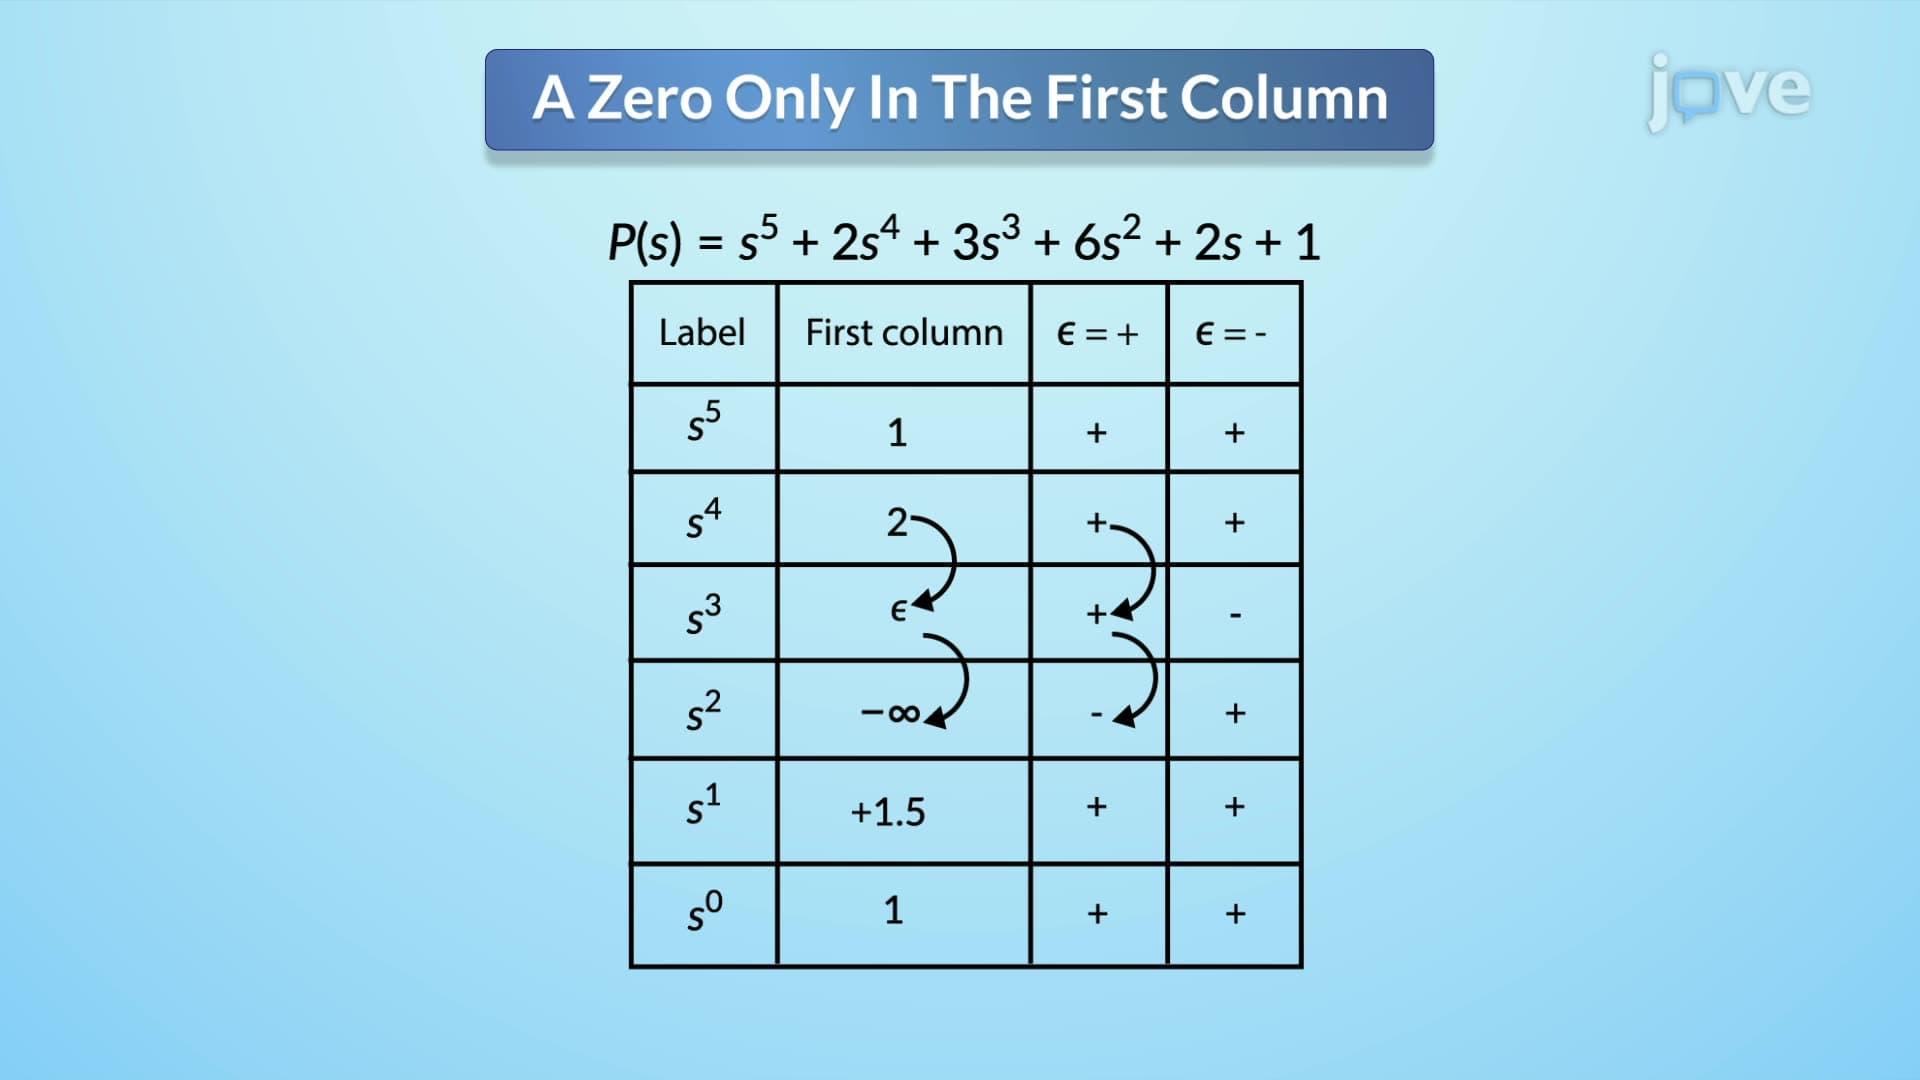
The first scenario occurs when a singular zero appears in the first column of the Routh table. This situation creates a division by zero issues. To resolve this, a small positive or negative number, denoted as epsilon (∈), is substituted for the zero. The stability analysis proceeds by assuming a sign for ∈. If ∈ is positive, any sign change in the first...