Related Experiment Video
Updated: Jun 23, 2026
09:28
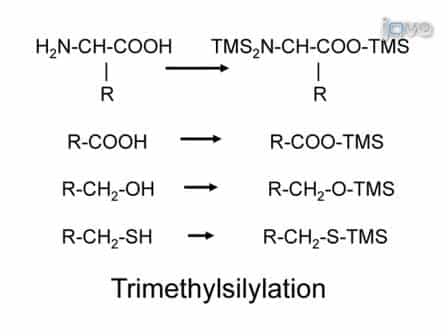
One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Correction: Spenger et al. Glutaric Aciduria Type I Missed by Newborn Screening: Report of Four Cases from Three
Johannes Spenger1, Esther M Maier2, Katharina Wechselberger3
1Department of Pediatrics, Salzburger Landeskliniken (SALK) and Paracelsus Medical University (PMU), 5020 Salzburg, Austria.
International Journal of Neonatal Screening
|January 25, 2022

Abstract:
There was an error in the original publication [...].
Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Cells contain membrane-bound organelles called peroxisomes that oxidize organic molecules by transferring hydrogen atoms to oxygen, producing hydrogen peroxide. Peroxisomes enzymatically convert the released hydrogen peroxide into water and oxygen.
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:27
Glucose Transporters
Glucose transporters facilitate the transport of glucose across the cell membrane. In addition to glucose, some glucose transporters can also aid the movement of other hexoses such as fructose, mannose, and galactose.
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:20
Inborn Errors of Metabolism
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...