Gene Therapy Potential for Genetic Disorders of Surfactant Dysfunction
Related Experiment Video
Updated: Oct 5, 2025
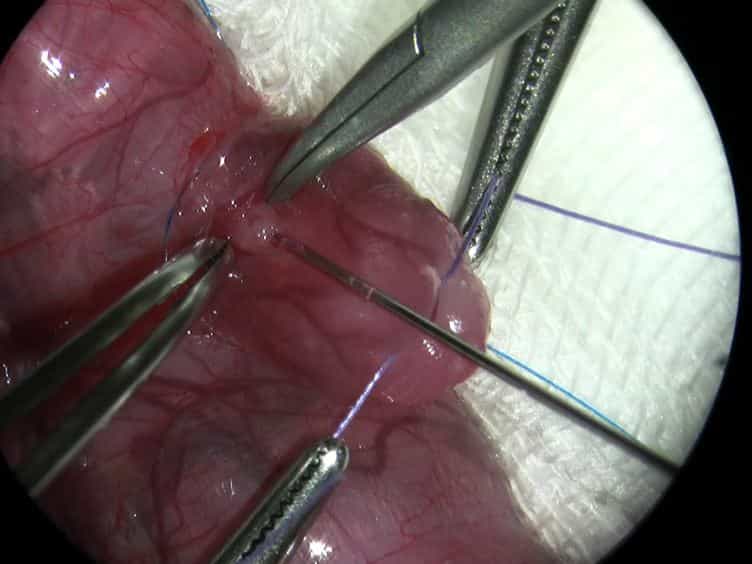
A Novel Surgical Approach for Intratracheal Administration of Bioactive Agents in a Fetal Mouse Model
Published on: October 31, 2012
Ashley L Cooney1,2, Jennifer A Wambach3, Patrick L Sinn1,2
1Department of Pediatrics, The University of Iowa, Iowa City, IA, United States.

Insights
Severe respiratory distress syndrome in infants can stem from genetic causes affecting pulmonary surfactant proteins SP-B, SP-C, and ABCA3. Gene therapy offers a promising treatment approach for these monogenic lung diseases.
Area of Science:
- Pulmonary medicine
- Genetics
- Molecular biology
Background:
- Pulmonary surfactant is crucial for lung function, preventing alveolar collapse.
- Severe respiratory distress syndrome (RDS) in term infants often indicates genetic causes related to surfactant dysfunction.
- Pathogenic variants in SFTPB, SFTPC, and ABCA3 genes lead to severe neonatal RDS or childhood interstitial lung disease (chILD).
Purpose of the Study:
- To review gene therapy strategies for monogenic lung diseases caused by surfactant dysfunction.
- To explore viral vector delivery systems for targeting alveolar epithelial type II cells (AEC2).
- To compare gene addition and gene editing approaches for treating SFTPB, SFTPC, and ABCA3 related lung disorders.
Main Methods:
- Review of adeno-associated virus (AAV), lentiviral, and adenoviral (Ad) vectors for gene delivery.
- Comparison of gene addition and gene editing therapeutic strategies.
- Focus on targeting alveolar epithelial type II cells (AEC2) for gene therapy.
Main Results:
- Gene therapy presents a promising avenue for treating genetic surfactant dysfunction disorders.
- Viral vector design and cell tropism are key challenges in developing effective lung gene therapies.
- Both gene addition and gene editing show potential for correcting mutations in SFTPB, SFTPC, and ABCA3.
Conclusions:
- Genetic disorders affecting pulmonary surfactant proteins (SP-B, SP-C, ABCA3) are significant causes of neonatal RDS and chILD.
- Gene therapy, utilizing various viral vectors and strategies, holds promise for treating these monogenic lung diseases.
- Further research into vector optimization and delivery is essential for successful clinical translation.
Abstract:
Pulmonary surfactant is critically important to prevent atelectasis by lowering the surface tension of the alveolar lining liquid. While respiratory distress syndrome (RDS) is common in premature infants, severe RDS in term and late preterm infants suggests an underlying genetic etiology. Pathogenic variants in the genes encoding key components of pulmonary surfactant including surfactant protein B (SP-B, SFTPB gene), surfactant protein C (SP-C, SFTPC gene), and the ATP-Binding Cassette transporter A3 (ABCA3, ABCA3 gene) result in severe neonatal RDS or childhood interstitial lung disease (chILD). These proteins play essential roles in pulmonary surfactant biogenesis and are expressed in alveolar epithelial type II cells (AEC2), the progenitor cell of the alveolar epithelium. SP-B deficiency most commonly presents in the neonatal period with severe RDS and requires lung transplantation for survival. SFTPC mutations act in an autosomal dominant fashion and more commonly presents with chILD or idiopathic pulmonary fibrosis than neonatal RDS. ABCA3 deficiency often presents as neonatal RDS or chILD. Gene therapy is a promising option to treat monogenic lung diseases. Successes and challenges in developing gene therapies for genetic disorders of surfactant dysfunction include viral vector design and tropism for target cell types. In this review, we explore adeno-associated virus (AAV), lentiviral, and adenoviral (Ad)-based vectors as delivery vehicles. Both gene addition and gene editing strategies are compared to best design treatments for lung diseases resulting from pathogenic variants in the SFTPB, SFTPC, and ABCA3 genes.
More Related Videos
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
 10:15
Conditional Reprogramming of Pediatric Human Esophageal Epithelial Cells for Use in Tissue Engineering and Disease Investigation
Published on: March 22, 2017
Related Concept Videos
 00:59
Gene Therapy
 01:05
Breathing
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 00:49
What is Genetic Engineering?
 01:20
Inborn Errors of Metabolism