Related Experiment Video
Updated: Oct 4, 2025


Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
ALDPI: adaptively learning importance of multi-scale topologies and multi-modality similarities for drug-protein
Kaimiao Hu1, Hui Cui2, Tiangang Zhang3
1School of Computer Science and Technology, Heilongjiang University, Harbin 150080, China.

Motivation:
Effective computational methods to predict drug-protein interactions (DPIs) are vital for drug discovery in reducing the time and cost of drug development. Recent DPI prediction methods mainly exploit graph data composed of multiple kinds of connections among drugs and proteins. Each node in the graph usually has topological structures with multiple scales formed by its first-order neighbors and multi-order neighbors. However, most of the previous methods do not consider the topological structures of multi-order neighbors. In addition, deep integration of the multi-modality similarities of drugs and proteins is also a challenging task.
Results:
We propose a model called ALDPI to adaptively learn the multi-scale topologies and multi-modality similarities with various significance levels. We first construct a drug-protein heterogeneous graph, which is composed of the interactions and the similarities with multiple modalities among drugs and proteins. An adaptive graph learning module is then designed to learn important kinds of connections in heterogeneous graph and generate new topology graphs. A module based on graph convolutional autoencoders is established to learn multiple representations, which imply the node attributes and multiple-scale topologies composed of one-order and multi-order neighbors, respectively. We also design an attention mechanism at neighbor topology level to distinguish the importance of these representations. Finally, since each similarity modality has its specific features, we construct a multi-layer convolutional neural network-based module to learn and fuse multi-modality features to obtain the attribute representation of each drug-protein node pair. Comprehensive experimental results show ALDPI's superior performance over six state-of-the-art methods. The results of recall rates of top-ranked candidates and case studies on five drugs further demonstrate the ability of ALDPI to discover potential drug-related protein candidates.
Contact:
zhang@hlju.edu.cn.
More Related Videos
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
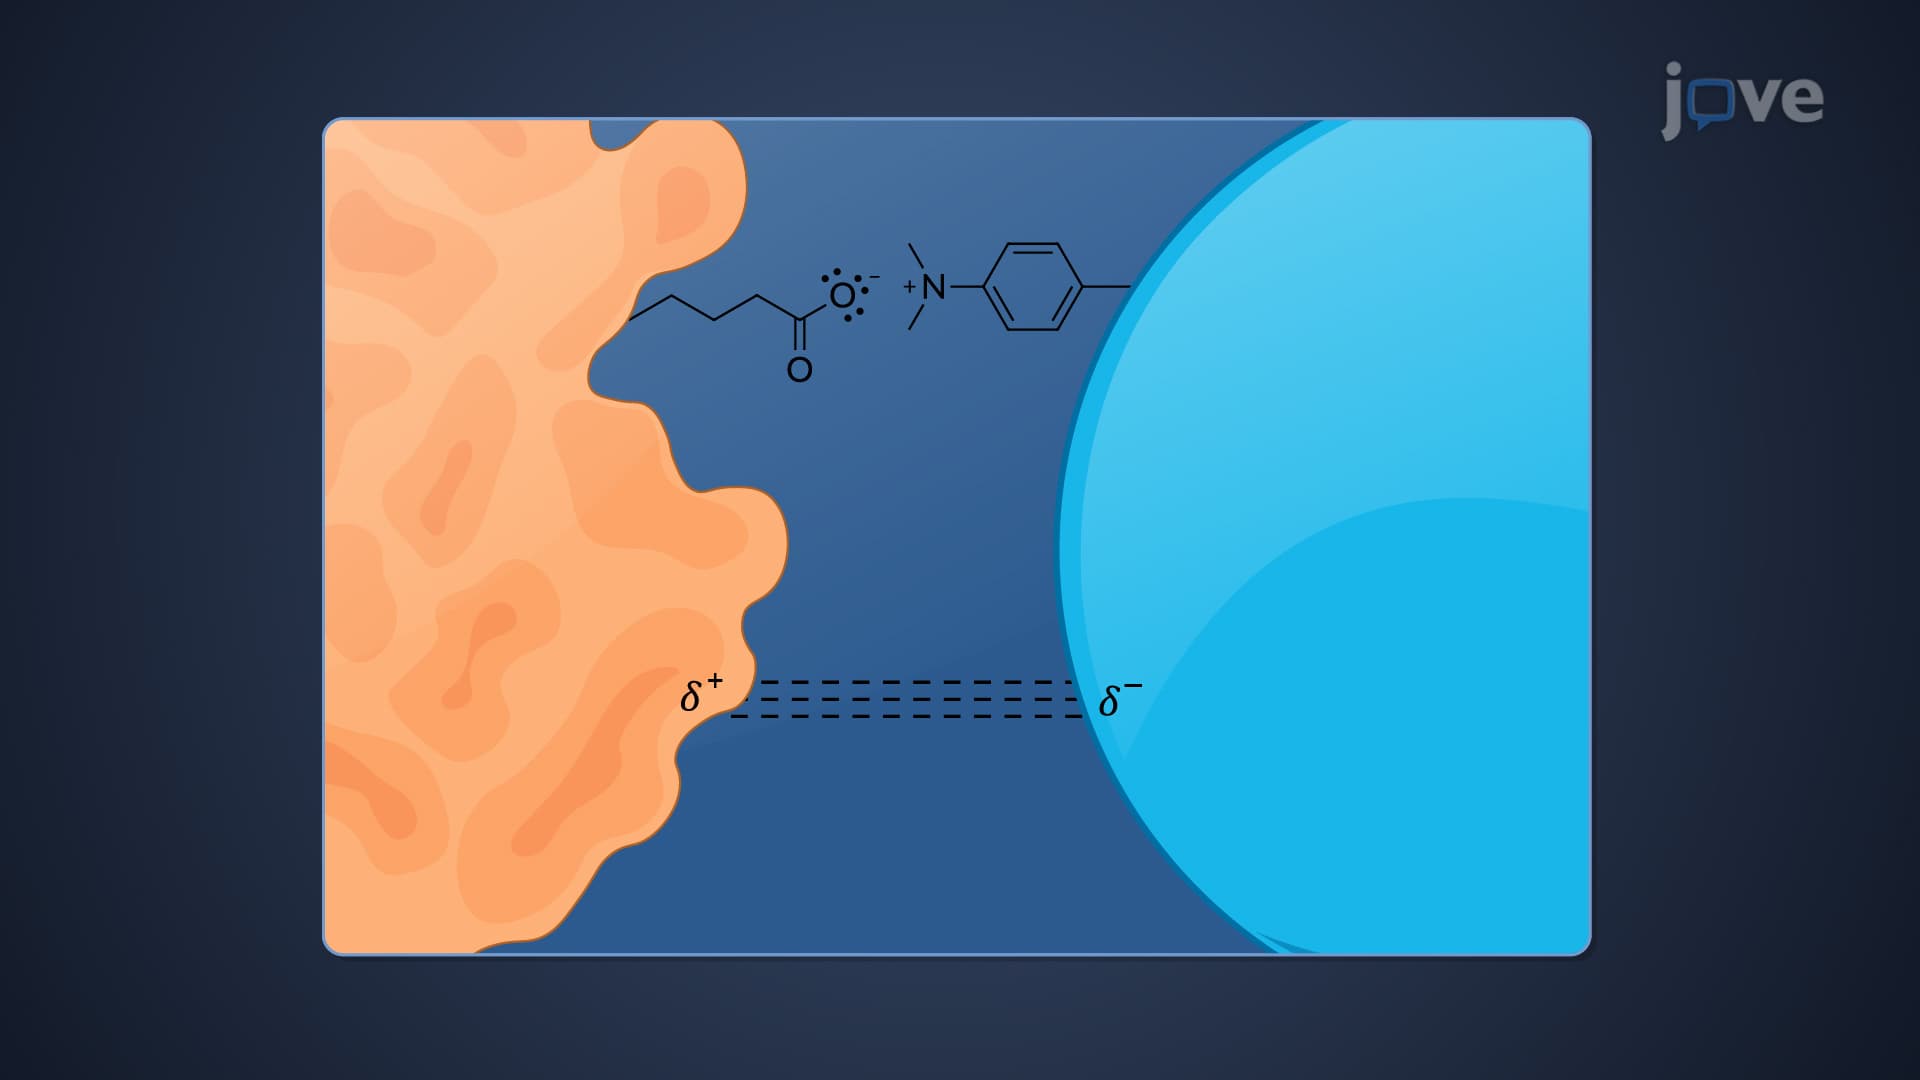
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...