Related Experiment Video
Updated: Oct 4, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Reconstruction of the unbinding pathways of noncovalent SARS-CoV and SARS-CoV-2 3CLpro inhibitors using unbiased
Fereshteh Noroozi Tiyoula1, Hassan Aryapour1
1Department of Biology, Faculty of Science, Golestan University, Gorgan, Iran.

Abstract:
The main protease (3CLpro) is one of the essential components of the SARS-CoVs viral life cycle, which makes it an interesting target for overpowering these viruses. Although many covalent and noncovalent inhibitors have been designed to inhibit this molecular target, none have gained FDA approval as a drug. Because of the high rate of COVID-19 pandemic development, in addition to laboratory research, we require in silico methods to accelerate rational drug design. The unbinding pathways of two SARS-CoV and SARS-CoV-2 3CLpro noncovalent inhibitors with the PDB IDs: 3V3M, 4MDS, 6W63, 5RF7 were explored from a comparative perspective using unbiased molecular dynamics (UMD) simulations. We uncovered common weak points for selected inhibitors that could not interact significantly with a binding pocket at specific residues by all their fragments. So water molecules entered the free binding S regions and weakened protein-inhibitor fundamental interactions gradually. N142, G143, and H163 are the essential residues, which cause key protein-ligand interactions in the binding pocket. We believe that these results will help design new potent inhibitors against SARS-CoV-2.
Related Concept Videos
 01:16
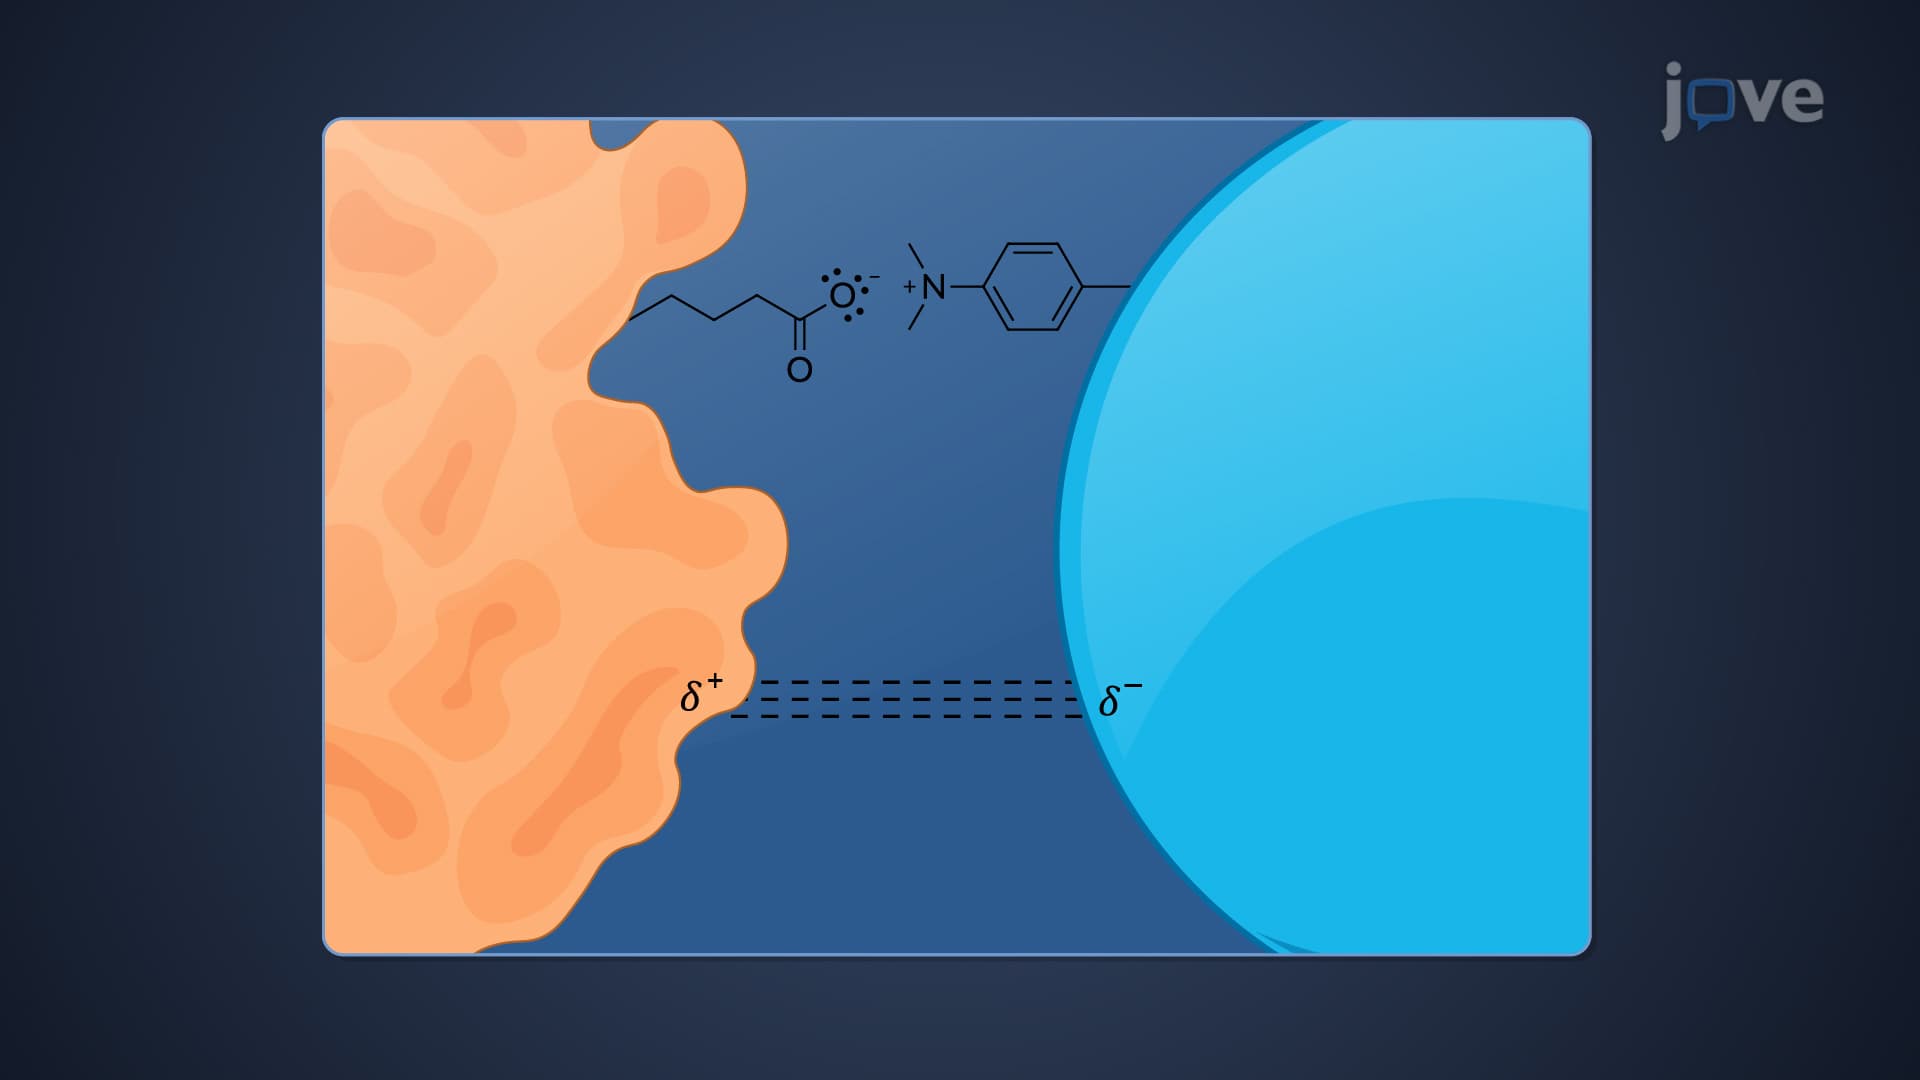
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...