Towards Elucidating Structure-Spectra Relationships in Rhamnogalacturonan II: Computational Protocols for Accurate 13C and 1H Shifts for Apiose and Its Borate Esters
Related Experiment Video
Updated: Oct 4, 2025
Biochemical and Structural Characterization of the Carbohydrate Transport Substrate-binding-protein SP0092
Published on: October 2, 2017
Vivek S Bharadwaj1, Luke P Westawker1, Michael F Crowley1
1Renewable Resources and Enabling Sciences Center, National Renewable Energy Laboratory, Golden, CO, United States.

Abstract:
Apiose is a naturally occurring, uncommon branched-chain pentose found in plant cell walls as part of the complex polysaccharide Rhamnogalacturonan II (RG-II). The structural elucidation of the three-dimensional structure of RG-II by nuclear magnetic resonance (NMR) spectroscopy is significantly complicated by the ability of apiose to cross-link via borate ester linkages to form RG-II dimers. Here, we developed a computational approach to gain insight into the structure-spectra relationships of apio-borate complexes in an effort to complement experimental assignments of NMR signals in RG-II. Our protocol involved structure optimizations using density functional theory (DFT) followed by isotropic magnetic shielding constant calculations using the gauge-invariant atomic orbital (GIAO) approach to predict chemical shifts. We evaluated the accuracy of 23 different functional-basis set (FBS) combinations with and without implicit solvation for predicting the experimental 1H and 13C shifts of a methyl apioside and its three borate derivatives. The computed NMR predictions were evaluated on the basis of the overall shift accuracy, relative shift ordering, and the ability to distinguish between dimers and monomers. We demonstrate that the consideration of implicit solvation during geometry optimizations in addition to the magnetic shielding constant calculations greatly increases the accuracy of NMR chemical shift predictions and can correctly reproduce the ordering of the 13C shifts and yield predictions that are, on average, within 1.50 ppm for 13C and 0.12 ppm for 1H shifts for apio-borate compounds.
Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 00:58
¹H NMR Chemical Shift Equivalence: Enantiotopic and Diastereotopic Protons
In chiral compounds such as 2-butanol, replacing the methylene hydrogens at C3 produces a pair of...
 01:02
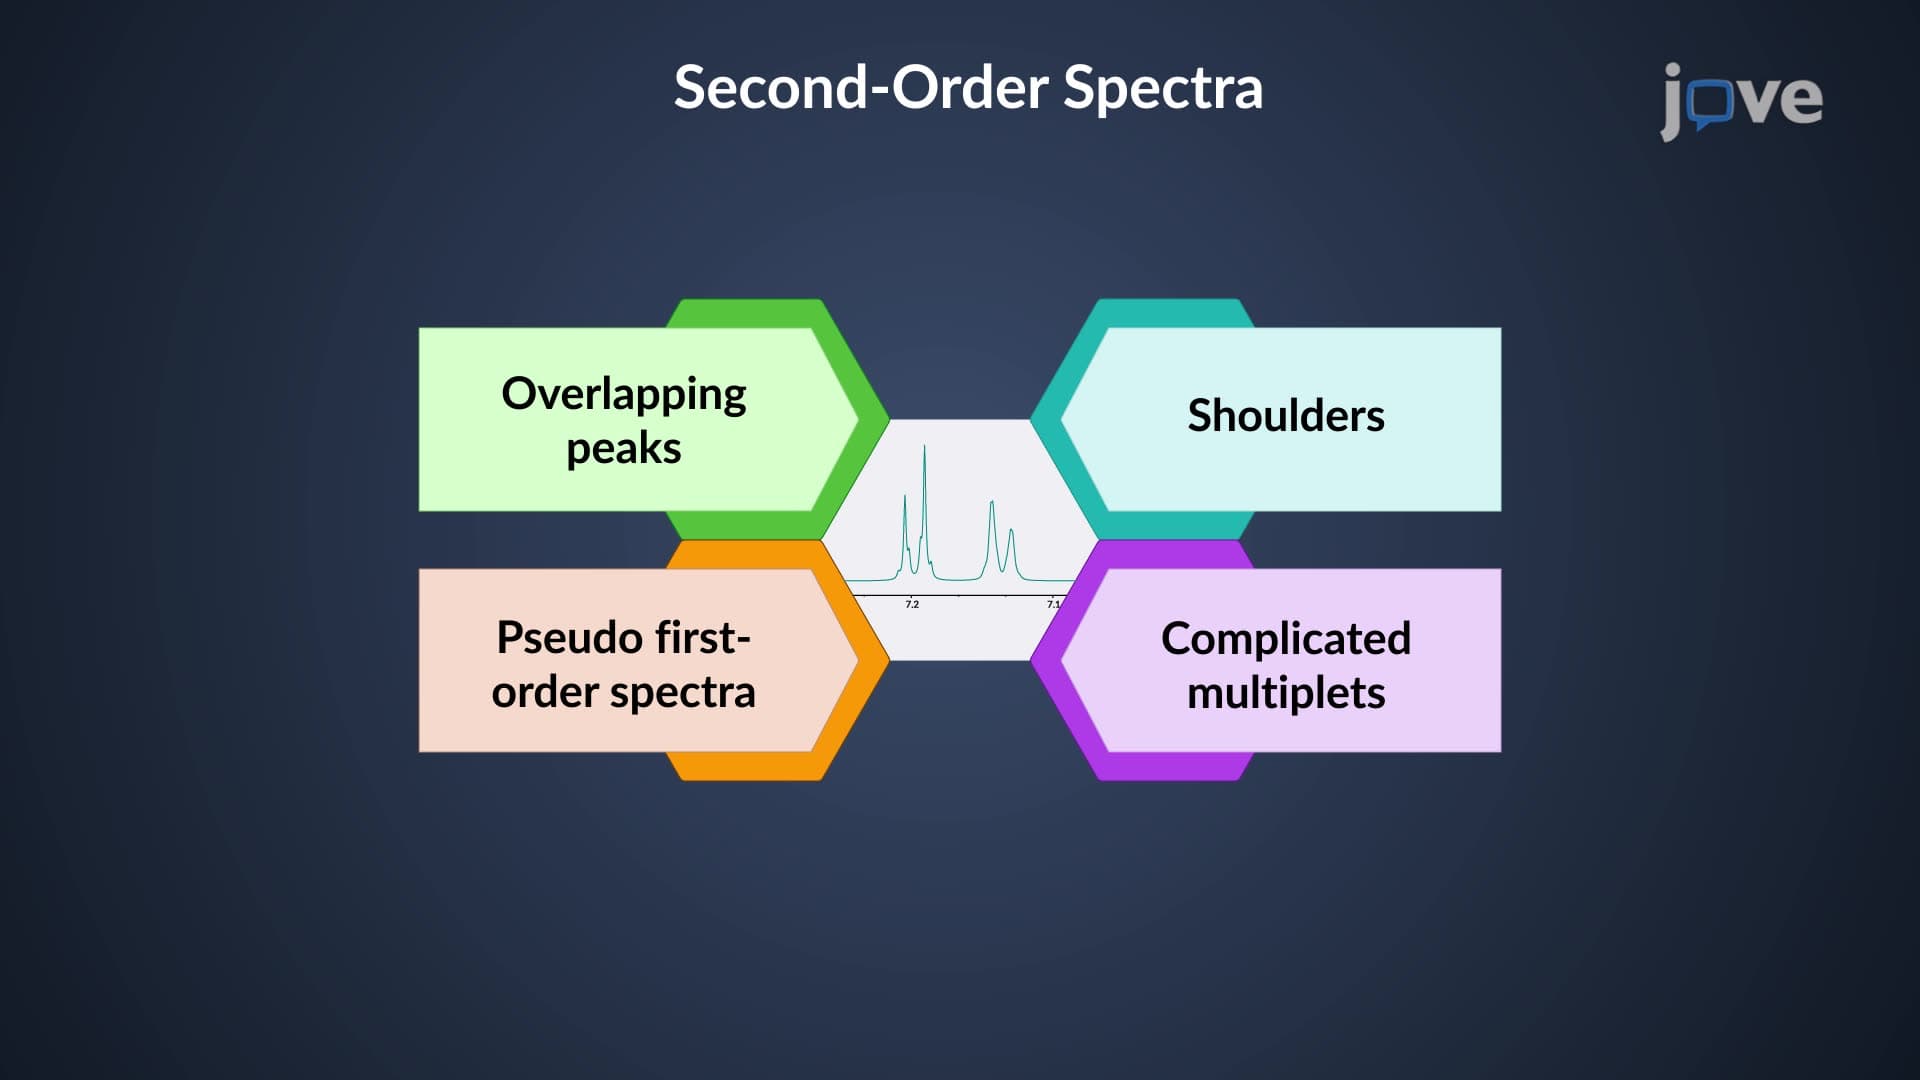
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:15
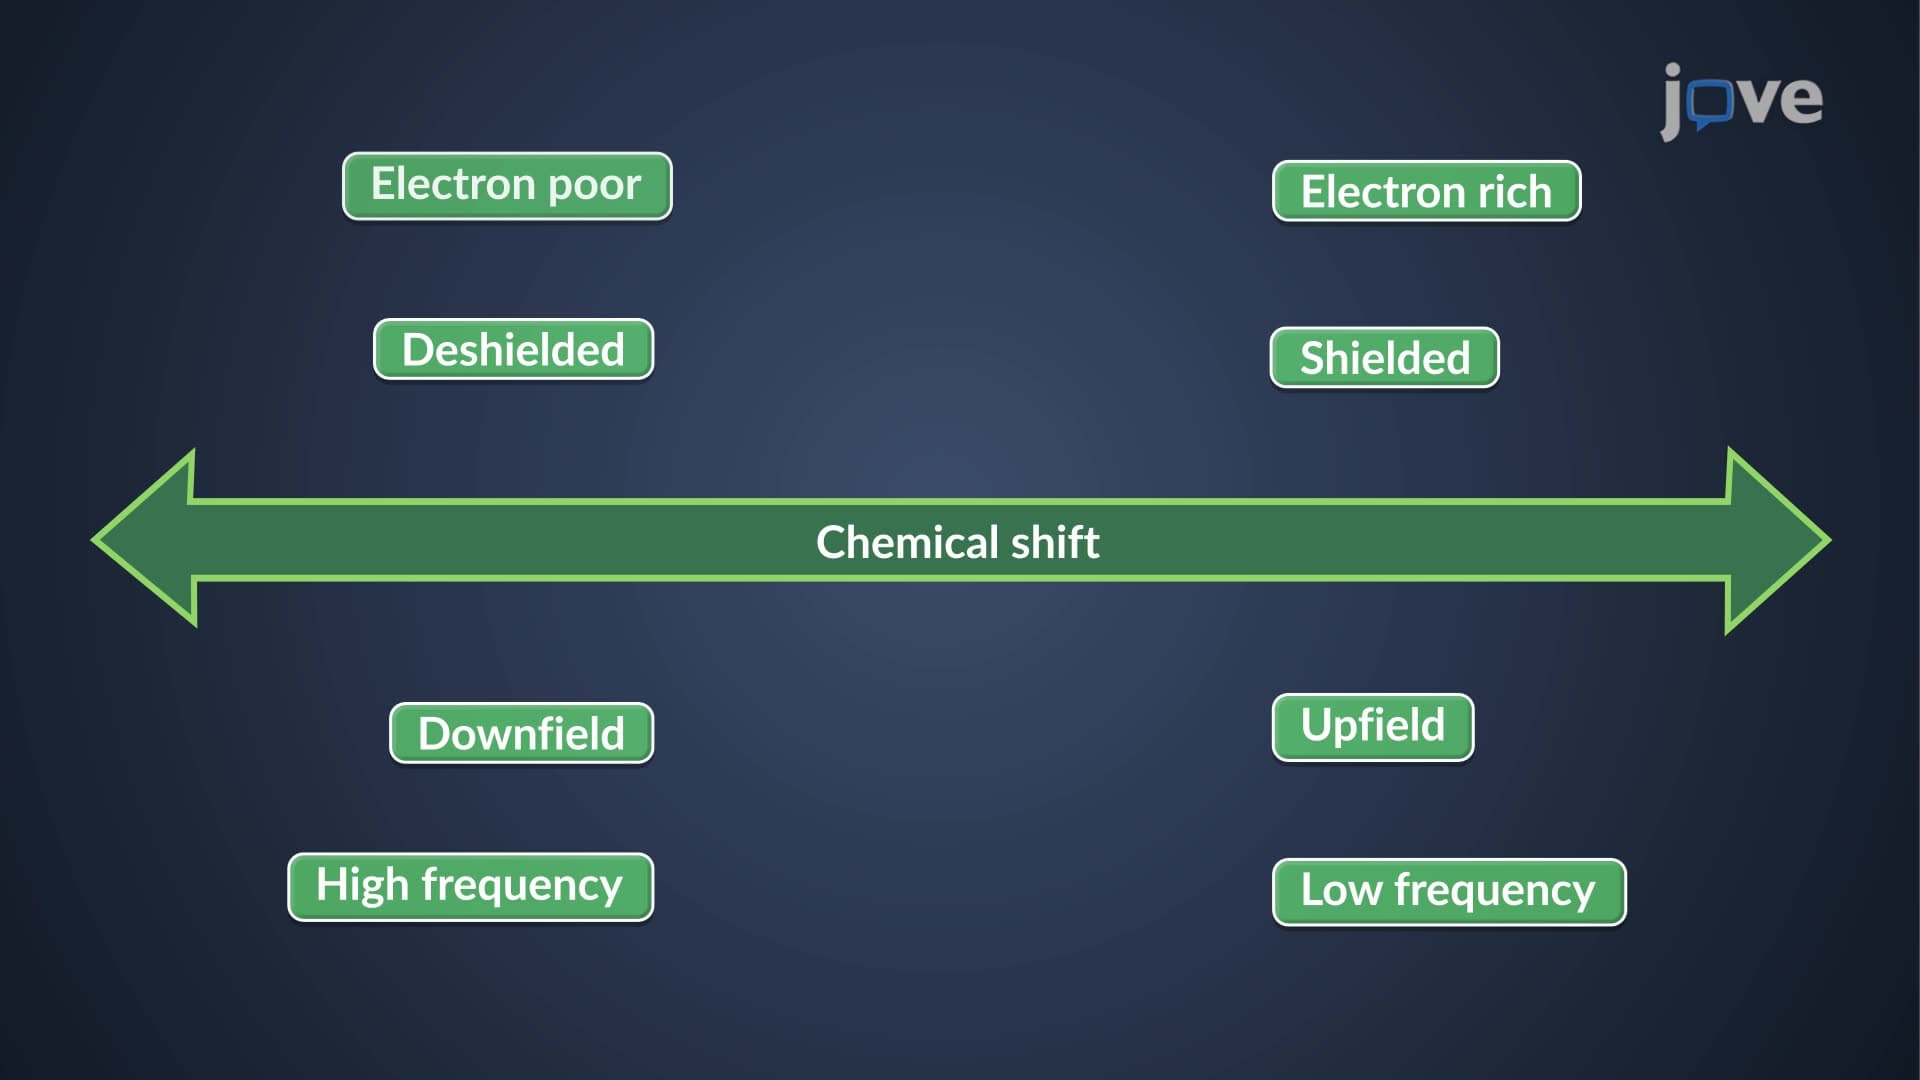
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:03
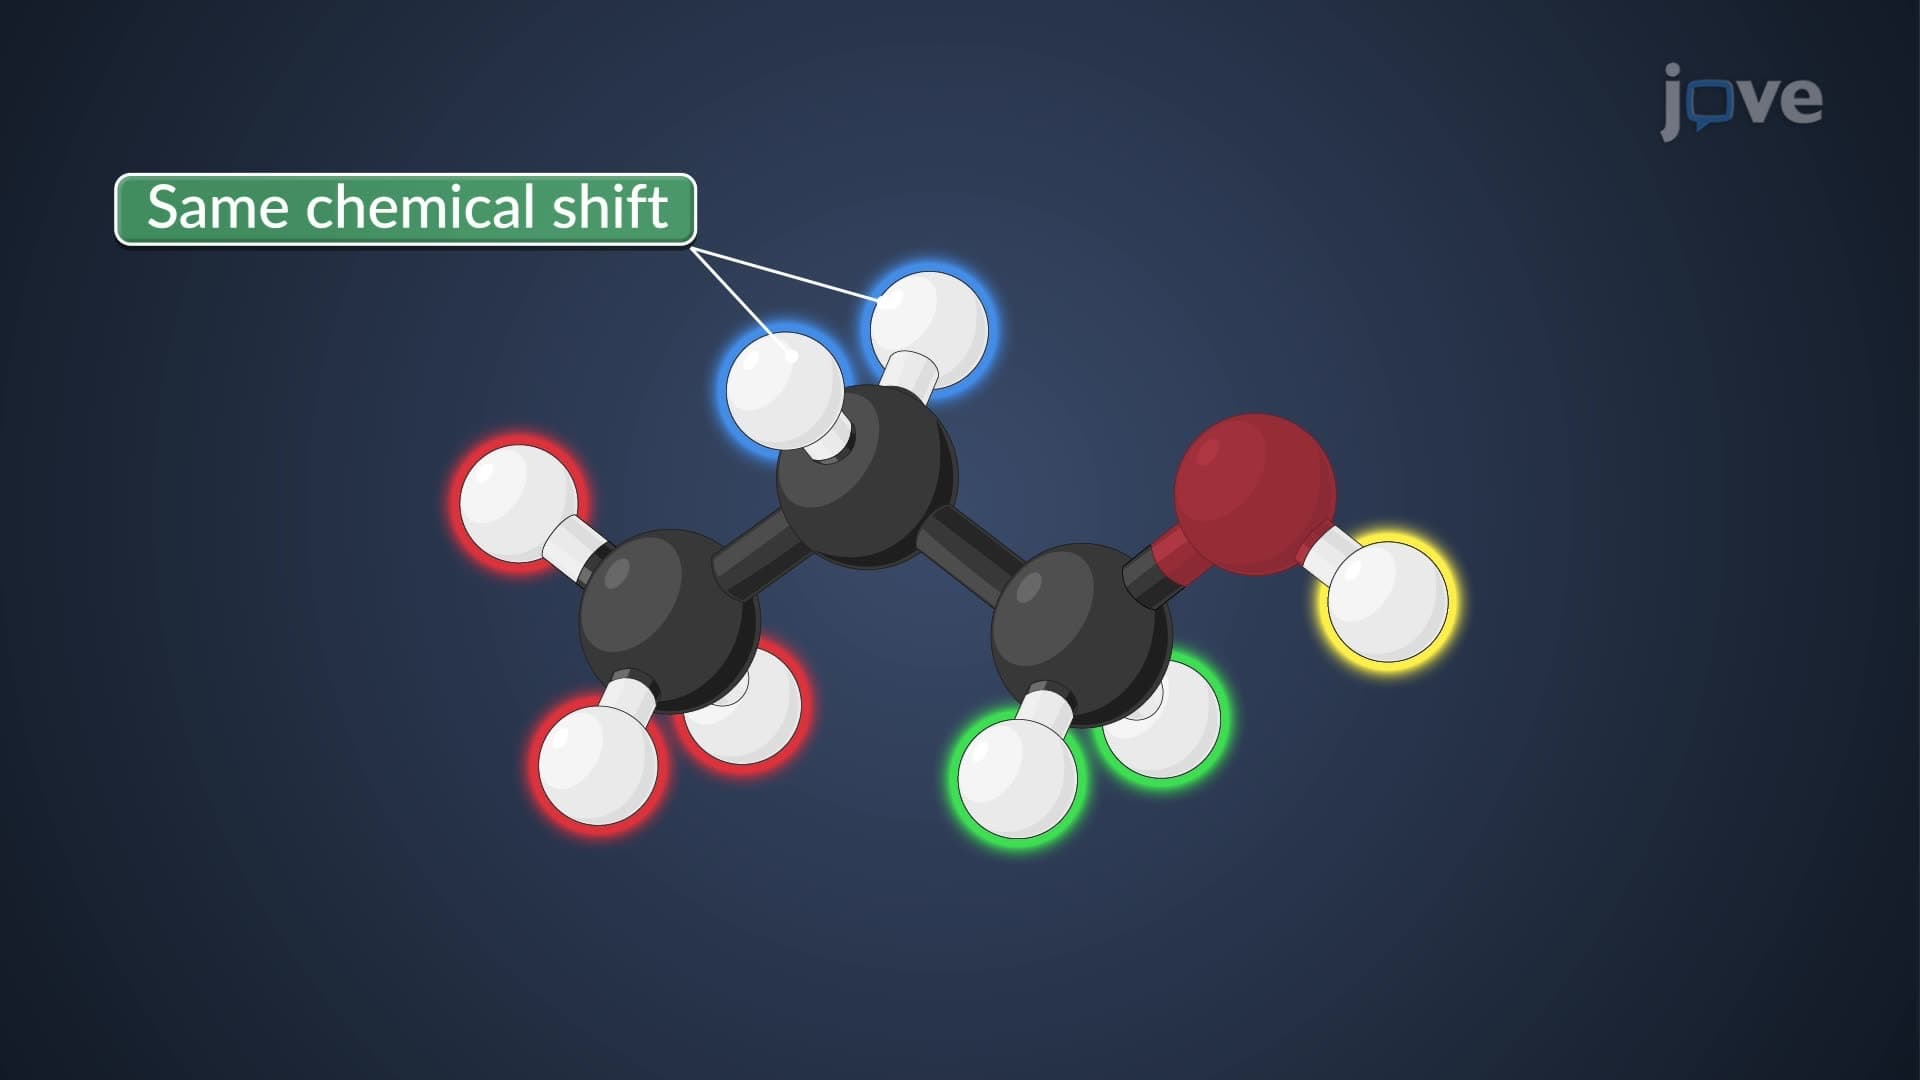
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...