Related Experiment Video
Updated: Oct 3, 2025

Surrogate Model Development for Digital Experiments in Welding
Published on: March 28, 2025
Predicting Segregation Energy in Single Atom Alloys Using Physics and Machine Learning
Maya Salem1, Michael J Cowan1, Giannis Mpourmpakis1
1Department of Chemical Engineering, University of Pittsburgh, Pittsburgh, Pennsylvania 15261, United States.

Abstract:
Single atom alloys (SAAs) show great promise as catalysts for a wide variety of reactions due to their tunable properties, which can enhance the catalytic activity and selectivity. To design SAAs, it is imperative for the heterometal dopant to be stable on the surface as an active catalytic site. One main approach to probe SAA stability is to calculate surface segregation energy. Density functional theory (DFT) can be applied to investigate the surface segregation energy in SAAs. However, DFT is computationally expensive and time-consuming; hence, there is a need for accelerated frameworks to screen metal segregation for new SAA catalysts across combinations of metal hosts and dopants. To this end, we developed a model that predicts surface segregation energy using machine learning for a series of SAA periodic slabs. The model leverages elemental descriptors and features inspired by the previously developed bond-centric model. The initial model accurately captures surface segregation energy across a diverse series of FCC-based SAAs with various surface facets and metal-host pairs. Following our machine learning methodology, we expanded our analysis to develop a new model for SAAs formed from FCC hosts with FCC, BCC, and HCP dopants. Our final, five-feature model utilizes second-order polynomial kernel ridge regression. The model is able to predict segregation energies with a high degree of accuracy, which is due to its physically motivated features. We then expanded our data set to test the accuracy of the five features used. We find that the retrained model can accurately capture E seg trends across different metal hosts and facets, confirming the significance of the features used in our final model. Finally, we apply our pretrained model to a series of Ir- and Pd-based SAA cuboctahedron nanoparticles (NPs), ranging in size and FCC dopants. Remarkably, our model (trained on periodic slabs) accurately predicts the DFT segregation energies of the SAA NPs. The results provide further evidence supporting the use of our model as a general tool for the rapid prediction of SAA segregation energies. By creating a framework to predict the metal segregation from bulk surfaces to NPs, we can accelerate the SAA catalyst design while simultaneously unraveling key physicochemical properties driving thermodynamic stabilization of SAAs.
More Related Videos
 12:02
Determination of Thermodynamic Properties of Alkaline Earth-liquid Metal Alloys Using the Electromotive Force Technique
Published on: November 3, 2017
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:54
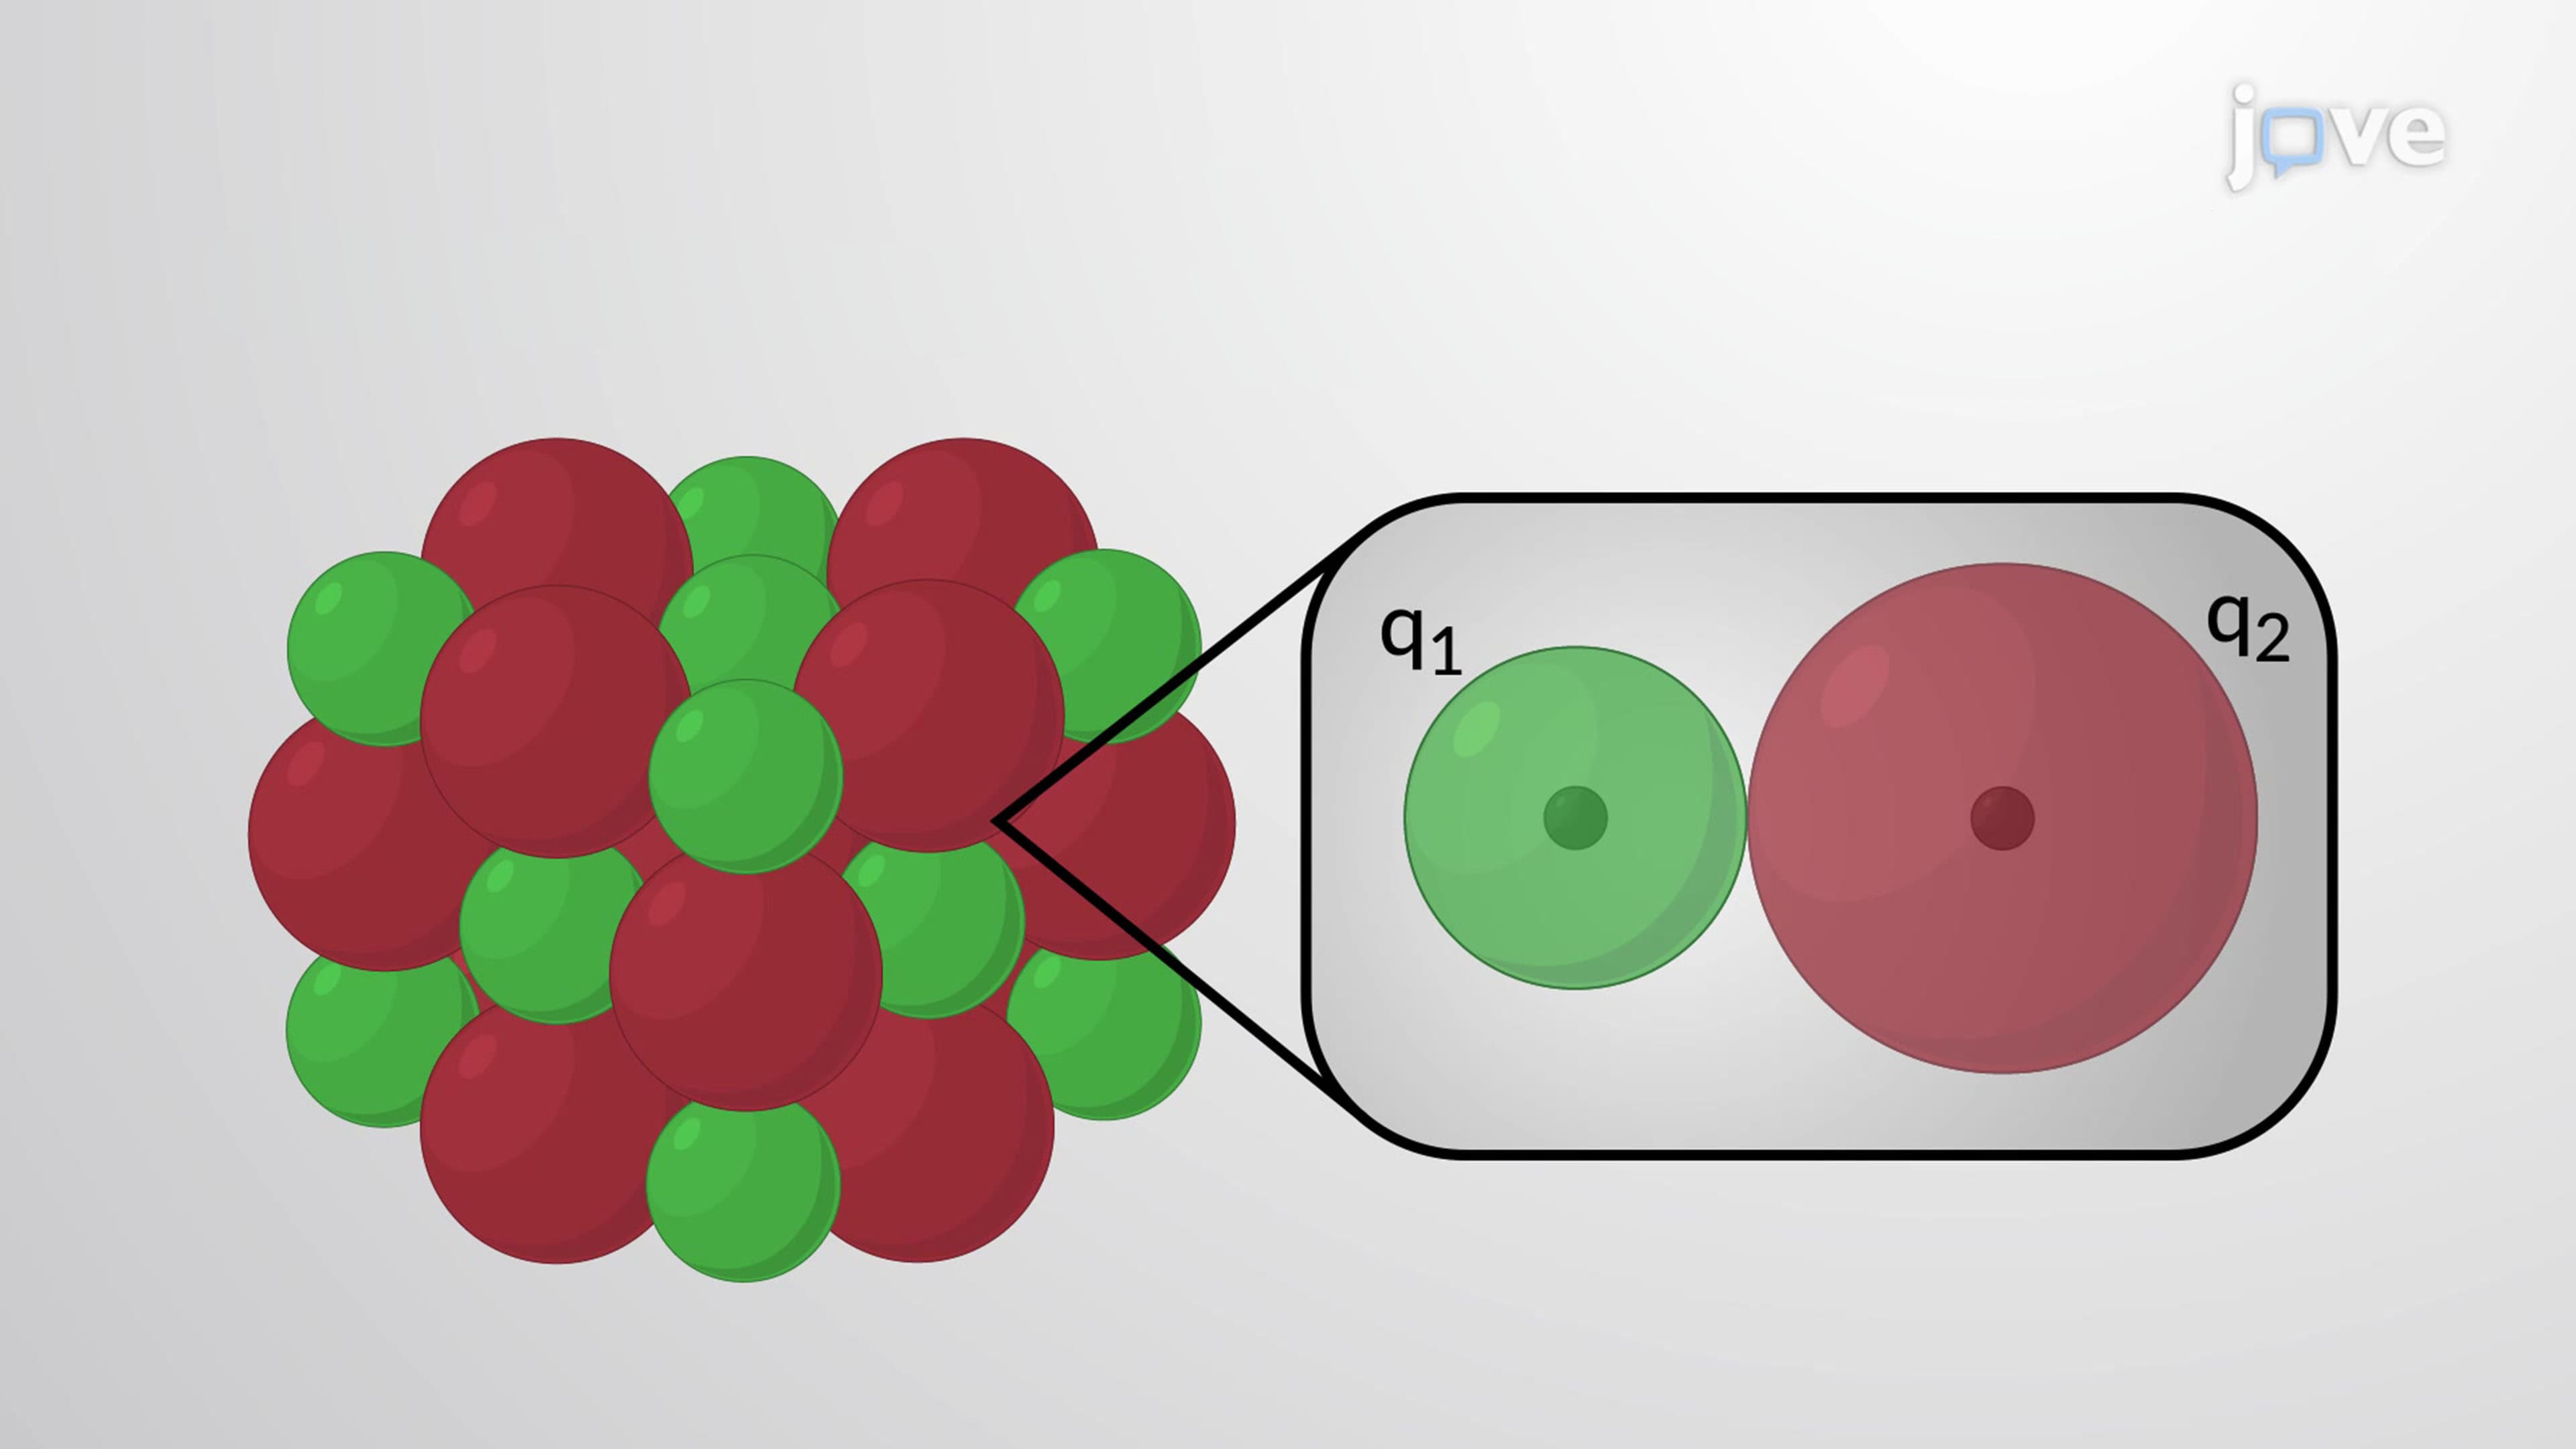
Trends in Lattice Energy: Ion Size and Charge
 01:49
Law of Segregation
 01:20
Atomic Emission Spectroscopy: Overview
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...