Related Experiment Video
Updated: Oct 3, 2025
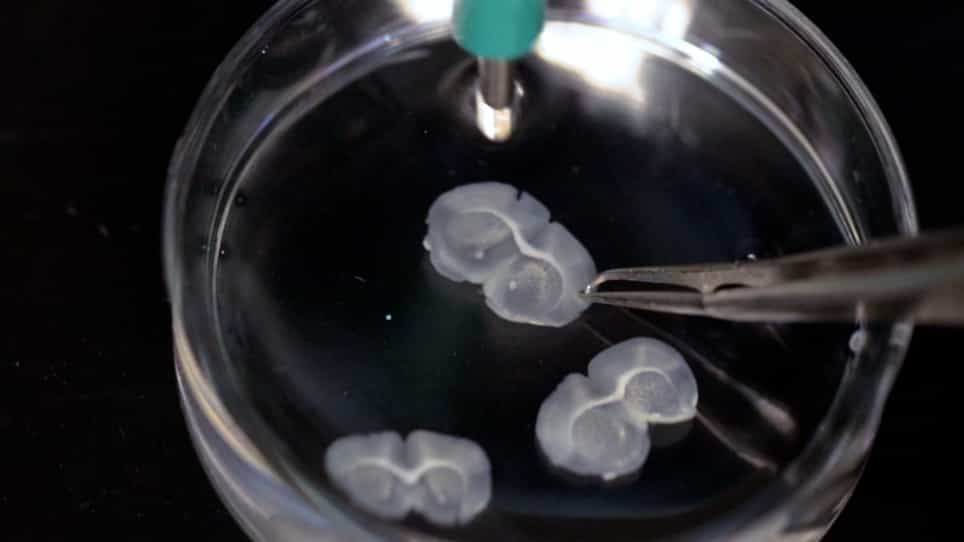

Measurement of Oxygen Consumption Rate in Acute Striatal Slices from Adult Mice
Published on: June 8, 2022
Mitochondrial dysfunction and oxidative stress contribute to cognitive and motor impairment in FOXP1 syndrome
Jing Wang1, Henning Fröhlich1, Felipe Bodaleo Torres2
1Department of Human Molecular Genetics, Institute of Human Genetics, Heidelberg University Hospital, D-69120 Heidelberg, Germany.

Abstract:
FOXP1 syndrome caused by haploinsufficiency of the forkhead box protein P1 (FOXP1) gene is a neurodevelopmental disorder that manifests motor dysfunction, intellectual disability, autism, and language impairment. In this study, we used a Foxp1+/- mouse model to address whether cognitive and motor deficits in FOXP1 syndrome are associated with mitochondrial dysfunction and oxidative stress. Here, we show that genes with a role in mitochondrial biogenesis and dynamics (e.g., Foxo1, Pgc-1α, Tfam, Opa1, and Drp1) were dysregulated in the striatum of Foxp1+/- mice at different postnatal stages. Furthermore, these animals exhibit a reduced mitochondrial membrane potential and complex I activity, as well as decreased expression of the antioxidants superoxide dismutase 2 (Sod2) and glutathione (GSH), resulting in increased oxidative stress and lipid peroxidation. These features can explain the reduced neurite branching, learning and memory, endurance, and motor coordination that we observed in these animals. Taken together, we provide strong evidence of mitochondrial dysfunction in Foxp1+/- mice, suggesting that insufficient energy supply and excessive oxidative stress underlie the cognitive and motor impairment in FOXP1 deficiency.
Related Concept Videos
 01:28
Disorders of the Nervous Tissue
Homeostatic Imbalances:
Alzheimer's disease manifests as a gradual decline in memory and cognitive abilities, attributed to the buildup of amyloid plaques and neurofibrillary tangles in the brain.
Parkinson's disease arises from the...
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...