Related Experiment Video
Updated: Oct 1, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Tension between predicting accurate ground state correlation energies and excitation energies from adiabatic
Lucas M Everhart1, Julio A Derteano1, Jefferson E Bates1
1Department of Chemistry and Fermentation Sciences, Appalachian State University, Boone, North Carolina 28607, USA.

Abstract:
The connection between the adiabatic excitation energy of time-dependent density functional theory and the ground state correlation energy from the adiabatic connection fluctuation-dissipation theorem (ACFDT) is explored in the limiting case of one excited state. An exact expression is derived for any adiabatic Hartree-exchange-correlation kernel that connects the excitation energy and the potential contribution to correlation. The resulting formula is applied to the asymmetric Hubbard dimer, a system where this limit is exact. Results from a hierarchy of approximations to the kernel, including the random phase approximation (RPA) with and without exchange and the adiabatically exact (AE) approximation, are compared to the exact ones. At full coupling, the numerical results indicate a tension between predicting an accurate excitation energy and an accurate potential contribution to correlation. The AE approximation is capable of making accurate predictions of both quantities, but only in parts of the parameter space that classify as weakly correlated, while RPA tends to be unable to accurately predict these properties simultaneously anywhere. For a strongly correlated dimer, the AE approximation greatly overestimates the excitation energy yet continues to yield an accurate ground state correlation energy due to its accurate prediction of the adiabatic connection integrand. If similar trends hold for real systems, the development of correlation kernels will be important for applications of the ACFDT in systems with large potential contributions to correlation.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:10
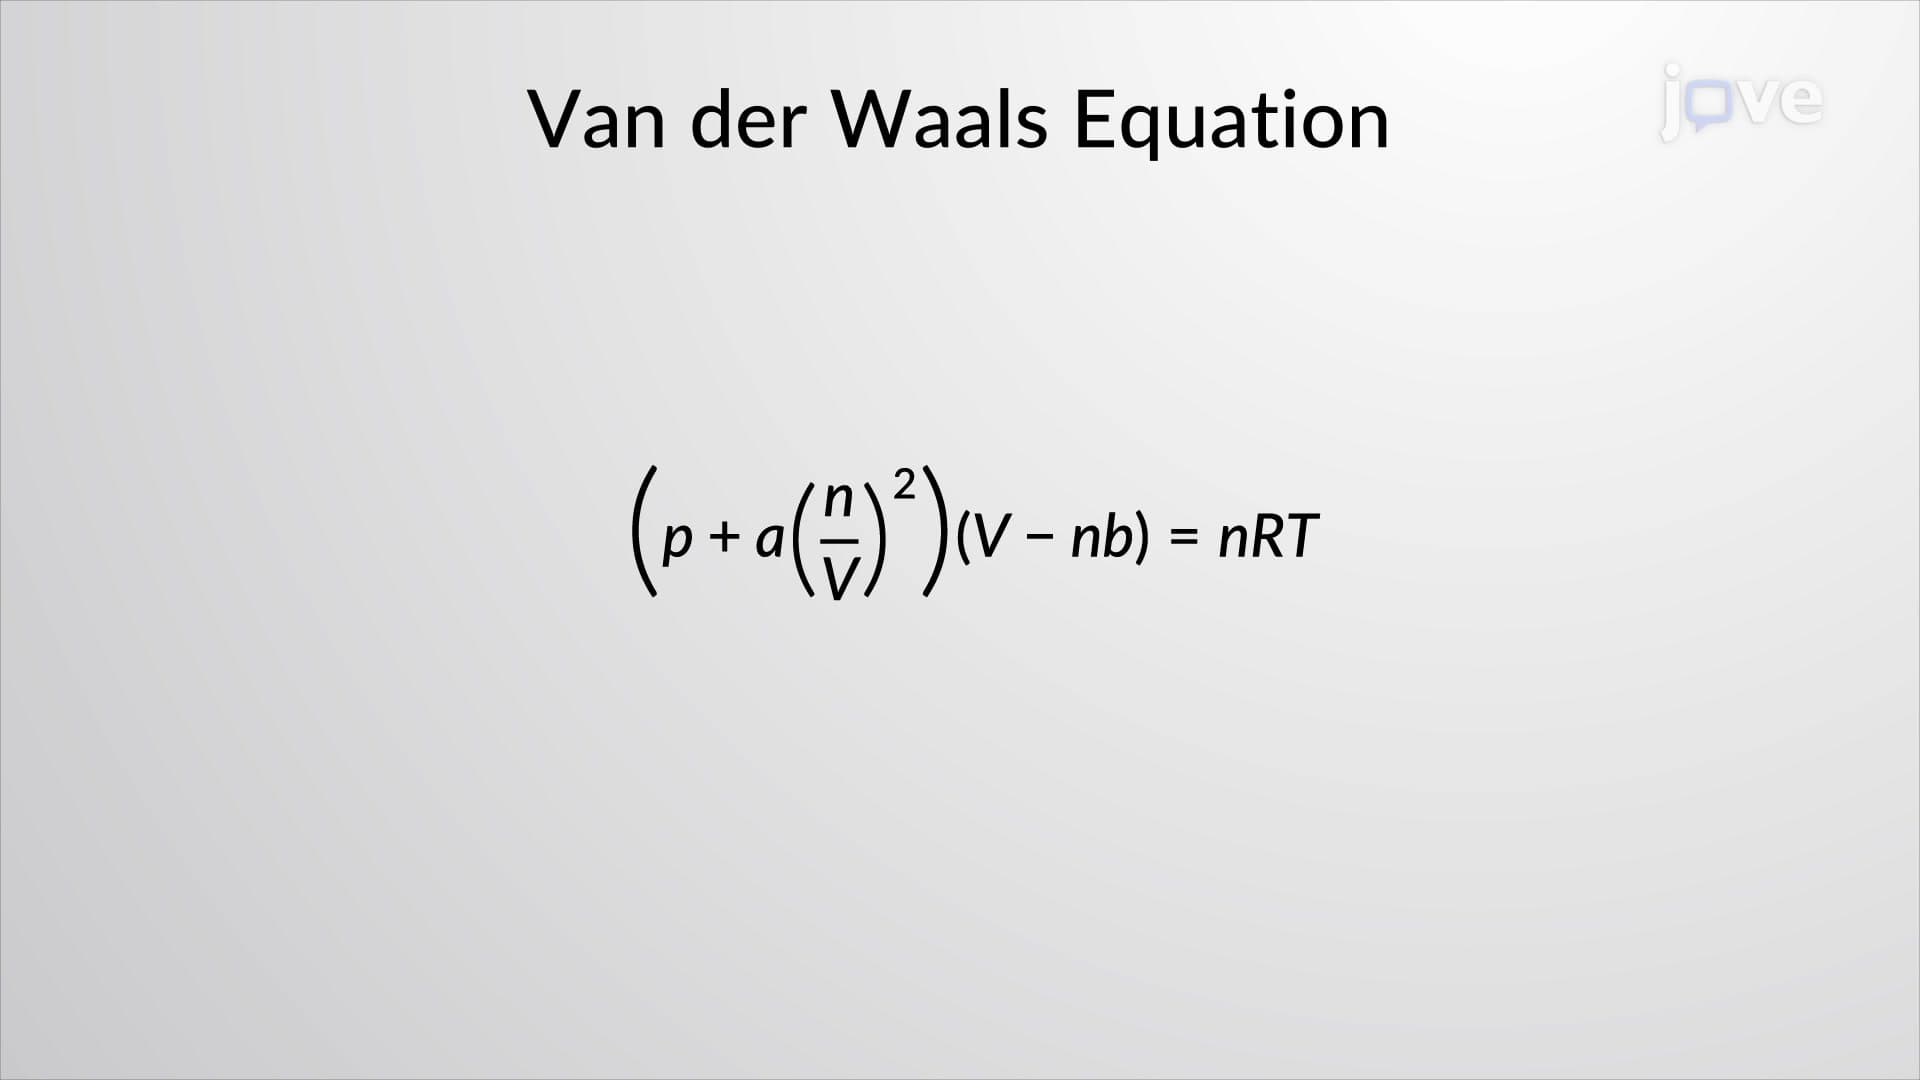
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:21
The Energies of Atomic Orbitals
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...