Related Experiment Video
Updated: Sep 30, 2025

Analyzing Large Protein Complexes by Structural Mass Spectrometry
Published on: June 19, 2010
A general procedure for rounding m/z values in low-resolution mass spectra
Mikhail Khrisanfov1, Andrey Samokhin1
1Chemistry Department, Lomonosov Moscow State University, Moscow, Russia.

Abstract:
It was revealed that nominal mass spectra extracted from the same NetCDF file using different gas chromatography/mass spectrometry (GC/MS) software products are not identical. This phenomenon is caused by differences in algorithms used for rounding floating-point m/z values to integers. It was found that all programs under consideration (AMDIS, ChemStation, ChromaTOF, MS Search, OpenChrom) use different procedures. It is necessary to know how fractional parts of accurate m/z values of ions are distributed to determine which algorithm yields more robust results. We estimated the respective distribution using two databases (PubChem and NIST). As a result, we came up with a procedure that minimizes the influence of random errors on rounding to integer m/z values. The procedure we suggest is to sum intensities of all floating-point m/z values in a bin [MZ - 0.38; MZ + 0.62] and assign MZ as a nominal m/z value, where MZ is an integer m/z value.
More Related Videos
Related Concept Videos
 01:15
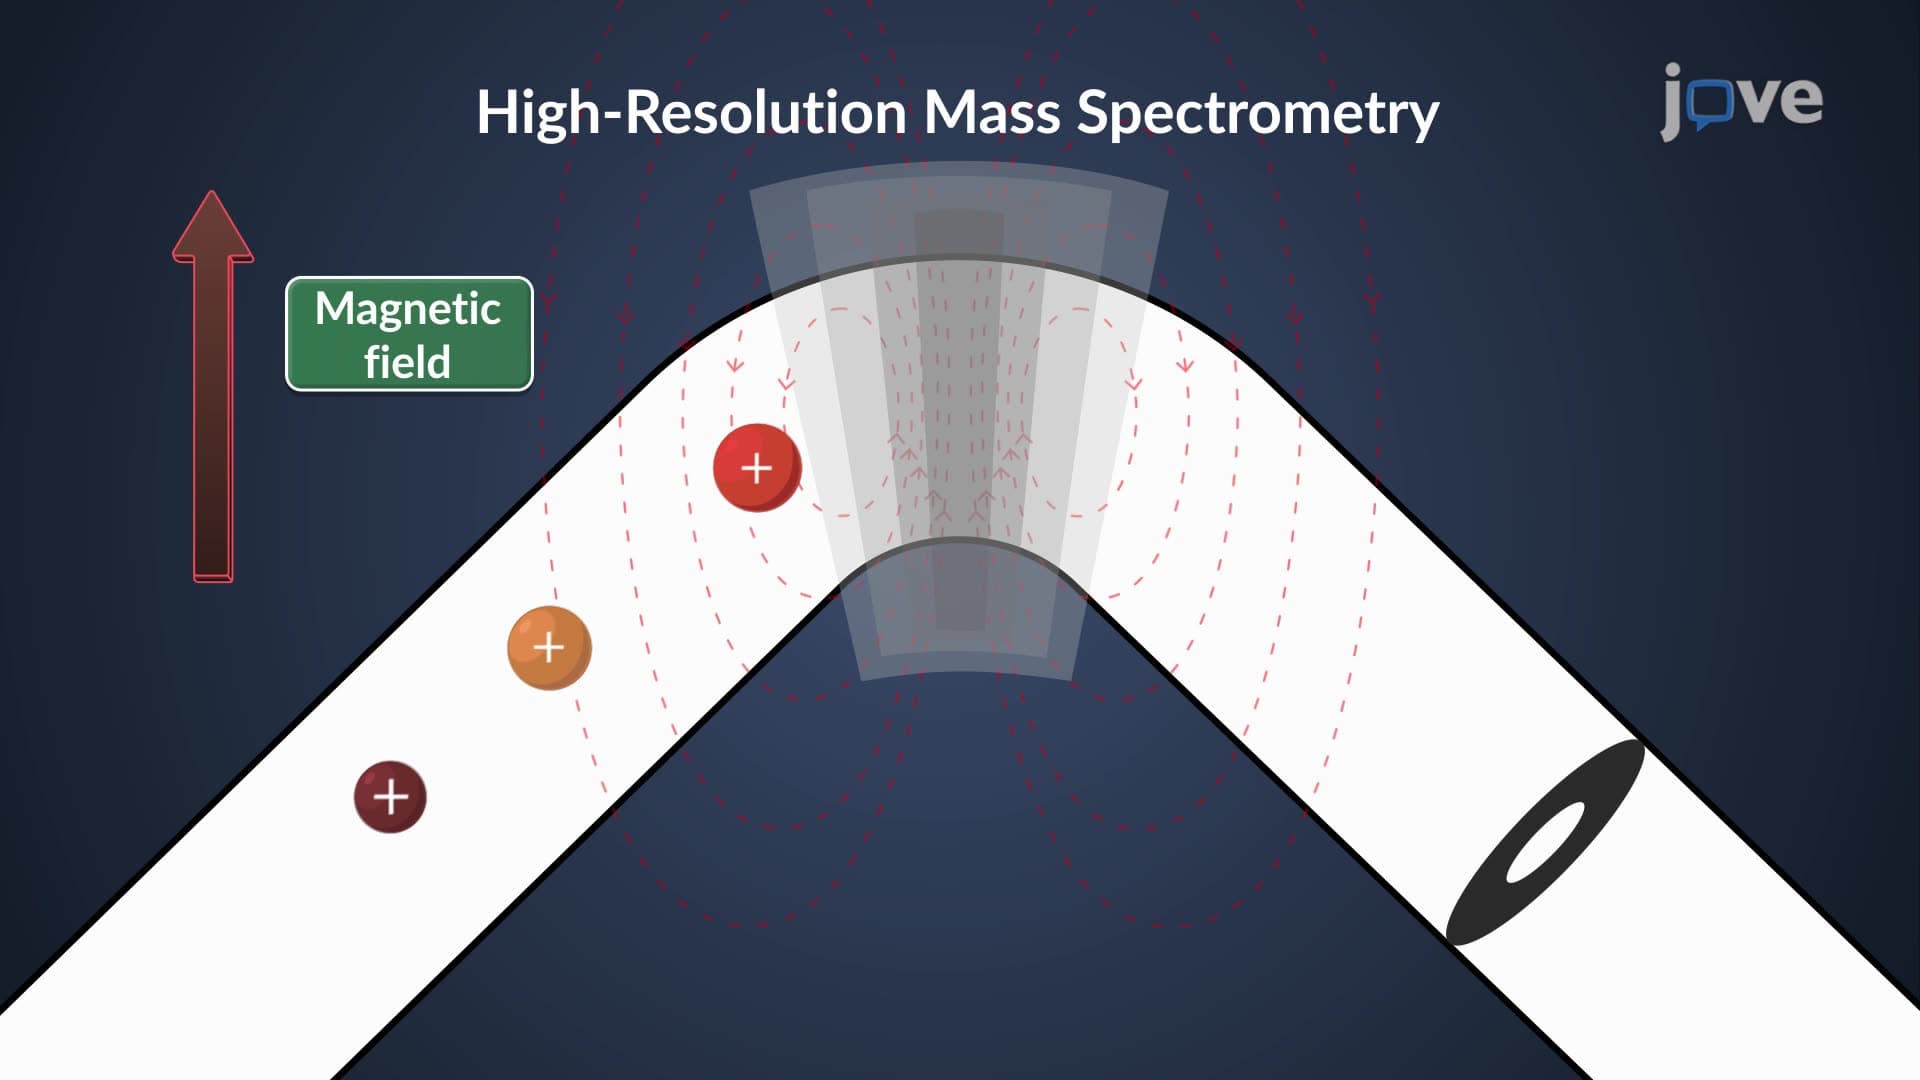
High-Resolution Mass Spectrometry (HRMS)
 01:13
Mass Analyzers: Overview
 01:24
Mass Spectrum: Interpretation
To...
 01:23
Mass Spectrum
 01:19
Mass Spectrometry: Overview
 01:16
Mass Spectrometers