Related Experiment Video
Updated: Sep 30, 2025
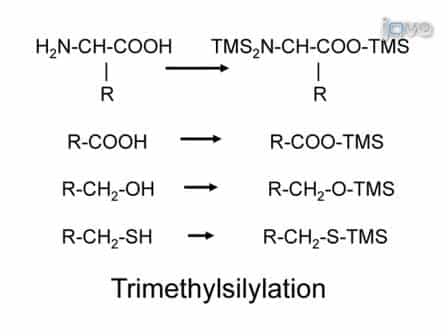

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Diagnosis of inborn errors of metabolism within the expanded newborn screening in the Madrid region
Álvaro Martín-Rivada1, Laura Palomino Pérez1, Pedro Ruiz-Sala2
1Sección de Gastroenterología y Nutrición Hospital Infantil Universitario Niño Jesús Madrid Spain.

Abstract:
We present the results of our experience in the diagnosis of inborn errors of metabolism (IEM) since the Expanded Newborn Screening was implemented in our Region. Dried blood samples were collected 48 h after birth. Amino acids and acylcarnitines were quantitated by mass spectrometry (MS)/MS. Newborns with alterations were referred to the clinical centers for follow-up. Biochemical and molecular genetic studies for confirmation of a disease were performed. In the period 2011 to 2019, 592 822 children were screened: 902 of them were referred for abnormal results. An IEM was confirmed in 222 (1/2670): aminoacidopathies: 89 hyperphenylalaninemia (HPA) (51 benign HPA, 32 phenylketonuria, 4 DNAJC12 defect, and 2 primapterinuria), 6 hypermethioninemia, 3 tyrosinemia type 1 (TYR-1), 1 TYR-3, 4 maple syrup urine disease (MSUD), 2 branched-chain amino acid transferase 2 deficiency, 2 homocystinuria, 1 cystinuria, 2 ornithine transcarbamylase (OTC) deficiency, 2 citrullinemia type I (CTLN1); FAO defects: 43 medium-chain acyl-CoA dehydrogenase deficiency (MCADD), 13 very long-chain acyl-CoA dehydrogenase deficiency, 2 long-chain 3-hydroxyacyl-CoA dehydrogenase deficiency (LCHADD), 1 multiple acyl-coA dehydrogenation deficiency, 11 systemic primary carnitine deficiency, 2 carnitine palmitoyltransferase type 2 (CPT-II) deficiency, 1 CPT-I deficiency; organic acidurias: 12 glutaric aciduria type 1 (GA-1), 4 methylmalonic acidemia (MMA), 7 MMA including combined cases with homocystinuria (MMAHC), 6 propionic acidemia (PA), 7 3-methylcrotonyl-CoA carboxylase, 1 3-hydroxy-3-methylglutaryl-CoA lyase deficiency lyase deficiency. Only 19 infants (8.5%) were symptomatic at newborn screening result (1 LCHADD, 5 PA, 1 CPT-II deficiency, 1 MMA, 3 MMAHC, 2 MSUD, 2 OTC deficiency, 1 CTLN1, 1 MCADD, 2 TYR-1). No false negative cases were identified. Genetic diagnosis was conclusive in all biochemically confirmed cases, except for two infants with HPA, identifying pathogenic variants in 32 different genes. The conditions with the highest incidence were HPA (1/6661) and MCAD deficiencies (1/13 787).
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:30
Serum Laboratory Studies, Stool Test, Breath Test
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes: