Clustering biological sequences with dynamic sequence similarity threshold
Related Experiment Video
Updated: Sep 28, 2025
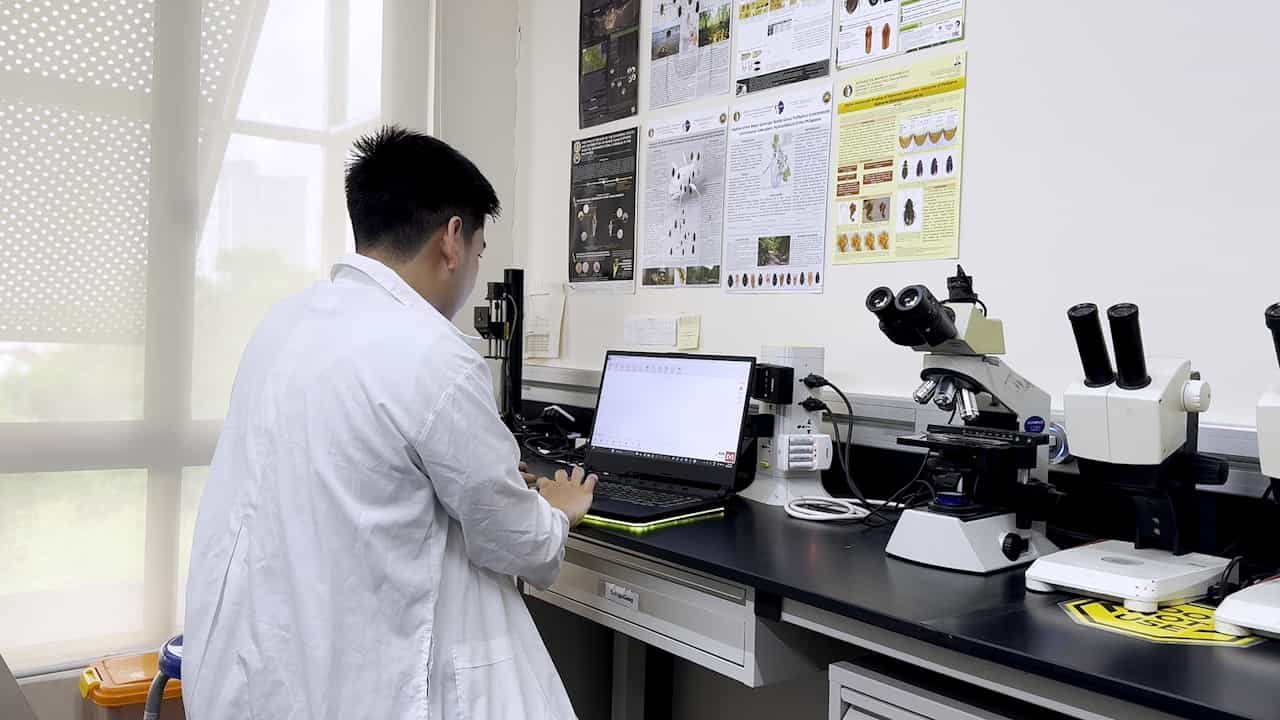
A Concoction Pipeline for Generating Molecular Operational Taxonomic Units (MOTUs) Among Riparian and Aquatic Beetles
Published on: July 11, 2025
Jimmy Ka Ho Chiu1, Rick Twee-Hee Ong2
1Saw Swee Hock School of Public Health, National University of Singapore and National University Health System, Singapore, 117549, Singapore.

Background:
Biological sequence clustering is a complicated data clustering problem owing to the high computation costs incurred for pairwise sequence distance calculations through sequence alignments, as well as difficulties in determining parameters for deriving robust clusters. While current approaches are successful in reducing the number of sequence alignments performed, the generated clusters are based on a single sequence identity threshold applied to every cluster. Poor choices of this identity threshold would thus lead to low quality clusters. There is however little support provided to users in selecting thresholds that are well matched with the input sequences.
Results:
We present a novel sequence clustering approach called ALFATClust that exploits rapid pairwise alignment-free sequence distance calculations and community detection in graph for clusters generation. Instead of a single threshold applied to every generated cluster, ALFATClust is capable of dynamically determining the cut-off threshold for each individual cluster by considering both cluster separation and intra-cluster sequence similarity. Benchmarking analysis shows that ALFATClust generally outperforms existing approaches by simultaneously maintaining cluster robustness and substantial cluster separation for the benchmark datasets. The software also provides an evaluation report for verifying the quality of the non-singleton clusters obtained.
Conclusions:
ALFATClust is able to generate sequence clusters having high intra-cluster sequence similarity and substantial separation between clusters without having users to decide precise similarity cut-off thresholds.

Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:29
Modern Molecular Taxonomy
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
 01:20
Applications of Molecular Taxonomy
 01:05
Maxam-Gilbert Sequencing
Challenges of the Maxam-Gilbert Method
The...