
Calculation of Metallocene Ionization Potentials via Auxiliary Field Quantum Monte Carlo: Toward Benchmark Quantum
Benjamin Rudshteyn1, John L Weber1, Dilek Coskun1
1Department of Chemistry, Columbia University, 3000 Broadway, New York, New York 10027, United States.

Abstract:
The accurate ab initio prediction of ionization energies is essential to understanding the electrochemistry of transition metal complexes in both materials science and biological applications. However, such predictions have been complicated by the scarcity of gas phase experimental data, the relatively large size of the relevant molecules, and the presence of strong electron correlation effects. In this work, we apply all-electron phaseless auxiliary-field quantum Monte Carlo (ph-AFQMC) utilizing multideterminant trial wave functions to six metallocene complexes to compare the computed adiabatic and vertical ionization energies with experimental results. We find that ph-AFQMC yields mean absolute errors (MAEs) of 1.69 ± 1.02 kcal/mol for the adiabatic energies and 2.85 ± 1.13 kcal/mol for the vertical energies. We also carry out density functional theory (DFT) calculations using a variety of functionals, which yields MAEs of 3.62-6.98 kcal/mol and 3.31-9.88 kcal/mol, as well as one variant of localized coupled cluster calculations (DLPNO-CCSD(T0) with moderate PNO cutoffs), which has MAEs of 4.96 and 6.08 kcal/mol, respectively. We also test the reliability of DLPNO-CCSD(T0) and DFT on acetylacetonate (acac) complexes for adiabatic energies measured in the same manner experimentally, and we find higher MAEs, ranging from 4.56 to 10.99 kcal/mol (with a different ordering) for DFT and 6.97 kcal/mol for DLPNO-CCSD(T0). Finally, by utilizing experimental solvation energies, we show that accurate reduction potentials in solution for the metallocene series can be obtained from the AFQMC gas phase results.
More Related Videos
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 01:18
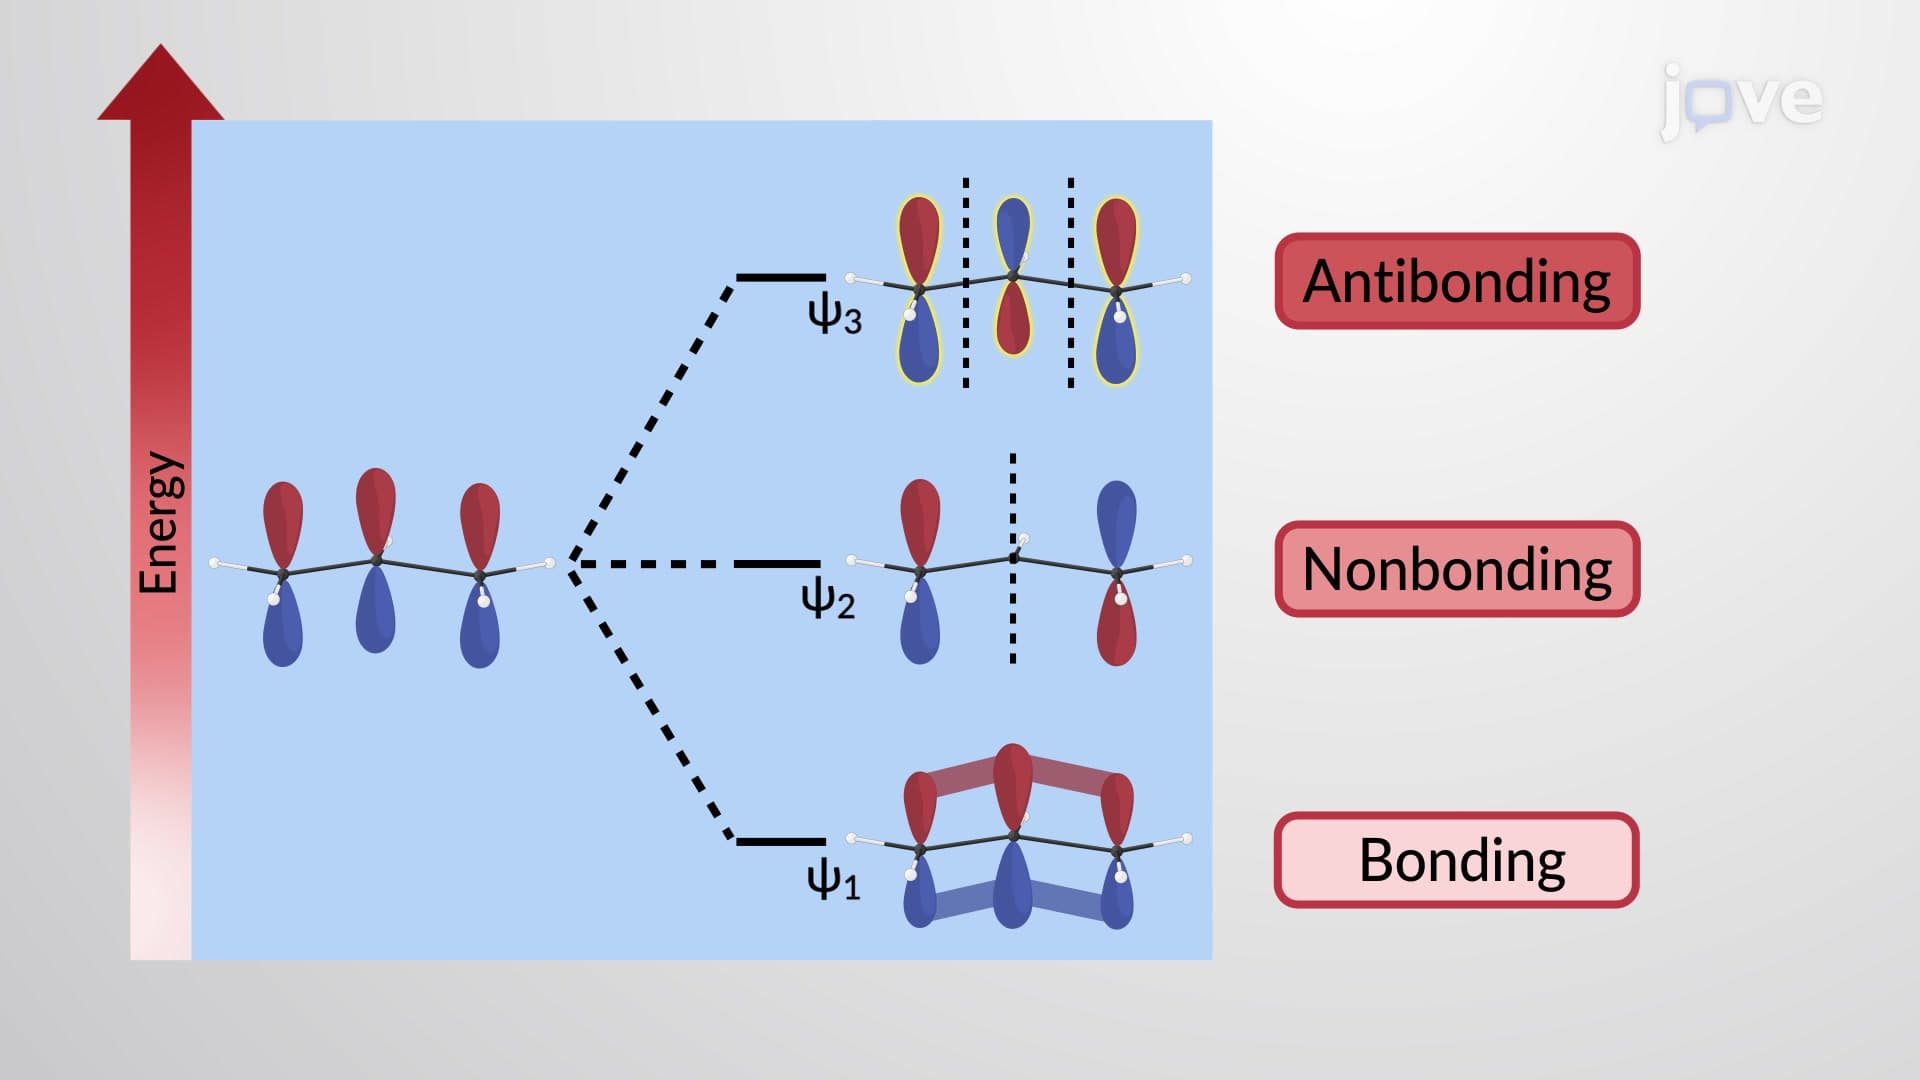
π Molecular Orbitals of the Allyl Cation and Anion
 03:12
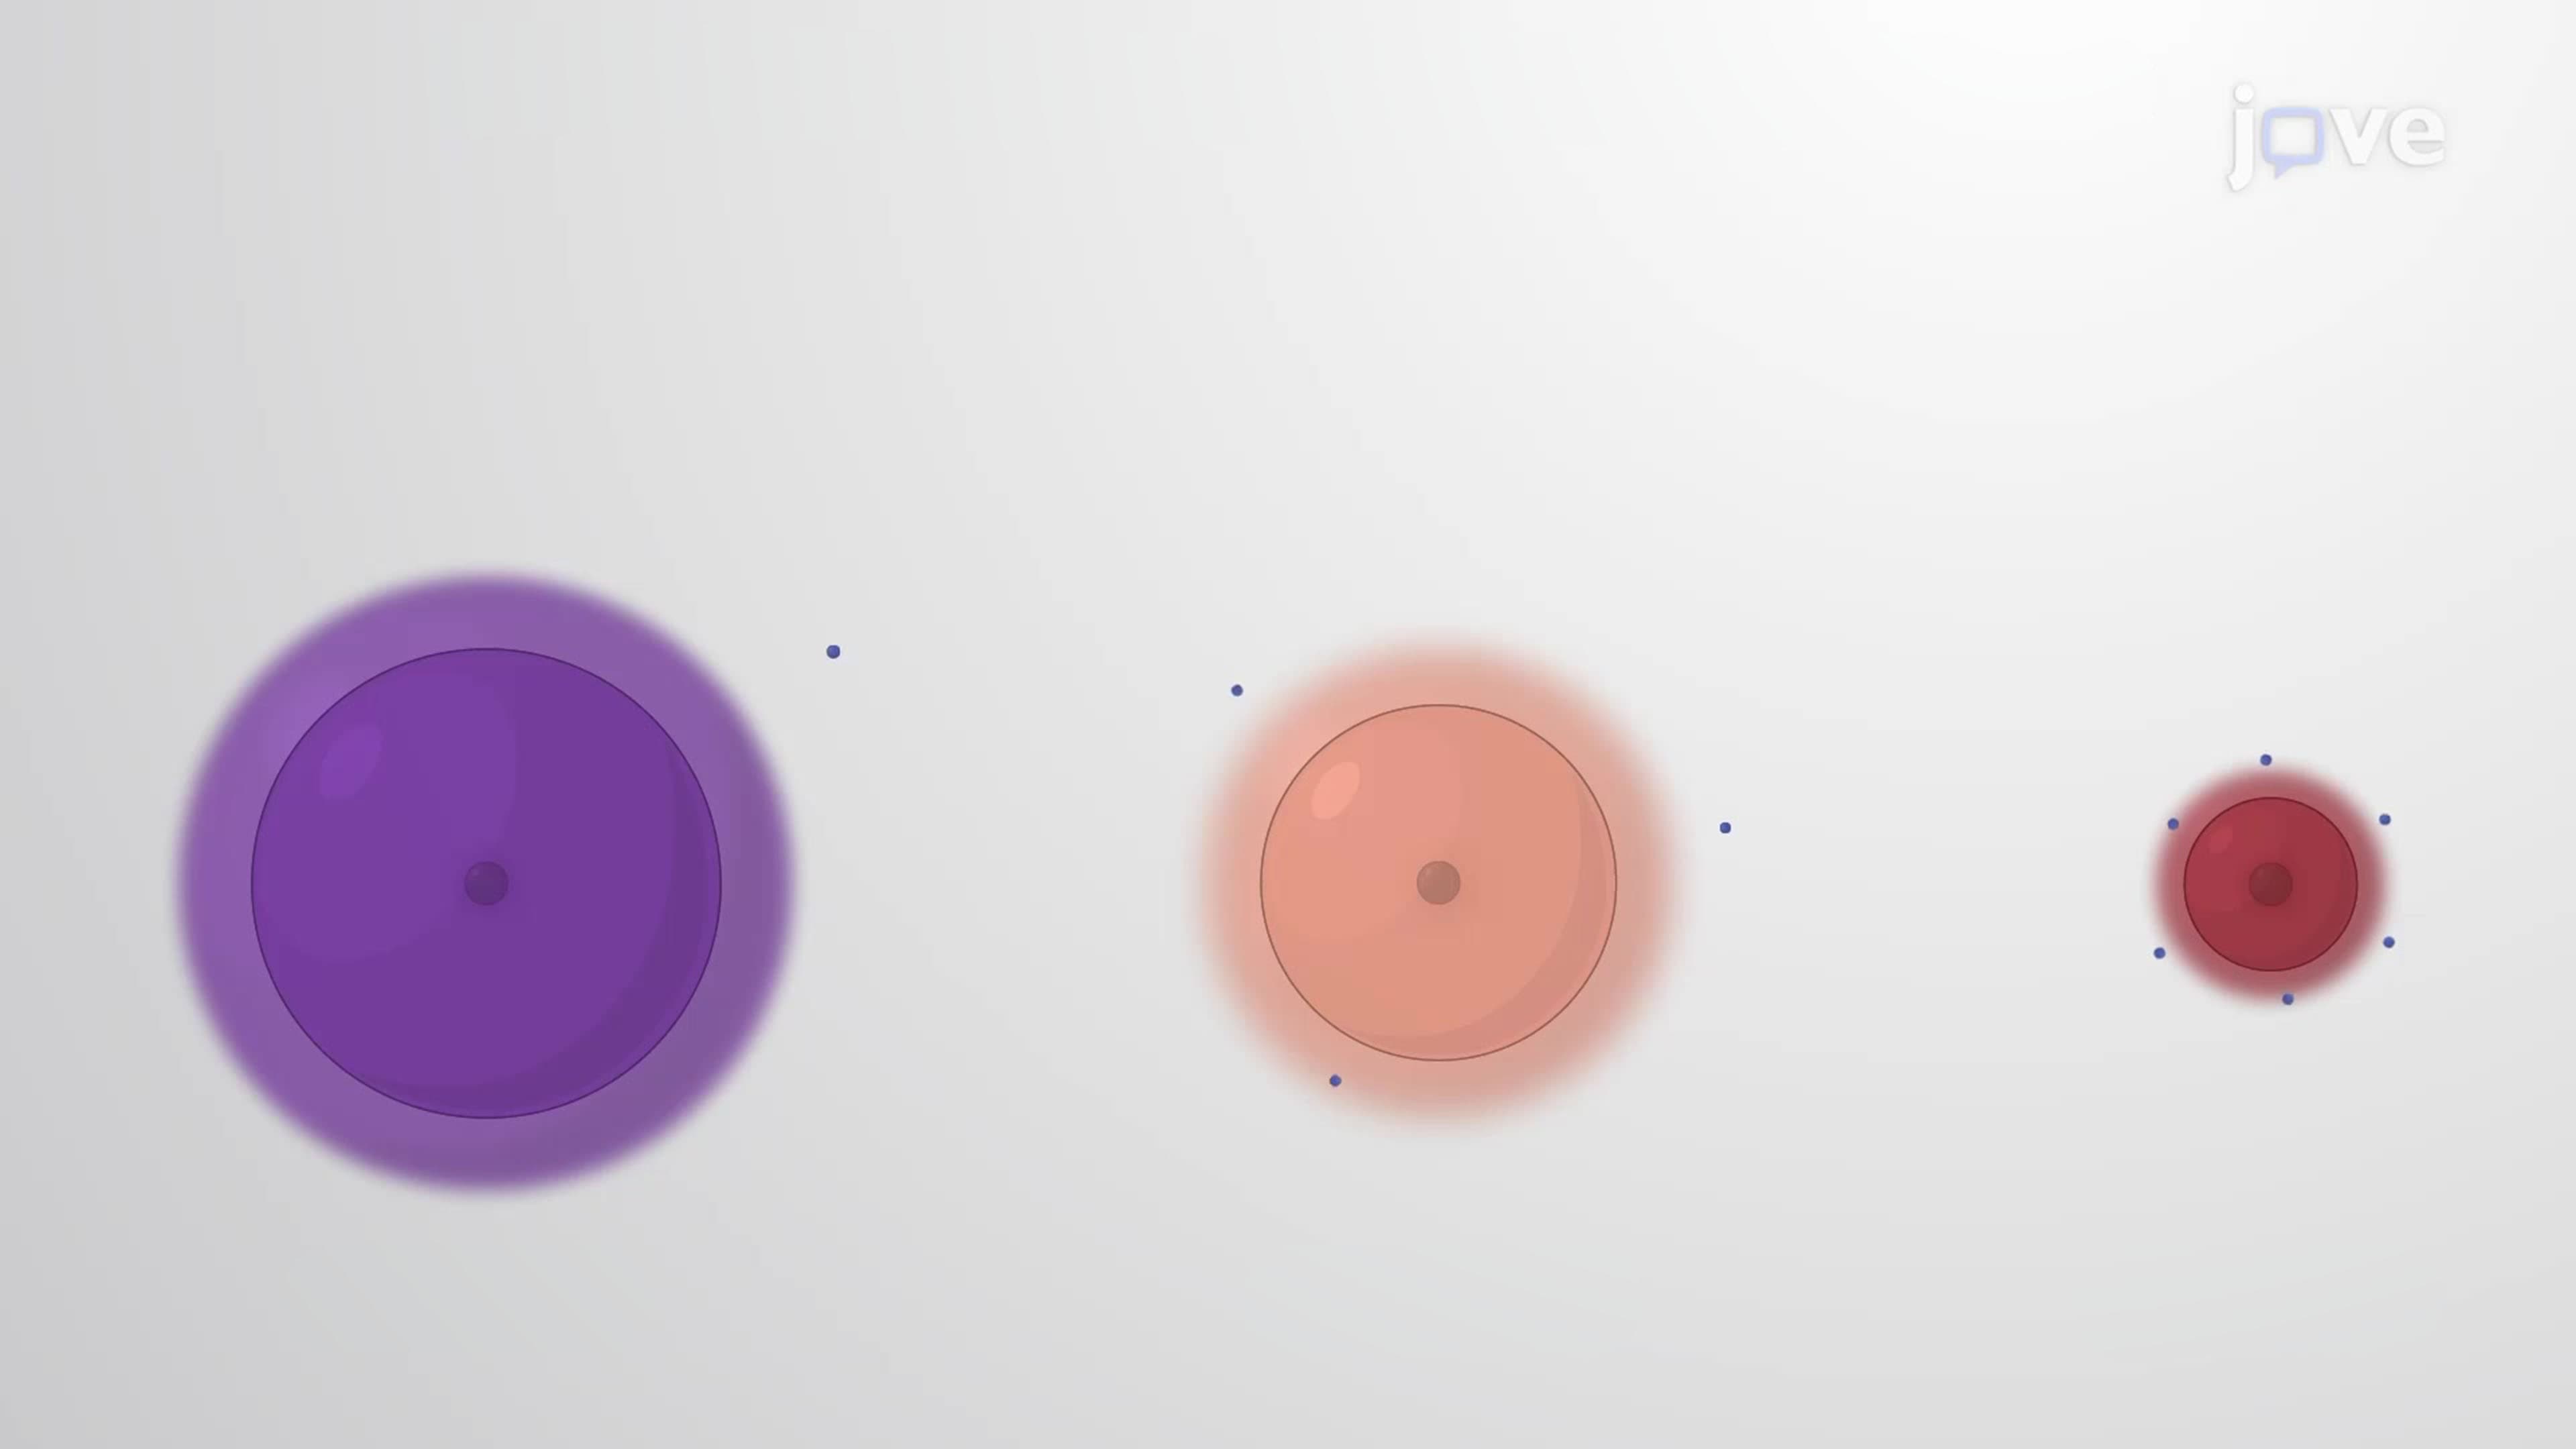
Ionization Energy
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...