Related Experiment Video
Updated: Sep 27, 2025
Author Spotlight: Decoding Mitochondrial Aging
Published on: June 30, 2023
Alteration of Mitochondrial Integrity as Upstream Event in the Pathophysiology of SOD1-ALS
René Günther1,2, Arun Pal1,3, Chloe Williams4
1Department of Neurology, University Hospital Carl Gustav Carus Dresden, Technische Universität Dresden, 01307 Dresden, Germany.

Abstract:
Little is known about the early pathogenic events by which mutant superoxide dismutase 1 (SOD1) causes amyotrophic lateral sclerosis (ALS). This lack of mechanistic understanding is a major barrier to the development and evaluation of efficient therapies. Although protein aggregation is known to be involved, it is not understood how mutant SOD1 causes degeneration of motoneurons (MNs). Previous research has relied heavily on the overexpression of mutant SOD1, but the clinical relevance of SOD1 overexpression models remains questionable. We used a human induced pluripotent stem cell (iPSC) model of spinal MNs and three different endogenous ALS-associated SOD1 mutations (D90Ahom, R115Ghet or A4Vhet) to investigate early cellular disturbances in MNs. Although enhanced misfolding and aggregation of SOD1 was induced by proteasome inhibition, it was not affected by activation of the stress granule pathway. Interestingly, we identified loss of mitochondrial, but not lysosomal, integrity as the earliest common pathological phenotype, which preceded elevated levels of insoluble, aggregated SOD1. A super-elongated mitochondrial morphology with impaired inner mitochondrial membrane potential was a unifying feature in mutant SOD1 iPSC-derived MNs. Impaired mitochondrial integrity was most prominent in mutant D90Ahom MNs, whereas both soluble disordered and detergent-resistant misfolded SOD1 was more prominent in R115Ghet and A4Vhet mutant lines. Taking advantage of patient-specific models of SOD1-ALS in vitro, our data suggest that mitochondrial dysfunction is one of the first crucial steps in the pathogenic cascade that leads to SOD1-ALS and also highlights the need for individualized medical approaches for SOD1-ALS.
Insights
Mitochondrial dysfunction, not protein aggregation, is an early event in mutant superoxide dismutase 1 (SOD1) causing amyotrophic lateral sclerosis (ALS). This finding in patient-derived cells suggests personalized therapies for SOD1-ALS.
Area of Science:
- Neuroscience
- Cell Biology
- Genetics
Background:
- Amyotrophic lateral sclerosis (ALS) pathogenesis linked to mutant superoxide dismutase 1 (SOD1) remains poorly understood.
- Current models often use SOD1 overexpression, questioning clinical relevance for studying early pathogenic events.
Purpose of the Study:
- Investigate early cellular disturbances in motoneurons (MNs) using patient-specific induced pluripotent stem cells (iPSCs) with endogenous SOD1 mutations.
- Determine the sequence of pathological events, focusing on mitochondrial integrity and SOD1 aggregation.
Main Methods:
- Utilized human iPSC-derived spinal MNs with three endogenous ALS-associated SOD1 mutations (D90A, R115G, A4V).
- Assessed cellular phenotypes including mitochondrial and lysosomal integrity, SOD1 misfolding, aggregation, and stress granule formation.
- Compared pathological events under conditions of proteasome inhibition and stress granule pathway activation.
Main Results:
- Loss of mitochondrial integrity, characterized by super-elongated morphology and impaired inner membrane potential, was the earliest common pathological phenotype.
- Mitochondrial dysfunction preceded elevated levels of insoluble, aggregated SOD1.
- Specific mutations showed distinct patterns: D90A primarily affected mitochondrial integrity, while R115G and A4V showed more prominent SOD1 misfolding and aggregation.
Conclusions:
- Mitochondrial dysfunction is a critical early step in the pathogenesis of SOD1-ALS.
- Patient-specific iPSC models reveal distinct cellular defects based on SOD1 mutation type.
- Findings underscore the need for individualized therapeutic strategies for SOD1-ALS.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:45
Mitochondrial Membranes
 02:59
Animal Mitochondrial Genetics
 01:37
Mitochondria
 01:19
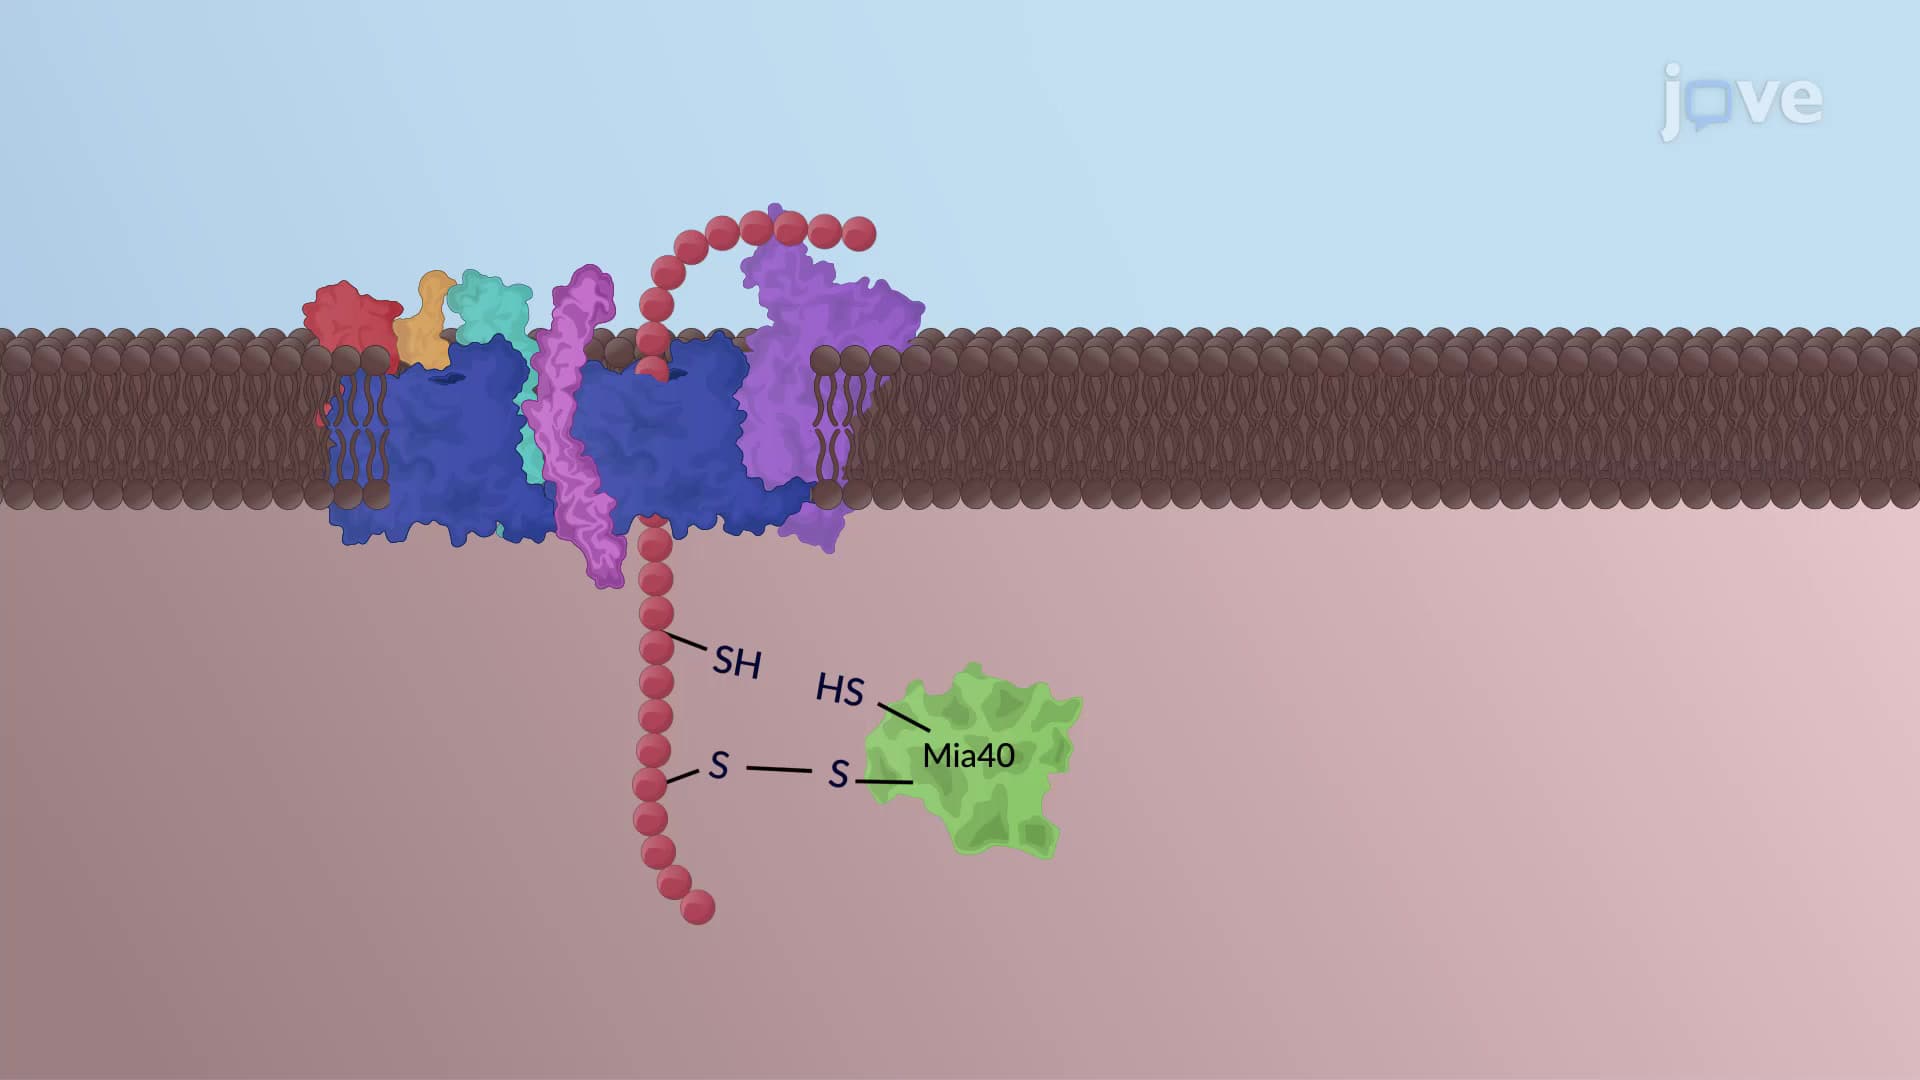
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mutations