Related Experiment Video
Updated: Sep 27, 2025

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Mining of molecular insights of CYP2A6 and its variants complex with coumarin (CYP2A6*-coumarin) using molecular
Anamika Yadav1, Akanksha Kesharwani1,2, Dheeraj Kumar Chaurasia2
1Computational omics Lab, Centre of Bioinformatics, University of Allahabad, Prayagraj, India.

Abstract:
CYP2A6 is a very important enzyme that plays a crucial role in nicotine compounds and is responsible for the metabolism of more than 3% drugs of total metabolized drugs by the CYP family and reported as one of very important pharmacogenes. CYP2A6 is highly polymorphic in nature and reported with more than 40 variants, most of these variants are SNPs originated and population specific. It has been well observed and reported that the presence of these population-specific non-synonymous SNPs in CYP2A6 alters the rate of drug metabolism and as a functional consequence, drugs produce an abnormal response. Though genomics and pharmacogenomics studies are there, very less is known about the structural effects of these SNPs on molecular-interaction and folding of CYP2A6. To fill the knowledge gap, SNPs based four variants, i.e., CYP2A6*2, CYP2A6*18, CYP2A6*21, and CYP2A6*35, which are frequently reported in the South Asian population, were considered for the study. Coumarin (DB04665), a well reported drug, is considered as a model substance, and the effect of all four variants on 'CYP2A6*-coumarin' complex was studied. MD simulation-based analysis (at 200 ns) was performed and comparative analysis with respect to wild type 'CYP2A6-coumarin' complex was done. Though observation didn't find any global effect on complete complex but found some crucial minor-local alteration in interaction and folding process. It is assumed that the change due to SNPs in the single amino acid did not bring global change in physiochemical properties of CYP2A6* but caused local-trivial changes which are very crucial for its metabolic activity.Communicated by Ramaswamy H. Sarma.
More Related Videos
Related Concept Videos
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:30
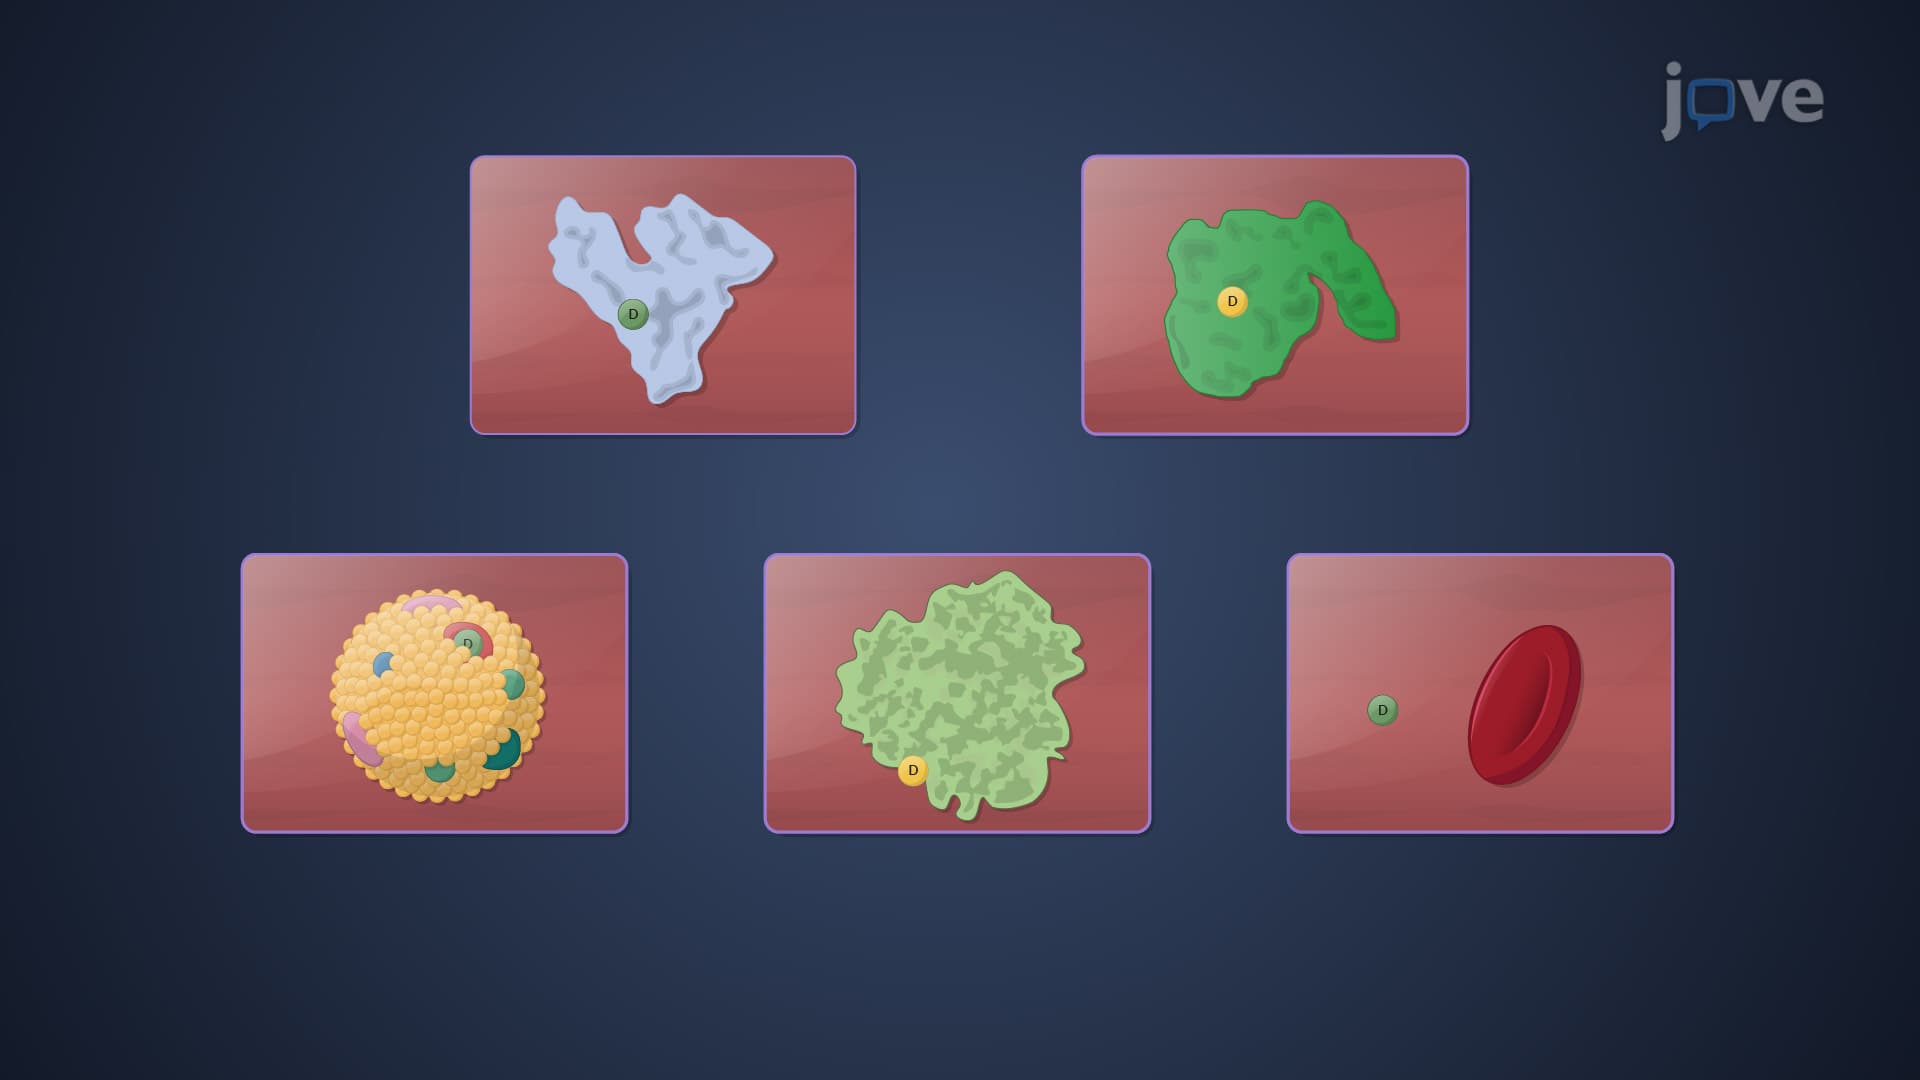
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are...