Related Experiment Video
Updated: Sep 25, 2025

Deacetylation Assays to Unravel the Interplay between Sirtuins SIRT2 and Specific Protein-substrates
Published on: February 27, 2016
Expanding the Accessible Chemical Space of SIRT2 Inhibitors through Exploration of Binding Pocket Dynamics
Nemanja Djokovic1, Dusan Ruzic1, Minna Rahnasto-Rilla2
1Department of Pharmaceutical Chemistry, Faculty of Pharmacy, University of Belgrade, Vojvode Stepe 450, 11221 Belgrade, Serbia.

Abstract:
Considerations of binding pocket dynamics are one of the crucial aspects of the rational design of binders. Identification of alternative conformational states or cryptic subpockets could lead to the discovery of completely novel groups of the ligands. However, experimental characterization of pocket dynamics, besides being expensive, may not be able to elucidate all of the conformational states relevant for drug discovery projects. In this study, we propose the protocol for computational simulations of sirtuin 2 (SIRT2) binding pocket dynamics and its integration into the structure-based virtual screening (SBVS) pipeline. Initially, unbiased molecular dynamics simulations of SIRT2:inhibitor complexes were performed using optimized force field parameters of SIRT2 inhibitors. Time-lagged independent component analysis (tICA) was used to design pocket-related collective variables (CVs) for enhanced sampling of SIRT2 pocket dynamics. Metadynamics simulations in the tICA eigenvector space revealed alternative conformational states of the SIRT2 binding pocket and the existence of a cryptic subpocket. Newly identified SIRT2 conformational states outperformed experimentally resolved states in retrospective SBVS validation. After performing prospective SBVS, compounds from the under-represented portions of the SIRT2 inhibitor chemical space were selected for in vitro evaluation. Two compounds, NDJ18 and NDJ85, were identified as potent and selective SIRT2 inhibitors, which validated the in silico protocol and opened up the possibility for generalization and broadening of its application. The anticancer effects of the most potent compound NDJ18 were examined on the triple-negative breast cancer cell line. Results indicated that NDJ18 represents a promising structure suitable for further evaluation.

Related Concept Videos
 00:49
Ligand Binding and Linkage
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
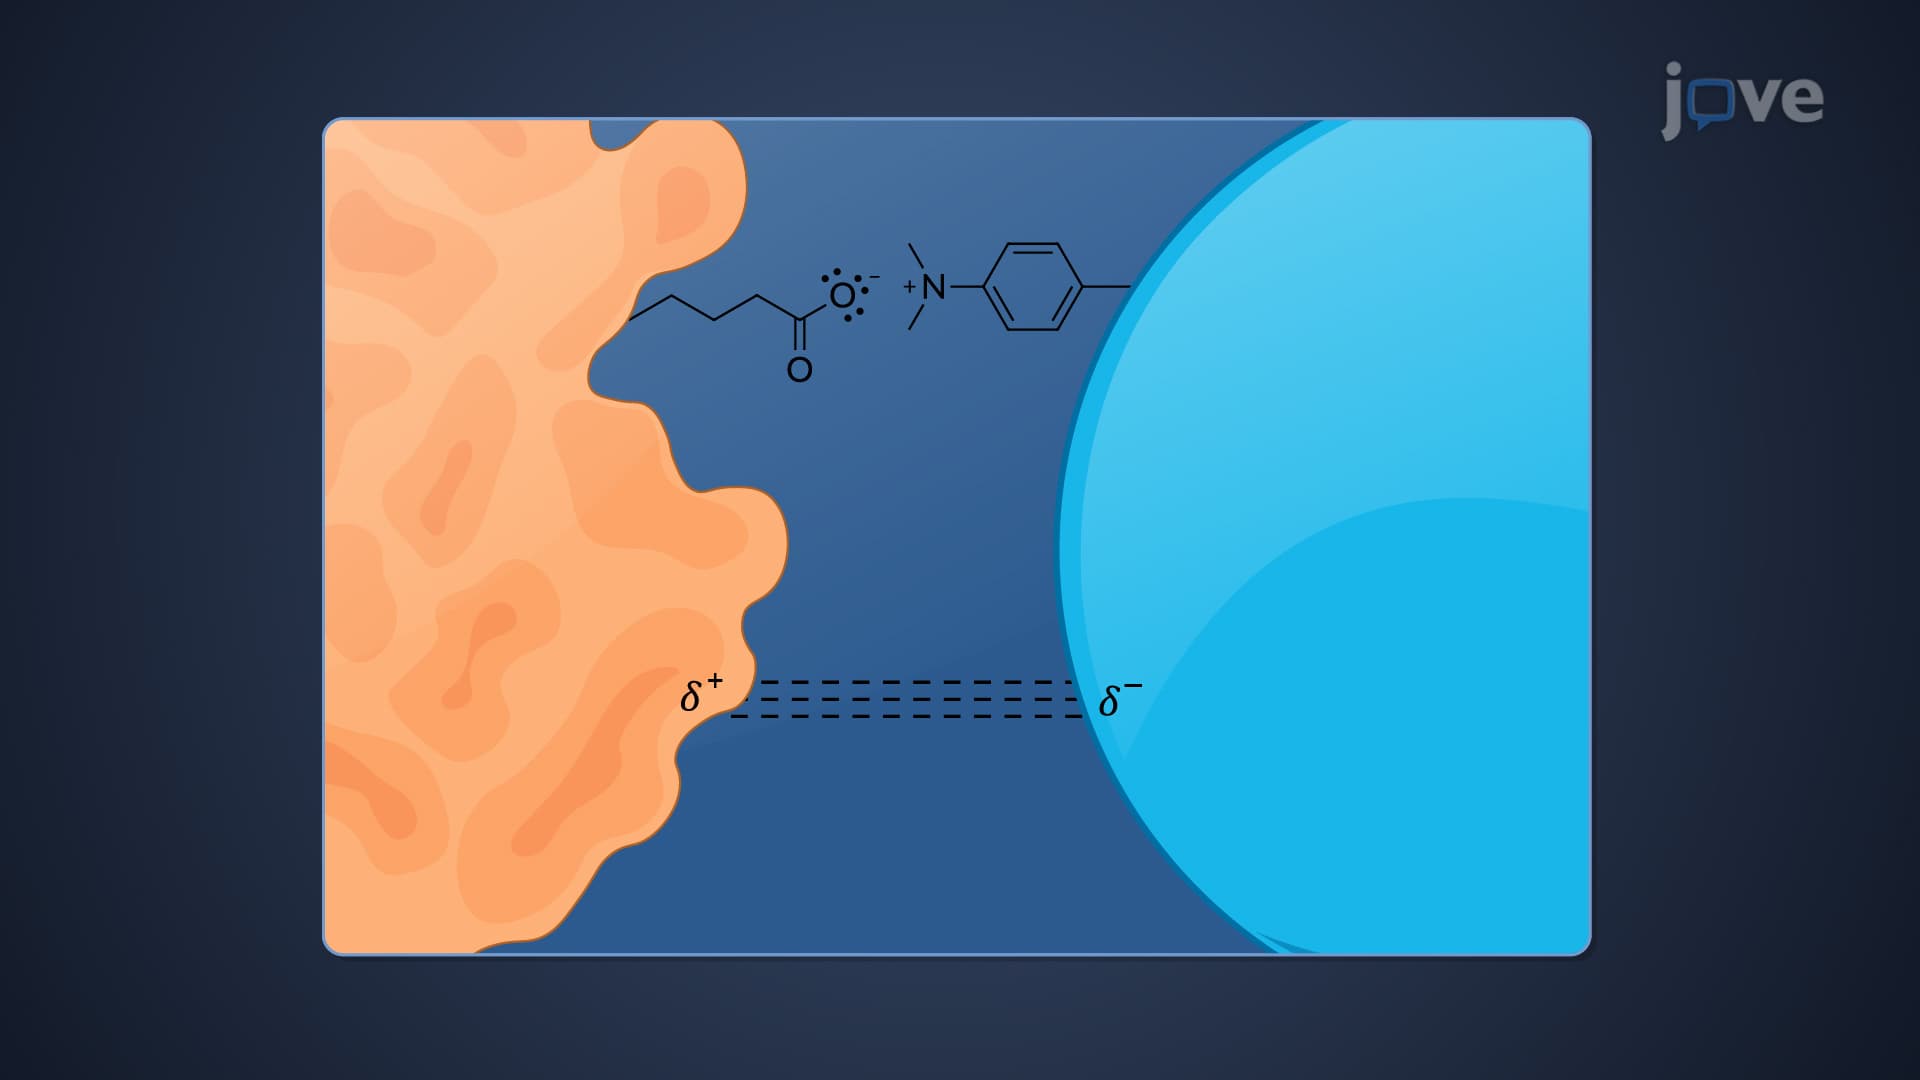
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:58
Cooperative Allosteric Transitions