Related Experiment Video
Updated: Sep 25, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Variational Density Functional Calculations of Excited States: Conical Intersection and Avoided Crossing in Ethylene
Yorick L A Schmerwitz1, Aleksei V Ivanov1, Elvar Ö Jónsson1
1Science Institute of the University of Iceland, VR-III, 107 Reykjavík, Iceland.

Abstract:
Theoretical studies of photochemical processes require a description of the energy surfaces of excited electronic states, especially near degeneracies, where transitions between states are most likely. Systems relevant to photochemical applications are typically too large for high-level multireference methods, and while time-dependent density functional theory (TDDFT) is efficient, it can fail to provide the required accuracy. A variational, time-independent density functional approach is applied to the twisting of the double bond and pyramidal distortion in ethylene, the quintessential model for photochemical studies. By allowing for symmetry breaking, the calculated energy surfaces exhibit the correct topology around the twisted-pyramidalized conical intersection even when using a semilocal functional approximation, and by including explicit self-interaction correction, the torsional energy curves are in close agreement with published multireference results. The findings of the present work point to the possibility of using a single determinant time-independent density functional approach to simulate nonadiabatic dynamics, even for large systems where multireference methods are impractical and TDDFT is often not accurate enough.

Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
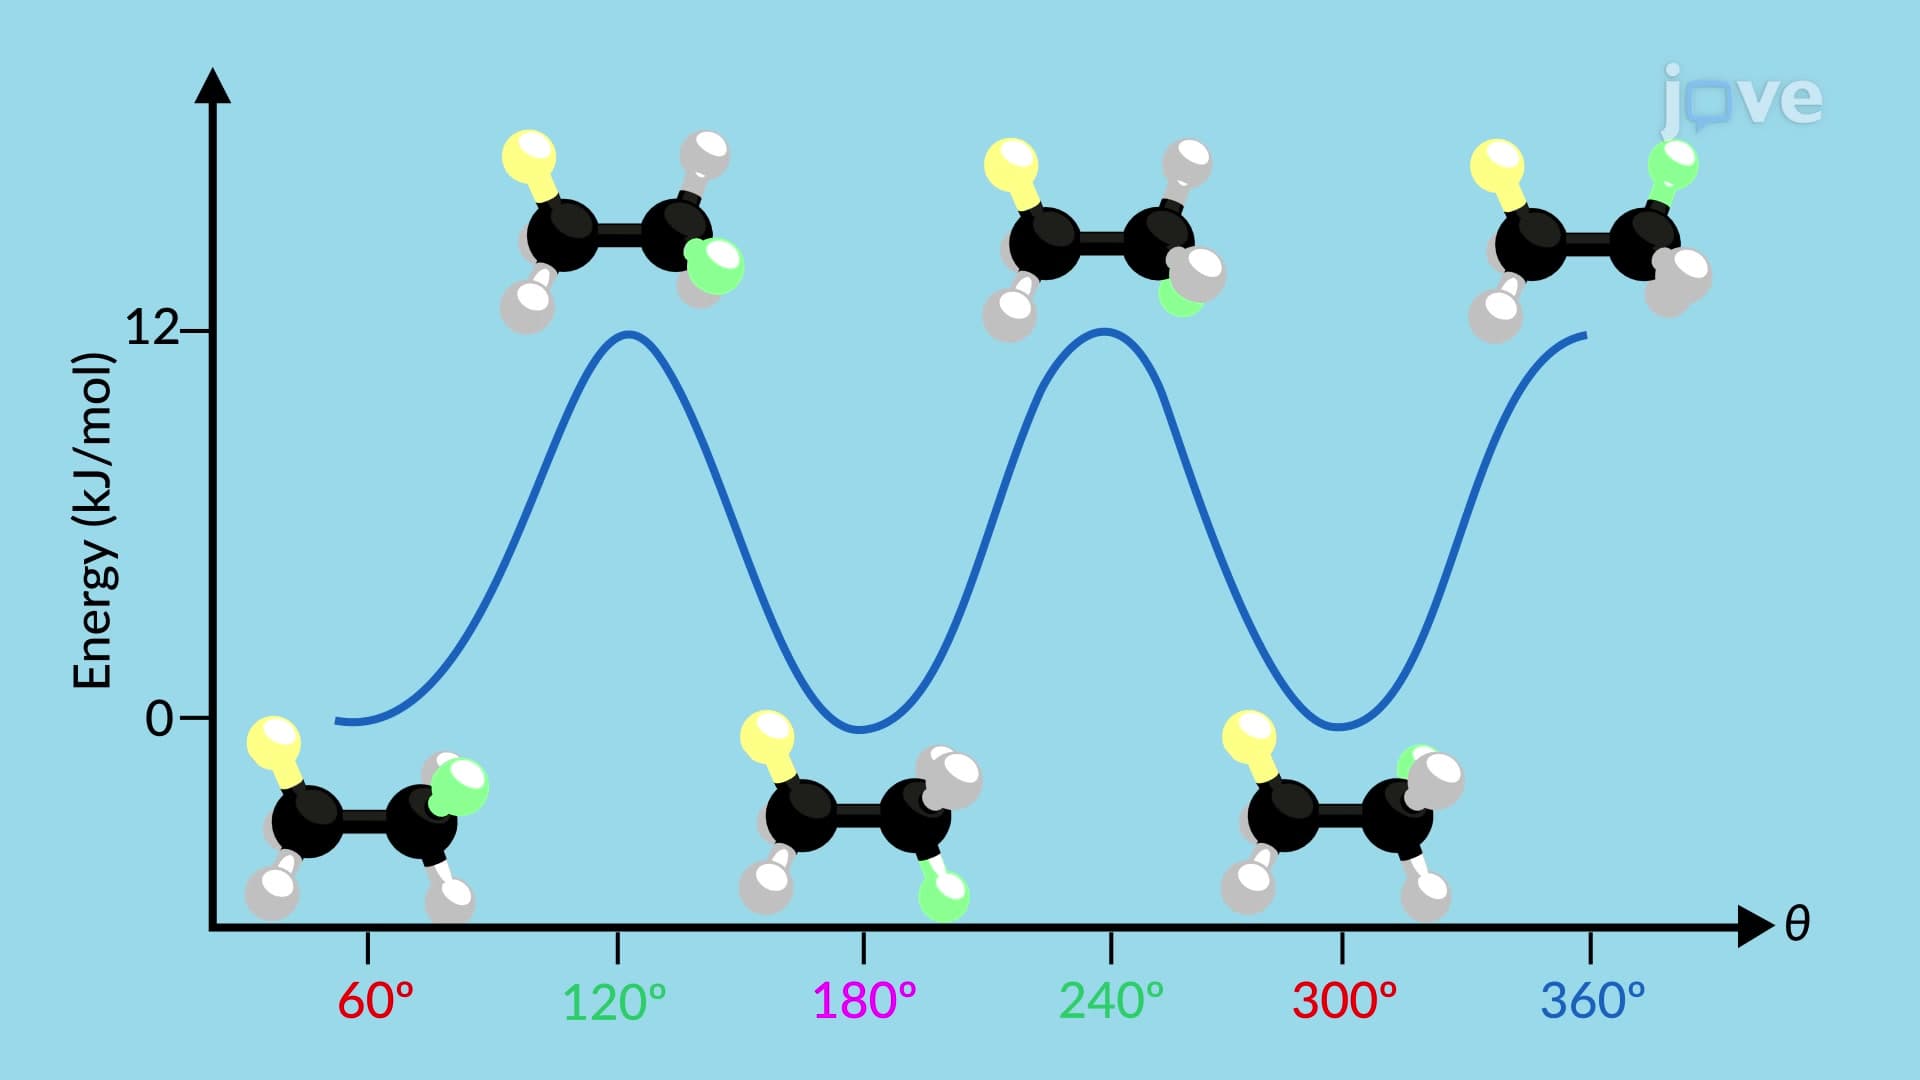 02:18
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
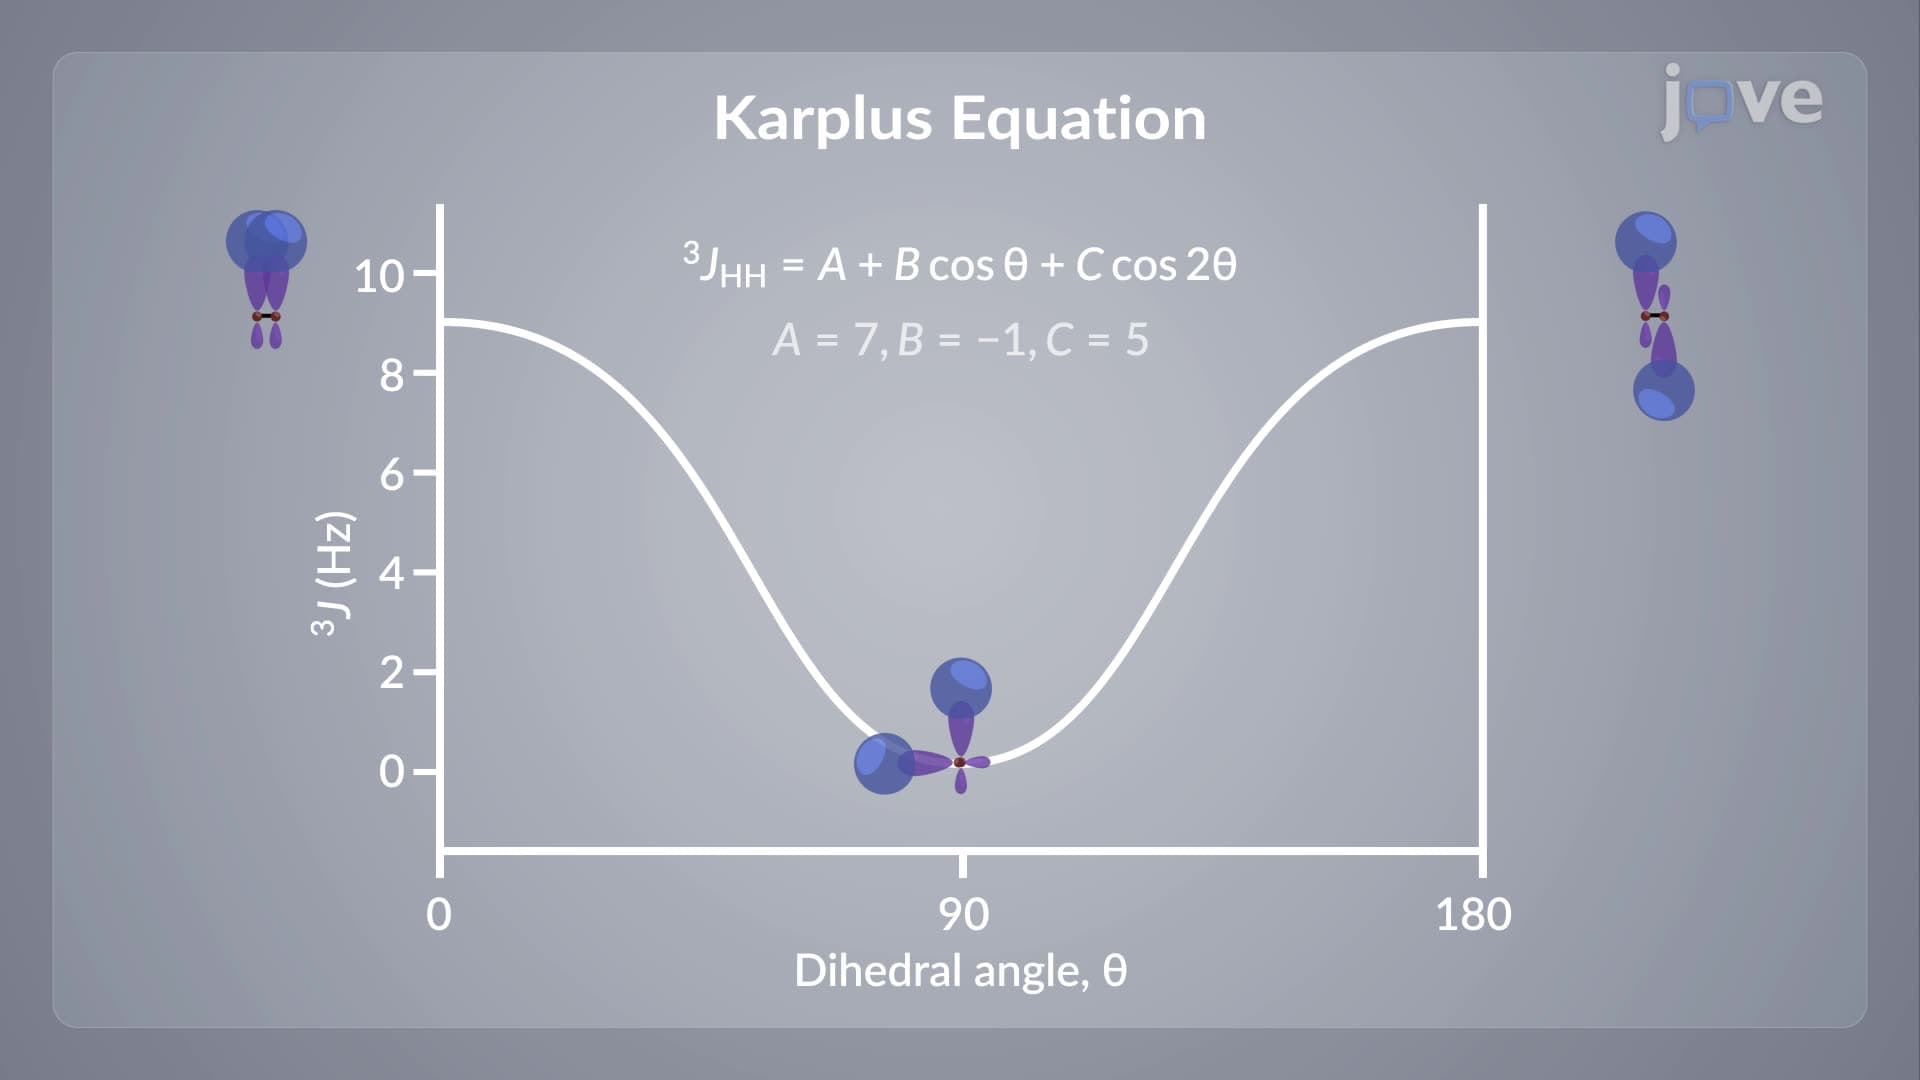 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:16
Structure of Conjugated Dienes
Conjugated dienes are compounds characterized by the presence of alternating double and single bonds. In a conjugated system like 1,3-butadiene, the unhybridized 2p orbital on each carbon overlaps continuously, allowing the π electrons to be delocalized across the entire molecule. In contrast, this type of overlap does not occur in cumulated and isolated dienes, such as 2,3-pentadiene and 1,4-pentadiene, respectively. Instead, the π electrons remain localized between the double...