A bioinformatic pipeline for simulating viral integration data
Related Experiment Video
Updated: Sep 25, 2025
Amplification, Next-generation Sequencing, and Genomic DNA Mapping of Retroviral Integration Sites
Published on: March 22, 2016
Suzanne Scott1,2,3, Susanna Grigson4, Felix Hartkopf5
1Australian e-Health Research Centre, Commonwealth Scientific and Industrial Research Organisation, North Ryde, Australia.

Abstract:
Viral integration is a complex biological process, and it is useful to have a reference integration dataset with known properties to compare experimental data against, or for comparing with the results from computational tools that detect integration. To generate these data, we developed a pipeline for simulating integrations of a viral or vector genome into a host genome. Our method reproduces more complex characteristics of vector and viral integration, including integration of sub-genomic fragments, structural variation of the integrated genomes, and deletions from the host genome at the integration site. Our method [1] takes the form of a snakemake [2] pipeline, consisting of a Python [3] script using the Biopython [4] module that simulates integrations of a viral reference into a host reference. This produces a reference containing integrations, from which sequencing reads are simulated using ART [5]. The IDs of the reads crossing integration junctions are then annotated using another python script to produce the final output, consisting of the simulated reads and a table of the locations of those integrations and the reads crossing each integration junction. To illustrate our method, we provide simulated reads, integration locations, as well as the code required to simulate integrations using any virus and host reference. This simulation method was used to investigate the performance of viral integration tools in our research [6].
More Related Videos
Related Concept Videos
 01:26
Size and Structure of Viral Genomes
 01:29
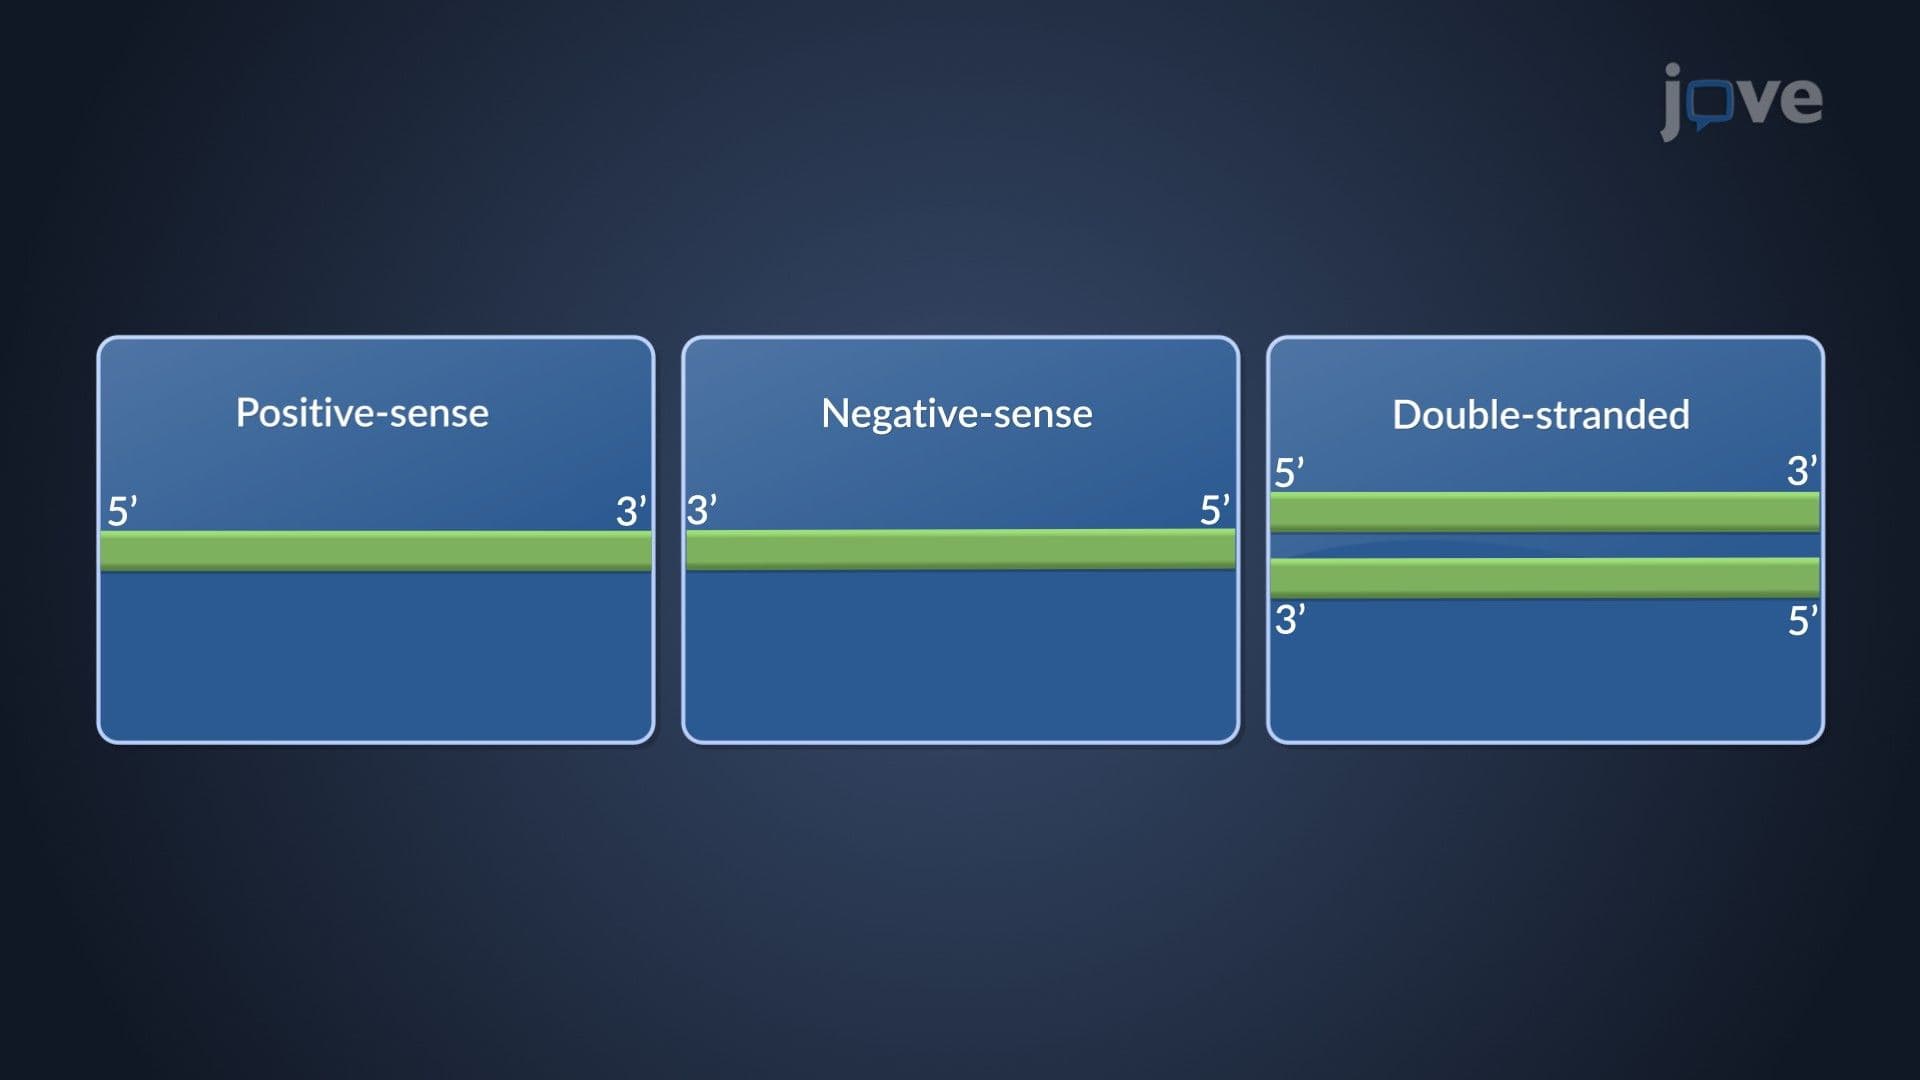
Viruses with RNA Genomes
 00:57
Viral Recombination
 00:36
Viral Mutations