Related Experiment Video
Updated: Sep 24, 2025


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Nature of intramolecular O-H⋯π interactions as elucidated by QTAIM dual functional analysis with QC calculations
Satoko Hayashi1, Taro Nishide1, Waro Nakanishi1
1Faculty of Systems Engineering, Wakayama University 930 Sakaedani Wakayama 640-8510 Japan hayashi3@sys.wakayama-u.ac.jp nakanisi@sys.wakayama-u.ac.jp +81 73 457 8253 +81 73 457 8252.

Abstract:
The intrinsic dynamic and static nature of intramolecular OH-*-π interactions is elucidated using a QTAIM dual functional analysis (QTAIM-DFA) after clarifying the structural features. Asterisks (*) are employed to emphasize the presence of bond critical points (BCPs) on the bond paths (BPs), which correspond to the interactions in question. Data from the fully optimized structures correspond to the static nature of the interactions. In our treatment, data from the perturbed structures, which are based around the fully optimized structure, are employed for the analysis in addition to those from the fully optimized structure, which represent the dynamic nature of the interaction. Seven intramolecular OH-*-C(π) interactions were detected in six-membered rings, with six BPs and BCPs for each, among the 72 conformers of the species examined here (1-15). The interactions are predicted to have a vdW or t-HBnc (typical hydrogen bonds with no covalency) nature, which appeared in the pure closed shell region. They appear to be stronger than the corresponding intermolecular interactions. Nine BPs with BCPs were also detected for the intramolecular O-*-X interactions (X = C(π) and H(π), joined to C(π)) in the 5-7-membered rings. The E(2) values of the interactions, as obtained by NBO, are discussed in relation to the stabilities of the conformers and the BPs with BCPs.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:51
Molecular Orbital Theory II
 03:00
Intermolecular vs Intramolecular Forces
 02:35
Molecular Orbital Theory I
 03:35
Hybridization of Atomic Orbitals II
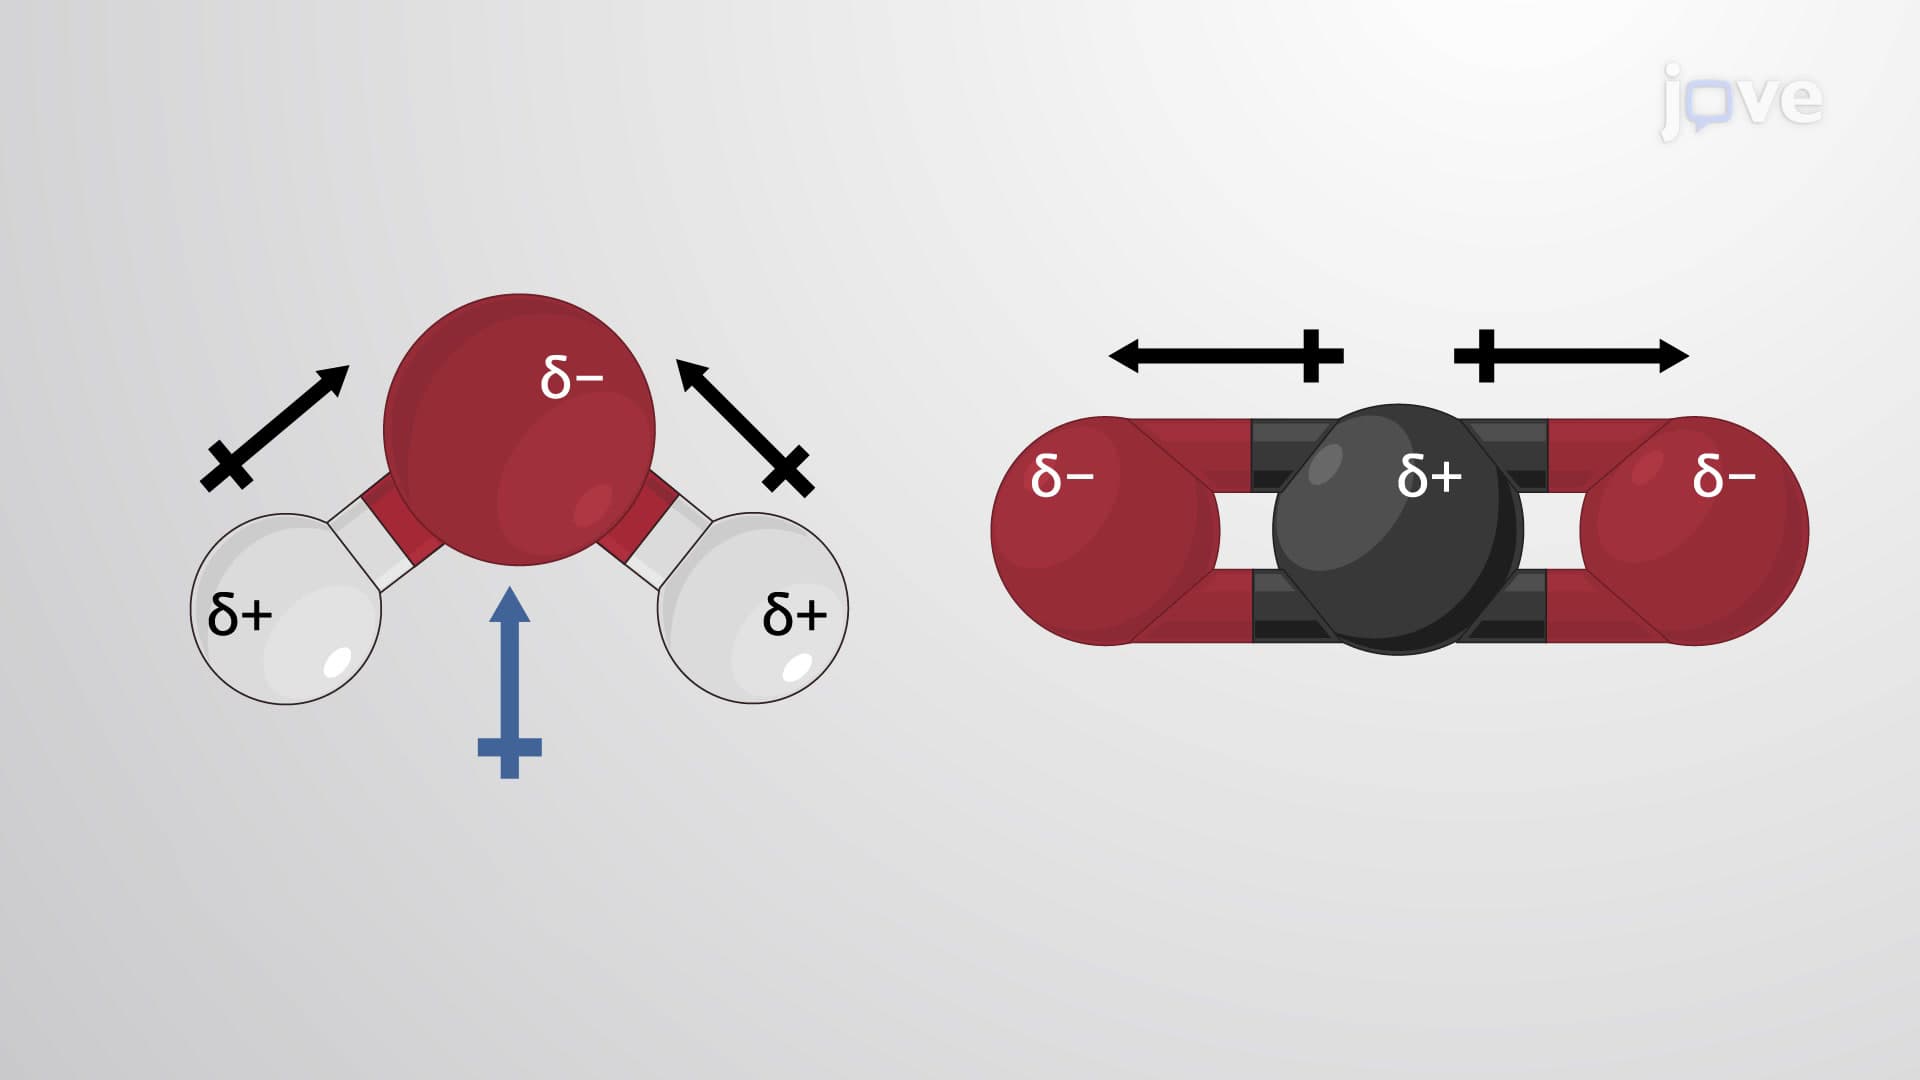 02:36
Molecular Geometry and Dipole Moments
 03:24
Hybridization of Atomic Orbitals I