Related Experiment Video
Updated: Sep 24, 2025

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Pathological remodeling of distal lung matrix in end-stage cystic fibrosis patients
Meghan R Pinezich1, Manuel A Tamargo2, Sharon Fleischer1
1Department of Biomedical Engineering, Columbia University, New York, NY, USA.

Background:
Manifestations of cystic fibrosis, although well-characterized in the proximal airways, are understudied in the distal lung. Characterization of the cystic fibrosis lung 'matrisome' (matrix proteome) has not been previously described, and could help identify biomarkers and inform therapeutic strategies.
Methods:
We performed liquid chromatography-mass spectrometry, gene ontology analysis, and multi-modal imaging, including histology, immunofluorescence, and electron microscopy for a comprehensive evaluation of distal human lung extracellular matrix (matrix) structure and composition in end-stage cystic fibrosis.
Results:
Quantitative proteomic profiling identified sixty-eight (68) matrix constituents with significantly altered expression in end-stage cystic fibrosis. Over 90% of significantly different matrix peptides detected, including structural and basement membrane proteins, were expressed at lower levels in cystic fibrosis. However, the total abundance of matrix in cystic fibrosis lungs was not significantly different from control lungs, suggesting that cystic fibrosis leads to loss of diversity among lung matrix proteins rather than an absolute loss of matrix. Visualization of distal lung matrix via immunofluorescence and electron microscopy revealed pathological remodeling of distal lung tissue architecture and loss of alveolar basement membrane, consistent with significantly altered pathways identified by gene ontology analysis.
Conclusions:
Dysregulation of matrix organization and aberrant wound healing pathways are associated with loss of matrix protein diversity and obliteration of distal lung tissue structure in end-stage cystic fibrosis. While many therapeutics aim to functionally restore defective cystic fibrosis transmembrane conductance regulator (CFTR), drugs that target dysregulated matrix pathways may serve as adjunct interventions to support lung recovery.
More Related Videos
 05:25
Author Spotlight: Dissection and Isolation of Region-Specific Decellularized Lung Tissue
Published on: September 29, 2023
 07:38
A Multimodal Imaging Approach Based on Micro-CT and Fluorescence Molecular Tomography for Longitudinal Assessment of Bleomycin-Induced Lung Fibrosis in Mice
Published on: April 13, 2018
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:40
Bone Remodeling
 01:26
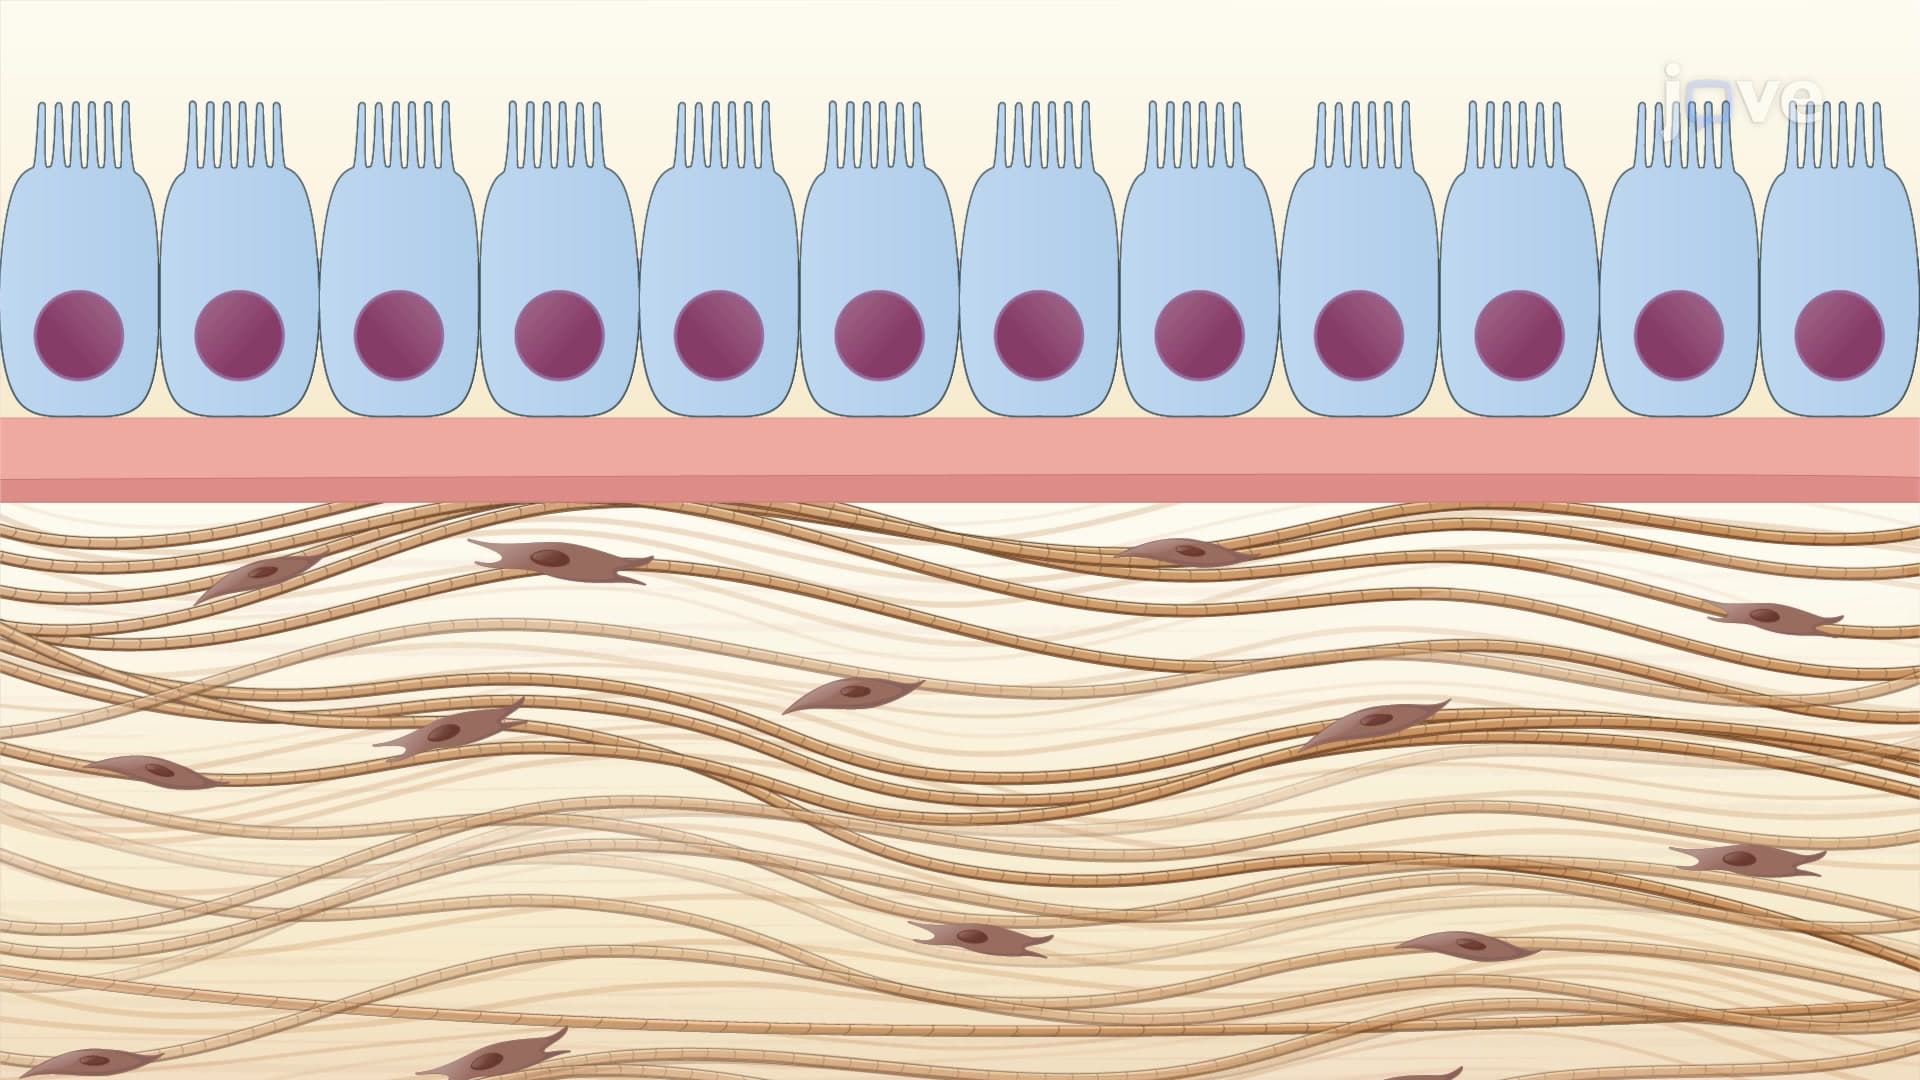
Extracellular Matrix