Related Experiment Video
Updated: Sep 24, 2025

Deep Proteome Profiling by Isobaric Labeling, Extensive Liquid Chromatography, Mass Spectrometry, and Software-assisted Quantification
Published on: November 15, 2017
Proteomic quantification of perturbation to pharmacokinetic target proteins in liver disease
Areti-Maria Vasilogianni1, Eman El-Khateeb2, Zubida M Al-Majdoub1
1Centre for Applied Pharmacokinetic Research, The University of Manchester, Manchester, UK.

Abstract:
Model-based assessment of drug pharmacokinetics in liver disease requires quantification of abundance and disease-related changes in hepatic enzymes and transporters. This study aimed to assess performance of three label-free methods [high N (HiN), intensity-based absolute quantification (iBAQ) and total protein approach (TPA)] against QconCAT-based targeted data in healthy and diseased (cancer and cirrhosis) liver tissue. Measurements were compared across methods and disease-to-control ratios provided a 'disease perturbation factor' (DPF) for each protein. Mean label-free measurements of targets correlated well (Pearson's coefficient, r = 0.91-0.98 p < 0.001) and with targeted data (r = 0.65-0.95, p < 0.001). Concordance with targeted data was generally moderate (Lin's concordance coefficient, ρc = 0.46-0.92), depending on methodology. Moderate precision and accuracy were observed for label-free methods (average fold error, AFE = 1.44-1.68; absolute average fold error, AAFE = 2.44-3.23). The DPF reconciled the data and indicated downregulated expression in cancer and cirrhosis, consistent with an inflammatory effect. HiN estimated perturbation consistently with targeted data (AFEHiN = 1.07, AAFEHiN = 1.57), whereas iBAQ overestimated (AFEiBAQ = 0.81, AAFEiBAQ = 1.67) and TPA underestimated (AFETPA = 1.37, AAFETPA = 1.65) disease effect. Progression from mild to severe cirrhosis was consistent with progressive decline in expression, reproduced by HiN but overestimated by iBAQ and underestimated by TPA (AFEHiN = 0.98, AFEiBAQ = 0.60, AFETPA = 1.24). DPF data confirmed non-uniform disease effect on drug-elimination pathways and progressive impact of disease severity. SIGNIFICANCE: This study demonstrated good correlation and moderate concordance between intensity-based label-free proteomic methods (HiN, iBAQ and TPA) and targeted data. Label-free measurements tended to overestimate abundance, but differences were reconciled using a disease perturbation factor (DPF) for each protein. With targeted data as a reference, HiN defined disease perturbation and the impact of disease progression consistently, indicating that the use of 'razor' peptides for quantification against an exogenous standard provides biologically sensible quantitative fingerprints of disease. Disease-driven perturbations in expression relative to healthy baseline are incorporated into drug kinetic models used to predict drug exposure in disease populations where clinical studies may not be feasible.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:09
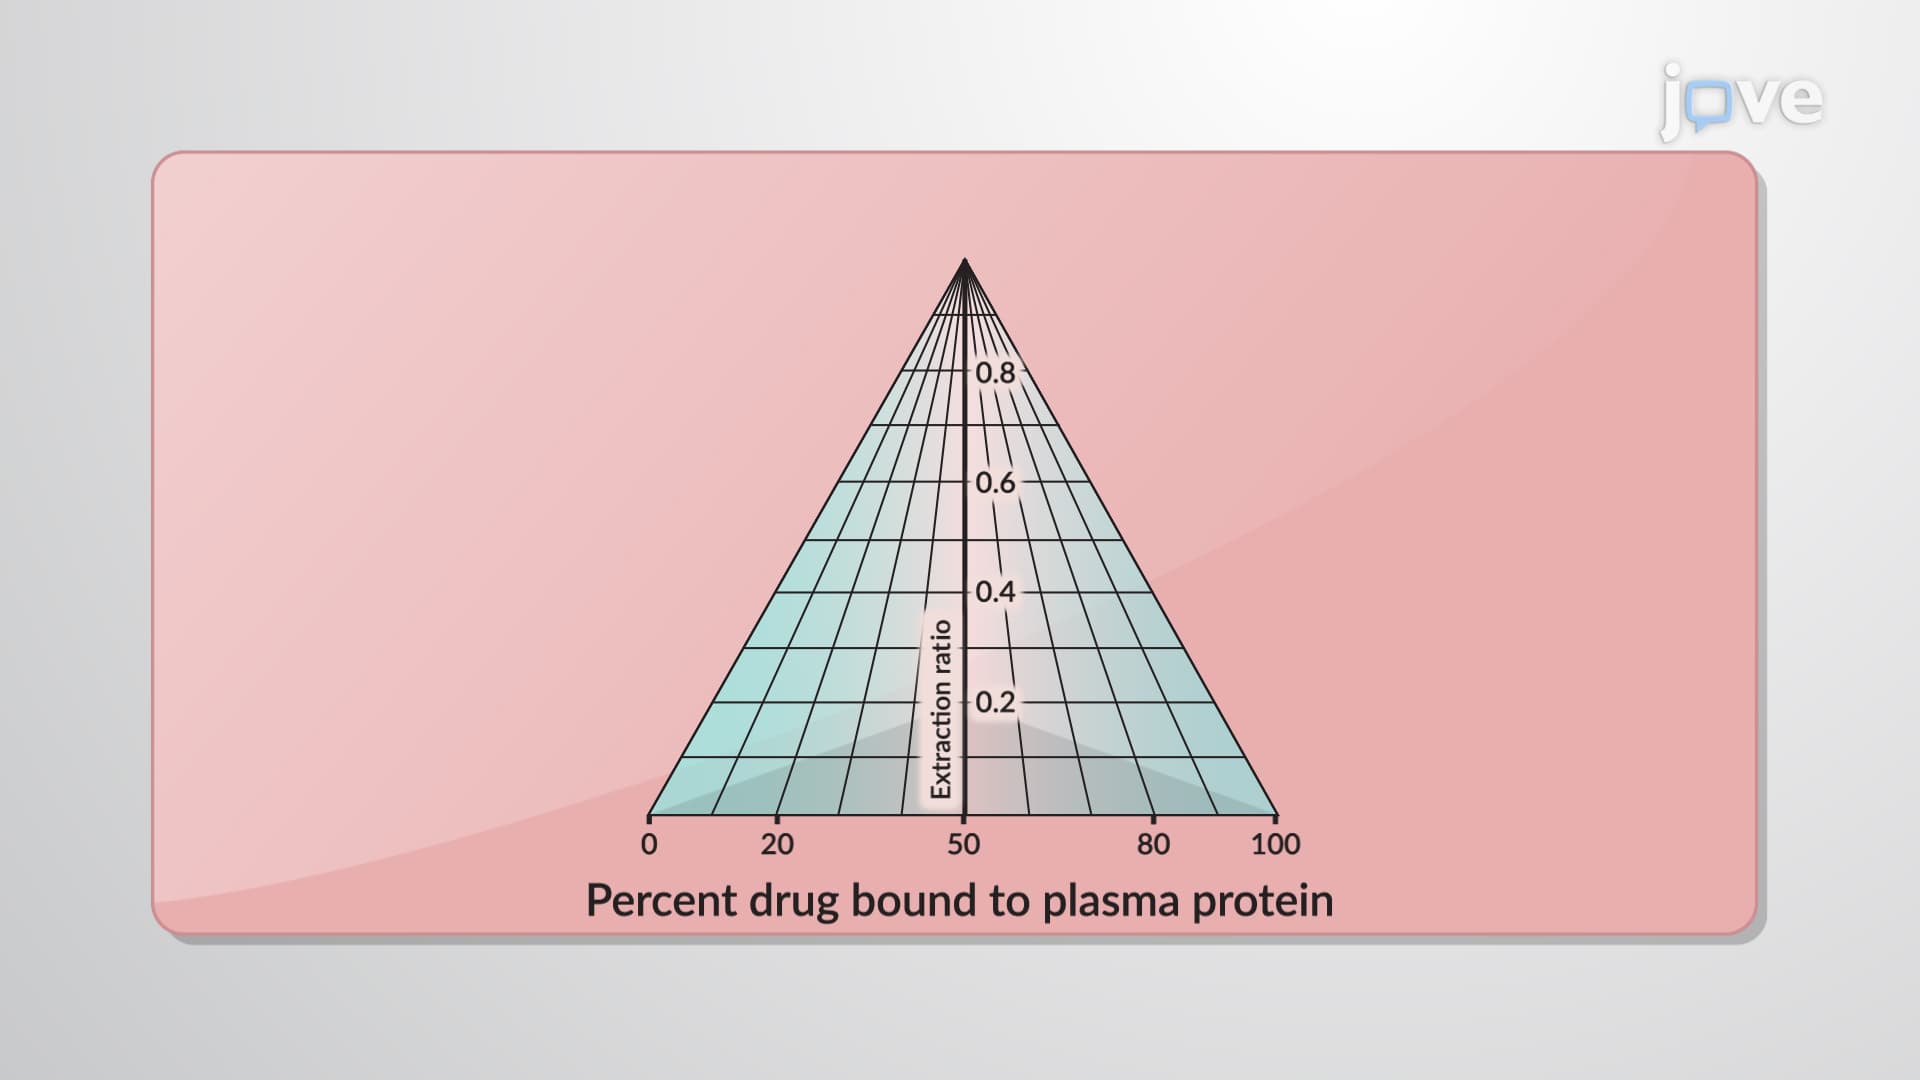
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:22
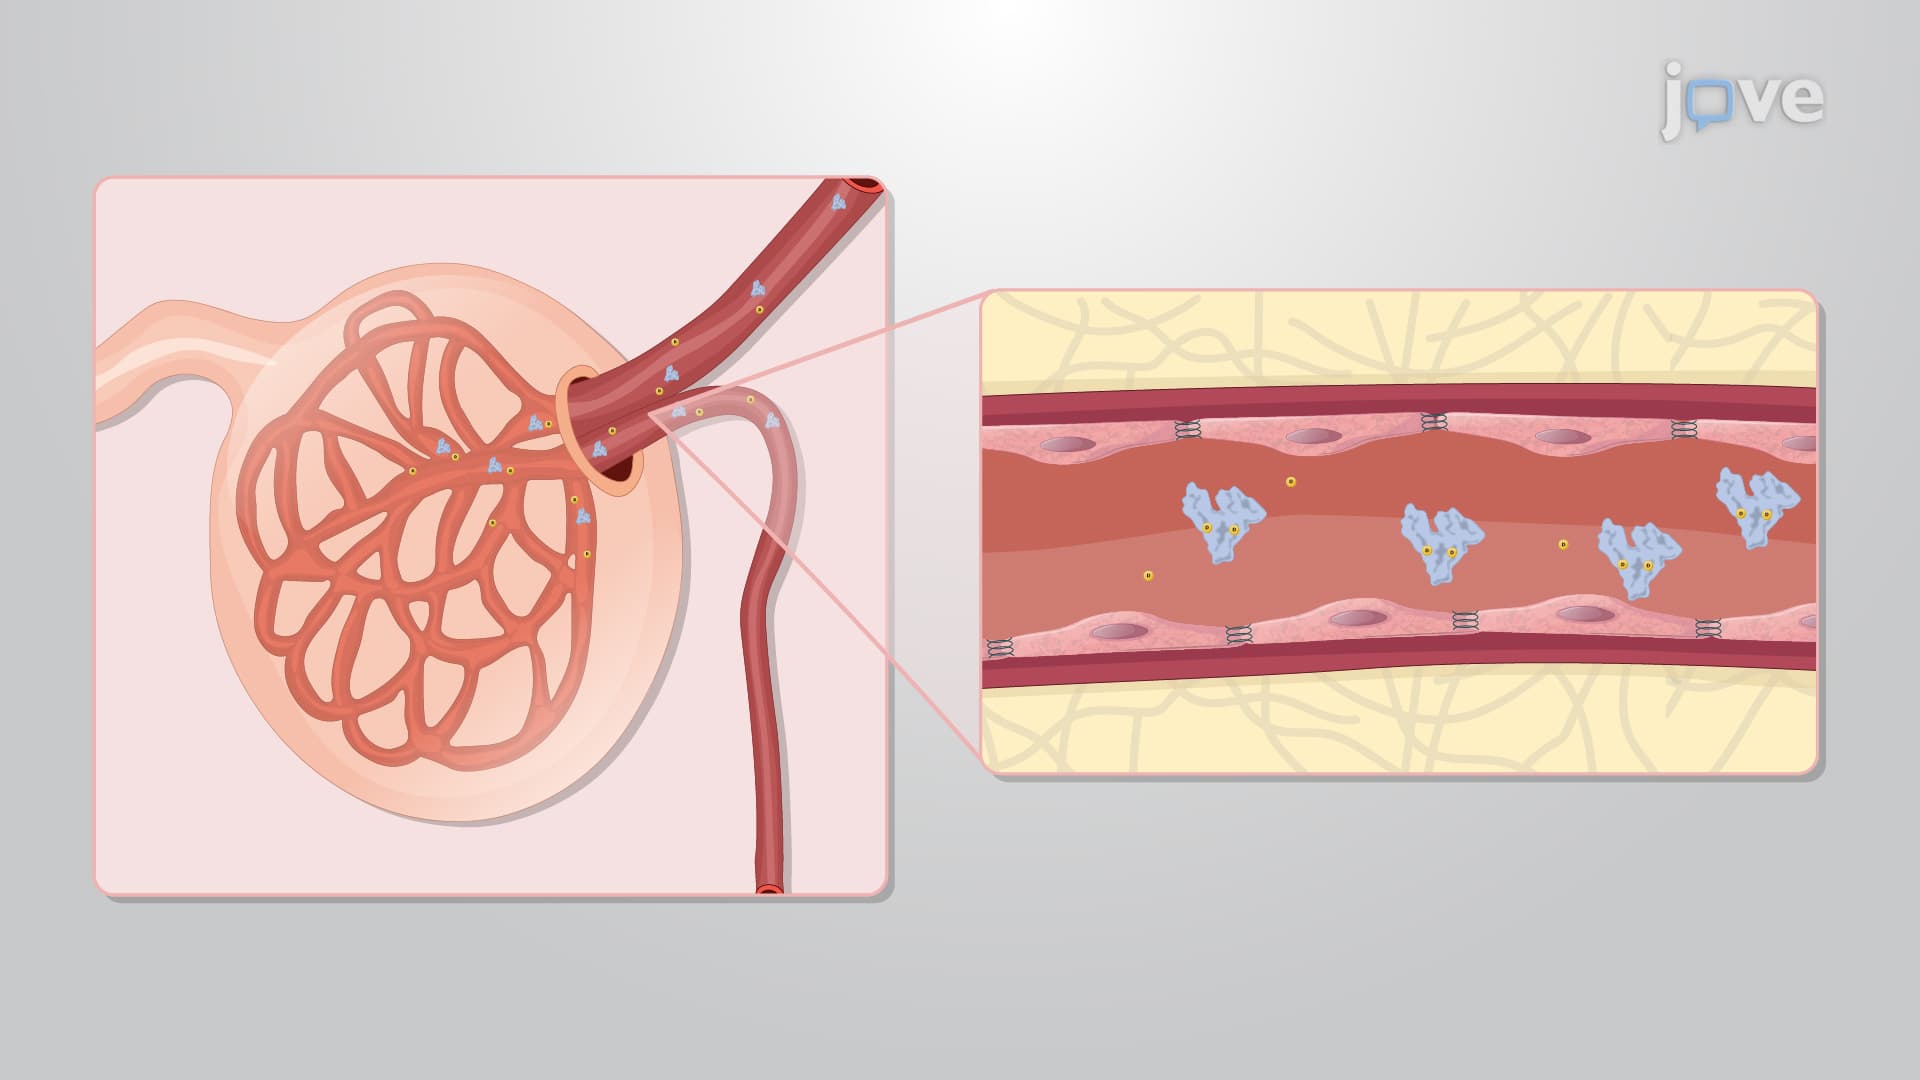
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:27
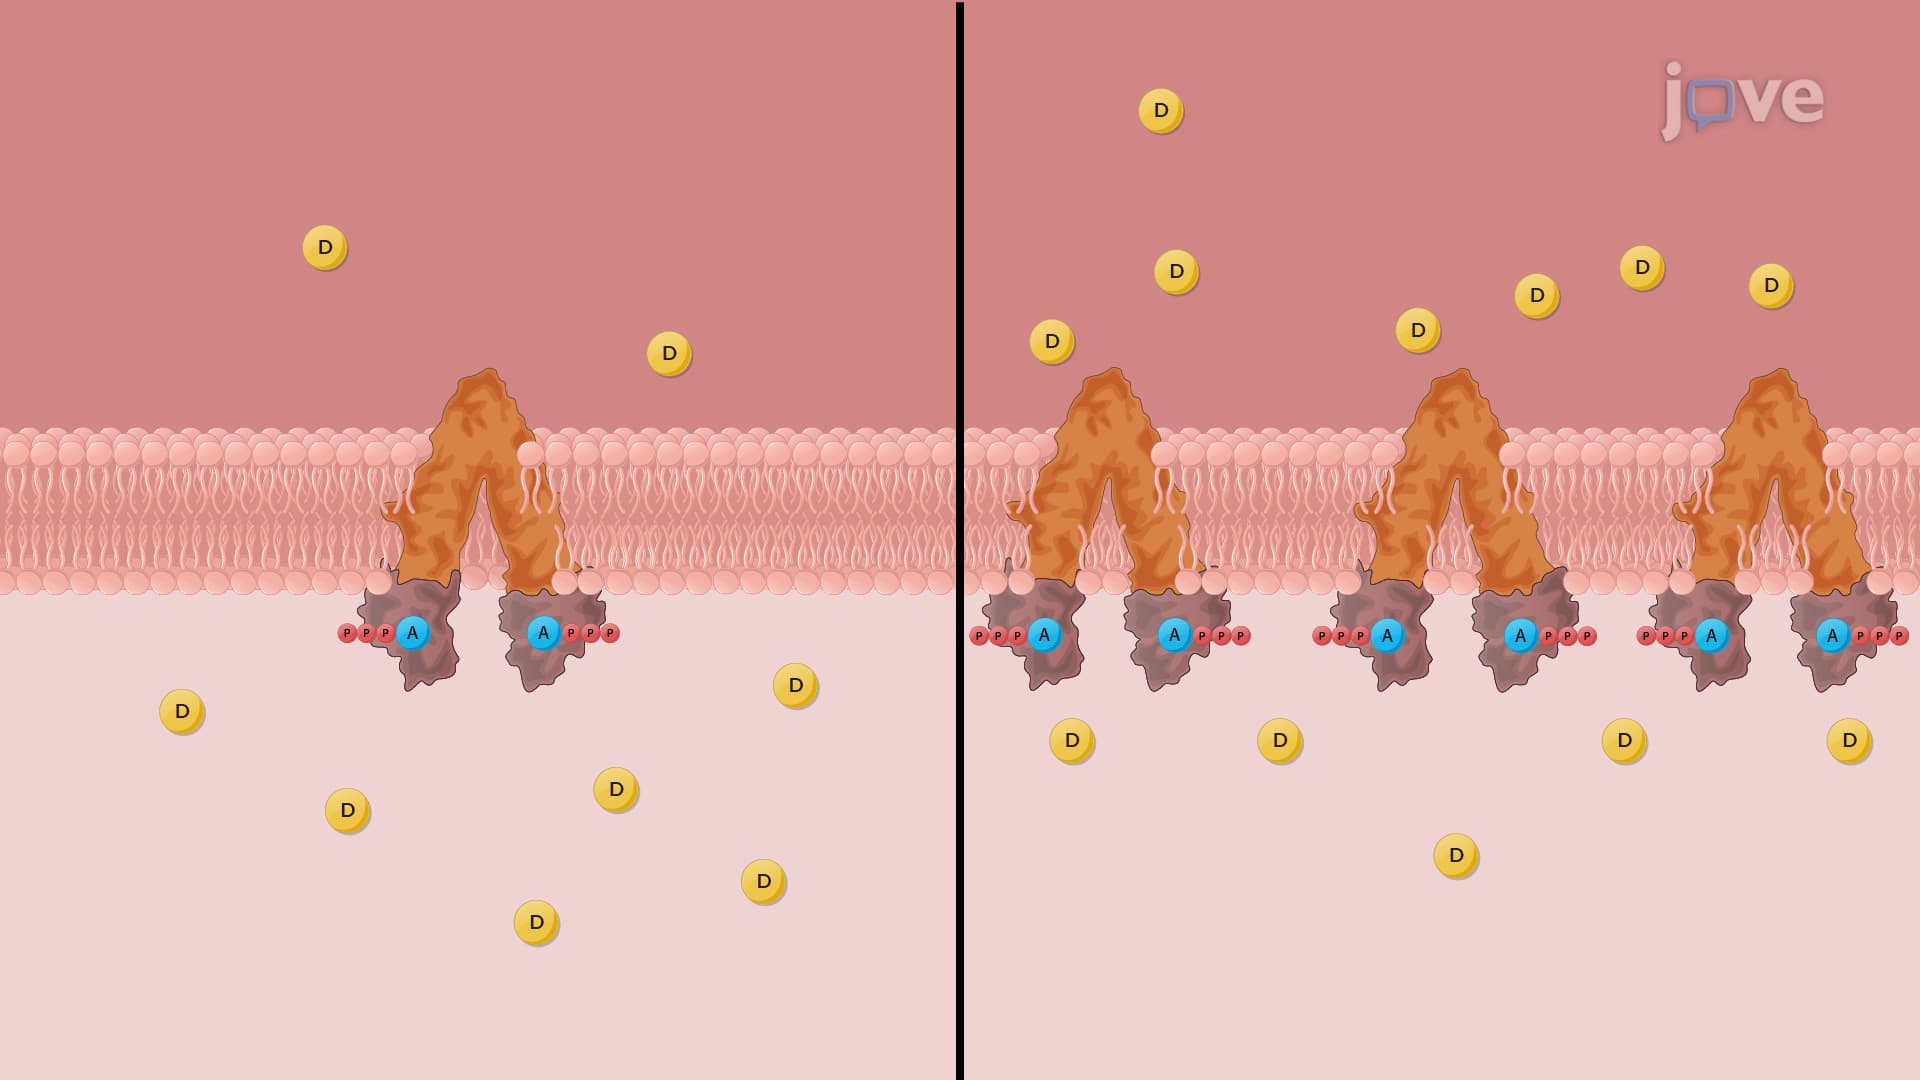
Nonlinear Pharmacokinetics: Role of Transporters
Polymorphisms occurring in drug transporters can alter...