Related Experiment Video
Updated: Sep 22, 2025

Author Spotlight: Advancing Cellular and Protein Engineering to Control Biological Functions and Develop Novel Therapies
Published on: September 27, 2024
8-azaadenosine and 8-chloroadenosine are not selective inhibitors of ADAR
Kyle A Cottrell1, Luisangely Soto Torres1, Michael G Dizon1
1Department of Medicine, Division of Molecular Oncology, Washington University School of Medicine, Saint Louis, Missouri, USA.

Abstract:
The RNA editing enzyme ADAR, is an attractive therapeutic target for multiple cancers. Through its deaminase activity, ADAR edits adenosine to inosine in dsRNAs. Loss of ADAR in some cancer cell lines causes activation of the type I interferon pathway and the PKR translational repressor, leading to inhibition of proliferation and stimulation of cell death. As such, inhibition of ADAR function is a viable therapeutic strategy for many cancers. However, there are no FDA approved inhibitors of ADAR. Two small molecules have been previously shown to inhibit ADAR or reduce its expression: 8-azaadenosine and 8-chloroadenosine. Here we show that neither molecule is a selective inhibitor of ADAR. Both 8-azaadenosine and 8-chloroadenosine show similar toxicity to ADAR-dependent and independent cancer cell lines. Furthermore, the toxicity of both small molecules is comparable between cell lines with either knockdown or overexpression of ADAR, and cells with unperturbed ADAR expression. Treatment with neither molecule causes activation of PKR. Finally, treatment with either molecule has no effect on A-to-I editing of multiple ADAR substrates. Together these data show that 8-azaadenosine and 8-chloroadenosine are not suitable small molecules for therapies that require selective inhibition of ADAR, and neither should be used in preclinical studies as ADAR inhibitors.
Insights
The RNA editing enzyme ADAR is a cancer target, but 8-azaadenosine and 8-chloroadenosine are not selective ADAR inhibitors. These compounds show similar toxicity across cancer cells, regardless of ADAR levels, making them unsuitable for ADAR-targeted therapies.
Area of Science:
- Biochemistry
- Molecular Biology
- Cancer Therapeutics
Background:
- Adenosine deaminase acting on RNA (ADAR) enzymes are crucial for RNA editing, converting adenosine to inosine in double-stranded RNAs.
- ADAR dysfunction is implicated in various cancers, making ADAR a promising therapeutic target.
- Inhibition of ADAR can trigger anti-cancer effects via the type I interferon pathway and PKR activation.
Purpose of the Study:
- To evaluate the selectivity and efficacy of 8-azaadenosine and 8-chloroadenosine as ADAR inhibitors for cancer therapy.
- To determine if these small molecules exhibit ADAR-dependent toxicity or modulate ADAR activity.
Main Methods:
- Assessing the toxicity of 8-azaadenosine and 8-chloroadenosine in cancer cell lines with varying ADAR expression levels (knockdown, overexpression, and wild-type).
- Monitoring the activation of the PKR pathway and A-to-I editing of ADAR substrates upon treatment with the small molecules.
- Comparing the effects of the compounds in ADAR-dependent versus ADAR-independent cancer cell lines.
Main Results:
- Neither 8-azaadenosine nor 8-chloroadenosine demonstrated selective inhibition of ADAR.
- Both compounds exhibited comparable toxicity in cancer cell lines irrespective of ADAR expression levels.
- Treatment with these molecules did not lead to PKR activation or affect A-to-I editing of ADAR substrates.
Conclusions:
- 8-azaadenosine and 8-chloroadenosine are not suitable for developing therapies that require selective ADAR inhibition.
- These compounds should not be utilized in preclinical studies as ADAR inhibitors due to their lack of specificity and efficacy.
- Further research is needed to identify potent and selective ADAR inhibitors for cancer treatment.
More Related Videos
 11:29
HPLC-based Assay to Monitor Extracellular Nucleotide/Nucleoside Metabolism in Human Chronic Lymphocytic Leukemia Cells
Published on: July 20, 2016
 10:24
NMR-Based Activity Assays for Determining Compound Inhibition, IC50 Values, Artifactual Activity, and Whole-Cell Activity of Nucleoside Ribohydrolases
Published on: June 30, 2019
Related Concept Videos
 01:17
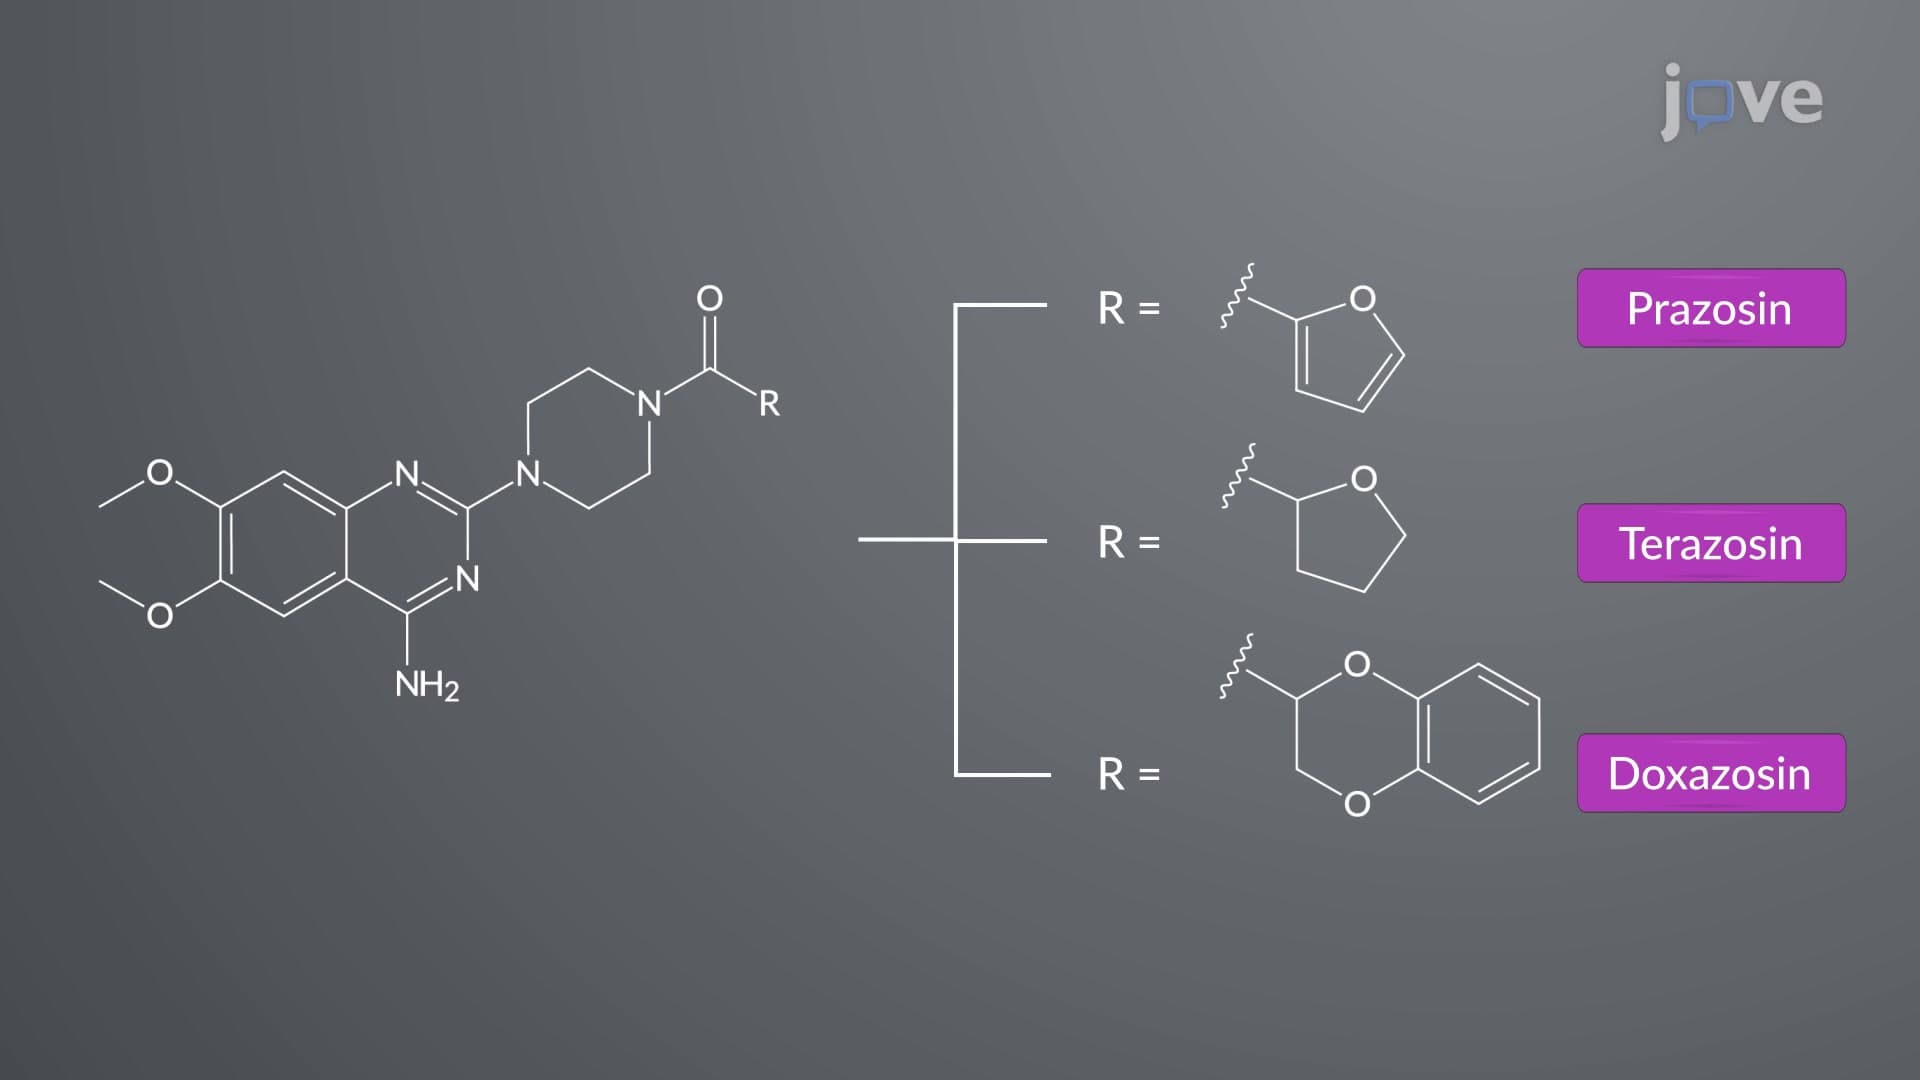
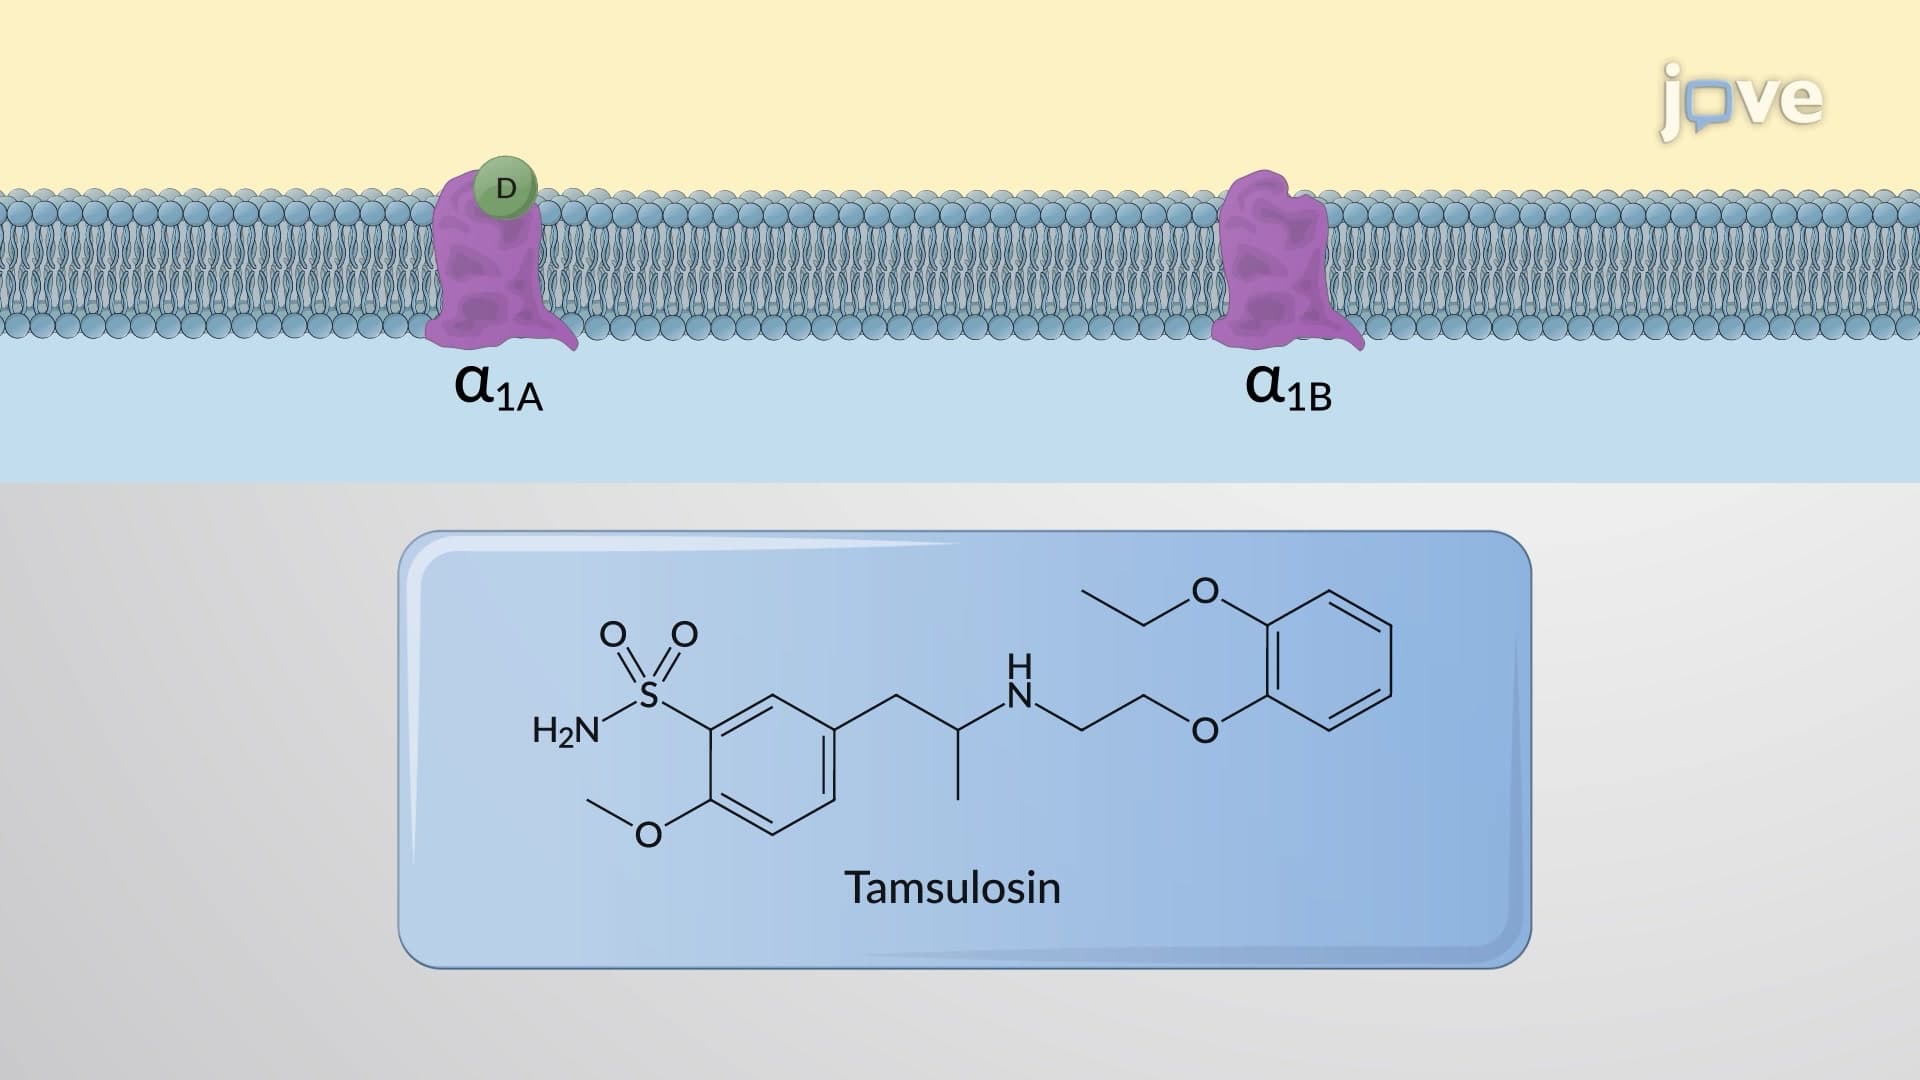
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:30
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally,...
 01:25
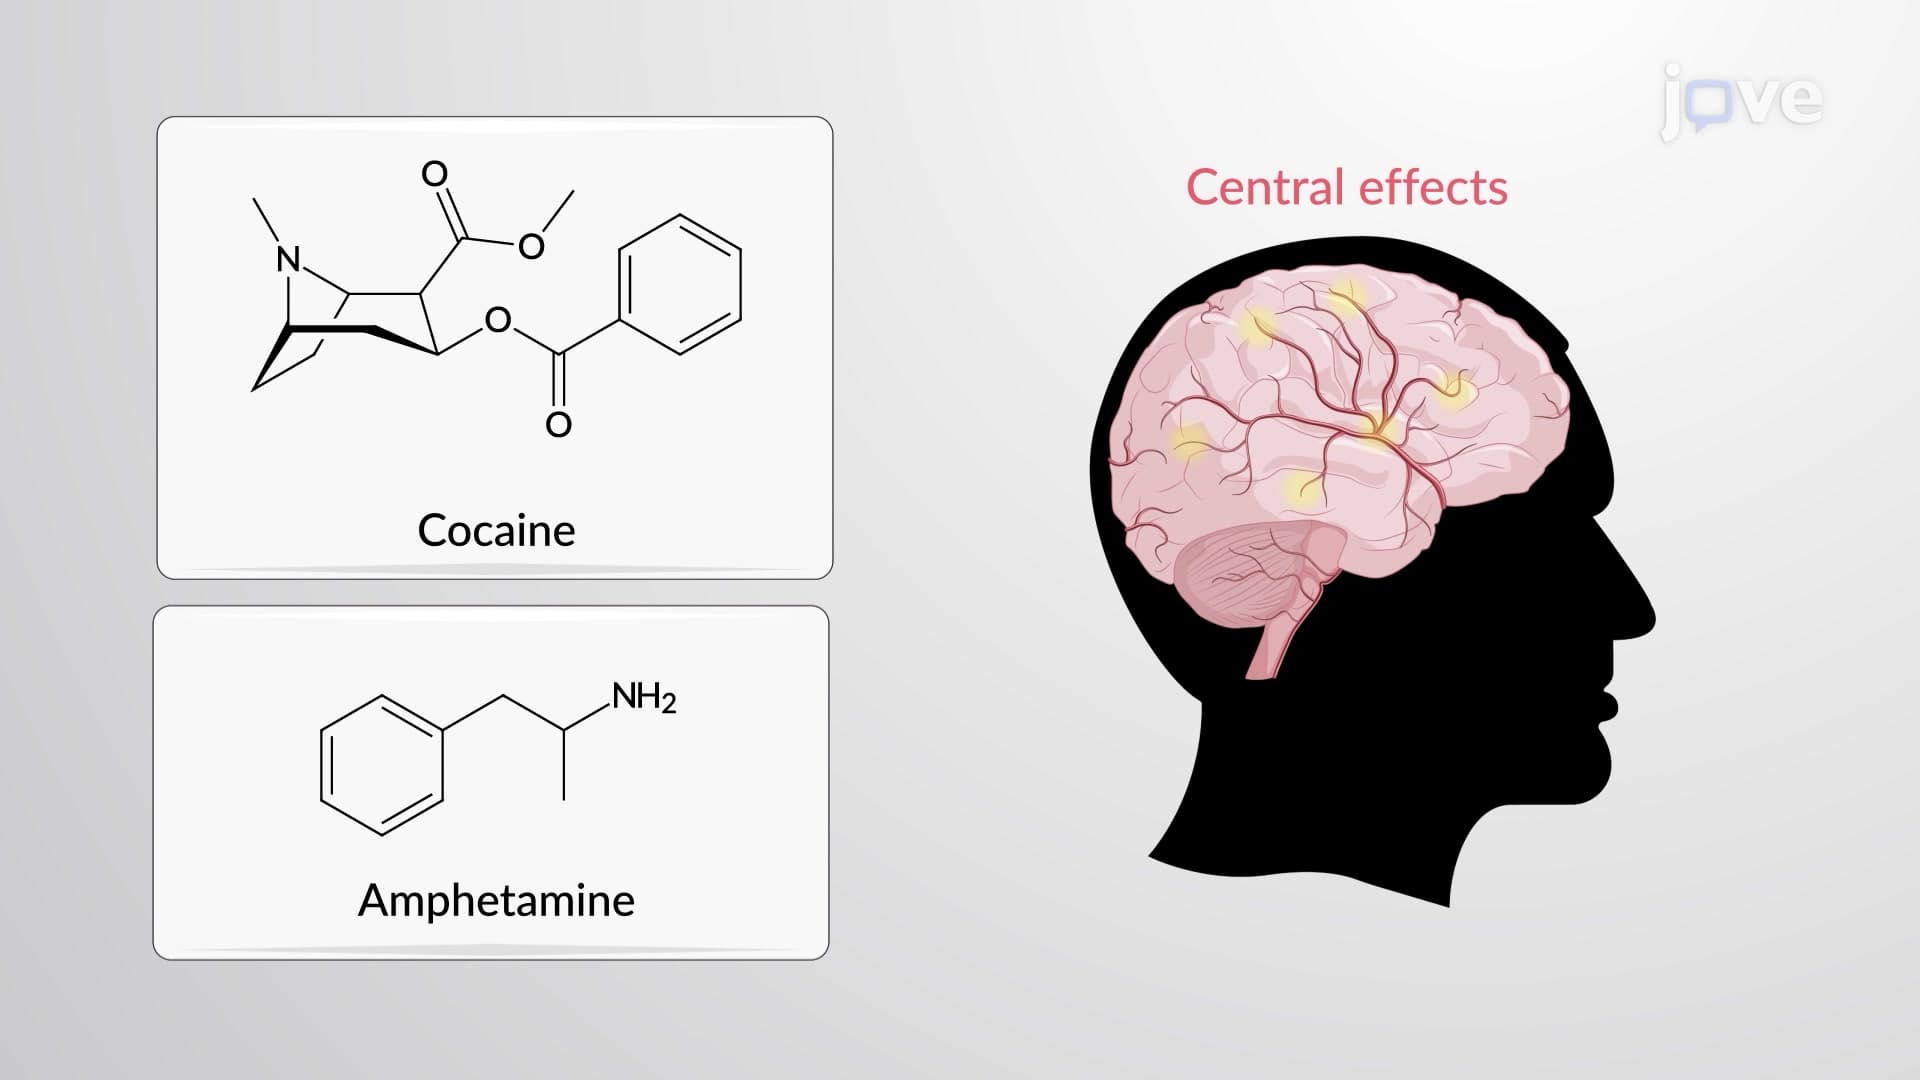
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral...