Related Experiment Video
Updated: Sep 20, 2025

Determination of the Transport Rate of Xenobiotics and Nanomaterials Across the Placenta using the ex vivo Human Placental Perfusion Model
Published on: June 18, 2013
Developing an In Vitro to In Vivo Extrapolation (IVIVE) Model to Predict Human Milk-to-Plasma Drug Concentration
Hong Yang1, Ivy Xue1, Qimei Gu1
1Office of Clinical Pharmacology, Center for Drug Evaluation and Research, U.S. Food and Drug Administration, 10903 New Hampshire Avenue, Silver Spring, Maryland 20993, United States.

Determining the amount of drug transferred into human milk is critical for benefit-risk analysis of taking medication while breastfeeding. In this study, we developed an in vitro and in vivo extrapolation (IVIVE) model to predict human milk/plasma (M/P) drug concentration ratios. Drug unionized fractions at pH 7.0 (Fni,7.0) and 7.4 (Fni,7.4), drug fractions unbound in human plasma (fup) and milk (fum), and in vitro cell permeability in both directions (efflux ratio, ER) were incorporated into the IVIVE model. A multiple regression Emax model was chosen to predict fum from fup and polar surface area (PSA). A total of 97 drugs with experimental ER from Caco-2 cells were used to test the IVIVE model. The M/P ratios predicted by the IVIVE model had a 1.93-fold geometric mean fold error (GMFE) and 72% of predictions were within two-fold error (Pw2FE), which were superior to the performance of previously reported five models. The IVIVE model showed a reasonable prediction accuracy for passive diffusion drugs (GMFE = 1.71-fold, Pw2FE = 82%, N = 50), BCRP substrates (BCRP: GMFE = 1.91-fold, Pw2FE = 60%, N = 5), and substrates of P-gp and BCRP (GMFE = 1.74-fold, Pw2FE = 75%, N = 8) and a lower prediction performance for P-gp substrates (GMFE = 2.51-fold, Pw2FE = 55%, N = 22). By fitting the observed M/P ratios of 39 P-gp substrates, an optimized ER (1.61) was generated to predict the M/P ratio of P-gp substrates using the developed IVIVE model. Compared with currently available in vitro models, the developed IVIVE model provides a more accurate prediction of the drug M/P ratio, especially for passive diffusion drugs. The model performance is expected to be further improved when more experimental fum and ER data are available.
Determining the amount of drug transferred into human milk is critical for benefit-risk analysis of taking medication while breastfeeding. In this study, we developed an in vitro and in vivo extrapolation (IVIVE) model to predict human milk/plasma (M/P) drug concentration ratios. Drug unionized fractions at pH 7.0 (Fni,7.0) and 7.4 (Fni,7.4), drug fractions unbound in human plasma (fup) and milk (fum), and in vitro cell permeability in both directions (efflux ratio, ER) were incorporated into the IVIVE model. A multiple regression Emax model was chosen to predict fum from fup and polar surface area (PSA). A total of 97 drugs with experimental ER from Caco-2 cells were used to test the IVIVE model. The M/P ratios predicted by the IVIVE model had a 1.93-fold geometric mean fold error (GMFE) and 72% of predictions were within two-fold error (Pw2FE), which were superior to the performance of previously reported five models. The IVIVE model showed a reasonable prediction accuracy for passive diffusion drugs (GMFE = 1.71-fold, Pw2FE = 82%, N = 50), BCRP substrates (BCRP: GMFE = 1.91-fold, Pw2FE = 60%, N = 5), and substrates of P-gp and BCRP (GMFE = 1.74-fold, Pw2FE = 75%, N = 8) and a lower prediction performance for P-gp substrates (GMFE = 2.51-fold, Pw2FE = 55%, N = 22). By fitting the observed M/P ratios of 39 P-gp substrates, an optimized ER (1.61) was generated to predict the M/P ratio of P-gp substrates using the developed IVIVE model. Compared with currently available in vitro models, the developed IVIVE model provides a more accurate prediction of the drug M/P ratio, especially for passive diffusion drugs. The model performance is expected to be further improved when more experimental fum and ER data are available.
Related Concept Videos
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:06
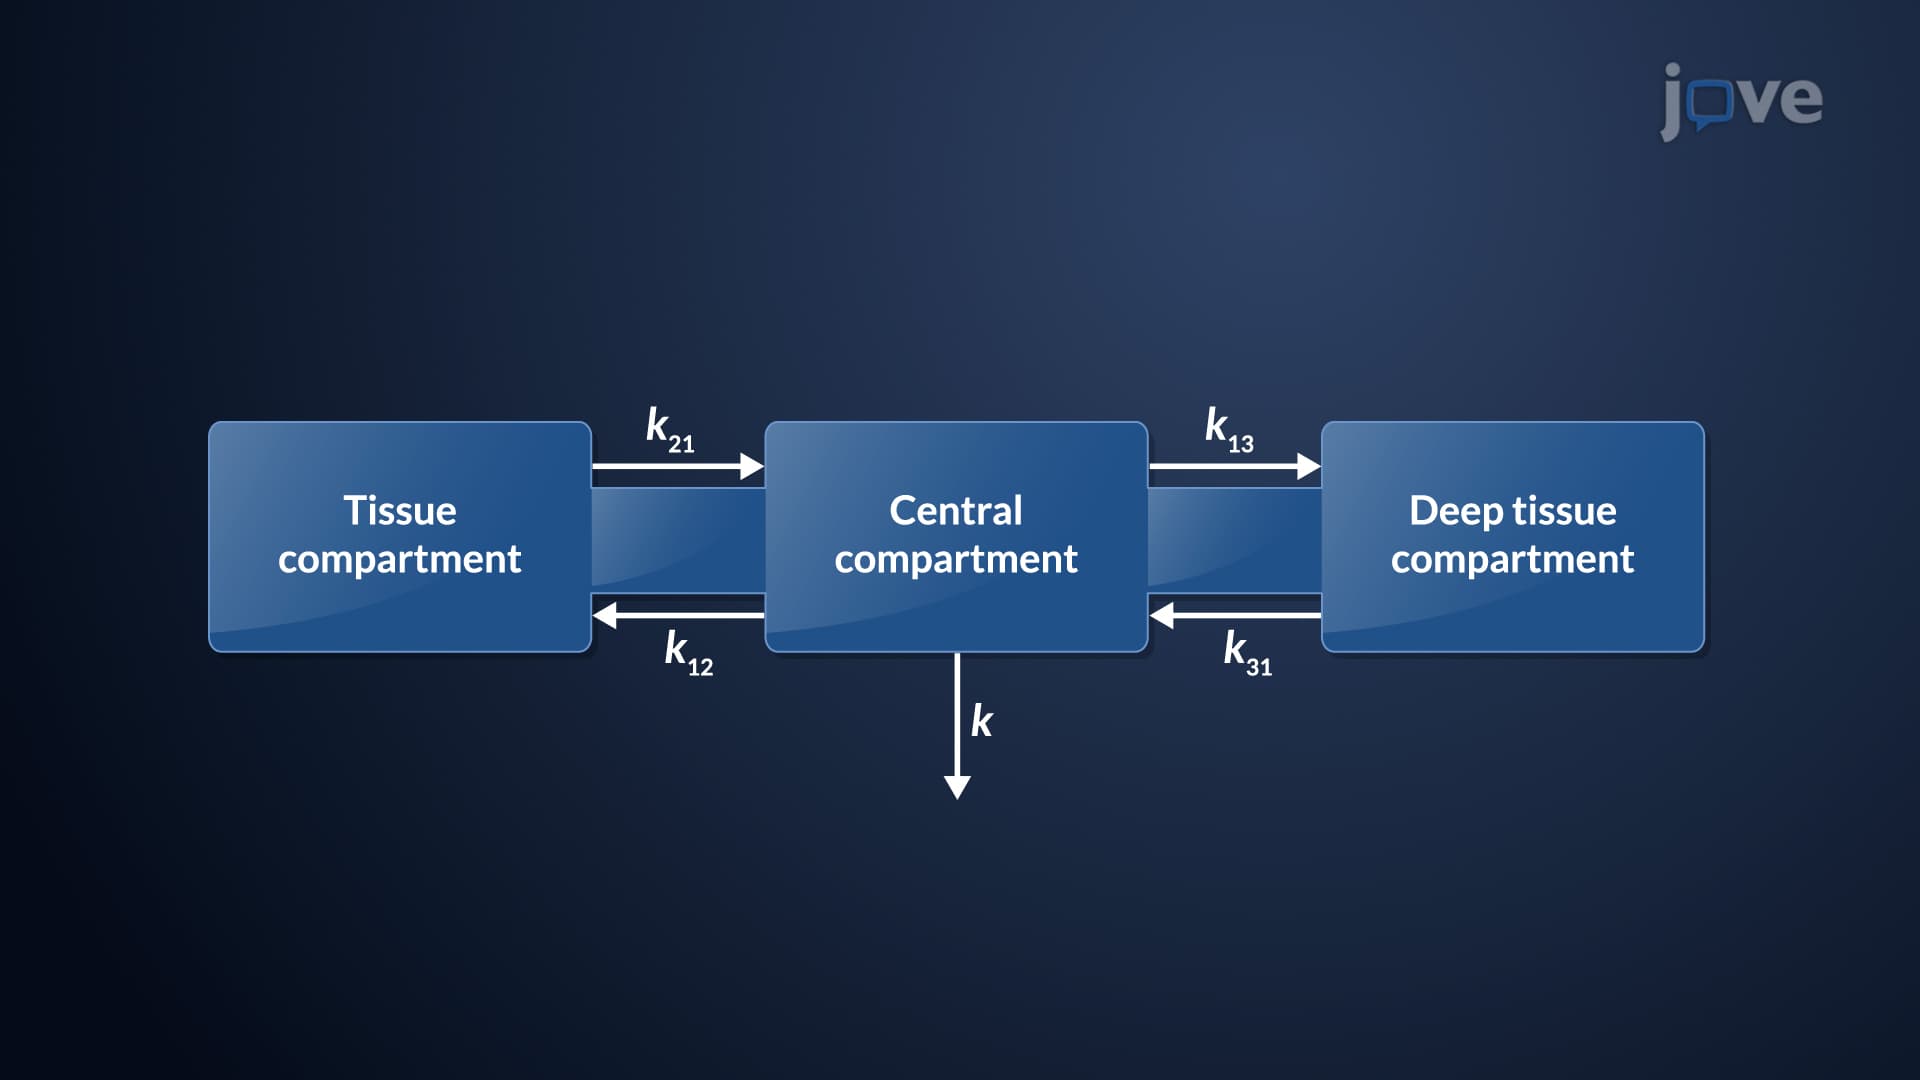
Three-Compartment Open Model
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:09
One-Compartment Open Model for IV Bolus Administration: Estimation of Elimination Rate Constant, Half-Life and Volume of Distribution
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal...