Related Experiment Video
Updated: Sep 7, 2025
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Explainable deep drug-target representations for binding affinity prediction
Nelson R C Monteiro1, Carlos J V Simões2, Henrique V Ávila3
1Univ Coimbra, Centre for Informatics and Systems of the University of Coimbra, Department of Informatics Engineering, Coimbra, Portugal. nelsonrcm@dei.uc.pt.

Background:
Several computational advances have been achieved in the drug discovery field, promoting the identification of novel drug-target interactions and new leads. However, most of these methodologies have been overlooking the importance of providing explanations to the decision-making process of deep learning architectures. In this research study, we explore the reliability of convolutional neural networks (CNNs) at identifying relevant regions for binding, specifically binding sites and motifs, and the significance of the deep representations extracted by providing explanations to the model's decisions based on the identification of the input regions that contributed the most to the prediction. We make use of an end-to-end deep learning architecture to predict binding affinity, where CNNs are exploited in their capacity to automatically identify and extract discriminating deep representations from 1D sequential and structural data.
Results:
The results demonstrate the effectiveness of the deep representations extracted from CNNs in the prediction of drug-target interactions. CNNs were found to identify and extract features from regions relevant for the interaction, where the weight associated with these spots was in the range of those with the highest positive influence given by the CNNs in the prediction. The end-to-end deep learning model achieved the highest performance both in the prediction of the binding affinity and on the ability to correctly distinguish the interaction strength rank order when compared to baseline approaches.
Conclusions:
This research study validates the potential applicability of an end-to-end deep learning architecture in the context of drug discovery beyond the confined space of proteins and ligands with determined 3D structure. Furthermore, it shows the reliability of the deep representations extracted from the CNNs by providing explainability to the decision-making process.
More Related Videos
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:25
Drug-Receptor Bonds
In...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:29
Drug-Receptor Interactions
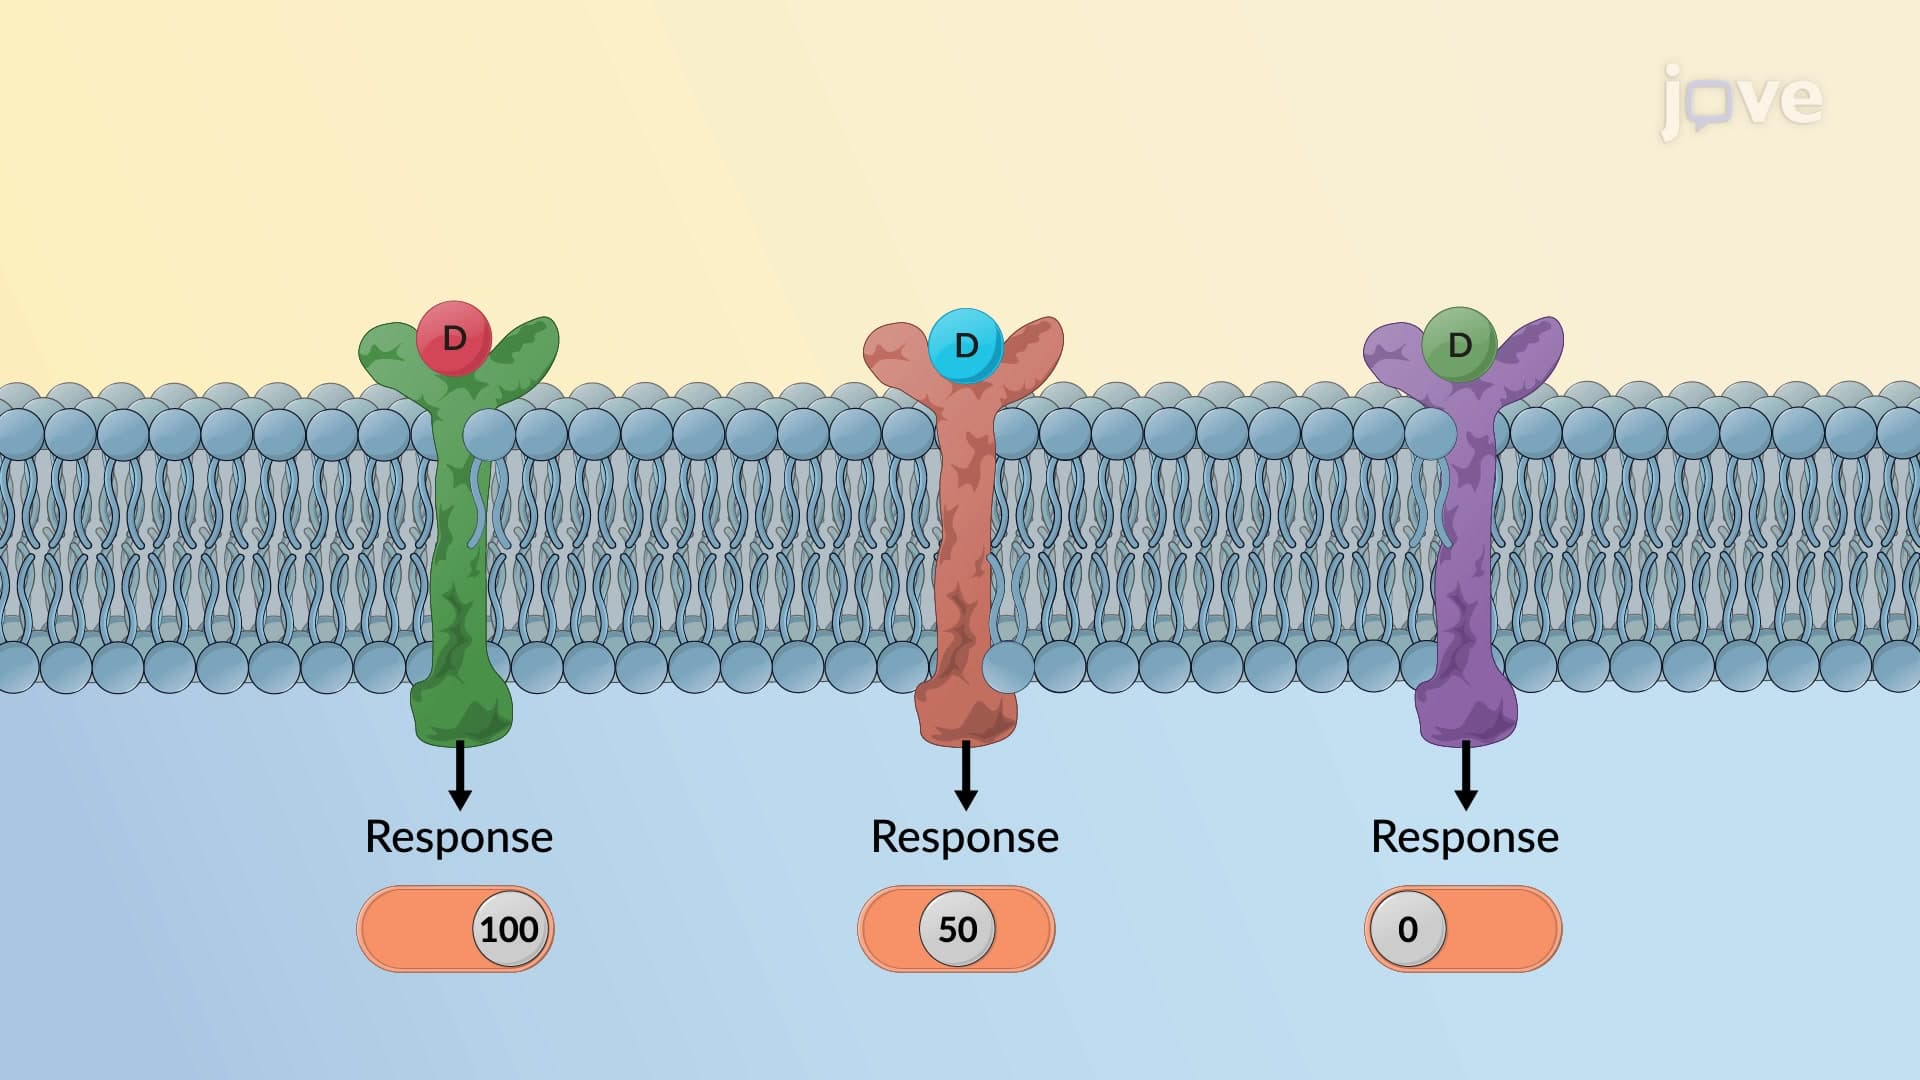
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...