
Superior Multielectron-Transferring Energy Storage by π-d Conjugated Frameworks
Dong Xia1, Ken Sakaushi1, Andrey Lyalin1,2
1Center for Green Research on Energy and Environmental Materials, National Institute for Materials Science 1-1 Namiki, Tsukuba, Ibaraki, 305-0044, Japan.

Abstract:
Reversible multielectron-transfer materials are of considerable interest because of the potential impact to advance present electrochemical energy storage technology by boosting energy density. To date, a few oxide-based materials can reach an electron-transfer number per metal-cation (eM ) larger than 2 upon a (de)intercalation mechanism. However, these materials suffer from degradation due to irreversible rearrangements of the cation-oxygen bonds, and are based on precious metals, for example, Ir and Ru. Hence, a design of the non-oxide-based reversible multielectron-transfer materials with abundant elements can provide a promising alternative. Herein, it is demonstrated that the bis(diimino)copper framework can show eM = 3.5 with cation/anion co-redox mechanism together with a dual-ion mechanism. In this study, the role of the cation-anion interactions is unveiled by using an experiment/theory collaboration applied to a series of the model non-oxide abundant electrode systems based on different metal-nitrogen bonds. These models provide designer multielectron-transfer due to the tunable π-d conjugated electronic structures. It is found that the Cu-nitrogen bonds show a unique reversible rearrangement upon Li-intercalation, and this process responds to acquire a significant reversible multielectron-transfer. This work provides new insights into the affordable multielectron-transfer electrodes and uncovers an alternative strategy to advance the electrochemical energy storage reactions.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
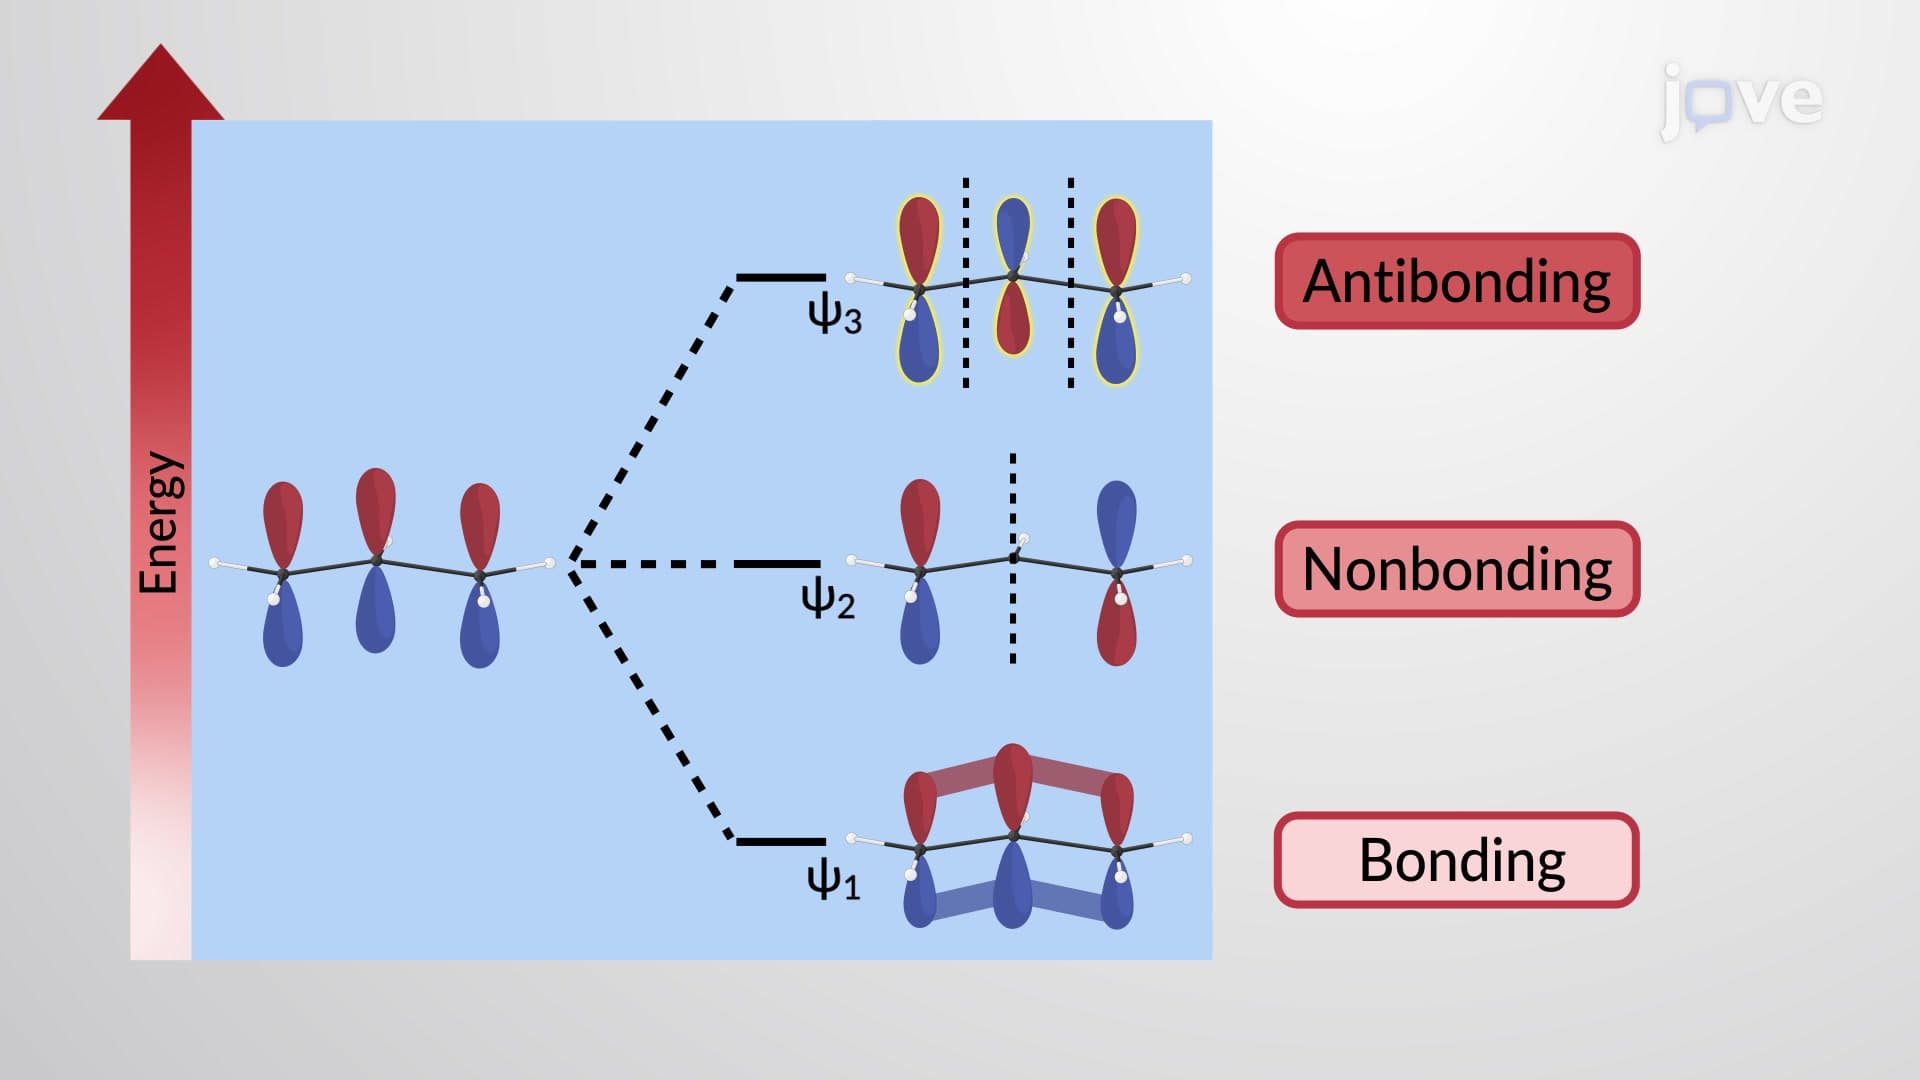
π Molecular Orbitals of the Allyl Cation and Anion
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.