Intense Molar Circular Dichroism in Fully Conjugated All-Carbon Macrocyclic 1,3-Butadiyne Linked pseudo-meta [2.2]Paracyclophanes
Related Experiment Video
Updated: Sep 6, 2025
Synthesis of Cyclic Polymers and Characterization of Their Diffusive Motion in the Melt State at the Single Molecule Level
Published on: September 26, 2016
Eric Sidler1, Patrick Zwick1, Charlotte Kress1
1Department of Chemistry, University of Basel, St. Johanns-Ring 19, 4056, Basel, Switzerland.

Abstract:
The synthetic access to macrocyclic molecular topologies with interesting photophysical properties has greatly improved thanks to the successful implementation of organic and inorganic corner units. Based on recent reports, we realized that pseudo-meta [2.2]paracyclophanes (PCPs) might serve as optimal corner units for constructing 3D functional materials, owing to their efficient electronic communication, angled substituents and planar chirality. Herein, we report the synthesis, characterization and optical properties of four novel all-carbon enantiopure macrocycles bearing three to six pseudo-meta PCPs linked by 1,3-butadiyne units. The macrocycles were obtained by a single step from enantiopure, literature-known dialkyne pseudo-meta PCP and were unambiguously identified and characterized by state of the art spectroscopic methods and in part even by x-ray crystallography. By comparing the optical properties to relevant reference compounds, it is shown that the pseudo-meta PCP subunit effectively elongates the conjugated system throughout the macrocyclic backbone, such that already the smallest macrocycle consisting of only three subunits reaches a polymer-like conjugation length. Additionally, it is shown that the chiral pseudo-meta PCPs induce a remarkable chiroptical response in the respective macrocycles, reaching unprecedented high molar circular dichroism values for all-carbon macrocycles of up to 1307 L mol-1 cm-1 .
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:33
Stereoisomerism of Cyclic Compounds
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:29
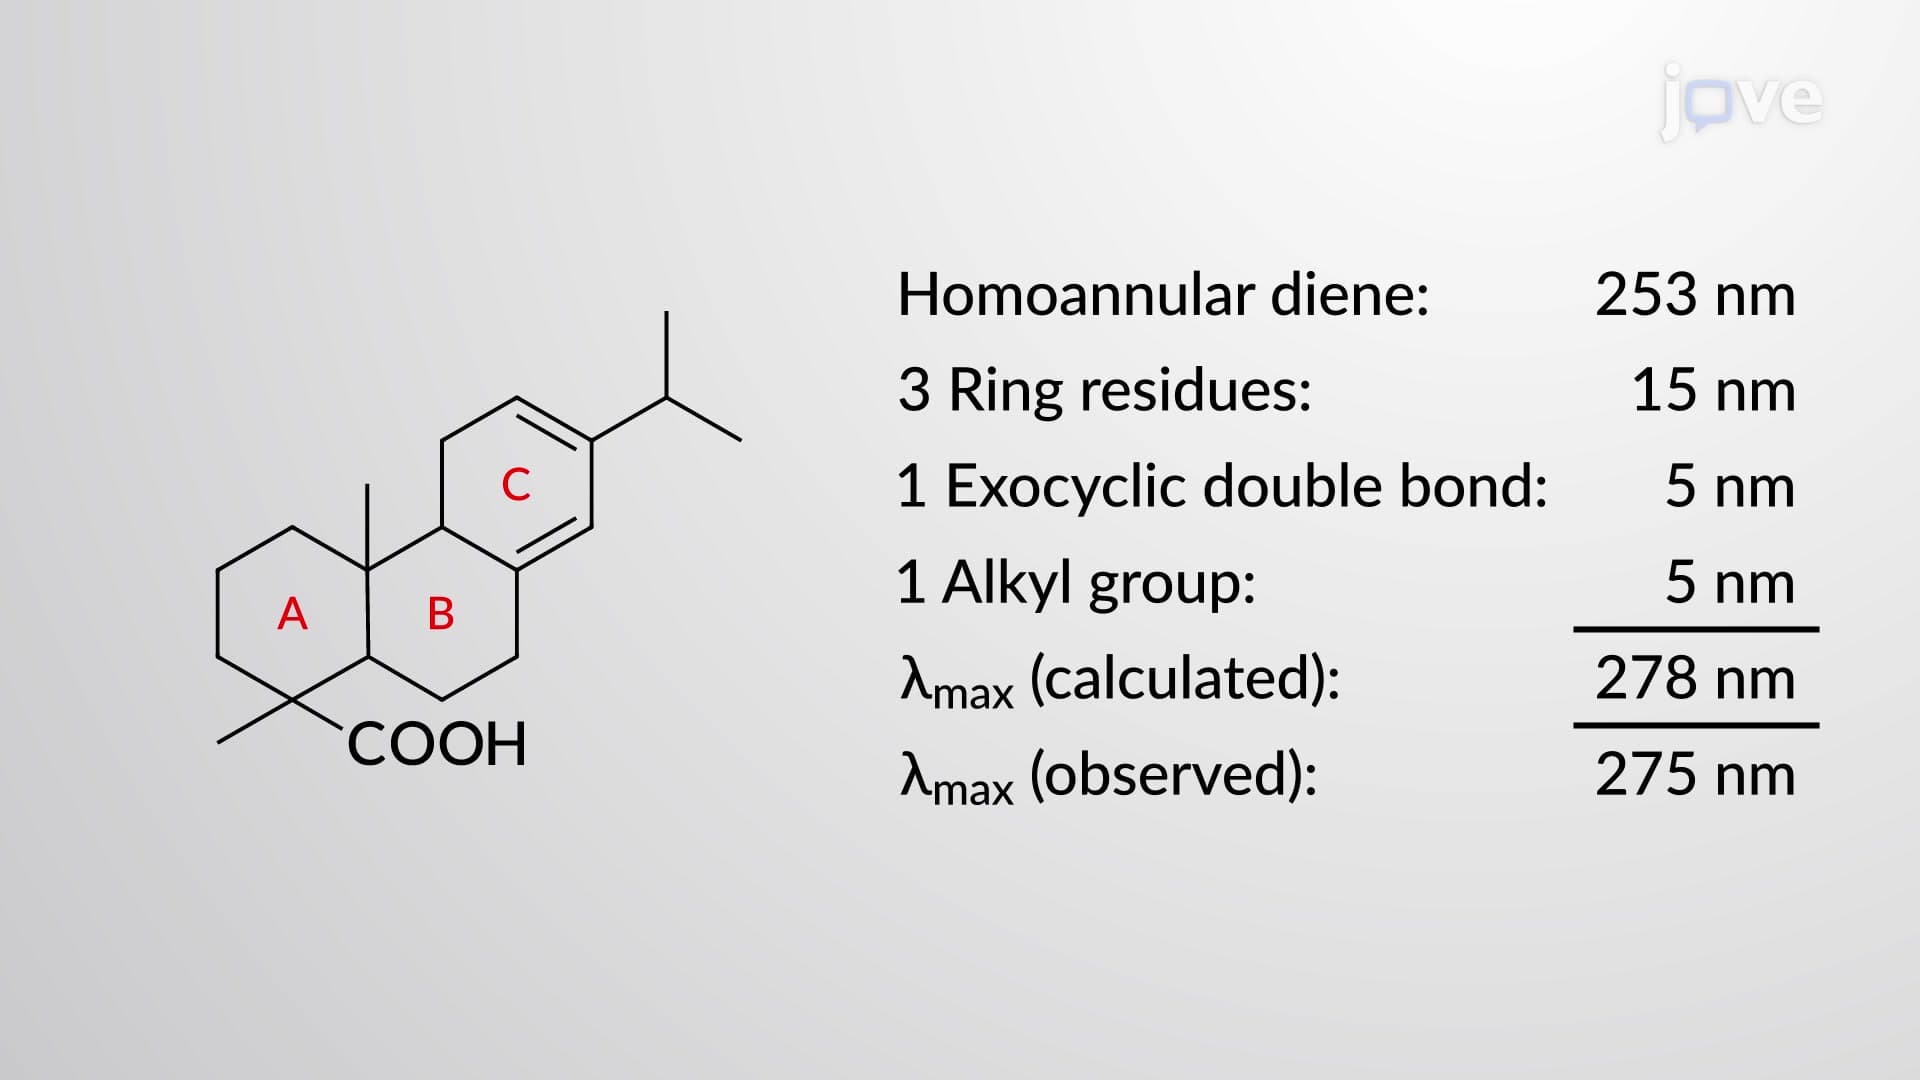
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds