Related Experiment Video
Updated: Sep 5, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
How Good Is the Density-Corrected SCAN Functional for Neutral and Ionic Aqueous Systems, and What Is So Right about
Saswata Dasgupta1, Chandra Shahi2, Pradeep Bhetwal2
1Department of Chemistry and Biochemistry, University of California San Diego, La Jolla, California 92093, United States.

Abstract:
Density functional theory (DFT) is the most widely used electronic structure method, due to its simplicity and cost effectiveness. The accuracy of a DFT calculation depends not only on the choice of the density functional approximation (DFA) adopted but also on the electron density produced by the DFA. SCAN is a modern functional that satisfies all known constraints for meta-GGA functionals. The density-driven errors, defined as energy errors arising from errors of the self-consistent DFA electron density, can hinder SCAN from achieving chemical accuracy in some systems, including water. Density-corrected DFT (DC-DFT) can alleviate this shortcoming by adopting a more accurate electron density which, in most applications, is the electron density obtained at the Hartree-Fock level of theory due to its relatively low computational cost. In this work, we present extensive calculations aimed at determining the accuracy of the DC-SCAN functional for various aqueous systems. DC-SCAN (SCAN@HF) shows remarkable consistency in reproducing reference data obtained at the coupled cluster level of theory, with minimal loss of accuracy. Density-driven errors in the description of ionic aqueous clusters are thoroughly investigated. By comparison with the orbital-optimized CCD density in the water dimer, we find that the self-consistent SCAN density transfers a spurious fraction of an electron across the hydrogen bond to the hydrogen atom (H*, covalently bound to the donor oxygen atom) from the acceptor (OA) and donor (OD) oxygen atoms, while HF makes a much smaller spurious transfer in the opposite direction, consistent with DC-SCAN (SCAN@HF) reduction of SCAN overbinding due to delocalization error. While LDA seems to be the conventional extreme of density delocalization error, and HF the conventional extreme of (usually much smaller) density localization error, these two densities do not quite yield the conventional range of density-driven error in energy differences. Finally, comparisons of the DC-SCAN results with those obtained with the Fermi-Löwdin orbital self-interaction correction (FLOSIC) method show that DC-SCAN represents a more accurate approach to reducing density-driven errors in SCAN calculations of ionic aqueous clusters. While the HF density is superior to that of SCAN for noncompact water clusters, the opposite is true for the compact water molecule with exactly 10 electrons.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:08
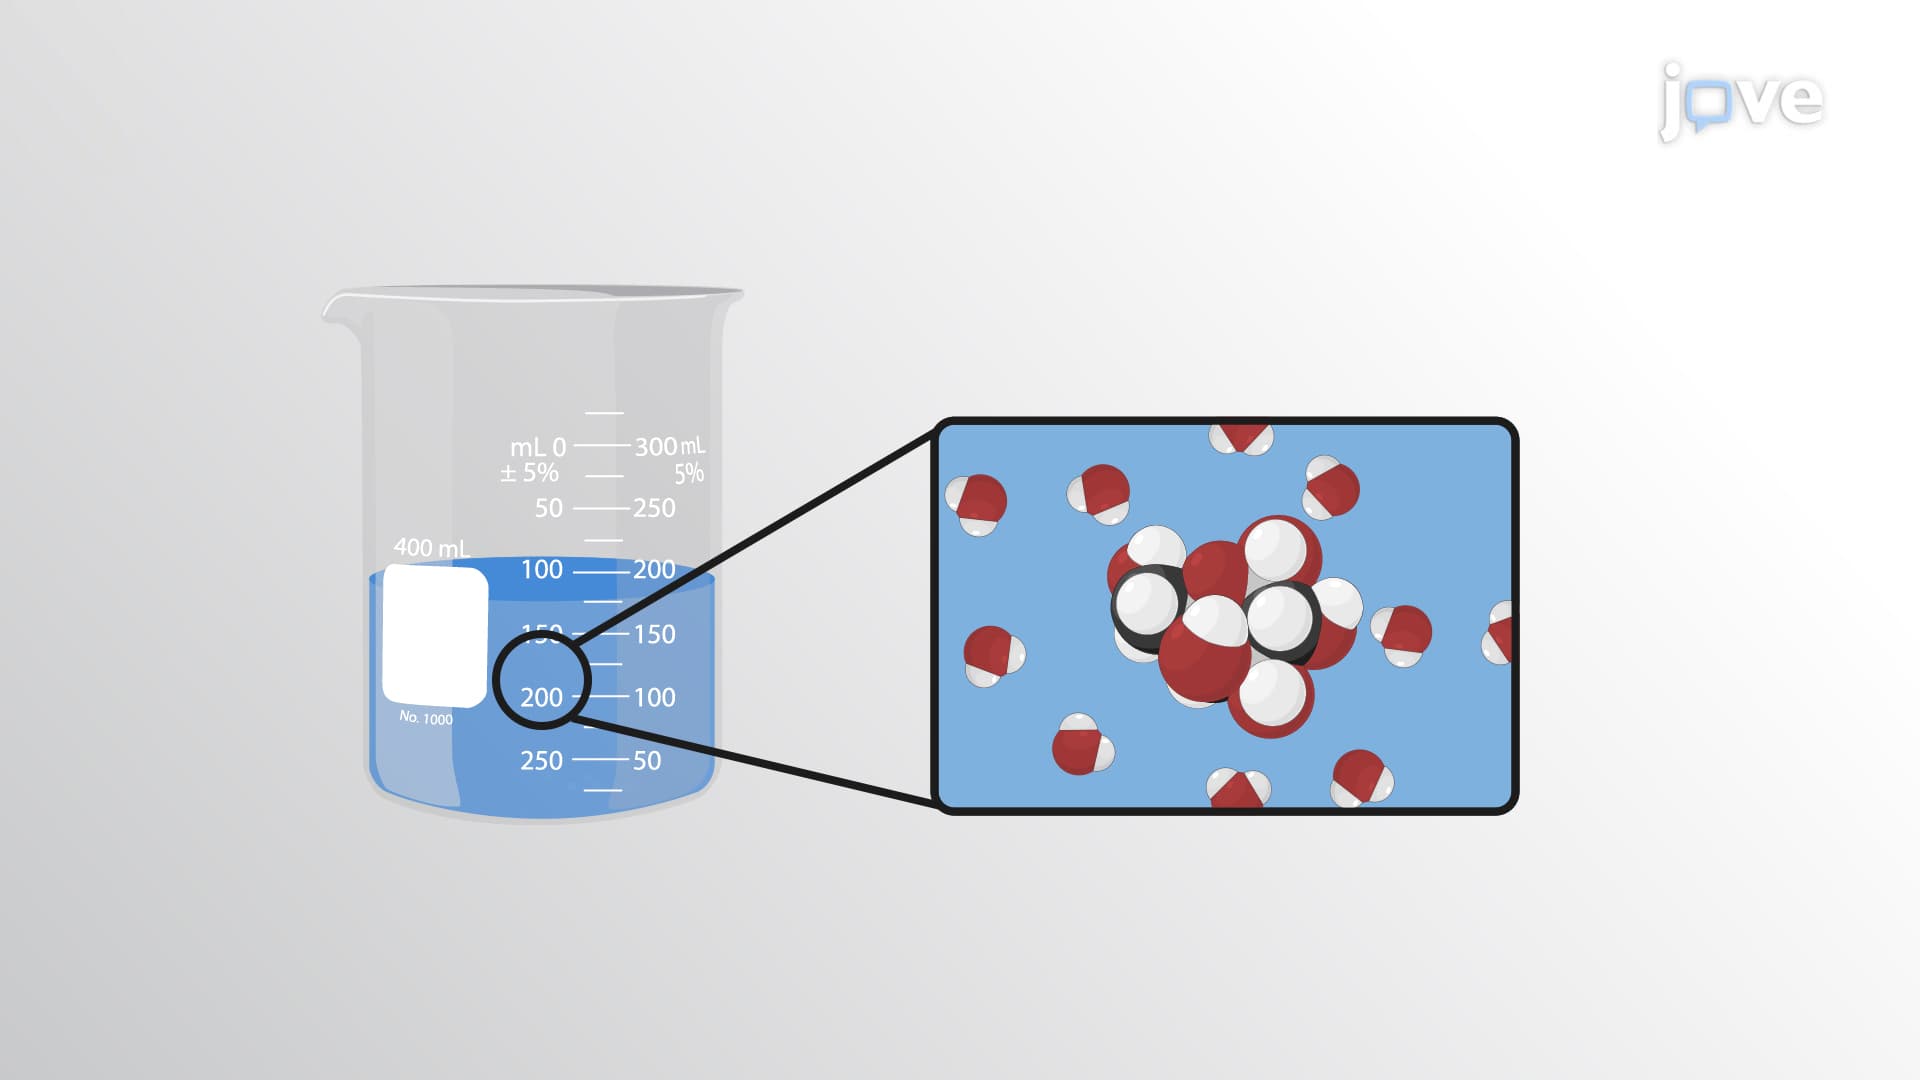
Electrolytes: van't Hoff Factor
The colligative properties of a solution depend only on the number, not on the identity, of solute species dissolved. The concentration terms in the equations for various colligative properties (freezing point depression, boiling point elevation, osmotic pressure) pertain to all solute species present in the solution. Nonelectrolytes dissolve physically without dissociation or any other accompanying process. Each molecule that dissolves yields one...
 00:56
Density
 02:30
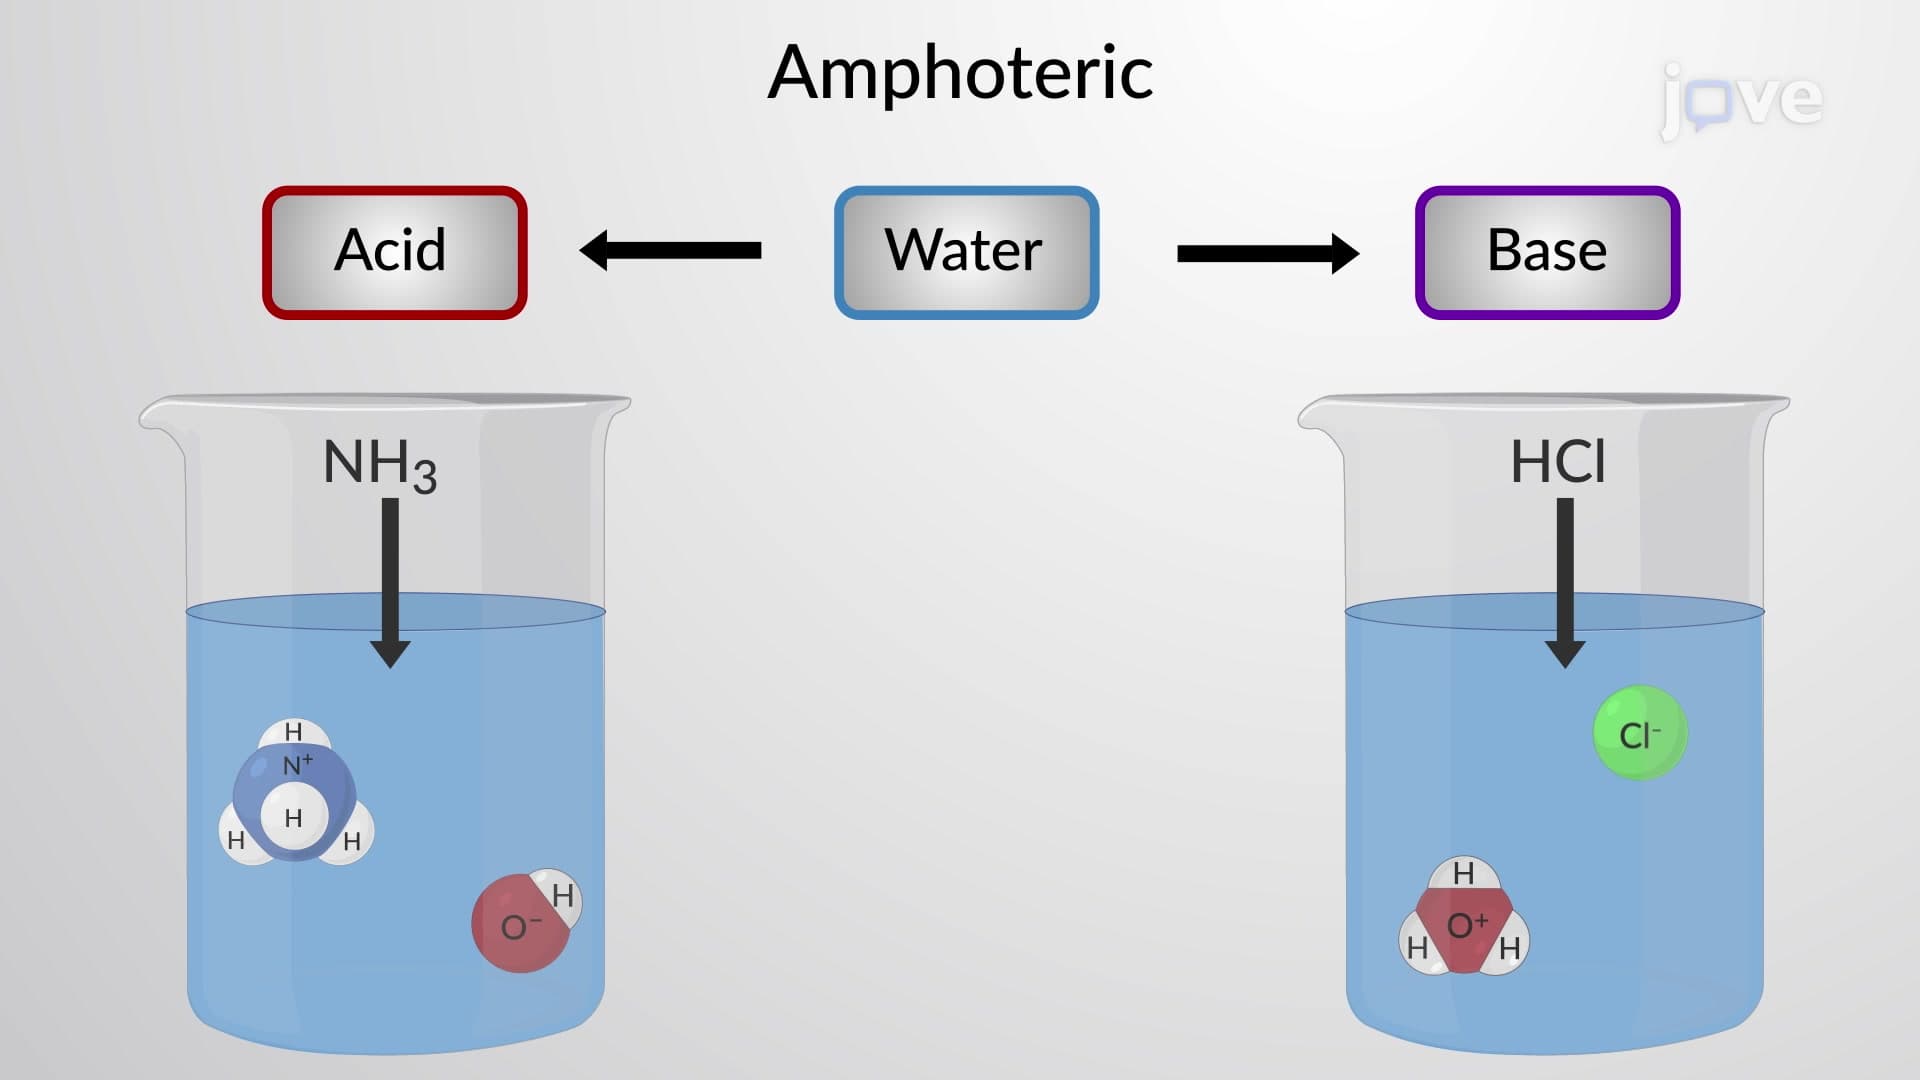
Water: A Bronsted-Lowry Acid and Base
 00:53
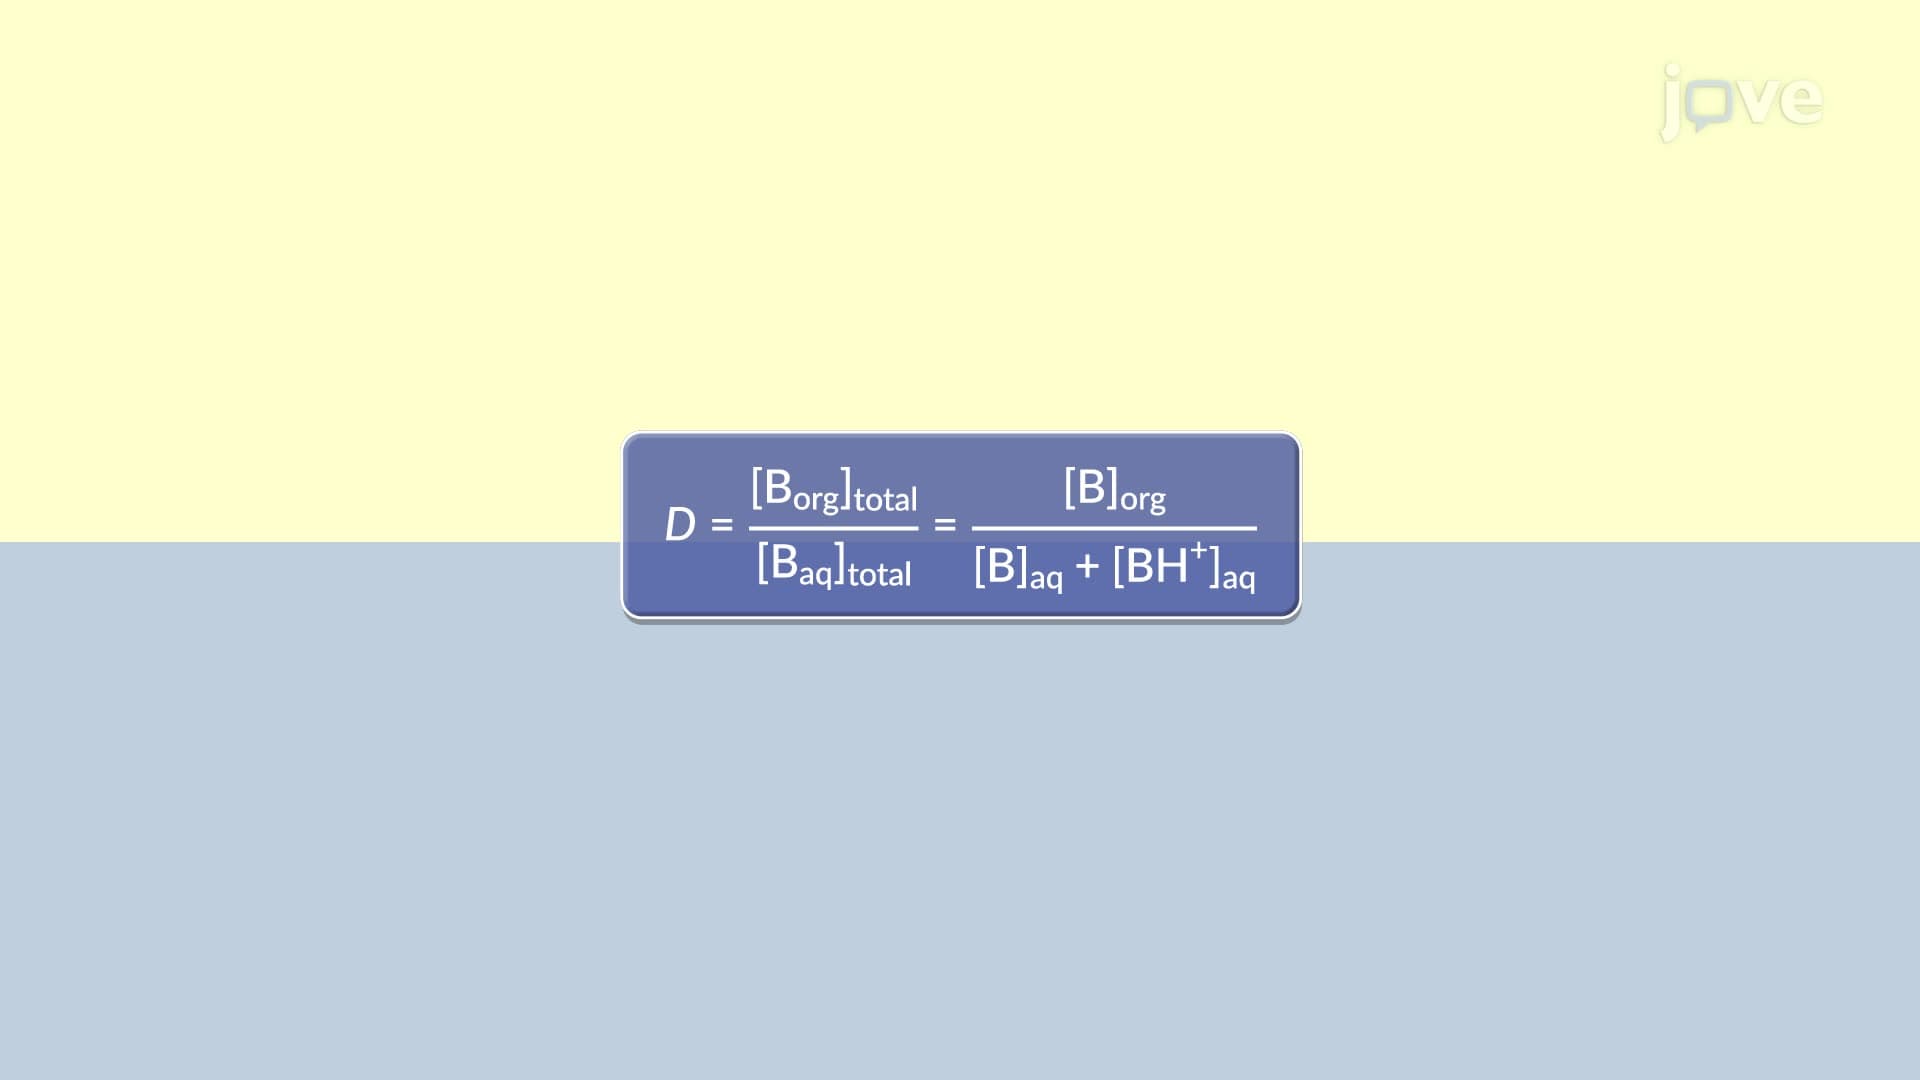
Extraction: Effects of pH
 01:17
Factors Affecting Activity Coefficient
The activity coefficient value for an ion is close to one when the solution has almost zero ionic strength, i.e., when the solution shows close to ideal behavior. As the ionic strength of the solution increases from 0 to 0.1 mol/L, a...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...